Related Experiment Video
Updated: Sep 2, 2025
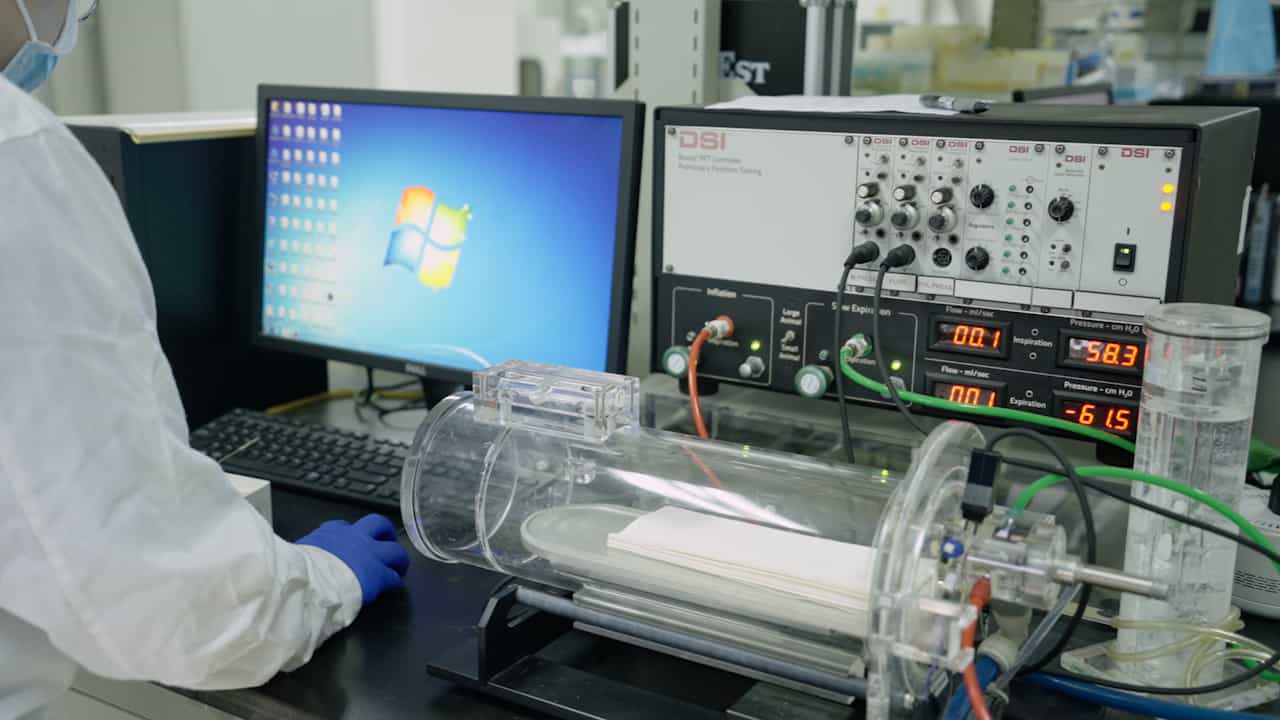

Establishment and Validation of a Rat Model of Pulmonary Arterial Hypertension Associated with Pulmonary Fibrosis
Published on: May 23, 2025
Aberrant Expression of ACO1 in Vasculatures Parallels Progression of Idiopathic Pulmonary Fibrosis
Jutaro Fukumoto1, Muling Lin1, Mudassir Meraj Banday2
1Division of Allergy and Immunology, Department of Internal Medicine, University of South Florida, Tampa, FL, United States.

Abstract:
Rationale: Idiopathic pulmonary fibrosis (IPF) is characterized by mitochondrial dysfunction. However, details about the non-mitochondrial enzymes that sustain the proliferative nature of IPF are unclear. Aconitases are a family of enzymes that sustain metabolism inside and outside mitochondria. It is hypothesized that aconitase 1 (ACO1) plays an important role in the pathogenesis of IPF given that ACO1 represents an important metabolic hub in the cytoplasm. Objectives: To determine if ACO1 expression in IPF lungs shows specific patterns that may be important in the pathogenesis of IPF. To determine the similarities and differences in ACO1 expression in IPF, bleomycin-treated, and aging lungs. Methods: ACO1 expression in IPF lungs were characterized and compared to non-IPF controls by western blotting, immunostaining, and enzymatic activity assay. ACO1-expressing cell types were identified by multicolor immunostaining. Using similar methods, the expression profiles of ACO1 in IPF lungs versus bleomycin-treated and aged mice were investigated. Measurements and main results: Lower lobes of IPF lungs, unlike non-IPF controls, exhibit significantly high levels of ACO1. Most of the signals colocalize with von Willebrand factor (vWF), a lineage marker for vascular endothelial cells. Bleomycin-treated lungs also show high ACO1 expressions. However, most of the signals colocalize with E-cadherin and/or prosurfactant protein C, representative epithelial cell markers, in remodeled areas. Conclusions: A characteristic ACO1 expression profile observed in IPF vasculatures may be a promising diagnostic target. It also may give clues as to how de novo angiogenesis contributes to the irreversible nature of IPF.
Related Concept Videos
 01:30
Pulmonary Hypertension: Classification and Pathogenesis
There are various classifications for PH, each relating to different underlying causes and also...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:20
COPD: Pathogenesis and Clinical Features
The primary cause for the onset of COPD is cigarette smoking and exposure to air pollution. These hazardous factors initiate a chain reaction within the lungs, resulting in chronic inflammation, damage to the airways, and a...
 01:22
Chronic Obstructive Pulmonary Disease
Smoking is a primary risk factor for COPD, with over 80% of patients having a history of it. Patients typically experience progressive dyspnea or labored breathing, frequent coughing, and recurrent pulmonary infections. Many eventually succumb to respiratory failure, characterized by...
 01:20
Chronic Obstructive Pulmonary Disease-I: Introduction