Related Experiment Video
Updated: Sep 2, 2025

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Exhaustive Mapping of the Conformational Space of Natural Dipeptides by the DFT-D3//COSMO-RS Method
Tadeáš Kalvoda1, Martin Culka1, Lubomír Rulíšek1
1Institute of Organic Chemistry and Biochemistry of the Czech Academy of Sciences, Flemingovo náměstí 2, 166 10 Praha, Czech Republic.

Abstract:
We extensively mapped energy landscapes and conformations of 22 (including three His protonation states) proteinogenic α-amino acids in trans configuration and the corresponding 484 (222) dipeptides. To mimic the environment in a protein chain, the N- and C-termini of the studied systems were capped with acetyl and N-methylamide groups, respectively. We systematically varied the main chain dihedral angles (ϕ, ψ) by 40° steps and all side chain angles by 90° or 120° steps. We optimized the molecular geometries with the GFN2-xTB semiempirical (SQM) method and performed single point density functional theory calculations at the BP86-D3/DGauss-DZVP//COSMO-RS level in water, 1-octanol, N,N-dimethylformamide, and n-hexane. For each restrained (nonequilibrium) structure, we also calculated energy gradients (in water) and natural atomic charges. The exhaustive and unprecedented QM-based sampling enabled us to construct Ramachandran plots of quantum mechanical (QM(BP86-D3)//COSMO-RS) energies calculated on SQM structures, for all 506 (484 dipeptides and 22 amino acids) studied systems. We showed how the character of an amino acid side chain influences the conformational space of single amino acids and dipeptides. With clustering techniques, we were able to identify unique minima of amino acids and dipeptides (i.e., minima on the GFN2-xTB potential energy surfaces) and analyze the distribution of their BP86-D3//COSMO-RS conformational energies in all four solvents. We also derived an empirical formula for the number of unique minima based on the overall number of rotatable bonds within each peptide. The final peptide conformer data set (PeptideCs) comprises over 400 million structures, all of them annotated with QM(BP86-D3)//COSMO-RS energies. Thanks to its completeness and unbiased nature, the PeptideCs can serve, inter alia, as a data set for the validation of new methods for predicting the energy landscapes of protein structures. This data set may also prove to be useful in the development and reparameterization of biomolecular force fields. The data set is deposited at Figshare (10.25452/figshare.plus.19607172) and can be accessed using a simple web interface at http://peptidecs.uochb.cas.cz.
More Related Videos

Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this...
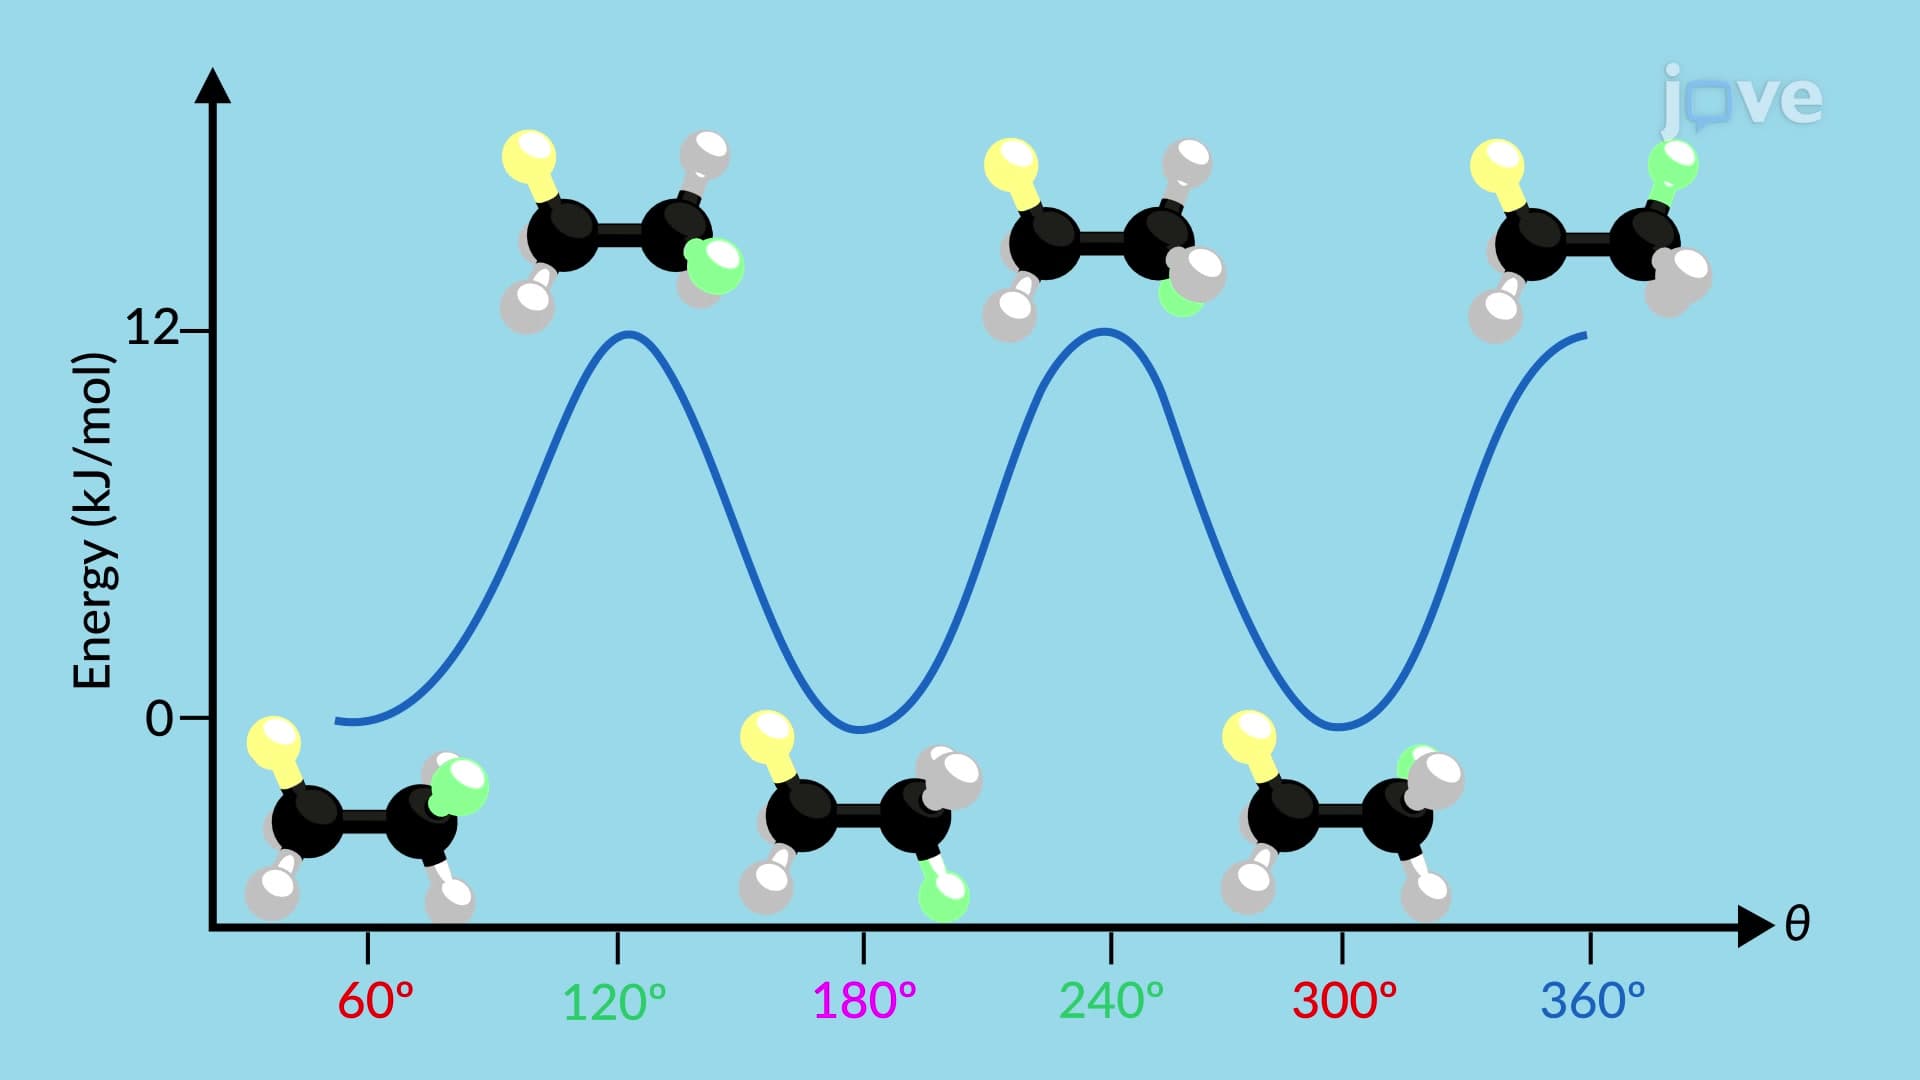 02:18
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR