Related Experiment Video
Updated: Sep 2, 2025
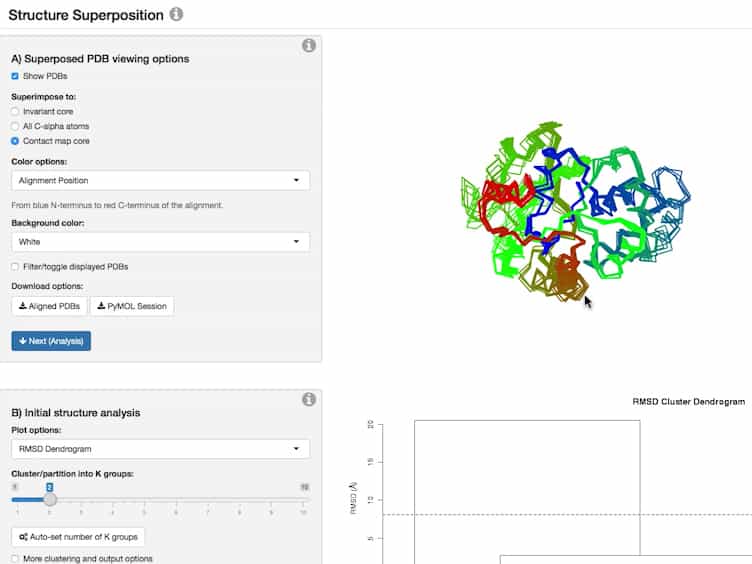

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Investigating the Bioactive Conformation of Angiotensin II Using Markov State Modeling Revisited with Web-Scale
Emmanouil Christoforou1,2, Hari Leontiadou2, Frank Noé3
1ITMB, Department of Informatics & Telecommunications, National and Kapodistrian University of Athens, Athens 15772, Greece.

Abstract:
Molecular dynamics simulation is a powerful technique for studying the structure and dynamics of biomolecules in atomic-level detail by sampling their various conformations in real time. Because of the long timescales that need to be sampled to study biomolecular processes and the big and complex nature of the corresponding data, relevant analyses of important biophysical phenomena are challenging. Clustering and Markov state models (MSMs) are efficient computational techniques that can be used to extract dominant conformational states and to connect those with kinetic information. In this work, we perform Molecular Dynamics simulations to investigate the free energy landscape of Angiotensin II (AngII) in order to unravel its bioactive conformations using different clustering techniques and Markov state modeling. AngII is an octapeptide hormone, which binds to the AT1 transmembrane receptor, and plays a vital role in the regulation of blood pressure, conservation of total blood volume, and salt homeostasis. To mimic the water-membrane interface as AngII approaches the AT1 receptor and to compare our findings with available experimental results, the simulations were performed in water as well as in water-ethanol mixtures. Our results show that in the water-ethanol environment, AngII adopts more compact U-shaped (folded) conformations than in water, which resembles its structure when bound to the AT1 receptor. For clustering of the conformations, we validate the efficiency of an inverted-quantized k-means algorithm, as a fast approximate clustering technique for web-scale data (millions of points into thousands or millions of clusters) compared to k-means, on data from trajectories of molecular dynamics simulations with reasonable trade-offs between time and accuracy. Finally, we extract MSMs using various clustering techniques for the generation of microstates and macrostates and for the selection of the macrostate representatives.
Related Concept Videos
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:06
Model Approaches for Pharmacokinetic Data: Distributed Parameter Models
The distributed parameter models are specifically designed to account for variations and differences in some drug classes. This model is particularly useful for assessing regional concentrations of anticancer or...
 01:23
Mechanistic Models: Compartment Models in Individual and Population Analysis
 01:14
Multicompartment Models: Overview
These models offer a more comprehensive representation of drug behavior in the body than one-compartment models. They accommodate the complexity of drug distribution,...