Two infants with mild, atypical clinical features of Kagami-Ogata syndrome caused by epimutation
Related Experiment Video
Updated: Sep 1, 2025
Array Comparative Genomic Hybridization Array CGH for Detection of Genomic Copy Number Variants
Published on: February 21, 2015
Hiroyuki Higashiyama1, Yoshiteru Ohsone2, Rieko Takatani2
1Department of Pediatrics, Toyama Prefectural Central Hospital, Toyama, Japan.

Insights
Kagami-Ogata syndrome (KOS) epimutation presents with milder symptoms and near-normal development in two infants. Aberrant methylation in specific regions was observed, suggesting tissue-specific mosaicism contributes to KOS phenotypes.
Area of Science:
- Genetics and Epigenetics
- Developmental Biology
- Pediatric Endocrinology
Background:
- Kagami-Ogata syndrome (KOS) is a rare imprinting disorder linked to chromosome 14q32.2 gene expression abnormalities.
- Epimutation, specifically aberrant hypermethylation of MEG3/DLK1:IG-DMR and MEG3:TSS-DMR, is a known cause of KOS.
- Typical KOS features include polyhydramnios, bell-shaped thorax, respiratory distress, abdominal defects, and developmental delay.
Observation:
- Two infants with KOS due to epimutation exhibited milder clinical courses and near-normal motor/intellectual development.
- Methylation analysis revealed increased MEG3/DLK1:IG-DMR and MEG3:TSS-DMR methylation in leukocytes, but lower than in upd(14)pat.
- Tissue-specific methylation analysis showed varying levels, with buccal cells closer to normal for MEG3:TSS-DMR.
Findings:
- Confirmed KOS in two infants caused by epimutation, distinct from paternal uniparental disomy 14 (upd(14)pat) and microdeletions.
- Demonstrated tissue-specific mosaicism in methylation patterns at the 14q32.2 imprinted region.
- Identified atypical clinical presentations and milder developmental outcomes associated with this specific epimutation.
Implications:
- Suggests that tissue-specific mosaicism in imprinting disorders like KOS can lead to atypical phenotypes.
- Highlights the importance of comprehensive methylation analysis across different tissues for accurate diagnosis.
- Expands the known phenotypic spectrum of Kagami-Ogata syndrome, emphasizing the role of epimutation.
Abstract:
Kagami-Ogata syndrome (KOS) is an imprinting disorder characterized by polyhydramnios, bell-shaped thorax with coat-hanger appearance (curved ribs), respiratory distress, abdominal wall defects, and distinct facial features, together with intellectual developmental delay with special needs. Abnormal expression of the imprinted genes on chromosome 14q32.2 causes KOS. Epimutation with aberrant hypermethylation of the MEG3/DLK1: intergenic differentially methylated region (MEG3/DLK1:IG-DMR) and the MEG3:TSS-DMR is one of the etiologies of KOS. We report two infants with KOS caused by epimutation presenting with some characteristic clinical features, mild clinical course, and almost normal motor and intellectual development. Methylation analysis for ten DMRs related to major imprinting disorders using pyrosequencing with genomic DNA (gDNA) extracted from leukocytes showed abnormally increased methylation levels of the MEG3/DLK1:IG-DMR and MEG3:TSS-DMR in both patients, but lower than those in patients with paternal uniparental disomy chromosome 14 (upd(14)pat). The methylation levels in the DMRs other than both DMRs were within normal range. We also conducted methylation analysis for the MEG3/DLK1:IG-DMR and MEG3:TSS-DMR with gDNA extracted from nails and buccal cells of both patients. Methylation levels in the MEG3:TSS-DMR, particularly in buccal cells, were closer to normal range compared to those in leukocytes. Microsatellite analysis for chromosome 14 and array comparative hybridization analysis showed no upd(14)pat or microdeletion involving the 14q32.2 imprinted region in either patient. A differential mosaic ratio of cells with aberrant methylation of DMRs at the 14q32.2 imprinted region among tissues (connective tissue, lung, and brain) might have led to their atypical clinical features. Further studies of patients with epimutation should further expand the phenotypic spectrum of KOS.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 08:22
A Robust Polymerase Chain Reaction-based Assay for Quantifying Cytosine-guanine-guanine Trinucleotide Repeats in Fragile X Mental Retardation-1 Gene
Published on: September 16, 2019
Related Concept Videos
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 03:09
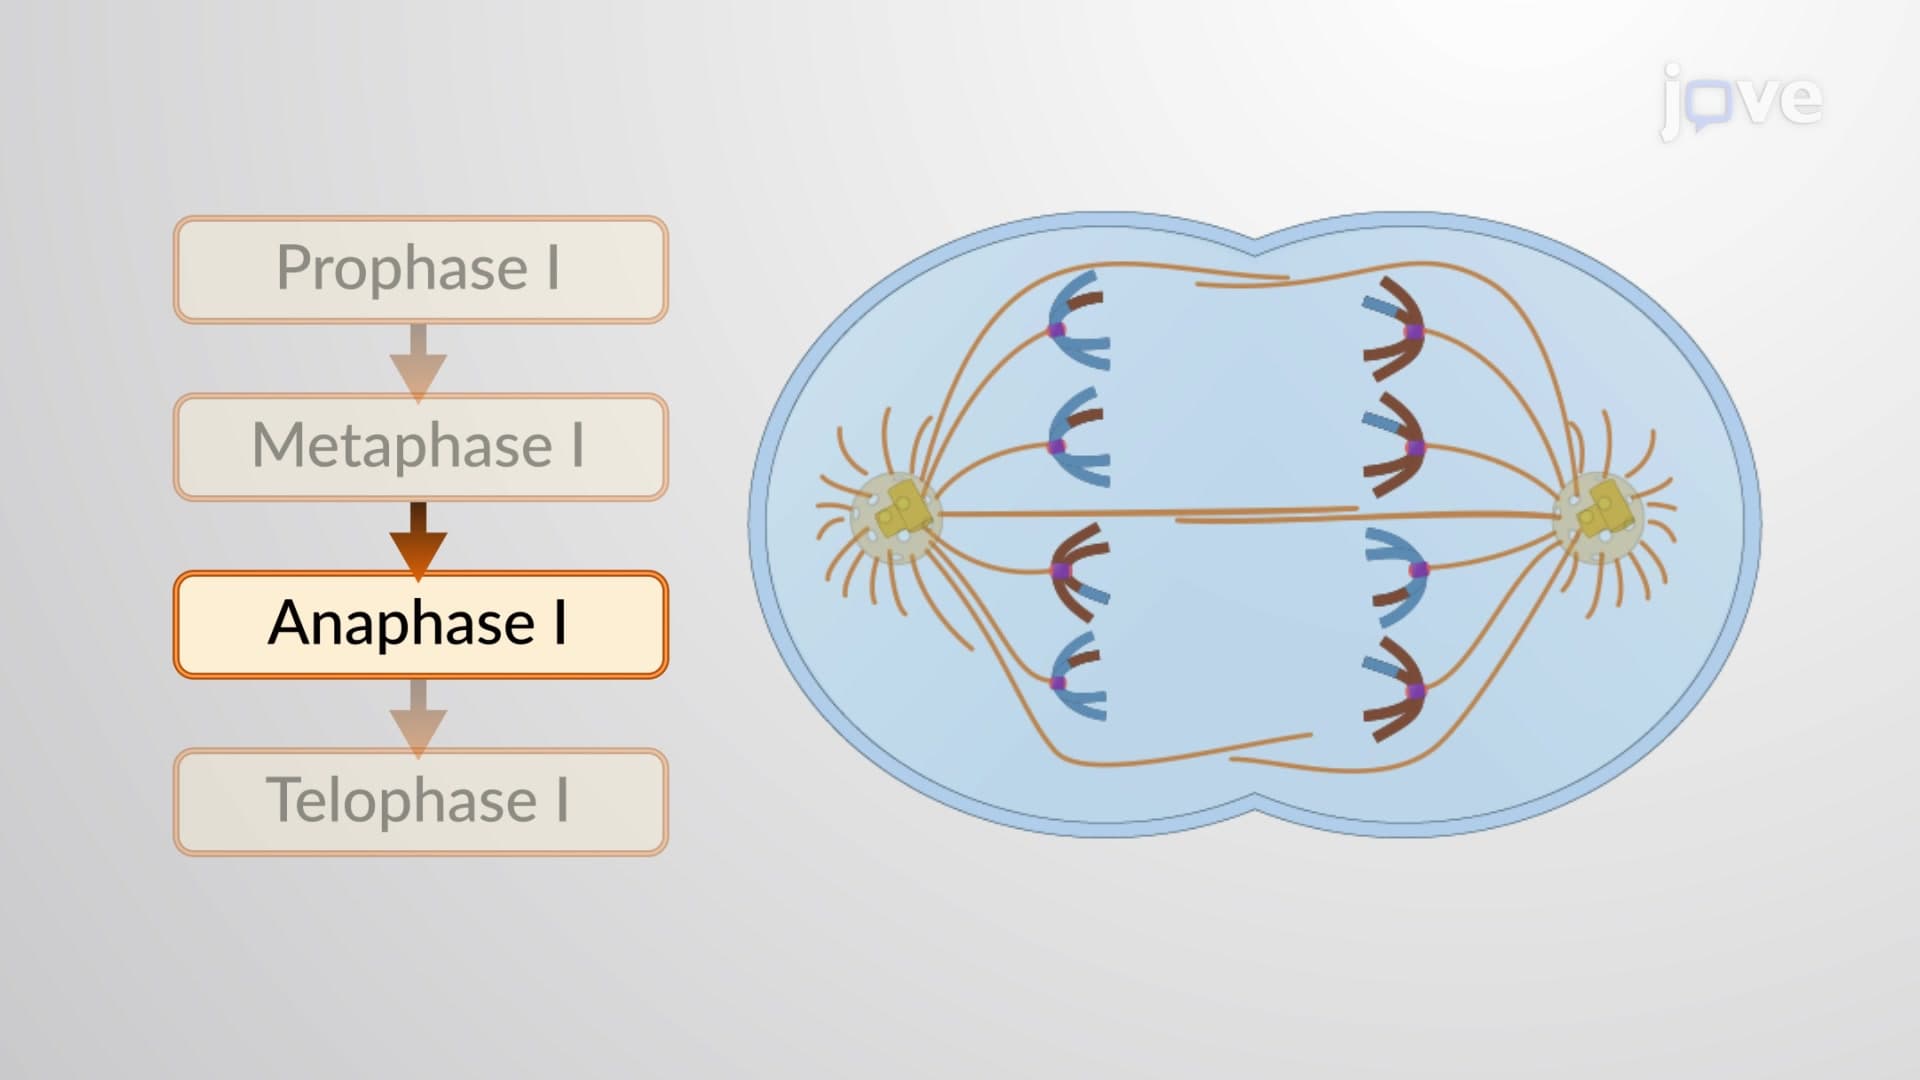
Meiosis I
Prophase I is the most extended and complex step of meiosis I characterized by synapsis, chromosome pairing, and recombination of the homologous chromosomes. This process is facilitated by a proteinaceous structure called the...
 01:21
Nondisjunction
 01:09
Epistasis Analysis
 01:20
Inborn Errors of Metabolism
 01:37
Epigenetic Regulation
X-chromosome...