Related Experiment Video
Updated: Sep 1, 2025

Specificity Analysis of Protein Lysine Methyltransferases Using SPOT Peptide Arrays
Published on: November 29, 2014
MLysPRED: graph-based multi-view clustering and multi-dimensional normal distribution resampling techniques to
Yun Zuo1, Yue Hong1, Xiangxiang Zeng2
1Department of Computer Science, Xiamen University, Xiamen 361005, China.

Abstract:
Posttranslational modification of lysine residues, K-PTM, is one of the most popular PTMs. Some lysine residues in proteins can be continuously or cascaded covalently modified, such as acetylation, crotonylation, methylation and succinylation modification. The covalent modification of lysine residues may have some special functions in basic research and drug development. Although many computational methods have been developed to predict lysine PTMs, up to now, the K-PTM prediction methods have been modeled and learned a single class of K-PTM modification. In view of this, this study aims to fill this gap by building a multi-label computational model that can be directly used to predict multiple K-PTMs in proteins. In this study, a multi-label prediction model, MLysPRED, is proposed to identify multiple lysine sites using features generated from human protein sequences. In MLysPRED, three kinds of multi-label sequence encoding algorithms (MLDBPB, MLPSDAAP, MLPSTAAP) are proposed and combined with three encoding strategies (CHHAA, DR and Kmer) to convert preprocessed lysine sequences into effective numerical features. A multidimensional normal distribution oversampling technique and graph-based multi-view clustering under-sampling algorithm were first proposed and incorporated to reduce the proportion of the original training samples, and multi-label nearest neighbor algorithm is used for classification. It is observed that MLysPRED achieved an Aiming of 92.21%, Coverage of 94.98%, Accuracy of 89.63%, Absolute-True of 81.46% and Absolute-False of 0.0682 on the independent datasets. Additionally, comparison of results with five existing predictors also indicated that MLysPRED is very promising and encouraging to predict multiple K-PTMs in proteins. For the convenience of the experimental scientists, 'MLysPRED' has been deployed as a user-friendly web-server at http://47.100.136.41:8181.
More Related Videos
 06:50
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
 12:49
Quantification of Site-specific Protein Lysine Acetylation and Succinylation Stoichiometry Using Data-independent Acquisition Mass Spectrometry
Published on: April 4, 2018
Related Concept Videos
 01:20
Cluster Sampling Method
To choose a cluster sample, divide the population into clusters (groups) and then randomly select some of the clusters. All the members from these clusters are in the cluster sample. For example, if you randomly sample four departments from your...
 01:06
Model Approaches for Pharmacokinetic Data: Distributed Parameter Models
The distributed parameter models are specifically designed to account for variations and differences in some drug classes. This model is particularly useful for assessing regional concentrations of anticancer or...
 01:24
One-Compartment Open Model: Wagner-Nelson and Loo Riegelman Method for ka Estimation
On...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:07
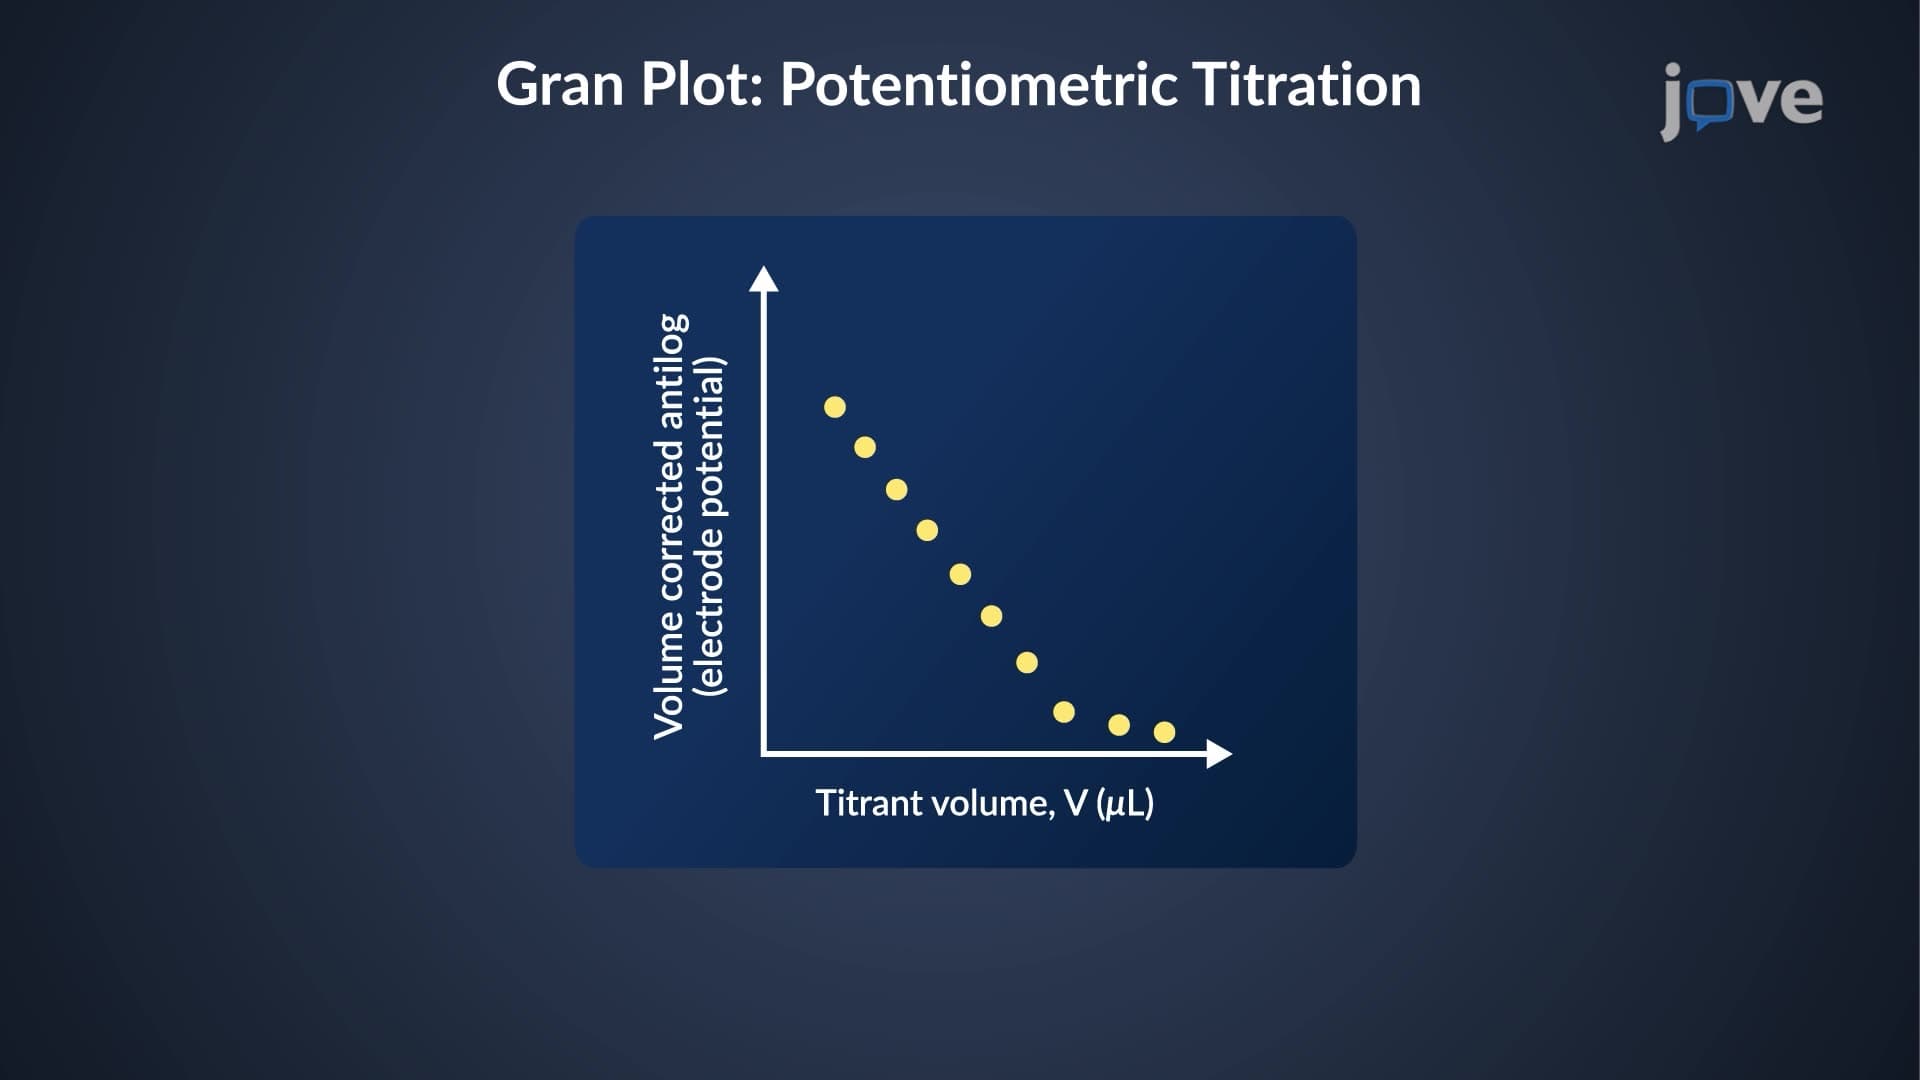
End Point Prediction: Gran Plot
For potentiometric titration, the Gran plot is created by plotting...