

Pharmaceutical Cocrystals of Ethenzamide: Molecular Structure Analysis Based on Vibrational Spectra and DFT
Mei Wan1, Jiyuan Fang1, Jiadan Xue2
1Centre for THz Research, China Jiliang University, Hangzhou 310018, China.

Abstract:
Pharmaceutical cocrystals can offer another advanced strategy for drug preparation and development and can facilitate improvements to the physicochemical properties of active pharmaceutical ingredients (APIs) without altering their chemical structures and corresponding pharmacological activities. Therefore, cocrystals show a great deal of potential in the development and research of drugs. In this work, pharmaceutical cocrystals of ethenzamide (ETZ) with 2,6-dihydroxybenzoic acid (26DHBA), 2,4-dihydroxybenzoic acid (24DHBA) and gallic acid (GA) were synthesized by the solvent evaporation method. In order to gain a deeper understanding of the structural changes after ETZ cocrystallization, terahertz time domain spectroscopy (THz-TDS) and Raman spectroscopy were used to characterize the single starting samples, corresponding physical mixtures and the cocrystals. In addition, the possible molecular structures of ETZ-GA, ETZ-26DHBA and ETZ-24DHBA cocrystals were optimized by density functional theory (DFT). The results of THz and Raman spectra with the DFT simulations for the three cocrystals revealed that the ETZ-GA cocrystal formed an O-H∙∙∙O hydrogen bond between the -OH of GA and oxygen of the amide group of the ETZ molecule, and it was also found that ETZ formed a dimer through a supramolecular amide-amide homosynthon; meanwhile, the ETZ-26DHBA cocrystal was formed by a powerful supramolecular acid-amide heterosynthon, and the ETZ-24DHBA cocrystal formed the O-H∙∙∙O hydrogen bond between the 4-hydroxy group of 24DHBA and oxygen of the amide group of the ETZ molecule. It could be seen that in the molecular structure analysis of the three cocrystals, the position and number of hydroxyl groups in the coformers play an essential role in guiding the formation of specific supramolecular synthons.
Related Concept Videos
 01:19
NMR Spectroscopy Of Amines
 01:14
NMR Spectroscopy of Aromatic Compounds
 01:26
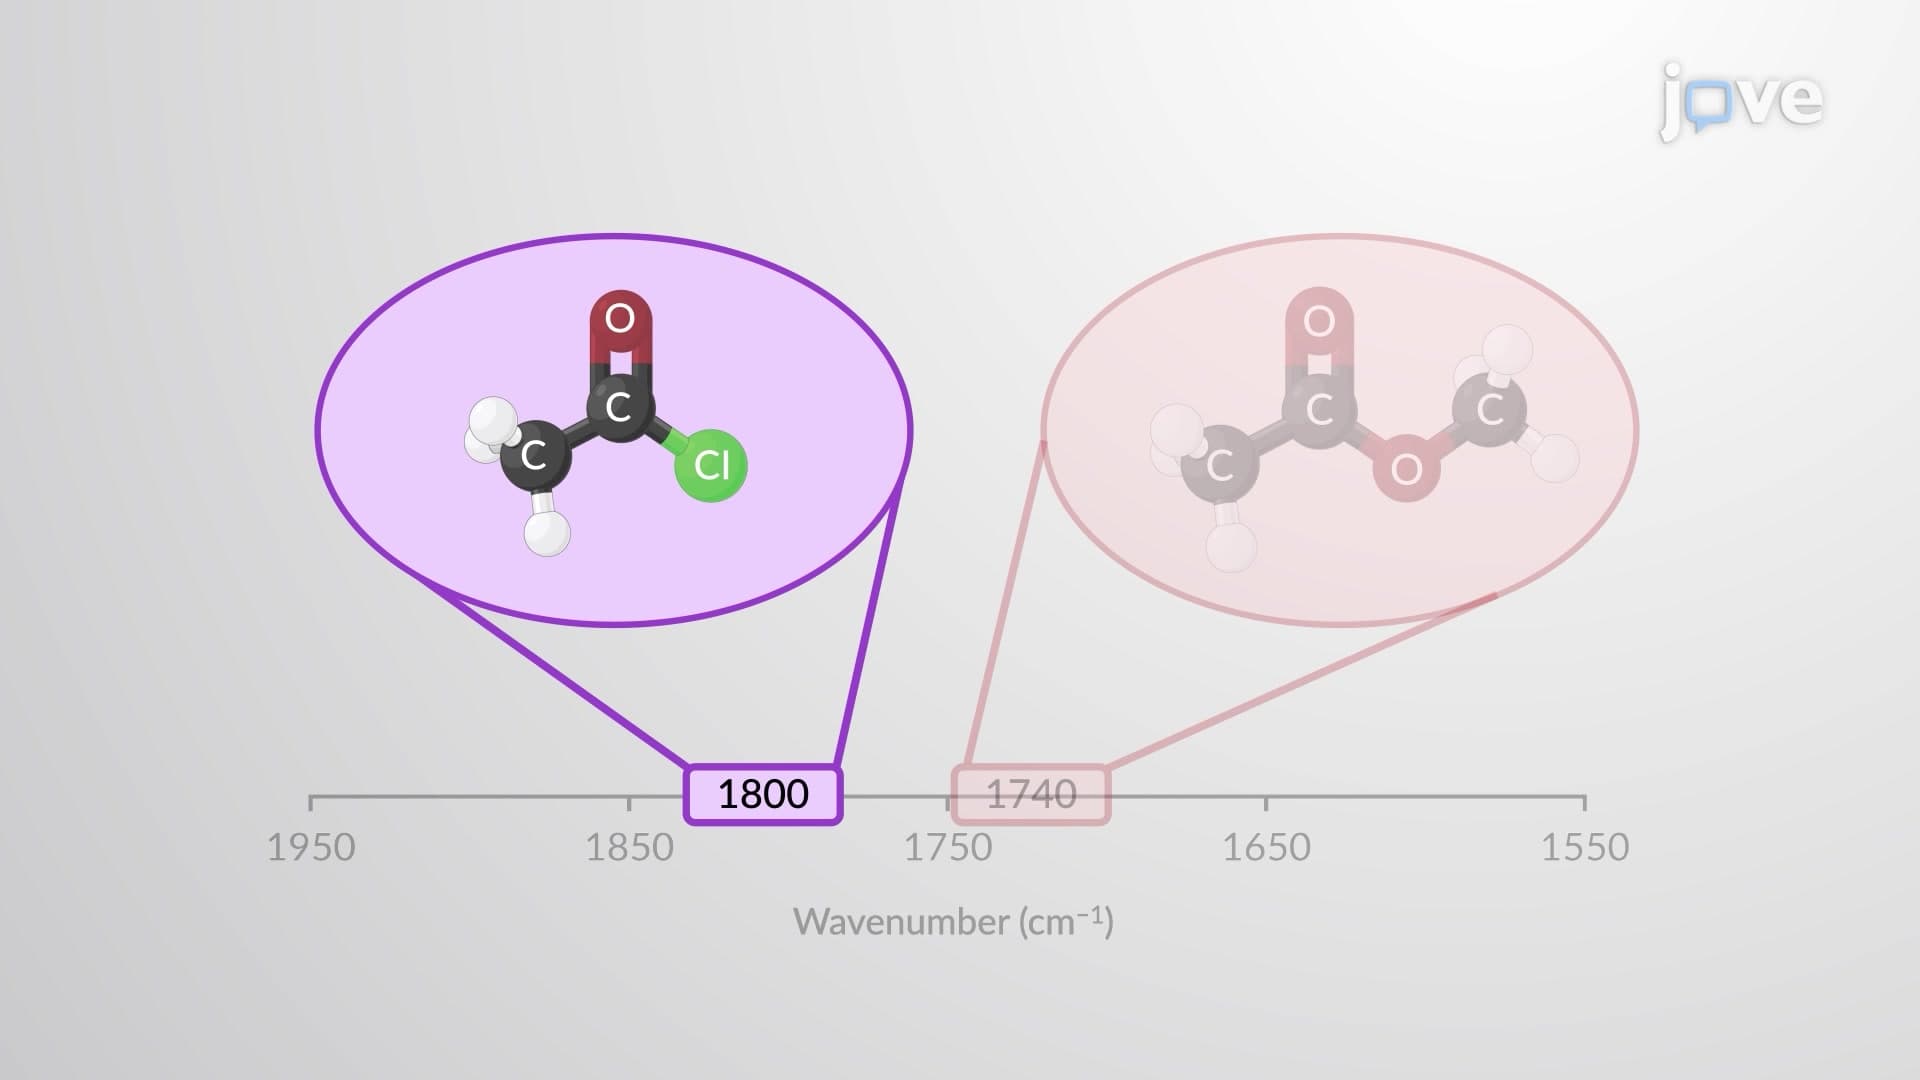
Spectroscopy of Carboxylic Acid Derivatives
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:24
Mass Spectrum: Interpretation
To...