Related Experiment Video
Updated: Sep 1, 2025

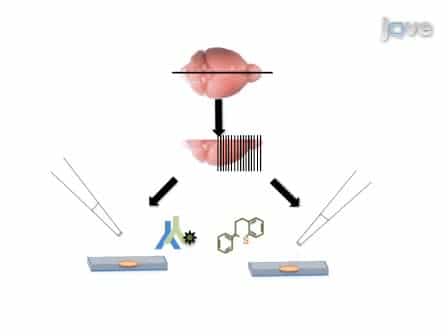
Detecting Amyloid-β Accumulation via Immunofluorescent Staining in a Mouse Model of Alzheimer's Disease
Published on: April 19, 2021
Adult-Onset Deficiency of Mitochondrial Complex III in a Mouse Model of Alzheimer's Disease Decreases Amyloid Beta
Milena Pinto1, Francisca Diaz2, Nadee Nissanka2
1Department of Neurology, University of Miami Miller School of Medicine, Miami, FL, USA. mpinto@med.miami.edu.

Abstract:
For decades, mitochondrial dysfunctions and the generation of reactive oxygen species have been proposed to promote the development and progression of the amyloid pathology in Alzheimer's disease, but this association is still debated. It is unclear whether different mitochondrial dysfunctions, such as oxidative phosphorylation deficiency and oxidative stress, are triggers or rather consequences of the formation of amyloid aggregates. Likewise, the role of the different mitochondrial oxidative phosphorylation complexes in Alzheimer's patients' brain remains poorly understood. Previous studies showed that genetic ablation of oxidative phosphorylation enzymes from early age decreased amyloid pathology, which were unexpected results. To better model oxidative phosphorylation defects in aging, we induced the ablation of mitochondrial Complex III (CIIIKO) in forebrain neurons of adult mice with amyloid pathology. We found that mitochondrial Complex III dysfunction in adult neurons induced mild oxidative stress but did not increase amyloid beta accumulation. On the contrary, CIIIKO-AD mice showed decreased plaque number, decreased Aβ42 toxic fragment, and altered amyloid precursor protein clearance pathway. Our results support the hypothesis that mitochondrial dysfunctions alone, caused by oxidative phosphorylation deficiency, is not the cause of amyloid accumulation.