Related Experiment Video
Updated: Sep 1, 2025
Visualizing the Interaction Between the Qdot-labeled Protein and Site-specifically Modified λ DNA at the Single Molecule Level
Published on: July 17, 2018
Mechanistic Investigation of Lysine-Targeted Covalent Inhibition of PI3Kδ via ONIOM QM:QM Computations
Volkan Fındık1,2, Betül Tuba Varınca Gerçik2, Öykü Sinek2
1LPCT, UMR 7019, University of Lorraine, CNRS, 54000, Nancy, France.

Abstract:
Phosphoinositide 3-kinase (PI3K) enzymes are important drug targets, especially in oncology, and several inhibitors are currently under investigation in clinical trials for the treatment of lymphocytic leukemia, follicular lymphoma, breast, thyroid, colorectal, and lung cancer. Targeted covalent inhibitors hold significant promise for drug discovery research especially for kinases. Targeting the lysine residues attracts attention as a new strategy in designing targeted covalent inhibitors, since the lysine residue provides several advantages over the traditional cysteine residue. Recently, new highly selective covalent inhibitors of PI3Kδ with activated ester warheads, targeting the conserved Lys779 residue, were reported. Based on the observed kinetics, a covalent inhibition mechanism was proposed, but the atomistic details of the reaction are still not understood. Therefore, in the present work, we have conducted quantum chemical ONIOM M06-2X/6-31+G(d,p):PM6 calculations on the active site cluster structure of PI3Kδ to elucidate the microscopic details of the mechanism of the aminolysis reaction between Lys779 and the ester inhibitors. Our calculations clearly discriminate the noncovalent methyl ester inhibitor and the covalent inhibitors with activated phenolic esters. For the representative p-NO2, p-F, p-H, and p-OCH3 phenolic esters, the Gibbs free energy profiles of alternative mechanistic paths through either Asp782 or Asp911 demonstrate the modulatory role of active site aspartate residues. The most plausible path alters depending on the electron-withdrawing/donating nature of the p-substituted phenolate leaving group. Inhibitors with sufficiently strong electron-withdrawing group prefer direct dissociation of the leaving group from the tetrahedral zwitterion intermediate, while the ones with electron-donating group favor the formation of a neutral tetrahedral intermediate prior to the dissociation. The relative Gibbs free energy barriers of p-NO2 < p-F < p-H < p-OCH3 substituted phenyl esters display the same qualitative trend as the experimentally measured kinact/KI values. Our results provide in depth insight into the mechanism, which can pave the way for optimizing the inhibitor efficiency.
Insights
Targeted covalent inhibitors targeting lysine residues in Phosphoinositide 3-kinase delta (PI3Kδ) show promise for cancer treatment. Quantum chemical calculations reveal the reaction mechanism, aiding in optimizing inhibitor efficiency for drug discovery.
Area of Science:
- Biochemistry and Medicinal Chemistry
- Computational Chemistry and Molecular Modeling
Background:
- Phosphoinositide 3-kinase (PI3K) enzymes are critical drug targets, particularly in oncology, with inhibitors in clinical trials for various cancers.
- Targeted covalent inhibitors offer advantages in drug discovery, and targeting lysine residues presents a novel strategy over traditional cysteine targeting.
- Recent development of selective covalent PI3Kδ inhibitors targeting Lys779 with activated ester warheads necessitates understanding their reaction mechanism.
Purpose of the Study:
- To elucidate the atomistic details of the covalent inhibition mechanism of PI3Kδ by ester inhibitors targeting Lys779.
- To investigate the role of active site aspartate residues (Asp782 and Asp911) in modulating the reaction pathway.
- To correlate computational findings with experimental kinetic data for optimizing inhibitor design.
Main Methods:
- Quantum chemical calculations using the ONIOM M06-2X/6-31+G(d,p):PM6 method were performed on a PI3Kδ active site cluster model.
- Analysis of Gibbs free energy profiles for alternative mechanistic pathways involving aminolysis reaction between Lys779 and various phenolic ester inhibitors.
- Comparison of calculated energy barriers with experimentally determined inactivation (k_inact/K_I) values.
Main Results:
- Calculations distinguished between noncovalent methyl ester and covalent activated phenolic ester inhibitors.
- The reaction mechanism and preferred pathway (via Asp782 or Asp911) are dependent on the electronic nature (electron-withdrawing/donating) of the phenolate leaving group.
- A clear correlation was observed between the calculated Gibbs free energy barriers and experimental kinetic data for differently substituted phenolic esters.
Conclusions:
- The study provides detailed microscopic insights into the covalent inhibition mechanism of PI3Kδ by activated ester inhibitors.
- The findings highlight the crucial role of the leaving group's electronic properties and active site aspartates in determining the reaction pathway.
- This mechanistic understanding can guide the rational design and optimization of more efficient PI3Kδ inhibitors for therapeutic applications.
Related Concept Videos
 01:22
PI3K/mTOR/AKT Signaling Pathway
 02:04
Covalently Linked Protein Regulators
These groups modify specific amino acids in a protein....
 01:42
Phosphoinositides and PIPs
Different phosphoinositides are synthesized and recruited on the cytosolic face of the plasma membrane. The localization of specific phosphoinositides concentrated in separate membrane...
 01:16
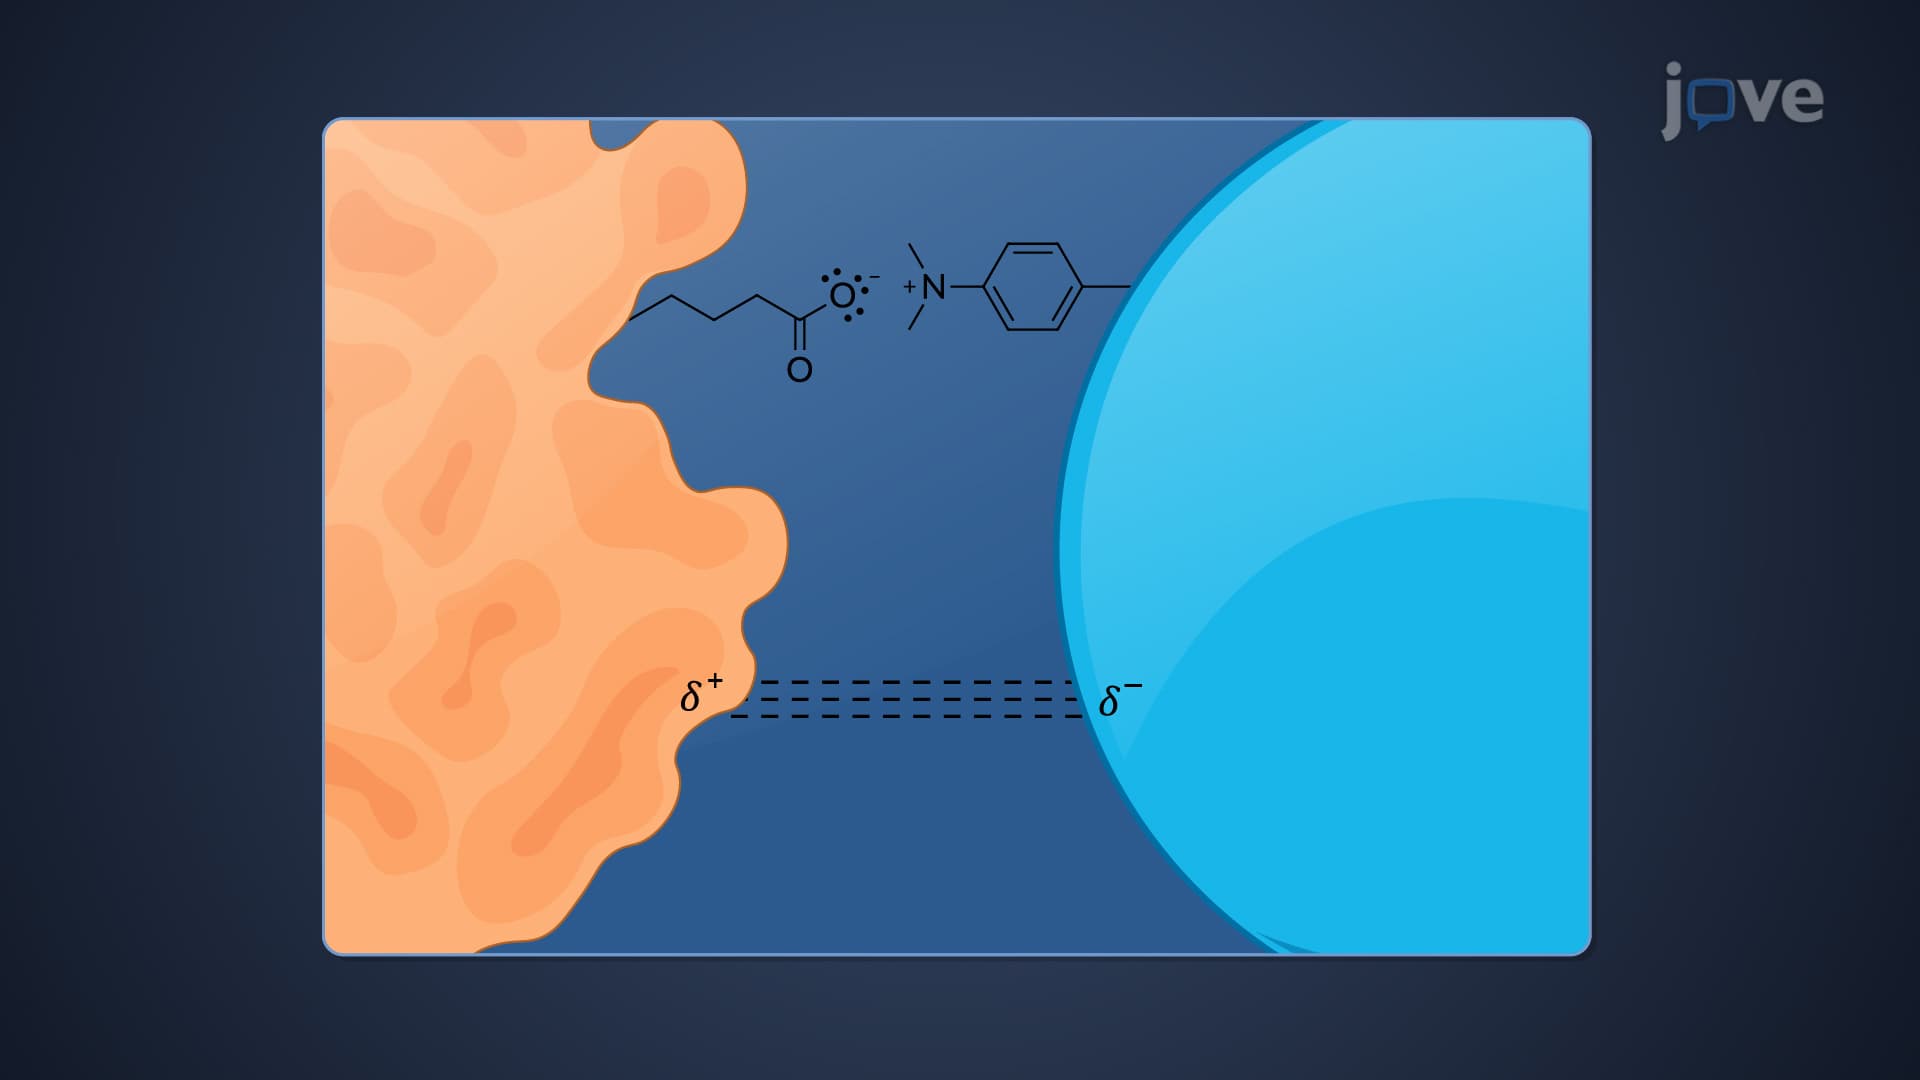
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...