Related Experiment Video
Updated: Aug 31, 2025

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Predicting the mutation effects of protein-ligand interactions via end-point binding free energy calculations:
Yang Yu1, Zhe Wang2, Lingling Wang1
1Department of Medicinal Chemistry, China Pharmaceutical University, Nanjing, 210009, Jiangsu, People's Republic of China.

Abstract:
Protein mutations occur frequently in biological systems, which may impact, for example, the binding of drugs to their targets through impairing the critical H-bonds, changing the hydrophobic interactions, etc. Thus, accurately predicting the effects of mutations on biological systems is of great interests to various fields. Unfortunately, it is still unavailable to conduct large-scale wet-lab mutation experiments because of the unaffordable experimental time and financial costs. Alternatively, in silico computation can serve as a pioneer to guide the experiments. In fact, numerous pioneering works have been conducted from computationally cheaper machine-learning (ML) methods to the more expensive alchemical methods with the purpose to accurately predict the mutation effects. However, these methods usually either cannot result in a physically understandable model (ML-based methods) or work with huge computational resources (alchemical methods). Thus, compromised methods with good physical characteristics and high computational efficiency are expected. Therefore, here, we conducted a comprehensive investigation on the mutation issues of biological systems with the famous end-point binding free energy calculation methods represented by MM/GBSA and MM/PBSA. Different computational strategies considering different length of MD simulations, different value of dielectric constants and whether to incorporate entropy effects to the predicted total binding affinities were investigated to provide a more accurate way for predicting the energetic change upon protein mutations. Overall, our result shows that a relatively long MD simulation (e.g. 100 ns) benefits the prediction accuracy for both MM/GBSA and MM/PBSA (with the best Pearson correlation coefficient between the predicted ∆∆G and the experimental data of ~ 0.44 for a challenging dataset). Further analyses shows that systems involving large perturbations (e.g. multiple mutations and large number of atoms change in the mutation site) are much easier to be accurately predicted since the algorithm works more sensitively to the large change of the systems. Besides, system-specific investigation reveals that conformational adjustment is needed to refine the micro-environment of the manually mutated systems and thus lead one to understand why longer MD simulation is necessary to improve the predicting result. The proposed strategy is expected to be applied in large-scale mutation effects investigation with interpretation.

Related Concept Videos
 02:18
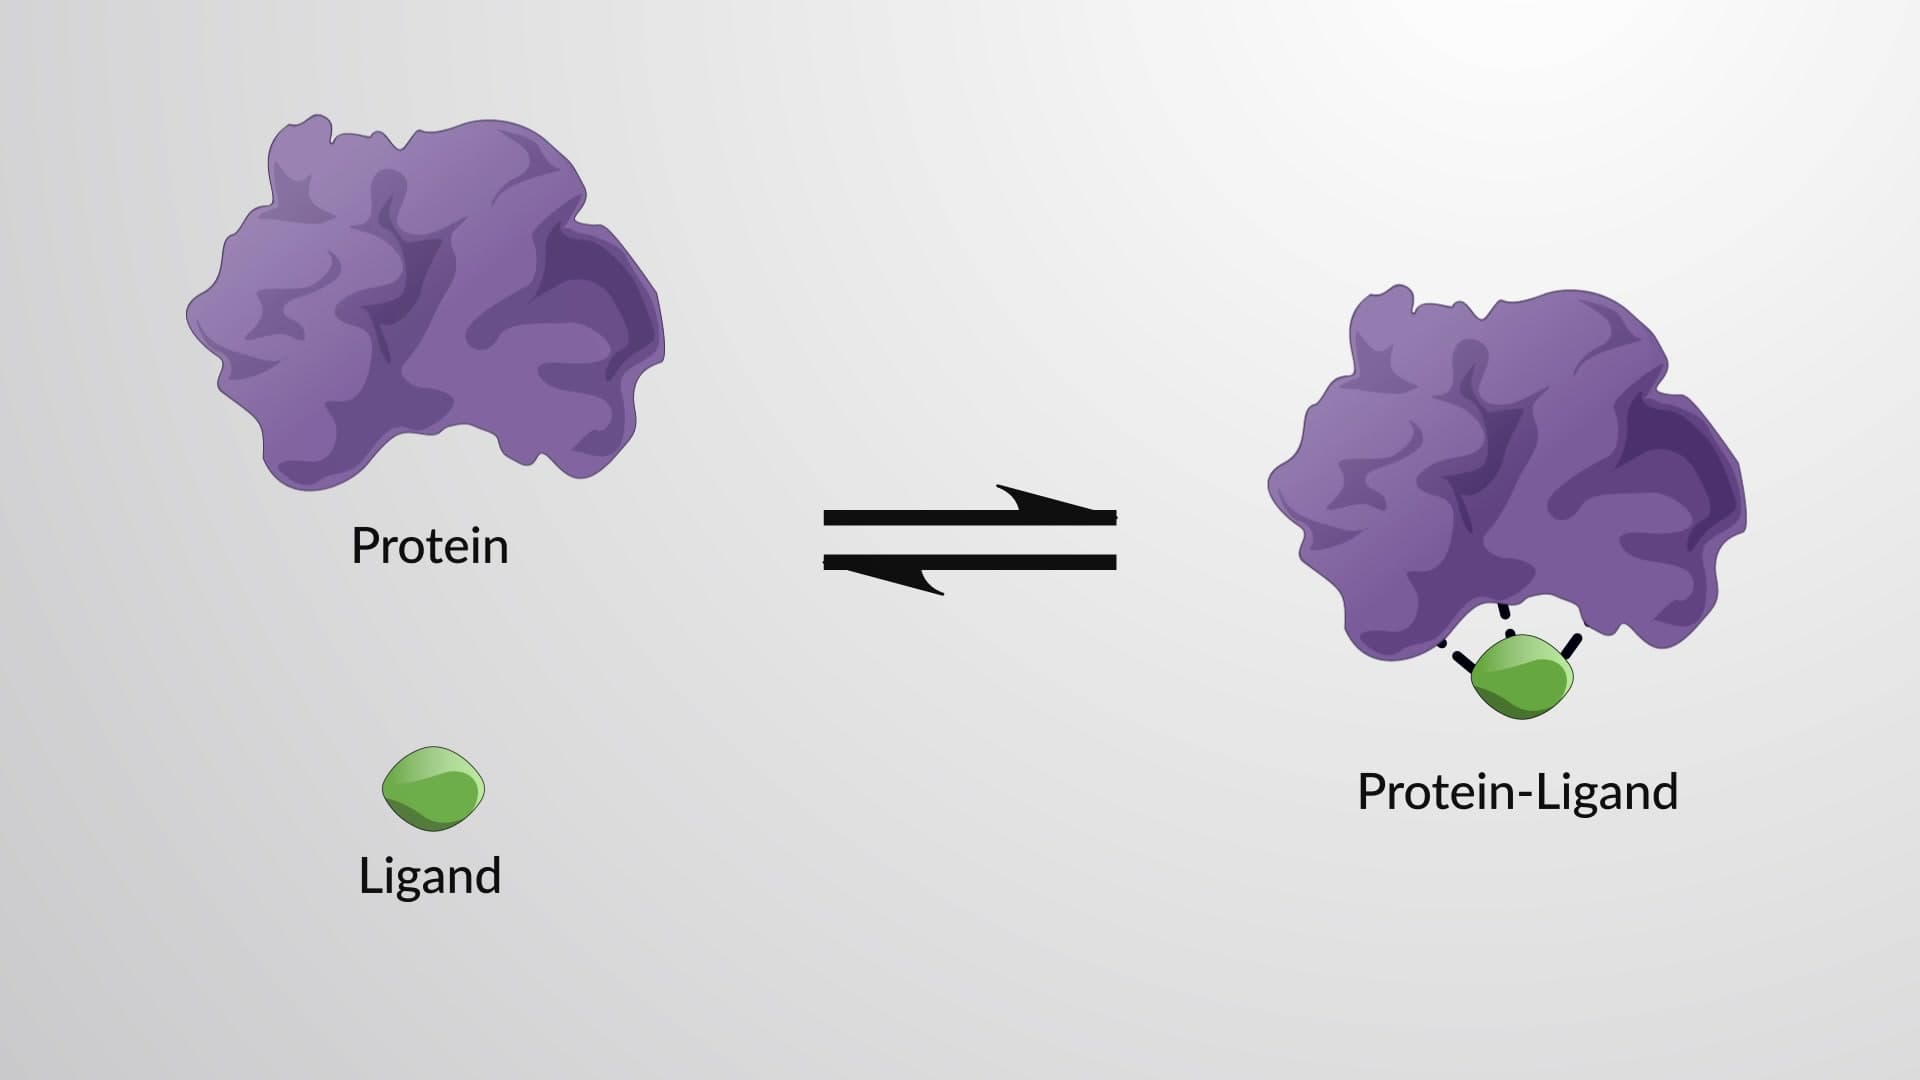
The Equilibrium Binding Constant and Binding Strength
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 00:49
Ligand Binding and Linkage