Related Experiment Video
Updated: Aug 31, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Phytochemical drug discovery for COVID-19 using high-resolution computational docking and machine learning assisted
Zirui Wang1,2, Theodore Belecciu1,2, Joelle Eaves1,2
1Institute for Quantitative Health Science and Engineering, Michigan State University, East Lansing, MI, USA.

Abstract:
The COVID-19 pandemic has resulted in millions of deaths around the world. Multiple vaccines are in use, but there are many underserved locations that do not have adequate access to them. Variants may emerge that are highly resistant to existing vaccines, and therefore cheap and readily obtainable therapeutics are needed. Phytochemicals, or plant chemicals, can possibly be such therapeutics. Phytochemicals can be used in a polypharmacological approach, where multiple viral proteins are inhibited and escape mutations are made less likely. Finding the right phytochemicals for viral protein inhibition is challenging, but in-silico screening methods can make this a more tractable problem. In this study, we screen a wide range of natural drug products against a comprehensive set of SARS-CoV-2 proteins using a high-resolution computational workflow. This workflow consists of a structure-based virtual screening (SBVS), where an initial phytochemical library was docked against all selected protein structures. Subsequently, ligand-based virtual screening (LBVS) was employed, where chemical features of 34 lead compounds obtained from the SBVS were used to predict 53 lead compounds from a larger phytochemical library via supervised learning. A computational docking validation of the 53 predicted leads obtained from LBVS revealed that 28 of them elicit strong binding interactions with SARS-CoV-2 proteins. Thus, the inclusion of LBVS resulted in a 4-fold increase in the lead discovery rate. Of the total 62 leads, 18 showed promising pharmacokinetic properties in a computational ADME screening. Collectively, this study demonstrates the advantage of incorporating machine learning elements into a virtual screening workflow.Communicated by Ramaswamy H. Sarma.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 08:15
Author Spotlight: Network Pharmacology and Molecular Docking to Decipher the Action of Jiawei Shengjiang San Against Diabetic Kidney Disease
Published on: May 10, 2024
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
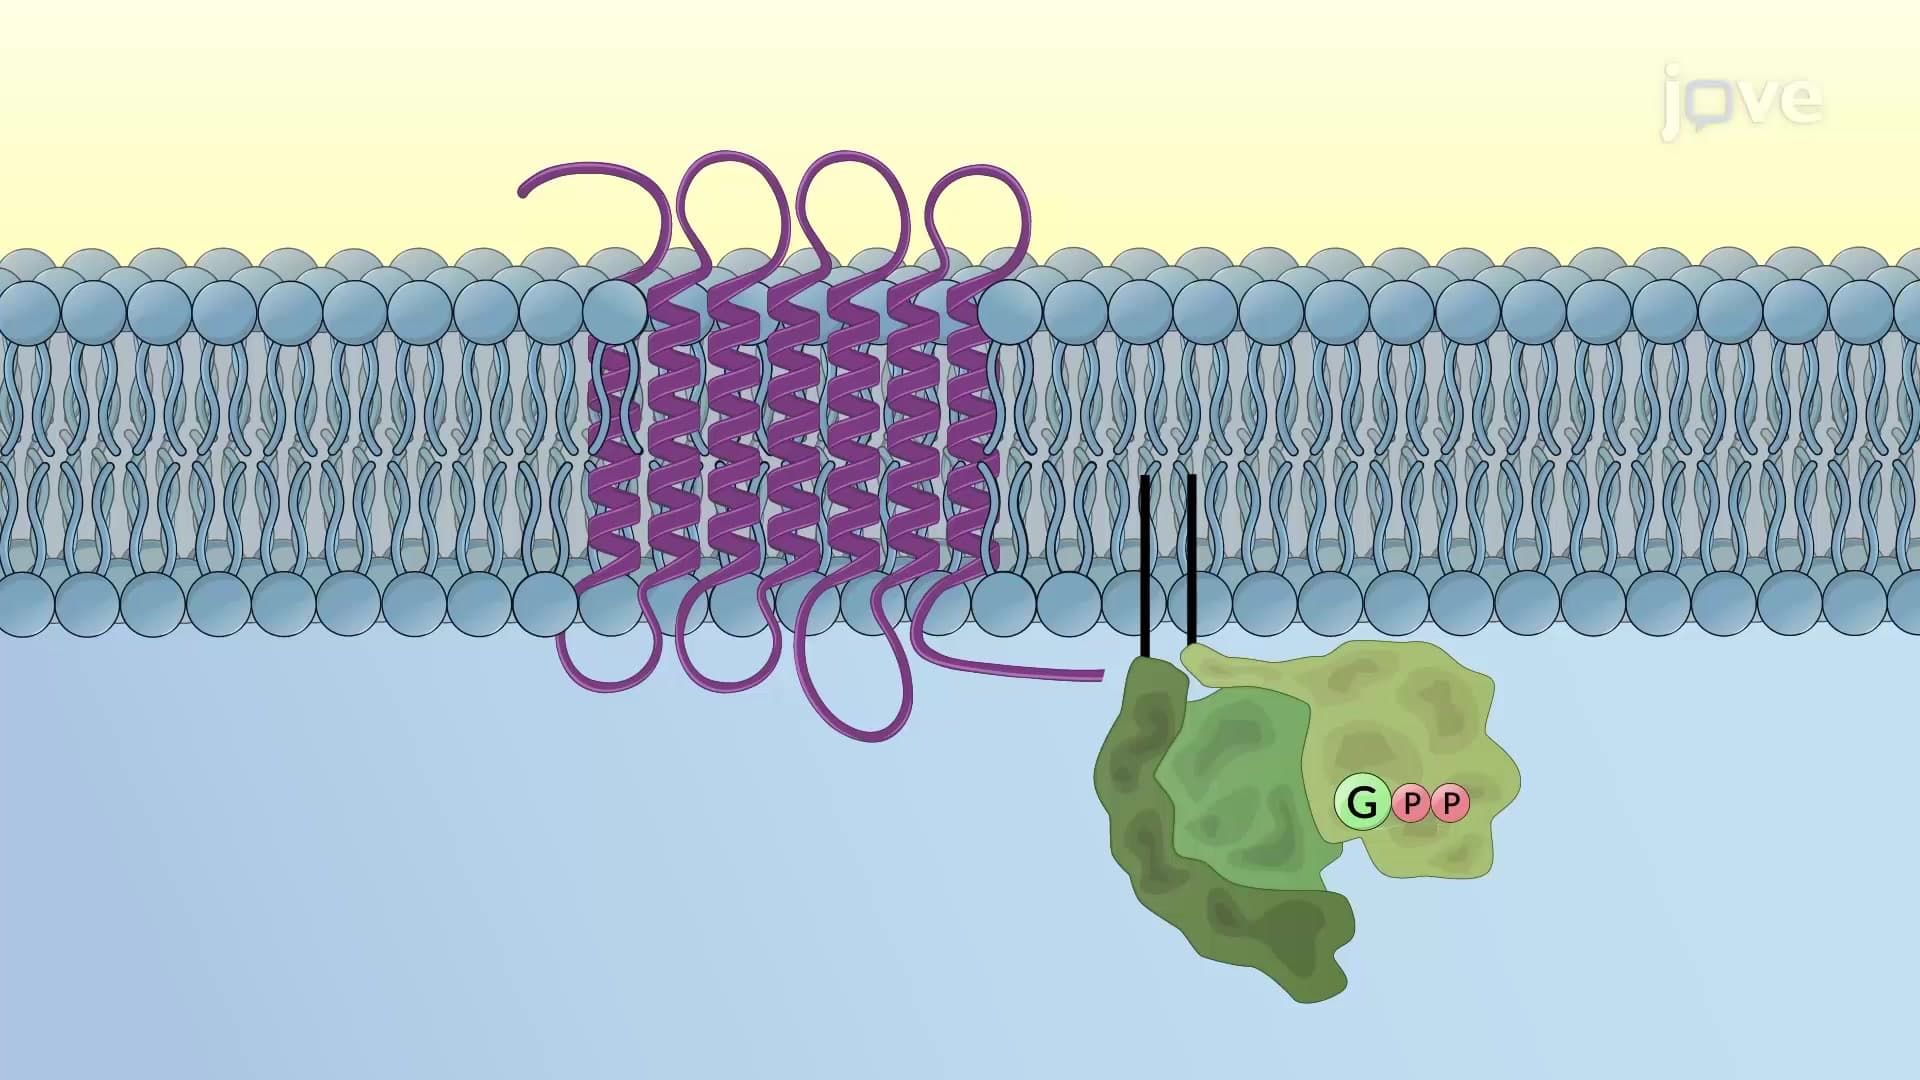
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...