Related Experiment Video
Updated: Aug 28, 2025
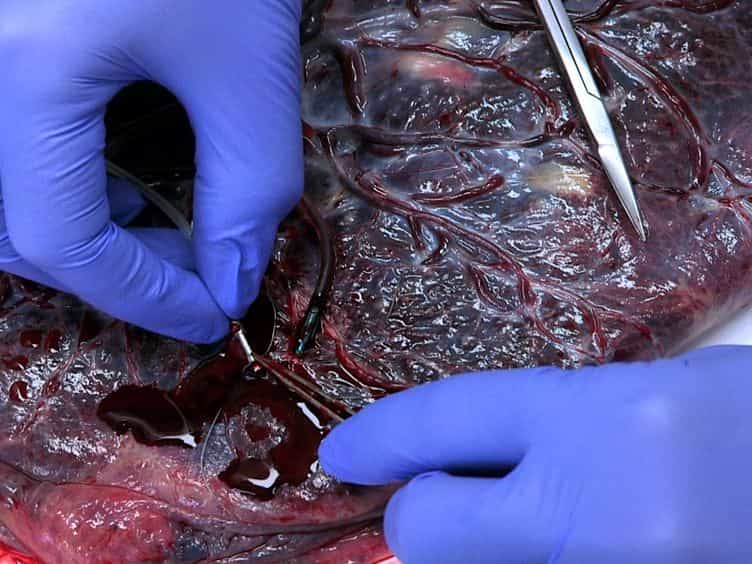
Determination of the Transport Rate of Xenobiotics and Nanomaterials Across the Placenta using the ex vivo Human Placental Perfusion Model
Published on: June 18, 2013
Predicting Human Fetal Drug Exposure Through Maternal-Fetal PBPK Modeling and In Vitro or Ex Vivo Studies
Ankit Balhara1, Aditya R Kumar1, Jashvant D Unadkat1
1Department of Pharmaceutics, University of Washington, Seattle, Washington, USA.

Abstract:
Medication (drug) use in human pregnancy is prevalent. Determining fetal safety and efficacy of drugs is logistically challenging. However, predicting (not measuring) fetal drug exposure (systemic and tissue) throughout pregnancy is possible through maternal-fetal physiologically based pharmacokinetic (PBPK) modeling and simulation. Such prediction can inform fetal drug safety and efficacy. Fetal drug exposure can be quantified in 2 complementary ways. First, the ratio of the steady-state unbound plasma concentration in the fetal plasma (or area under the plasma concentration-time curve) to the corresponding maternal plasma concentration (ie, Kp,uu ). Second, the maximum unbound peak (Cu,max,ss,f ) and trough (Cu,min,ss,f ) fetal steady-state plasma concentrations. We (and others) have developed a maternal-fetal PBPK model that can successfully predict maternal drug exposure. To predict fetal drug exposure, the model needs to be populated with drug specific parameters, of which transplacental clearances (active and/or passive) and placental/fetal metabolism of the drug are critical. Herein, we describe in vitro studies in cells/tissue fractions or the perfused human placenta that can be used to determine these drug-specific parameters. In addition, we provide examples whereby this approach has successfully predicted systemic fetal exposure to drugs that passively or actively cross the placenta. Apart from maternal-fetal PBPK models, animal studies also have the potential to estimate fetal drug exposure by allometric scaling. Whether such scaling will be successful is yet to be determined. Here, we review the above approaches to predict fetal drug exposure, outline gaps in our knowledge to make such predictions and map out future research directions that could fill these gaps.
Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:20
Pharmacokinetic Models: Overview
There are three primary types of models: empirical, compartment, and physiological. Empirical models, with minimal...
 01:12
Analysis of Population Pharmacokinetic Data
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:06
Model Approaches for Pharmacokinetic Data: Distributed Parameter Models
The distributed parameter models are specifically designed to account for variations and differences in some drug classes. This model is particularly useful for assessing regional concentrations of anticancer or...