Related Experiment Video
Updated: Aug 28, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Computational Protocol to Evaluate Electron-Phonon Interactions Within Density Matrix Perturbation Theory
Han Yang1,2, Marco Govoni2,3, Arpan Kundu2
1Department of Chemistry, University of Chicago, Chicago, Illinois 60637, United States.

Abstract:
We present a computational protocol, based on density matrix perturbation theory, to obtain non-adiabatic, frequency-dependent electron-phonon self-energies for molecules and solids. Our approach enables the evaluation of electron-phonon interaction using hybrid functionals, for spin-polarized systems, and the computational overhead to include dynamical and non-adiabatic terms in the evaluation of electron-phonon self-energies is negligible. We discuss results for molecules, as well as pristine and defective solids.


Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 01:20
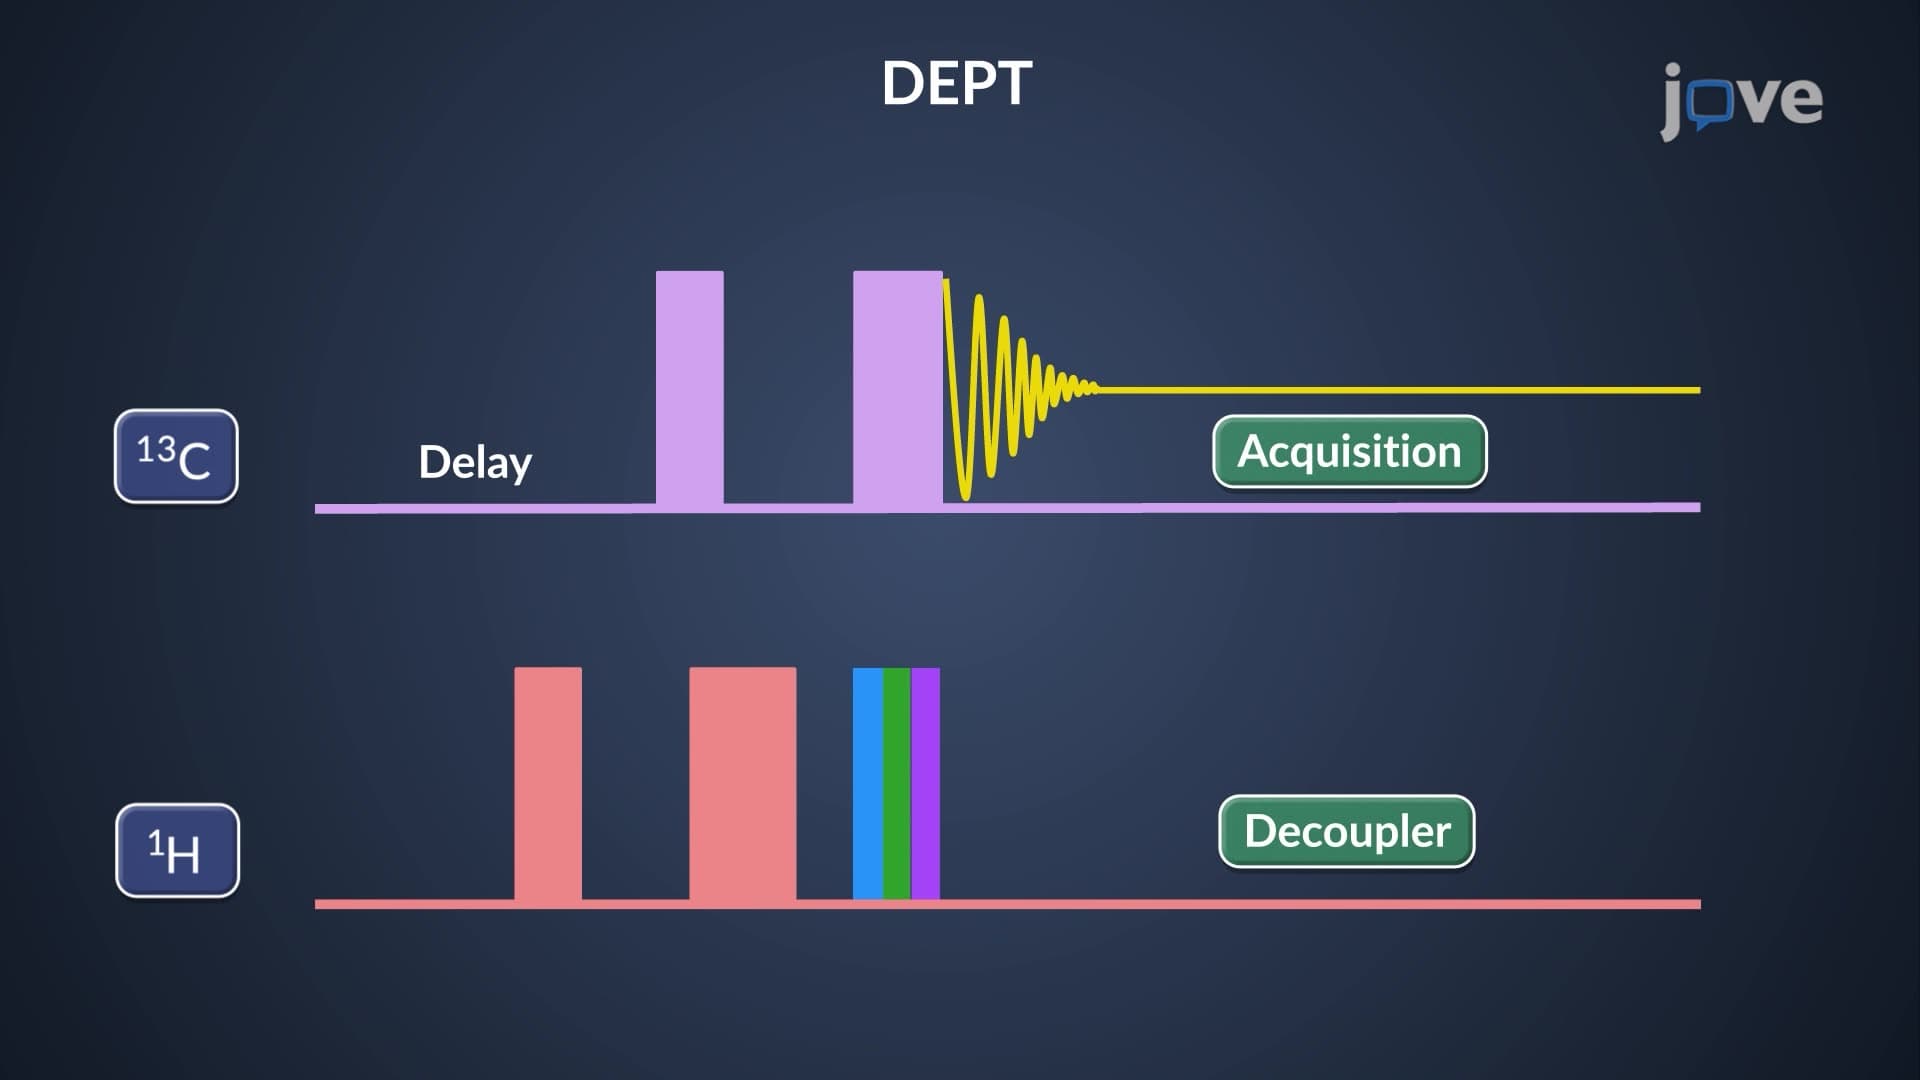
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 02:32
The de Broglie Wavelength
 02:45
The Quantum-Mechanical Model of an Atom
 02:40
MO Theory and Covalent Bonding
 02:35
Molecular Orbital Theory I