Related Experiment Video
Updated: Aug 27, 2025

Author Spotlight: Developing a Disposable Dosator for Preclinical Testing of Dry Powder Inhalers in Small Animal Models
Published on: August 18, 2023
Evaluation of Time Consolidation Effect of Pharmaceutical Powders
Prajwal Thool1, Chimara Stancill1, Minhthi Bui1
1Small Molecule Pharmaceutical Sciences, Genentech, Inc. 1 DNA Way, South San Francisco, CA, 94080, USA.

Purpose:
We aim to perform a systematic study of the time consolidation effect, i.e. the reduction of powder flowability resulting from at-rest storage, on a diverse array of pharmaceutical powders under different stress, humidity, and length of time.
Methods:
A ring shear cell-based methodology was employed. An instantaneous flow function was obtained, followed immediately by at-rest consolidation at precisely controlled humidity, stress, and duration. The consolidated powder was then subjected to shear-cell measurement. The difference in flowability between the immediate and consolidated specimens were attributed to the time consolidation effect.
Results:
Among the six excipients tested, three exhibited time consolidation at varying extents. Citric acid and starch underwent time consolidation only at high relative humidity (RH = 75%), promoted by vapor condensation and liquid bridge formation. For both materials, the flowability decreased with time, and the extent of time consolidation was not sensitive to the stress applied (0.4-2 kPa). Importantly, mannitol was found to time consolidate under both 50% and 75% RH. Given time, mannitol formed cake, giving rise to flow function below unity. Inverse gas chromatography analysis indicated that mannitol possesses high total surface energy among known pharmaceutical powders.
Conclusion:
While time consolidation is prevalent among pharmaceutical powders, most can be mitigated by controlling the RH to below 75%. Notably, for materials possessing high surface energy, such as mannitol, time consolidation could occur at ambient humidity. Therefore, it is desirable to consider in-depth time consolidation evaluation for high surface energy powders, e.g. those subjected to milling or of amorphous nature.
Related Concept Videos
 01:28
Factors Influencing Drug Absorption: Pharmaceutical Parameters
 01:23
Factors Affecting Dissolution: Particle Size and Effective Surface Area
 01:09
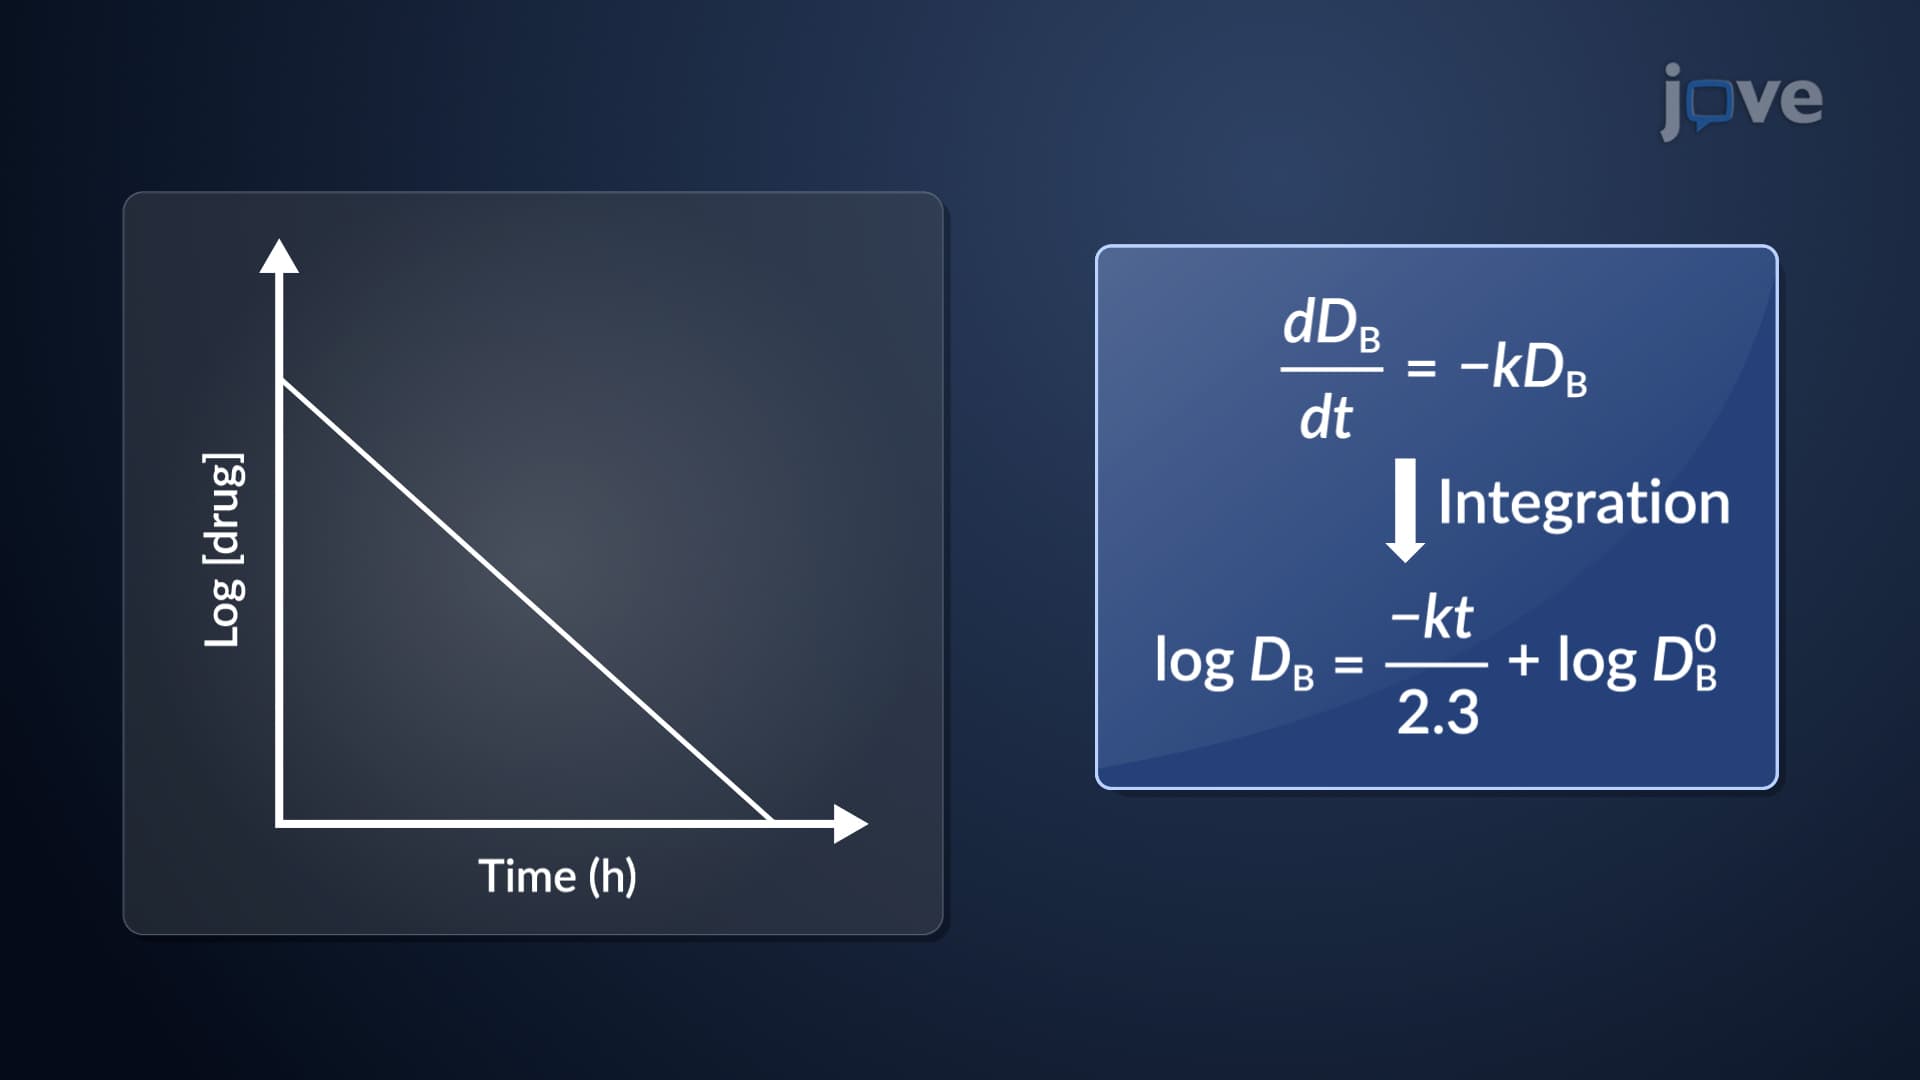
One-Compartment Open Model for IV Bolus Administration: Estimation of Elimination Rate Constant, Half-Life and Volume of Distribution
 01:02
Noncompartmental Analysis: Mean Transit, Absorption and Dissolution Time
One of the key parameters is the mean transit time (MTT), which refers to the total duration required for drug molecules to transit through the body. MTT is determined by calculating the ratio of the area under the moment curve to the area...
 01:14
Time Course of Drug Effect
 01:21
Factors Affecting Dissolution: Polymorphism, Amorphism and Pseudopolymorphism
Some polymorphic crystals possess lower aqueous solubility than their amorphous counterparts, leading to incomplete absorption. For instance, the oral suspension of Chloramphenicol, which...