Related Experiment Video
Updated: Aug 27, 2025

Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Multiconfiguration Pair-Density Functional Theory for Chromium(IV) Molecular Qubits
Arturo Sauza-de la Vega1, Riddhish Pandharkar1,2, Gautam D Stroscio1
1Department of Chemistry, Pritzker School of Molecular Engineering, James Franck Institute, Chicago Center for Theoretical Chemistry, University of Chicago, Chicago, Illinois 60637, United States.

Abstract:
Pseudotetrahedral organometallic complexes containing chromium(IV) and aryl ligands have been experimentally identified as promising molecular qubit candidates. Here we present a computational protocol based on multiconfiguration pair-density functional theory for computing singlet-triplet gaps and zero-field splitting (ZFS) parameters in Cr(IV) aryl complexes. We find that two multireference methods, multistate complete active space second-order perturbation theory (MS-CASPT2) and hybrid multistate pair-density functional theory (HMS-PDFT), perform better than Kohn-Sham density functional theory for singlet-triplet gaps. Despite the very small values of the ZFS parameters, both multireference methods performed qualitatively well. MS-CASPT2 and HMS-PDFT performed particularly well for predicting the trend in the ratio of the rhombic and axial ZFS parameters, |E/D|. We have also investigated the dependence and sensitivity of the calculated ZFS parameters on the active space and the molecular geometry. The methodologies outlined here can guide future prediction of ZFS parameters in molecular qubit candidates.

Related Concept Videos
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 03:26
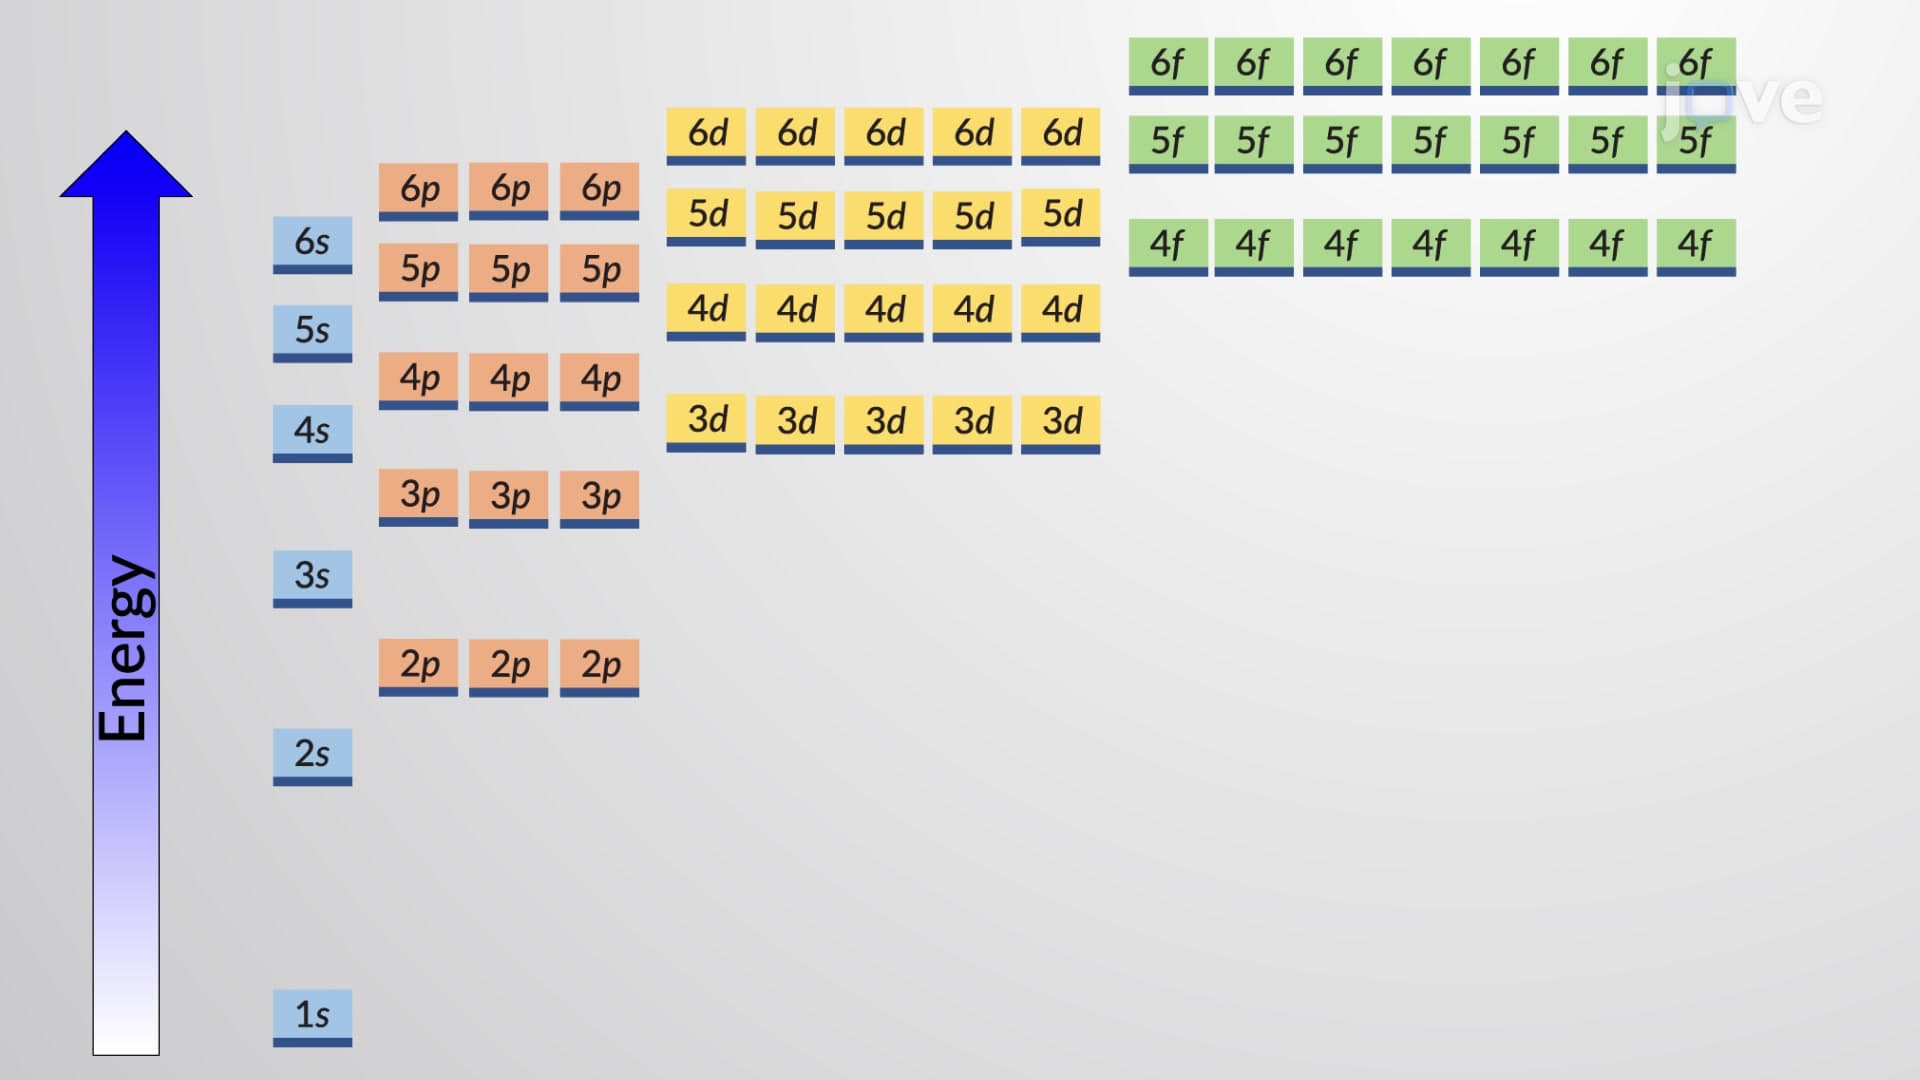
Electron Configuration of Multielectron Atoms
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...