Related Experiment Video
Updated: Aug 26, 2025

Author Spotlight: Functionalizing Metal-Organic Frameworks: Advancements, Challenges, and the Power of Post-Synthetic Ligand Exchange
Published on: June 23, 2023
Structural and Energetic Properties of Weak Noncovalent Interactions in Two Closely Related
Lamya H Al-Wahaibi1, Sekar Karthikeyan2, Olivier Blacque3
1Department of Chemistry, College of Sciences, Princess Nourah bint Abdulrahman University, Riyadh 11671, Saudi Arabia.

Abstract:
Two 3,6-disubstituted-[1,2,4]triazolo[3,4-b][1,3,4]thiadiazole derivatives, namely, 3-(adamantan-1-yl)-6-(2-chloro-6-fluorophenyl)-[1,2,4]triazolo[3,4-b][1,3,4]thiadiazole 1 and 6-(2-chloro-6-fluorophenyl)-3-phenyl-[1,2,4]triazolo[3,4-b][1,3,4]thiadiazole 2, were prepared, and the detailed analysis of the weak intermolecular interactions responsible for the supramolecular self-assembly was performed using X-ray diffraction and theoretical tools. Analyses of Hirshfeld surface and 2D fingerprint plot demonstrated the effect of adamant-1-yl/phenyl moieties on intermolecular interactions in solid-state structures. The effect of these substituents on H···H/Cl/N contacts was more specific. The CLP-PIXEL and density functional theory methods provide information on the energetics of molecular dimers observed in these compounds. The crystal structure of compound 1 stabilizes with a variety of weak intermolecular interactions, including C-H···N, C-H···π, and C-H···Cl hydrogen bonds, a directional C-S···π chalcogen bond, and unconventional short F···C/N contacts. The crystal structure of compound 2 is stabilized by π-stacking interactions, C-H···N, C-H···π, and C-H···Cl hydrogen bonds, and highly directional attractive σ-hole interactions such as the C-Cl···N halogen bond and the C-S···N chalcogen bond. In addition, S(lp)···C(π) and short N···N contacts play a supportive role in the stabilization of certain molecular dimers. The final supramolecular architectures resulting from the combination of different intermolecular interactions are observed in both the crystal packing. The molecular electrostatic potential map reveals complementary electrostatic potentials of the interacting atoms. The quantum theory of atoms in molecules approach was used to delineate the nature and strength of different intermolecular interactions present in different dimers of compounds 1 and 2. The in vitro experiments suggest that both compounds showed selectivity against COX-2 targets rather than COX-1. Molecular docking analysis showed the binding pose of the compounds at the active sites of COX-1/2 enzymes.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 01:16
Cycloaddition Reactions: MO Requirements for Thermal Activation
 02:29
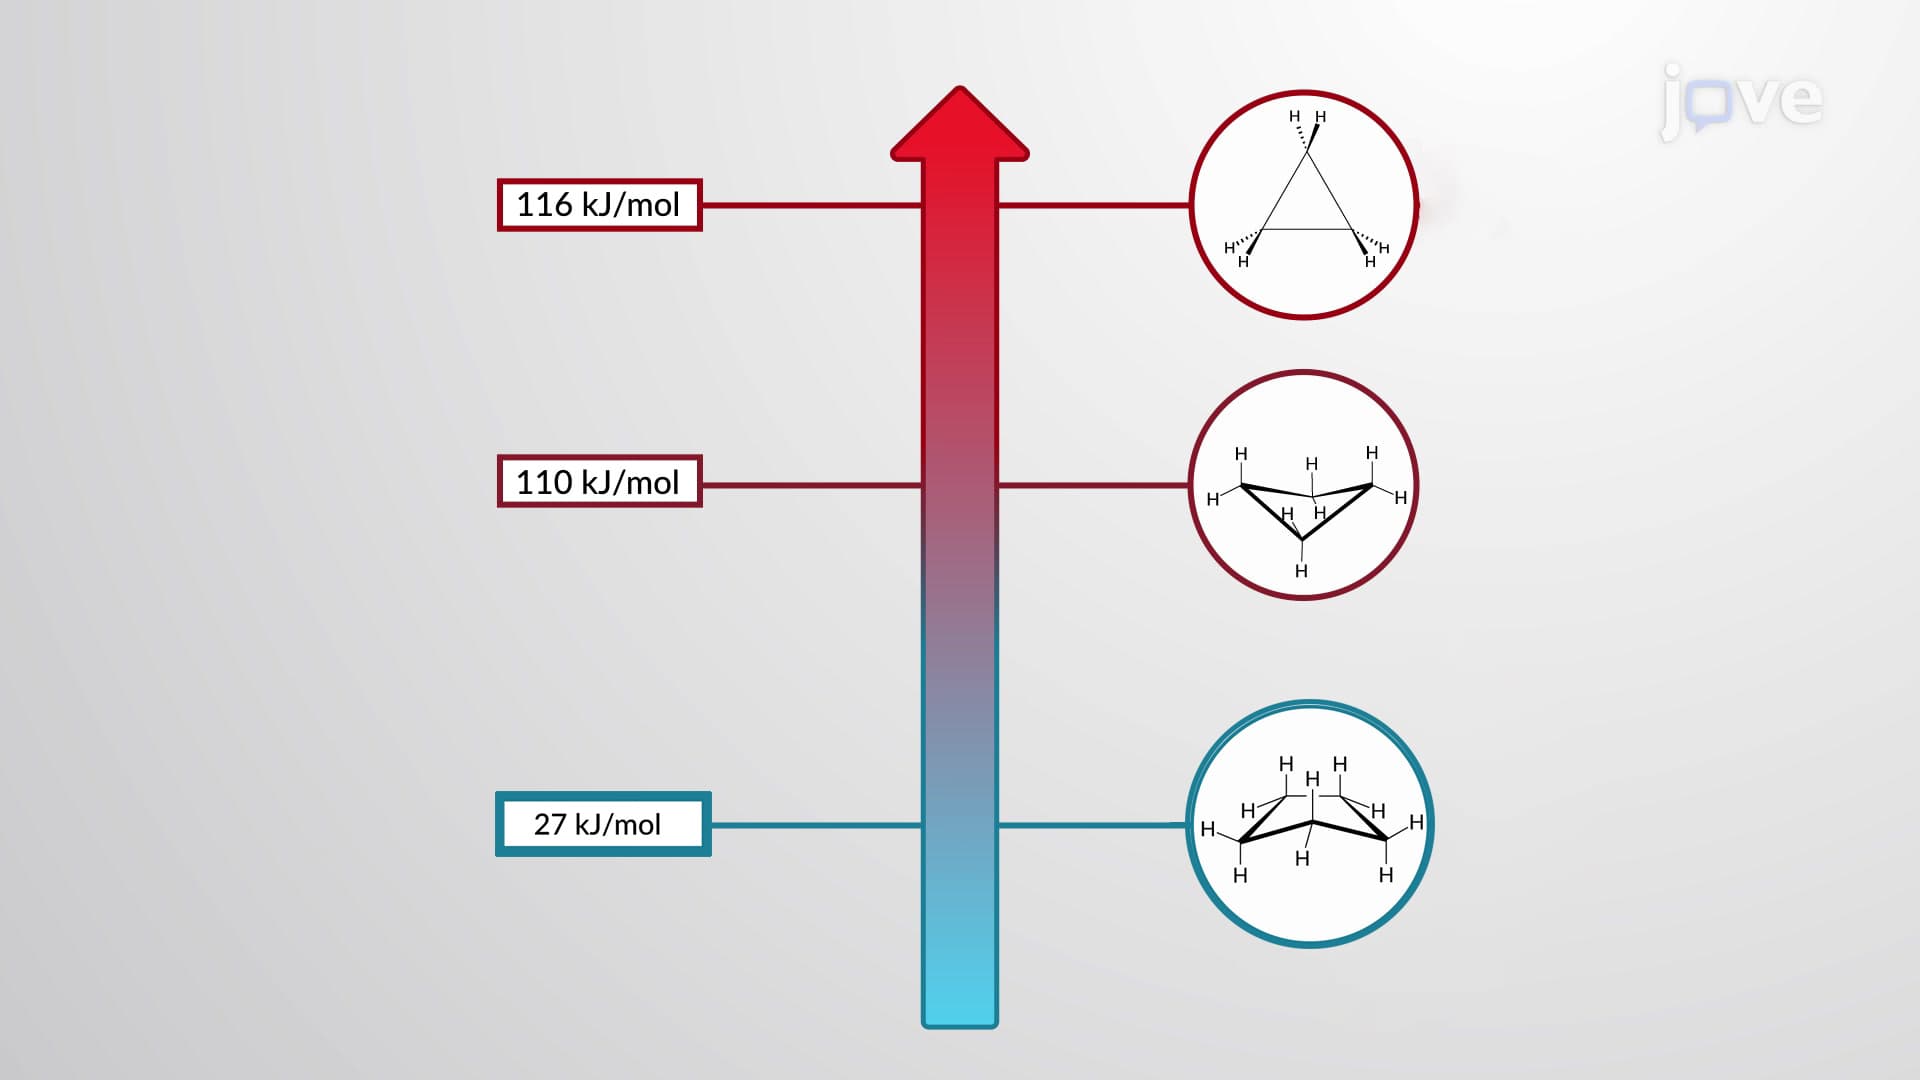
Conformations of Cycloalkanes
 02:37
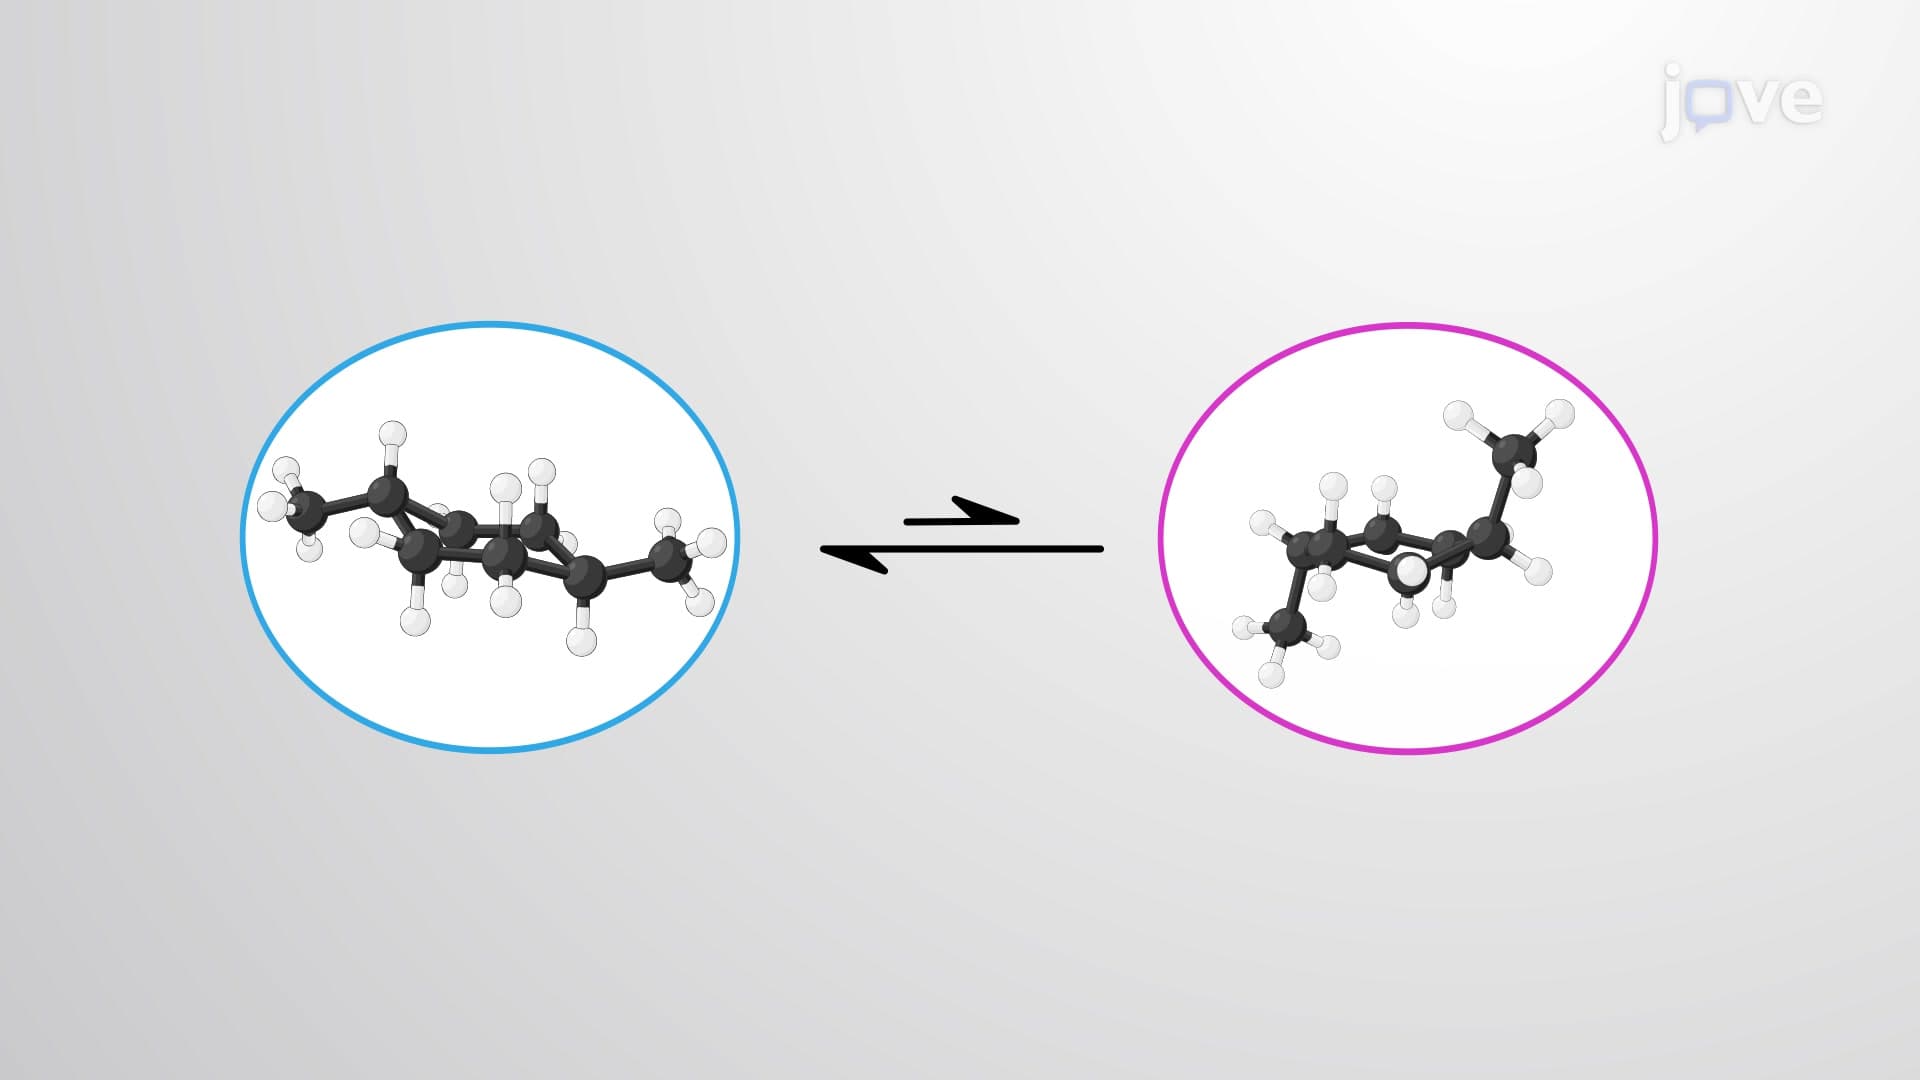
Disubstituted Cyclohexanes: cis-trans Isomerism
In cyclohexane, the substituents can occupy different positions generating distinct isomers....