Related Experiment Video
Updated: Aug 18, 2026

The Murine Choline-Deficient, Ethionine-Supplemented (CDE) Diet Model of Chronic Liver Injury
Published on: October 21, 2017
[Acute encephalopathy and recurrent hepatic steatosis with normal long and medium chain fatty acyl-CoA-dehydrogenase

Insights
A boy experienced recurrent encephalopathy and liver steatosis, suggesting a metabolic disorder affecting lipid metabolism. Despite normal tests for fatty acid oxidation, autopsy revealed significant steatosis, pointing to an unknown metabolic defect.
Area of Science:
- Biochemistry
- Pediatric Medicine
- Metabolic Disorders
Background:
- Investigating a rare pediatric case of recurrent acute encephalopathy and hepatic steatosis.
- Focusing on metabolic pathways, particularly lipid metabolism, as a potential cause.
Observation:
- Patient presented with recurrent episodes of acute encephalopathy and hepatic steatosis from 13 months to 40 months of age.
- Intermittent normal clinical status between episodes.
- Biologic findings included labile hypoglycemia, hyperammonemia without correlation to neurologic symptoms, and failure to produce ketones during fasting.
Findings:
- Normal plasma carnitine levels and normal long/medium-chain acyl-CoA dehydrogenase activity in fibroblasts.
- Autopsy revealed marked hepatic and renal tubular steatosis and foamy histiocytes in bone marrow.
- Absence of organic aciduria during fasting.
Implications:
- Suggests a potential novel defect in lipid metabolism not detectable by standard biochemical assays.
- Highlights the importance of considering metabolic derangements in unexplained pediatric encephalopathy and steatosis.
- Further research needed to identify the specific metabolic pathway affected.
Abstract:
A boy suffering from recurrent episodes of acute encephalopathy and hepatic steatosis died at 40 months of age. The symptoms started when he was 13 months old and he appeared completely normal in the intervals. Pertinent biologic findings were as follows: slight labile hypoglycemia and hyperammonemia having no direct correlation with neurologic derangement, no elevation of ammonia levels in loading tests, complete failure to generate ketones and the absence of organic aciduria during a fast, normal plasma carnitine levels and normal activity of long and medium chain acyl-CoA-dehydrogenase in skin fibroblasts. Pertinent autopsy findings were marked steatosis of liver and renal tubular cells with many foamy histiocytes in bone marrow. An error in metabolic pathways, particularly a derangement in lipid metabolism, was considered.
Related Concept Videos
 01:28
Overview of Fatty Acid Metabolism
Fatty acids are catabolized in a process called beta-oxidation, which takes place in the matrix of the mitochondria and converts their fatty acid chains into two-carbon units of acetyl groups. The acetyl...
 01:24
Chronic Pancreatitis I: Introduction
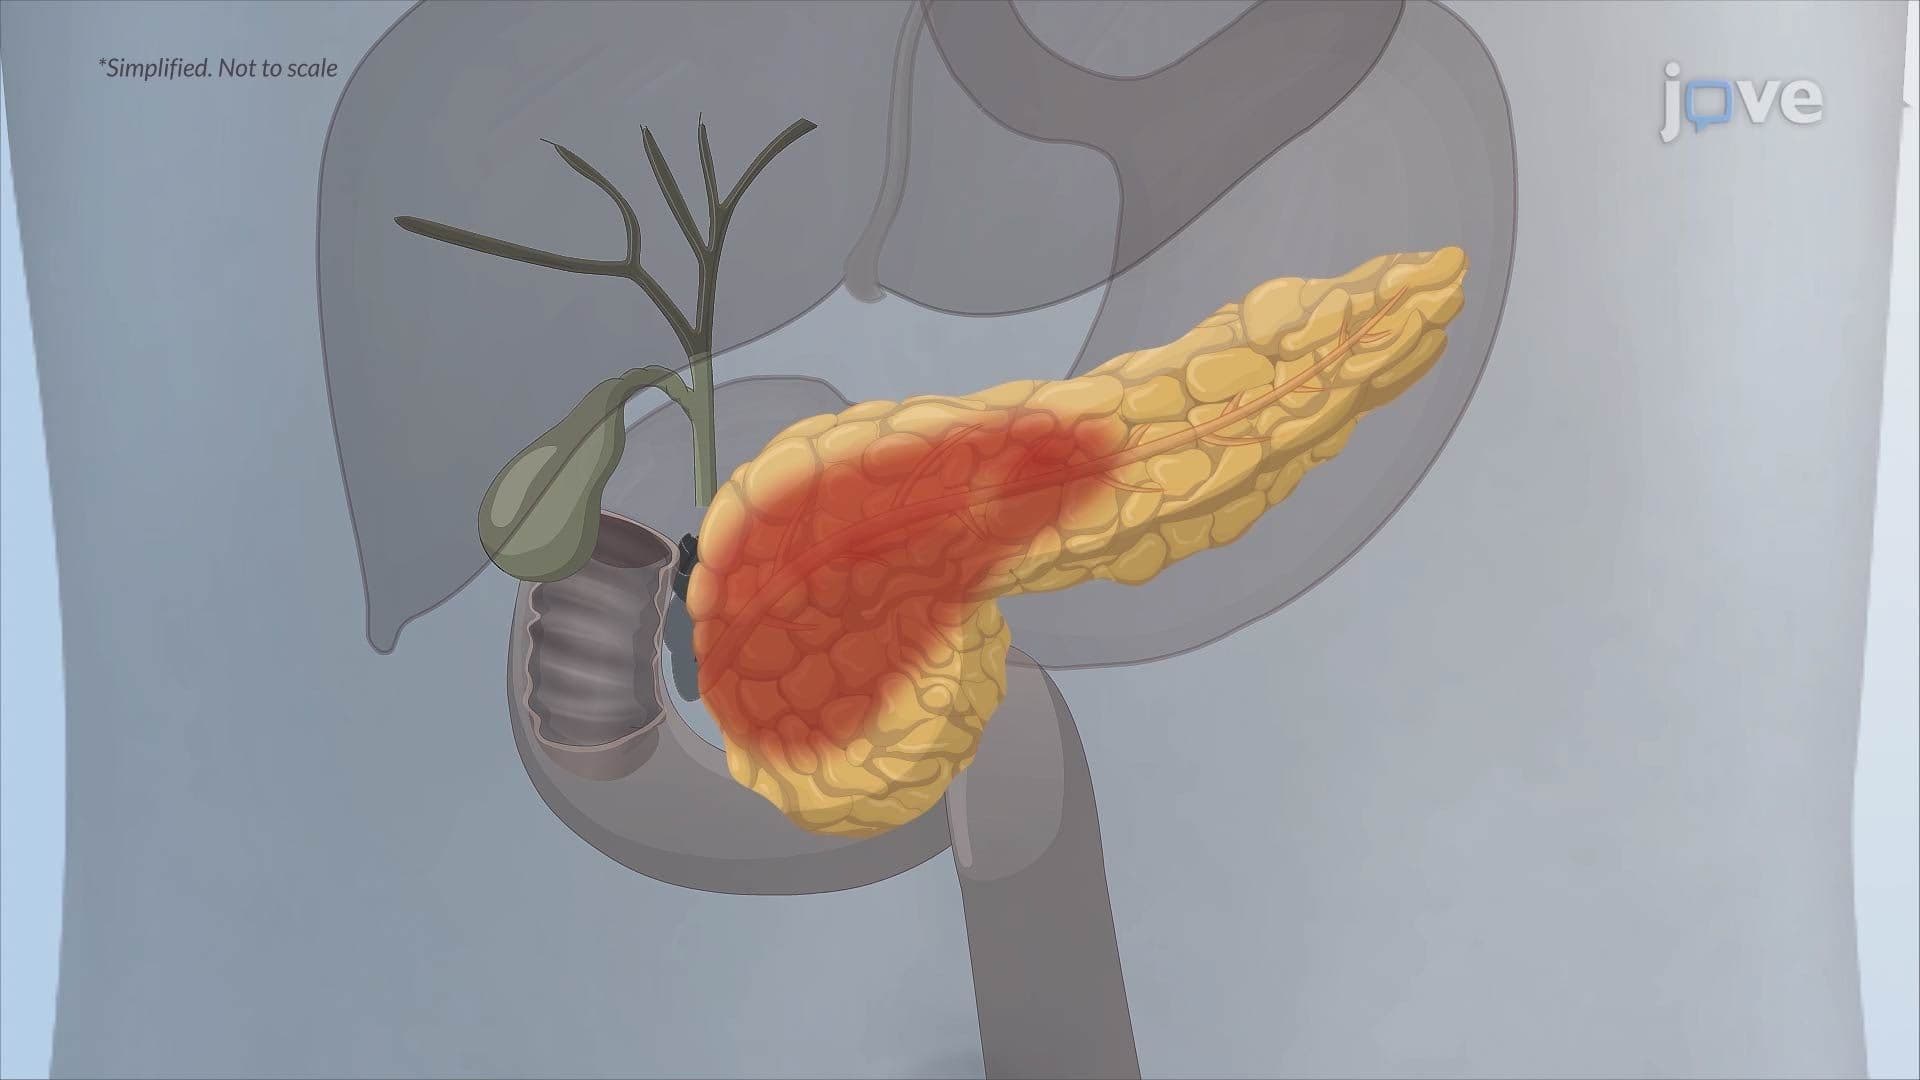
Pancreatitis is the inflammation of the pancreas, which occurs when the immune system becomes active and causes swelling, pain, and disruptions in organ function. Pancreatitis can manifest as either an acute or chronic condition.
Acute pancreatitis arises suddenly and lasts for a brief duration, while chronic pancreatitis is a long-term affliction...
 01:29
Hepatic Encephalopathy
 01:21
Acute Pancreatitis II: Pathophysiology
 01:25
Chronic Pancreatitis I: Introduction
 01:21
Chronic Pancreatitis II: Pathophysiology