Related Experiment Video
Updated: Aug 26, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Improving the accuracy of GIPAW chemical shielding calculations with cluster and fragment corrections
Joshua D Hartman1, James K Harper2
1Department of Chemistry, University of California, Riverside, Riverside, CA, United States.

Abstract:
Ab initio methods for predicting NMR parameters in the solid state are an essential tool for assigning experimental spectra and play an increasingly important role in structural characterizations. Recently, a molecular correction (MC) technique has been developed which combines the strengths of plane-wave methods (GIPAW) with single molecule calculations employing Gaussian basis sets. The GIPAW + MC method relies on a periodic calculation performed at a lower level of theory to model the crystalline environment. The GIPAW result is then corrected using a single molecule calculation performed at a higher level of theory. The success of the GIPAW + MC method in predicting a range of NMR parameters is a result of the highly local character of the tensors underlying the NMR observable. However, in applications involving strong intermolecular interactions we find that expanding the region treated at the higher level of theory more accurately captures local many-body contributions to the N15 NMR chemical shielding (CS) tensor. We propose alternative corrections to GIPAW which capture interactions between adjacent molecules at a higher level of theory using either fragment or cluster-based calculations. Benchmark calculations performed on N15 and C13 data sets show that these advanced GIPAW-corrected calculations improve the accuracy of chemical shielding tensor predictions relative to existing methods. Specifically, cluster-based N15 corrections show a 24% and 17% reduction in RMS error relative to GIPAW and GIPAW + MC calculations, respectively. Comparing the benchmark data sets using multiple computational models demonstrates that N15 CS tensor calculations are significantly more sensitive to intermolecular interactions relative to C13. However, fragment and cluster-based corrections that include direct hydrogen bond partners are sufficient for optimizing the accuracy of GIPAW-corrected methods. Finally, GIPAW-corrected methods are applied to the particularly challenging NMR spectral assignment of guanosine dihydrate which contains two guanosine molecules in the asymmetric unit.
Related Concept Videos
 01:20
Inductively Coupled Plasma-Mass Spectrometry (ICP-MS): Interferences
 01:27
π Electron Effects on Chemical Shift: Overview
 01:25
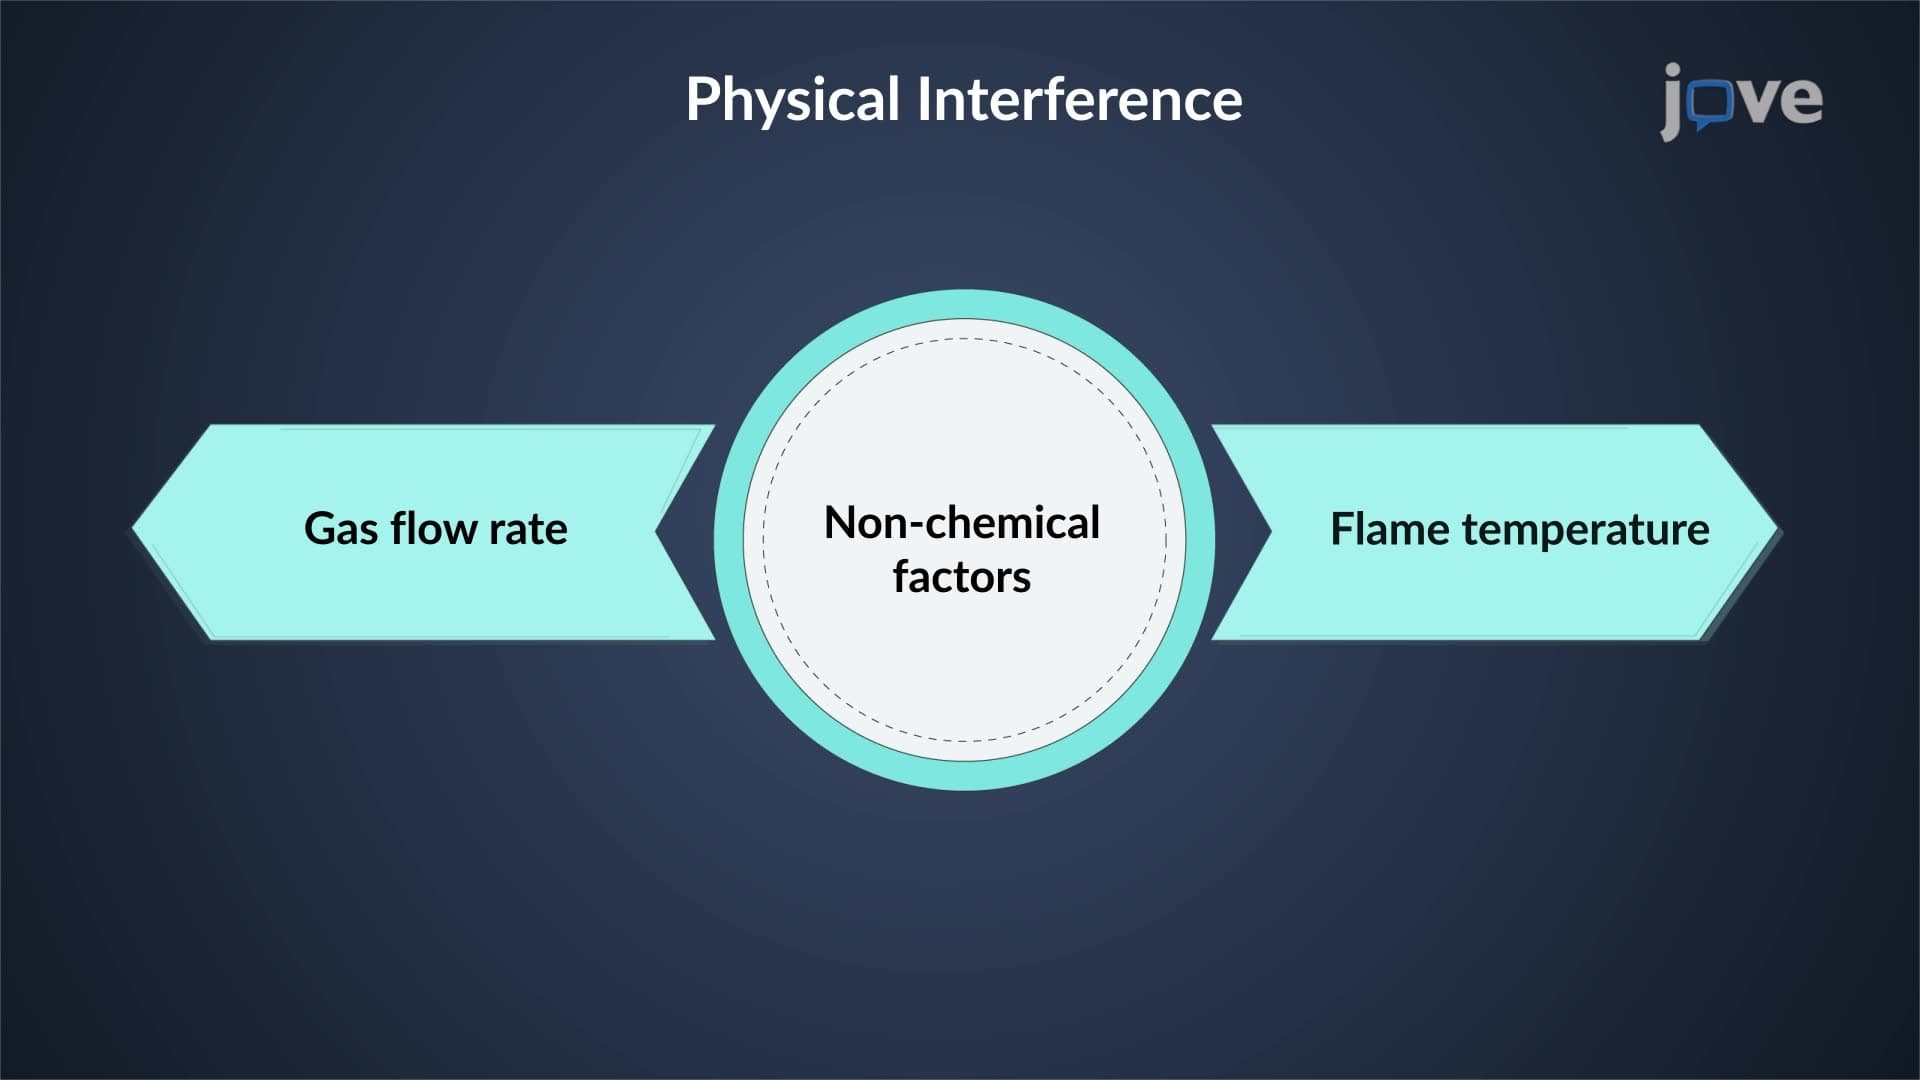
Atomic Absorption Spectroscopy: Interference
Spectral interference occurs when signals from other elements or molecules overlap with the analyte signal, falsely elevating or masking the analyte's absorbance. This interference can be corrected using Zeeman,...