Related Experiment Video
Updated: Aug 24, 2025


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Free Energy Calculations Using the Movable Type Method with Molecular Dynamics Driven Protein-Ligand Sampling
Wenlang Liu1, Zhenhao Liu1, Hao Liu2
1School of Chemistry, Chemical Engineering and Life Science, Wuhan University of Technology, 122 Luoshi Road, Wuhan430070, PR China.

Abstract:
Fast and accurate biomolecular free energy estimation has been a significant interest for decades, and with recent advances in computer hardware, interest in new method development in this field has even grown. Thorough configurational state sampling using molecular dynamics (MD) simulations has long been applied to the estimation of the free energy change corresponding to the receptor-ligand complexing process. However, performing large-scale simulation is still a computational burden for the high-throughput hit screening. Among molecular modeling tools, docking and scoring methods are widely used during the early stages of the drug discovery process in that they can rapidly generate discrete receptor-ligand binding modes and their individual binding affinities. Unfortunately, the lack of thorough conformational sampling in docking and scoring protocols leads to difficulty discovering global minimum binding modes on a complicated energy landscape. The Movable Type (MT) method is a novel absolute binding free energy approach which has demonstrated itself to be robust across a wide range of targets and ligands. Traditionally, the MT method is used with protein-ligand binding modes generated with rigid-receptor or flexible-receptor (induced fit) docking protocols; however, these protocols are by their nature less likely to be effective with more highly flexible targets or with those situations in which binding involves multiple step pathways. In these situations, more thorough samplings are required to better explain the free energy of binding. Therefore, to explore the prediction capability and computational efficiency of the MT method when using more thorough protein-ligand conformational sampling protocols, in the present work, we introduced a series of binding mode modeling protocols ranging from conventional docking routines to single-trajectory conventional molecular dynamics (cMD) and parallel Monte Carlo molecular dynamics (MCMD). Through validation against several structurally and mechanistically diverse protein-ligand test sets, we explore the performance of the MT method as a virtual screening tool to work with the docking protocols and as an MD simulation-based binding free energy tool.
More Related Videos
 05:56
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
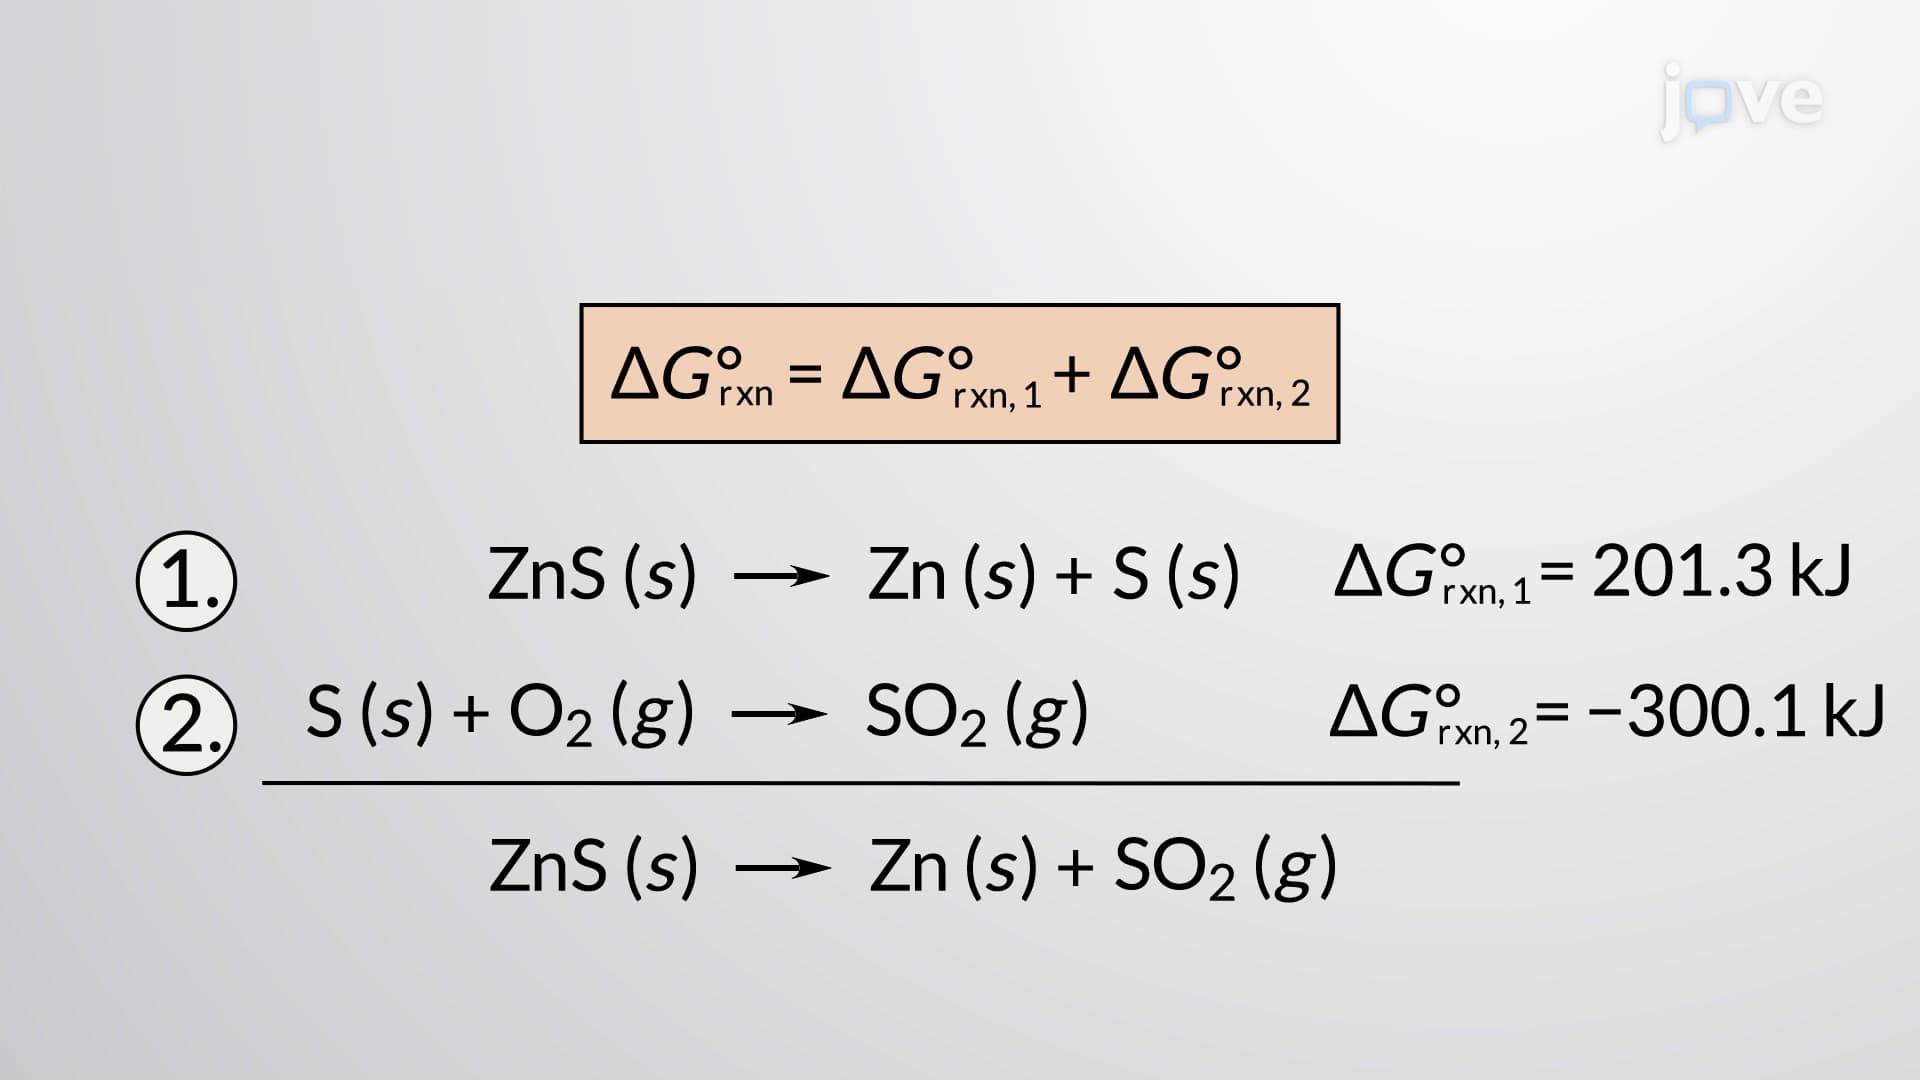 02:49
Calculating Standard Free Energy Changes
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...