Comparison of Density-Matrix Corrections to Density Functional Theory
Related Experiment Video
Updated: Aug 24, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Daniel Gibney1, Jan-Niklas Boyn1, David A Mazziotti1
1Department of Chemistry and The James Franck Institute, The University of Chicago, Chicago, Illinois 60637 United States.

Abstract:
Density functional theory (DFT), one of the most widely utilized methods available to computational chemistry, fails to describe systems with statically correlated electrons. To address this shortcoming, in previous work, we transformed DFT into a one-electron reduced density matrix theory (1-RDMFT) via the inclusion of a quadratic one-electron reduced density matrix (1-RDM) correction. Here, we combine our 1-RDMFT approach with different DFT functionals as well as Hartree-Fock to elucidate the method's dependence on the underlying functional selection. Furthermore, we generalize the information density matrix functional theory (iDMFT), recently developed as a correction to the Hartree-Fock method, by incorporating density functionals in place of the Hartree-Fock functional. We relate iDMFT mathematically to our approach and benchmark the two with a common set of functionals and systems.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:40
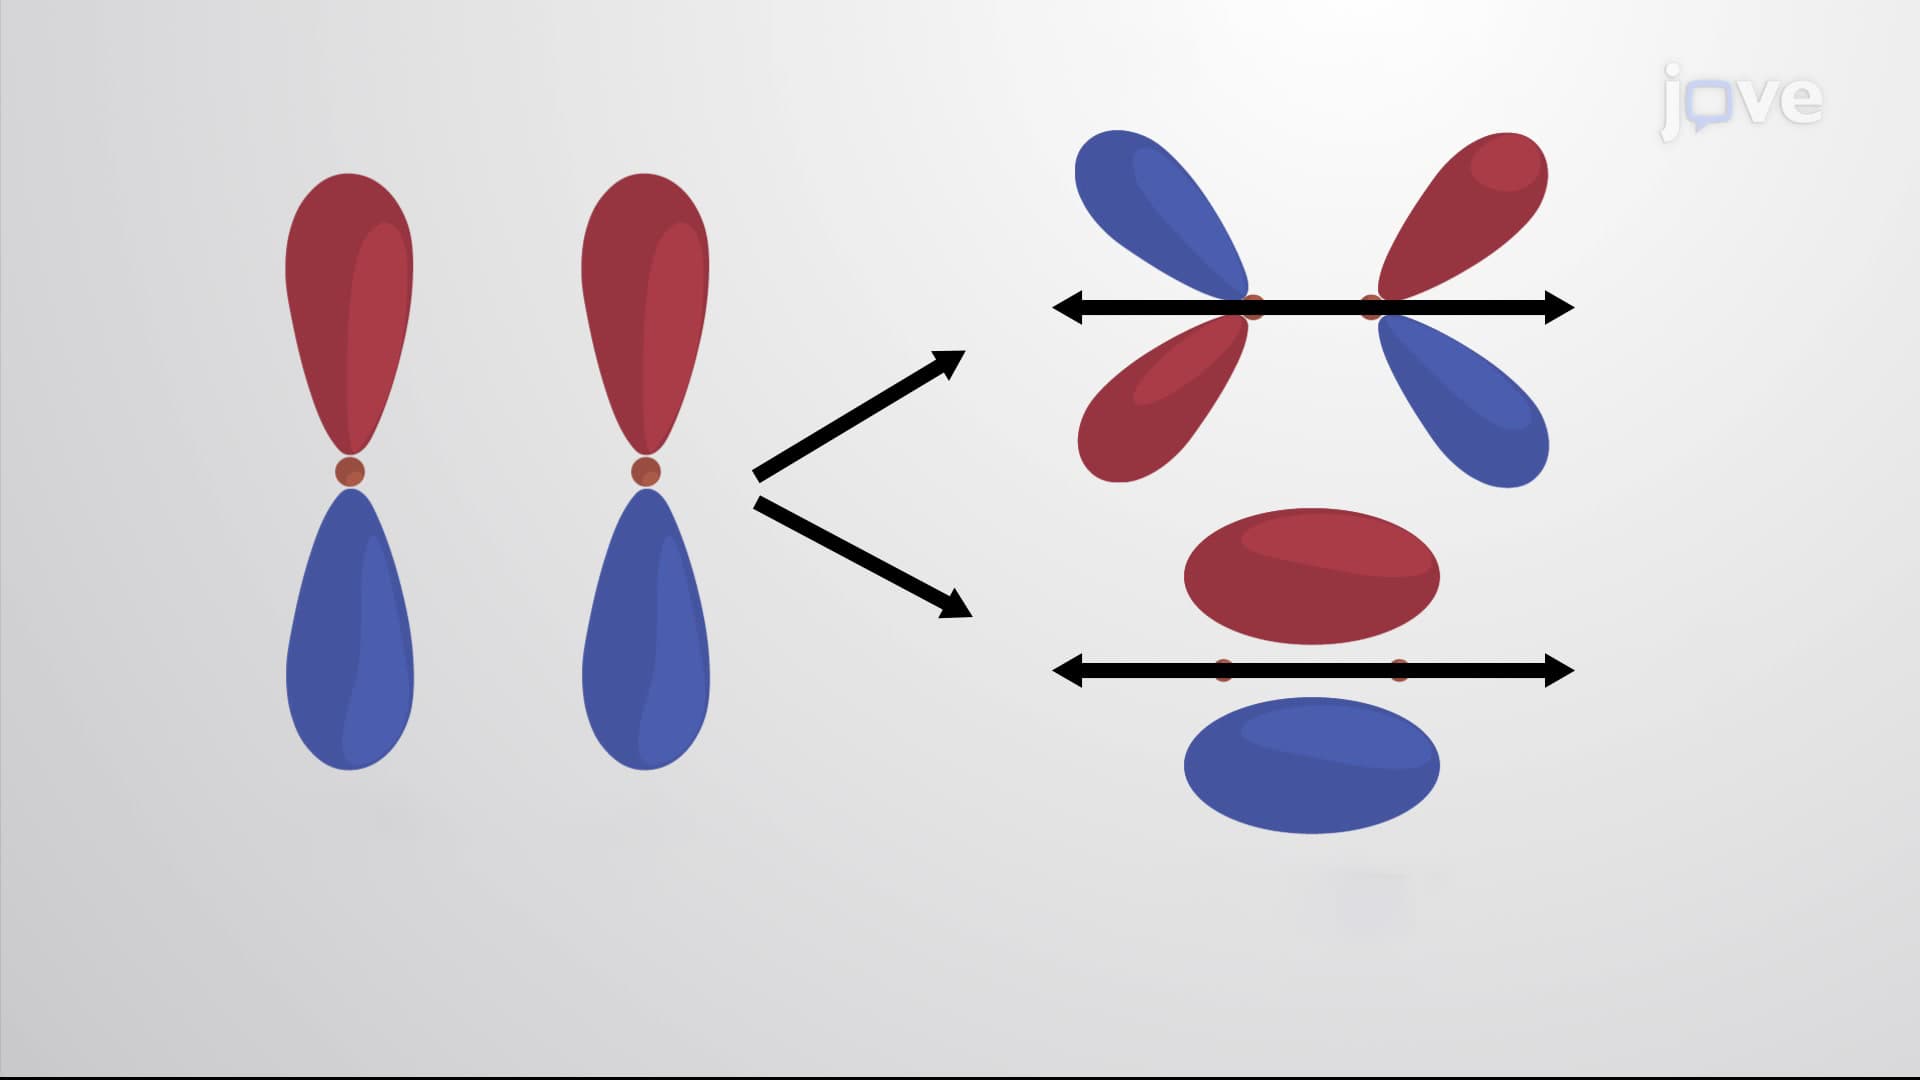
MO Theory and Covalent Bonding
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:51
Molecular Orbital Theory II
 00:56
Density
 01:10
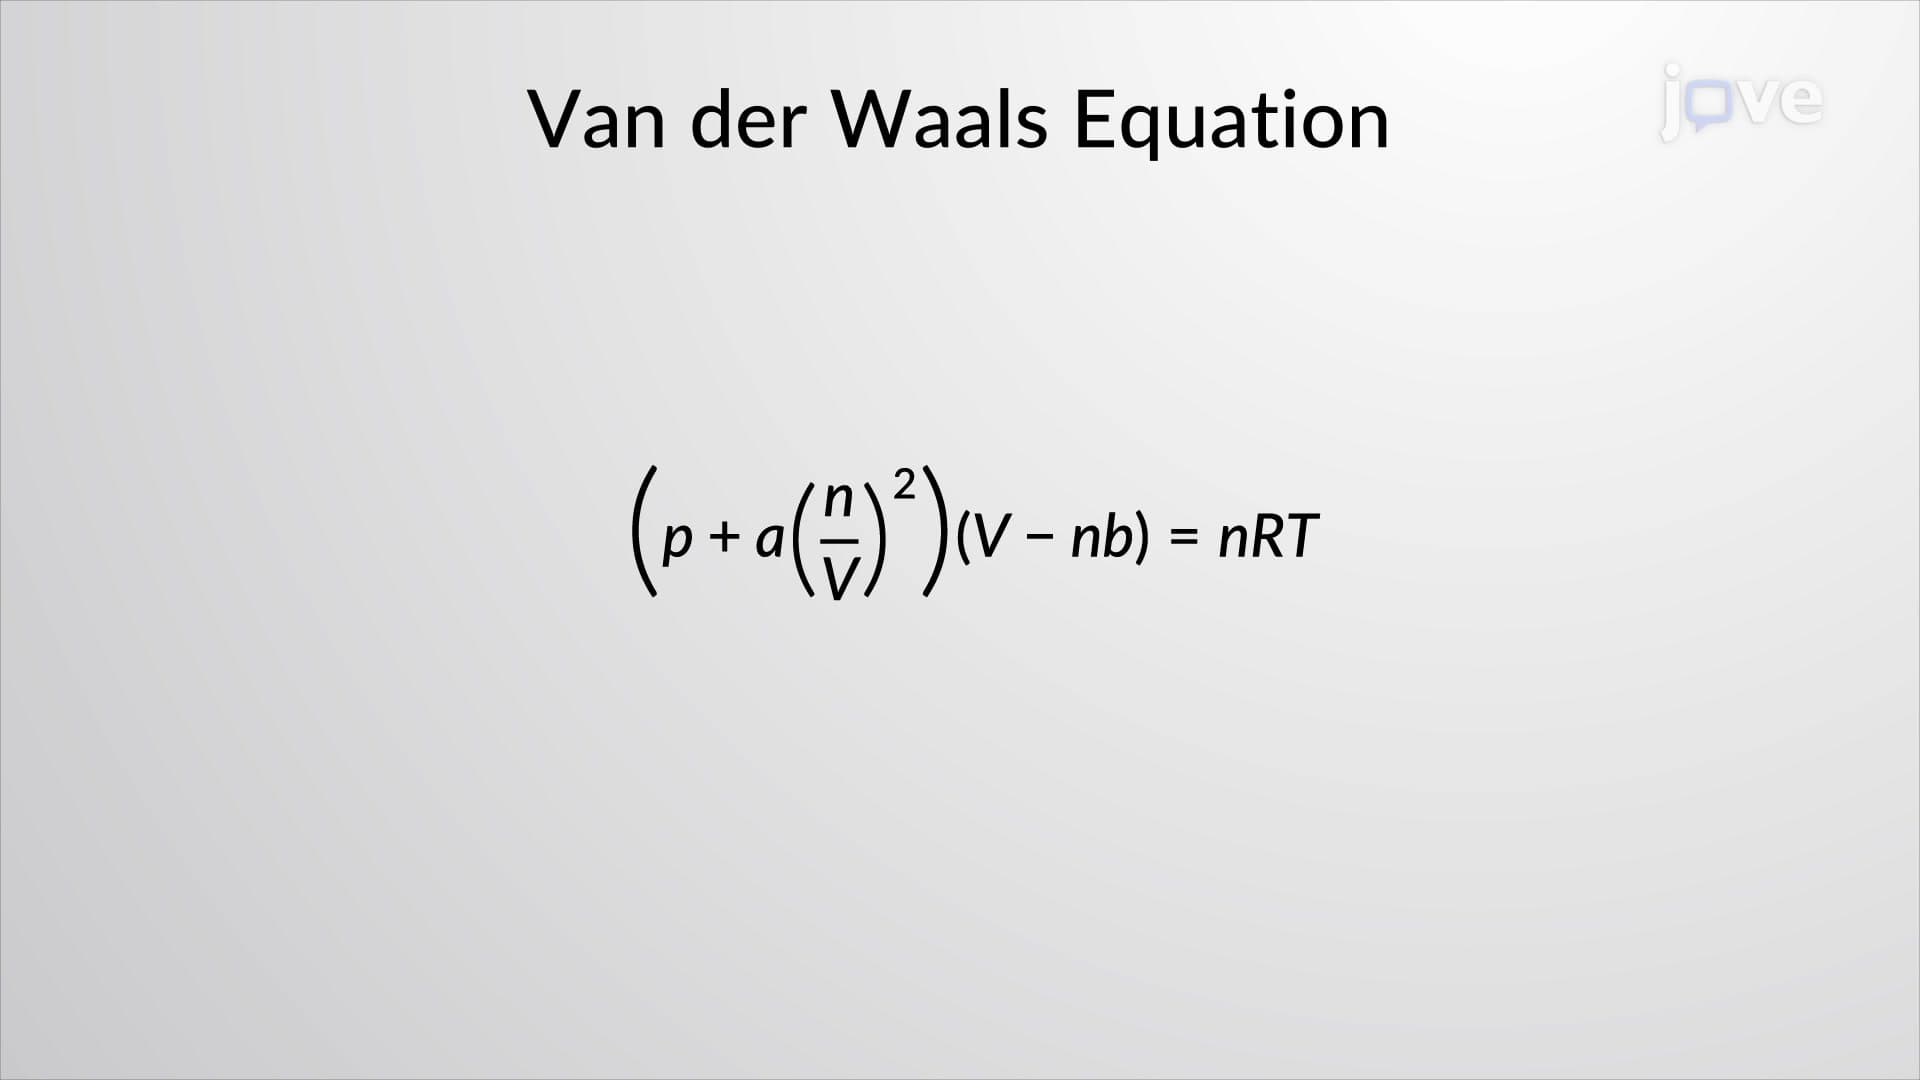
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...