In Silico molecular docking and dynamic analysis of natural compounds against major non-structural proteins of SARS-COV-2
Related Experiment Video
Updated: Aug 23, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Muneeb U Rehman1, Aarif Ali2, Ruhban Ansar2
1Department of Clinical Pharmacy, College of Pharmacy, King Saud University, Riyadh, Saudi Arabia.

Abstract:
COVID-19 has infected millions and significantly affected the global economy and healthcare systems. Despite continuous lockdowns, symptomatic management with currently available medications, and numerous vaccination drives, it is still far more difficult to control. Against COVID-19 infection, the pressure to develop vaccines and drugs has led to using some currently available medications like remdesivir, azithromycin, hydroxychloroquine and ritonavir. Understanding the importance and potential of harmless molecules to tackle SARS-COV-2, we designed the present study to identify potential natural phytocompounds. In the present study, we docked natural compounds and standard drugs against SARS-COV-2 proteins: papain-like protease, main protease and helicase. ADME/T and ProTox-II analyses were used to determine the toxicity of phytocompounds and drugs. The docking analysis revealed that podophyllotoxin gave the highest binding affinity scores of -8.1, -7.1 and -7.4 kcal/mol against PLpro, Mpro and helicase, respectively. Among the control drugs, doxycycline hydrochloride showed the highest binding affinity of -10.5, -8.4 and -8.8 kcal/mol against PLpro, Mpro and helicase. The results of this study revealed that podophyllotoxin and doxycycline hydrochloride could be promising inhibitors against SARS-Cov-2. Molecular dynamic simulations were executed for the best docked (PLpro-podophyllotoxin) complex, and the results displayed stable conformation and convergence. Energy plot results predicted a global minima average energy of -95 kcal/mol and indicated podophyllotoxin's role in stabilizing protein and making it compact and complex. FarPPI server used MM/GBSA approach to determine free binding affinity, and helicase-gallic acid complex showed the highest affinity, respectively. Therefore, it can be concluded that there is still a need for in vitro and in vivo studies to support further and validate these findings and validate these findings.Communicated by Ramaswamy H. Sarma.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 02:50
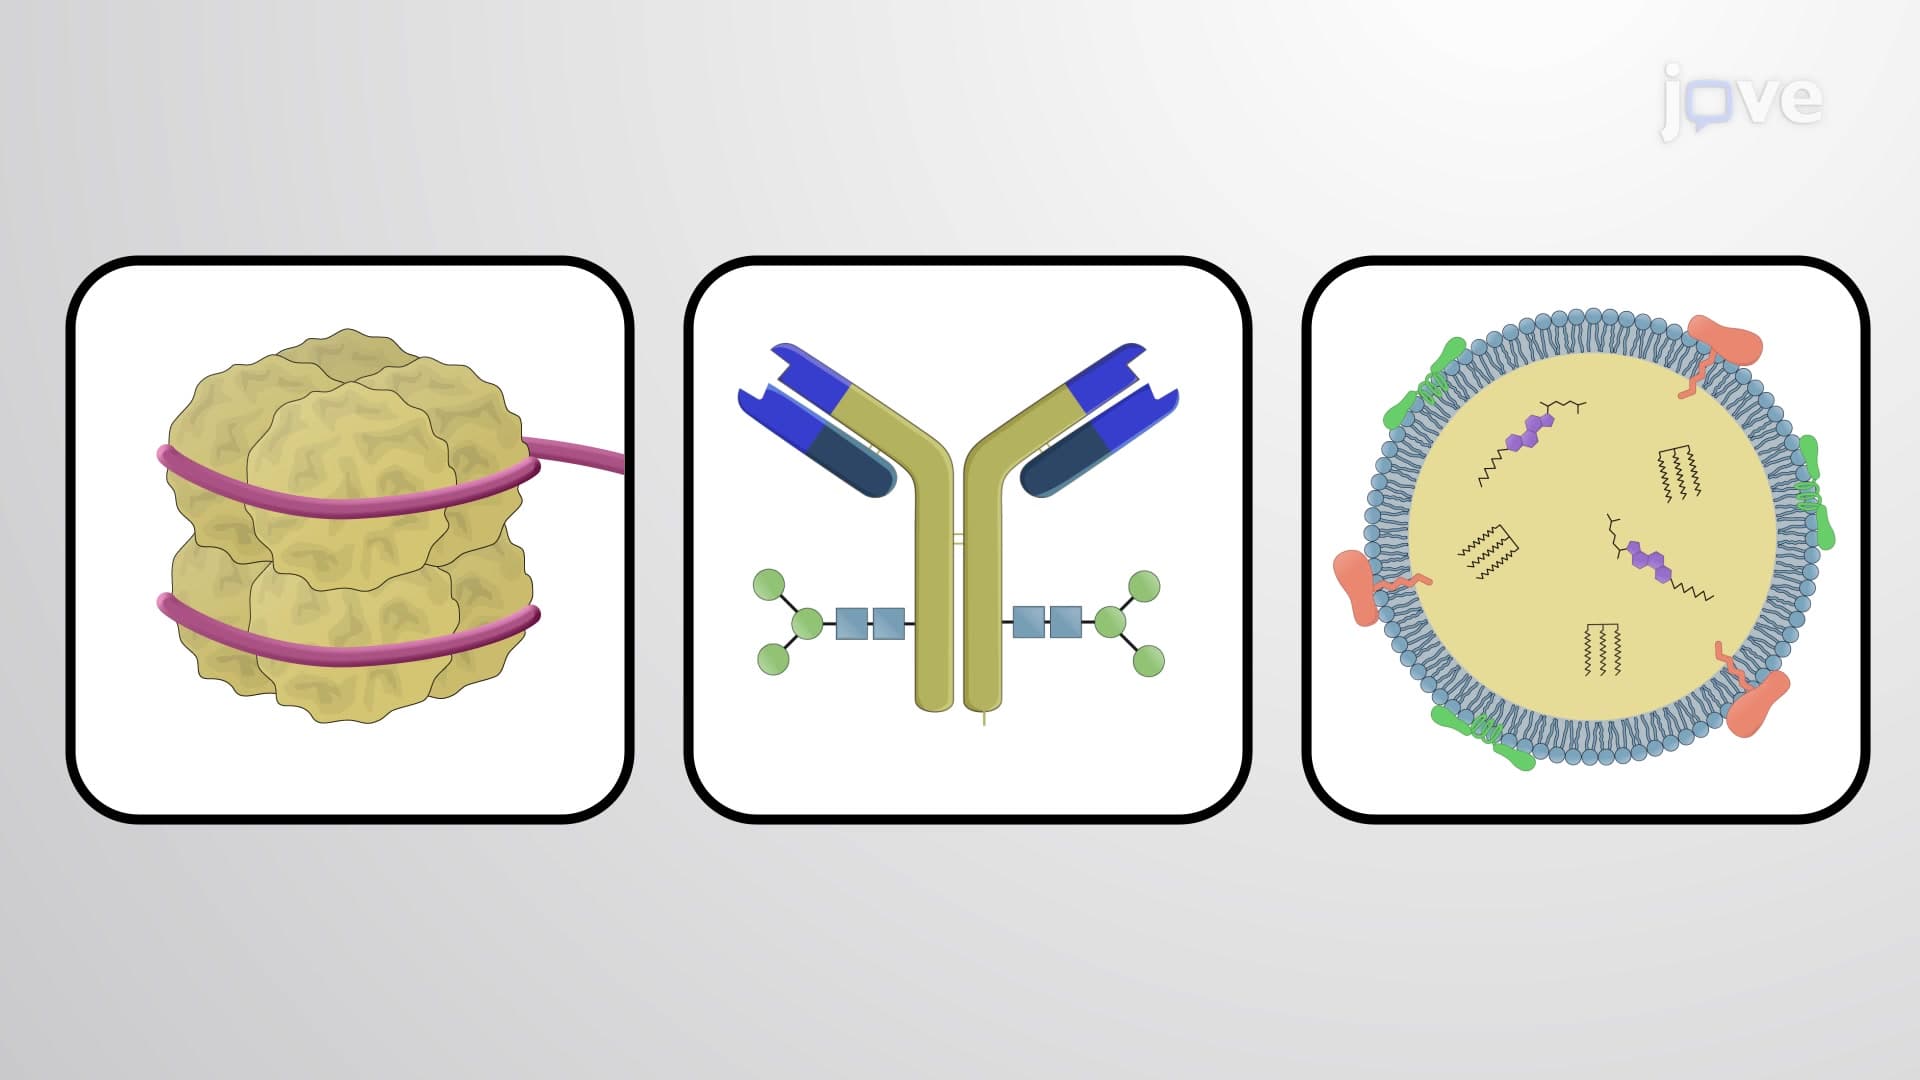
Conjugated Proteins
Nucleoproteins are protein complexes that contain nucleic acids, categorized as deoxyribonucleoproteins (DNPs) or ribonucleoproteins (RNPs) respectively. The nucleosome is a typical example of a DNP where nuclear DNA is associated with histone proteins. The major antigen for the Covid-19 virus SARS-CoV is an RNP that is critical...
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...