Related Experiment Video
Updated: Aug 22, 2025
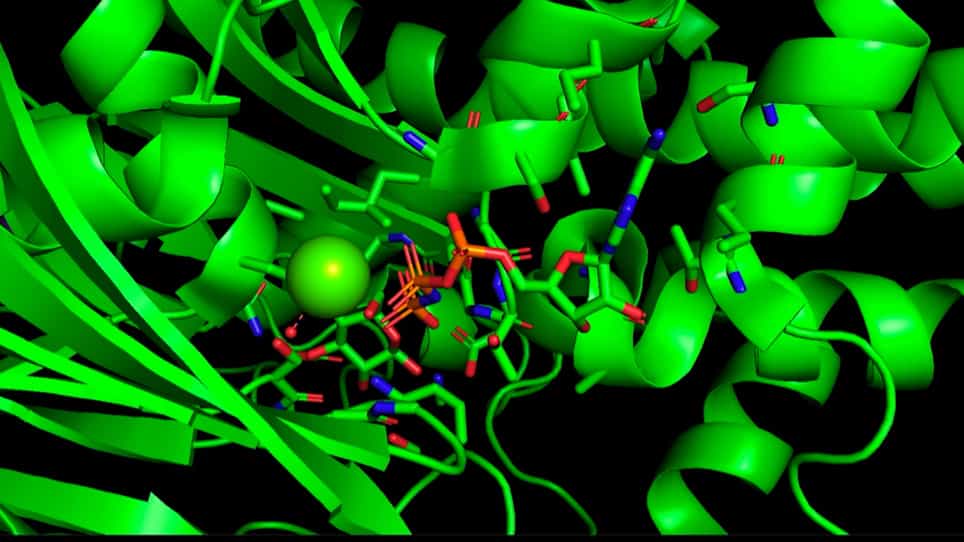

Modeling an Enzyme Active Site using Molecular Visualization Freeware
Published on: December 25, 2021
Molecular Modeling of Allosteric Site of Isoform-Specific Inhibition of the Peroxisome Proliferator-Activated
Suliman Almahmoud1, Haizhen A Zhong2
1Department of Medicinal Chemistry and Pharmacognosy, College of Pharmacy, Qassim University, Buraidah 51452, Saudi Arabia.

Abstract:
The peroxisome proliferator-activated receptor gamma (PPARγ) is a nuclear receptor and controls a number of gene expressions. The ligand binding domain (LBD) of PPARγ is large and involves two binding sites: orthosteric and allosteric binding sites. Increased evidence has shown that PPARγ is an oncogene and thus the PPARγ antagonists have potential as anticancer agents. In this paper, we use Glide Dock approach to determine which binding site, orthosteric or allosteric, would be a preferred pocket for PPARγ antagonist binding, though antidiabetic drugs such as thiazolidinediones (TZDs) bind to the orthosteric site. The Glide Dock results show that the binding of PPARγ antagonists at the allosteric site yielded results that were much closer to the experimental data than at the orthosteric site. The PPARγ antagonists seem to selectively bind to residues Lys265, Ser342 and Arg288 at the allosteric binding site, whereas PPARγ agonists would selectively bind to residues Leu228, Phe363, and His449, though Phe282 and Lys367 may also play a role for agonist binding at the orthosteric binding pocket. This finding will provide new perspectives in the design and optimization of selective and potent PPARγ antagonists or agonists.
Insights
Peroxisome proliferator-activated receptor gamma (PPARγ) antagonists show potential as anticancer agents. Computational docking suggests allosteric sites are preferred for PPARγ antagonist binding, offering new drug design perspectives.
Area of Science:
- Molecular Biology
- Pharmacology
- Computational Chemistry
Background:
- Peroxisome proliferator-activated receptor gamma (PPARγ) is a nuclear receptor regulating gene expression.
- PPARγ's ligand-binding domain (LBD) features distinct orthosteric and allosteric sites.
- Emerging evidence implicates PPARγ as an oncogene, highlighting antagonists as potential anticancer therapeutics.
Purpose of the Study:
- To computationally determine the preferred binding site (orthosteric vs. allosteric) for PPARγ antagonists.
- To investigate the potential of PPARγ antagonists as anticancer agents.
- To provide insights for designing selective PPARγ modulators.
Main Methods:
- Utilized the Glide Dock computational approach for molecular docking simulations.
- Analyzed binding interactions at both orthosteric and allosteric sites of PPARγ.
- Compared docking results with existing experimental data.
Main Results:
- PPARγ antagonist binding at the allosteric site showed better correlation with experimental data compared to the orthosteric site.
- Identified key residues (Lys265, Ser342, Arg288) involved in allosteric antagonist binding.
- Differentiated binding residues for agonists (Leu228, Phe363, His449) at the orthosteric site.
Conclusions:
- The allosteric binding site is a more favorable target for PPARγ antagonist design.
- Understanding selective binding residues can guide the development of potent and specific PPARγ antagonists and agonists.
- This study offers new perspectives for anticancer drug discovery targeting PPARγ.
Related Concept Videos
 01:08
Allosteric Regulation
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...
 01:58
Cooperative Allosteric Transitions
 00:49
Ligand Binding and Linkage
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:09
GPCRs Regulate Adenylyl Cylase Activity