Related Experiment Video
Updated: Aug 19, 2025

Intracellular Phosphoflow Cytometry of Acute Myeloid Leukemia Patient-Derived Xenotransplants
Published on: June 6, 2025
Disrupting PTPRJ transmembrane-mediated oligomerization counteracts oncogenic receptor tyrosine kinase FLT3 ITD
Marie Schwarz1, Sophie Rizzo2, Walter Espinoza Paz2
1Institute for Molecular Cell Biology, CMB - Center for Molecular Biomedicine, University Hospital Jena, Jena, Germany.

Abstract:
The receptor protein tyrosine phosphatase (RPTP) PTPRJ (also known as DEP-1) has been identified as a negative regulator of the receptor tyrosine kinase FLT3 signalling in vitro. The inactivation of the PTPRJ gene in mice expressing the constitutively active, oncogenic receptor tyrosine kinase FLT3 ITD aggravated known features of leukaemogenesis, revealing PTPRJ's antagonistic role. FLT3 ITD mutations resulting in constitutively kinase activity and cell transformation frequently occur in patients with acute myeloid leukaemia (AML). Thus, in situ activation of PTPRJ could be used to abrogate oncogenic FLT3 signalling. The activity of PTPRJ is suppressed by homodimerization, which is mediated by transmembrane domain (TMD) interactions. Specific Glycine-to-Leucine mutations in the TMD disrupt oligomerization and inhibit the Epidermal Growth Factor Receptor (EGFR) and EGFR-driven cancer cell phenotypes. To study the effects of PTPRJ TMD mutant proteins on FLT3 ITD activity in cell lines, endogenous PTPRJ was inactivated and replaced by stable expression of PTPRJ TMD mutants. Autophosphorylation of wild-type and ITD-mutated FLT3 was diminished in AML cell lines expressing the PTPRJ TMD mutants compared to wild-type-expressing cells. This was accompanied by reduced FLT3-mediated global protein tyrosine phosphorylation and downstream signalling. Further, PTPRJ TMD mutant proteins impaired the proliferation and in vitro transformation of leukemic cells. Although PTPRJ's TMD mutant proteins showed impaired self-association, the specific phosphatase activity of immunoprecipitated proteins remained unchanged. In conclusion, this study demonstrates that the destabilization of PTPRJ TMD-mediated self-association increases the activity of PTPRJ in situ and impairs FLT3 activity and FLT3-driven cell phenotypes of AML cells. Thus, disrupting the oligomerization of PTPRJ in situ could prove a valuable therapeutic strategy to restrict oncogenic FLT3 activity in leukemic cells.
Insights
Disrupting PTPRJ self-association enhances its activity, inhibiting oncogenic FLT3 signaling and leukemic cell growth in acute myeloid leukemia (AML). This offers a potential therapeutic strategy for AML.
Area of Science:
- Molecular Biology
- Cell Biology
- Cancer Research
Background:
- PTPRJ (DEP-1) negatively regulates FLT3 signaling, acting as a tumor suppressor.
- FLT3 ITD mutations are common in acute myeloid leukemia (AML) and drive leukemogenesis.
- PTPRJ activity is suppressed by homodimerization via its transmembrane domain (TMD).
Purpose of the Study:
- To investigate the therapeutic potential of disrupting PTPRJ TMD-mediated homodimerization.
- To determine the effect of PTPRJ TMD mutants on FLT3 ITD activity in AML cells.
- To assess the impact of PTPRJ TMD mutants on leukemic cell proliferation and transformation.
Main Methods:
- Generated AML cell lines with inactivated endogenous PTPRJ and stable expression of PTPRJ TMD mutants.
- Assessed FLT3 autophosphorylation and downstream signaling in cells expressing PTPRJ TMD mutants.
- Evaluated leukemic cell proliferation and in vitro transformation assays.
Main Results:
- PTPRJ TMD mutants diminished FLT3 autophosphorylation and downstream signaling in AML cells.
- Expression of PTPRJ TMD mutants reduced global protein tyrosine phosphorylation.
- PTPRJ TMD mutant proteins impaired leukemic cell proliferation and in vitro transformation.
- Mutant PTPRJ proteins showed impaired self-association but retained phosphatase activity.
Conclusions:
- Destabilizing PTPRJ TMD-mediated self-association enhances PTPRJ activity in situ.
- This disruption inhibits FLT3 activity and FLT3-driven phenotypes in AML cells.
- Targeting PTPRJ oligomerization is a potential therapeutic strategy for AML.
Related Concept Videos
 01:26
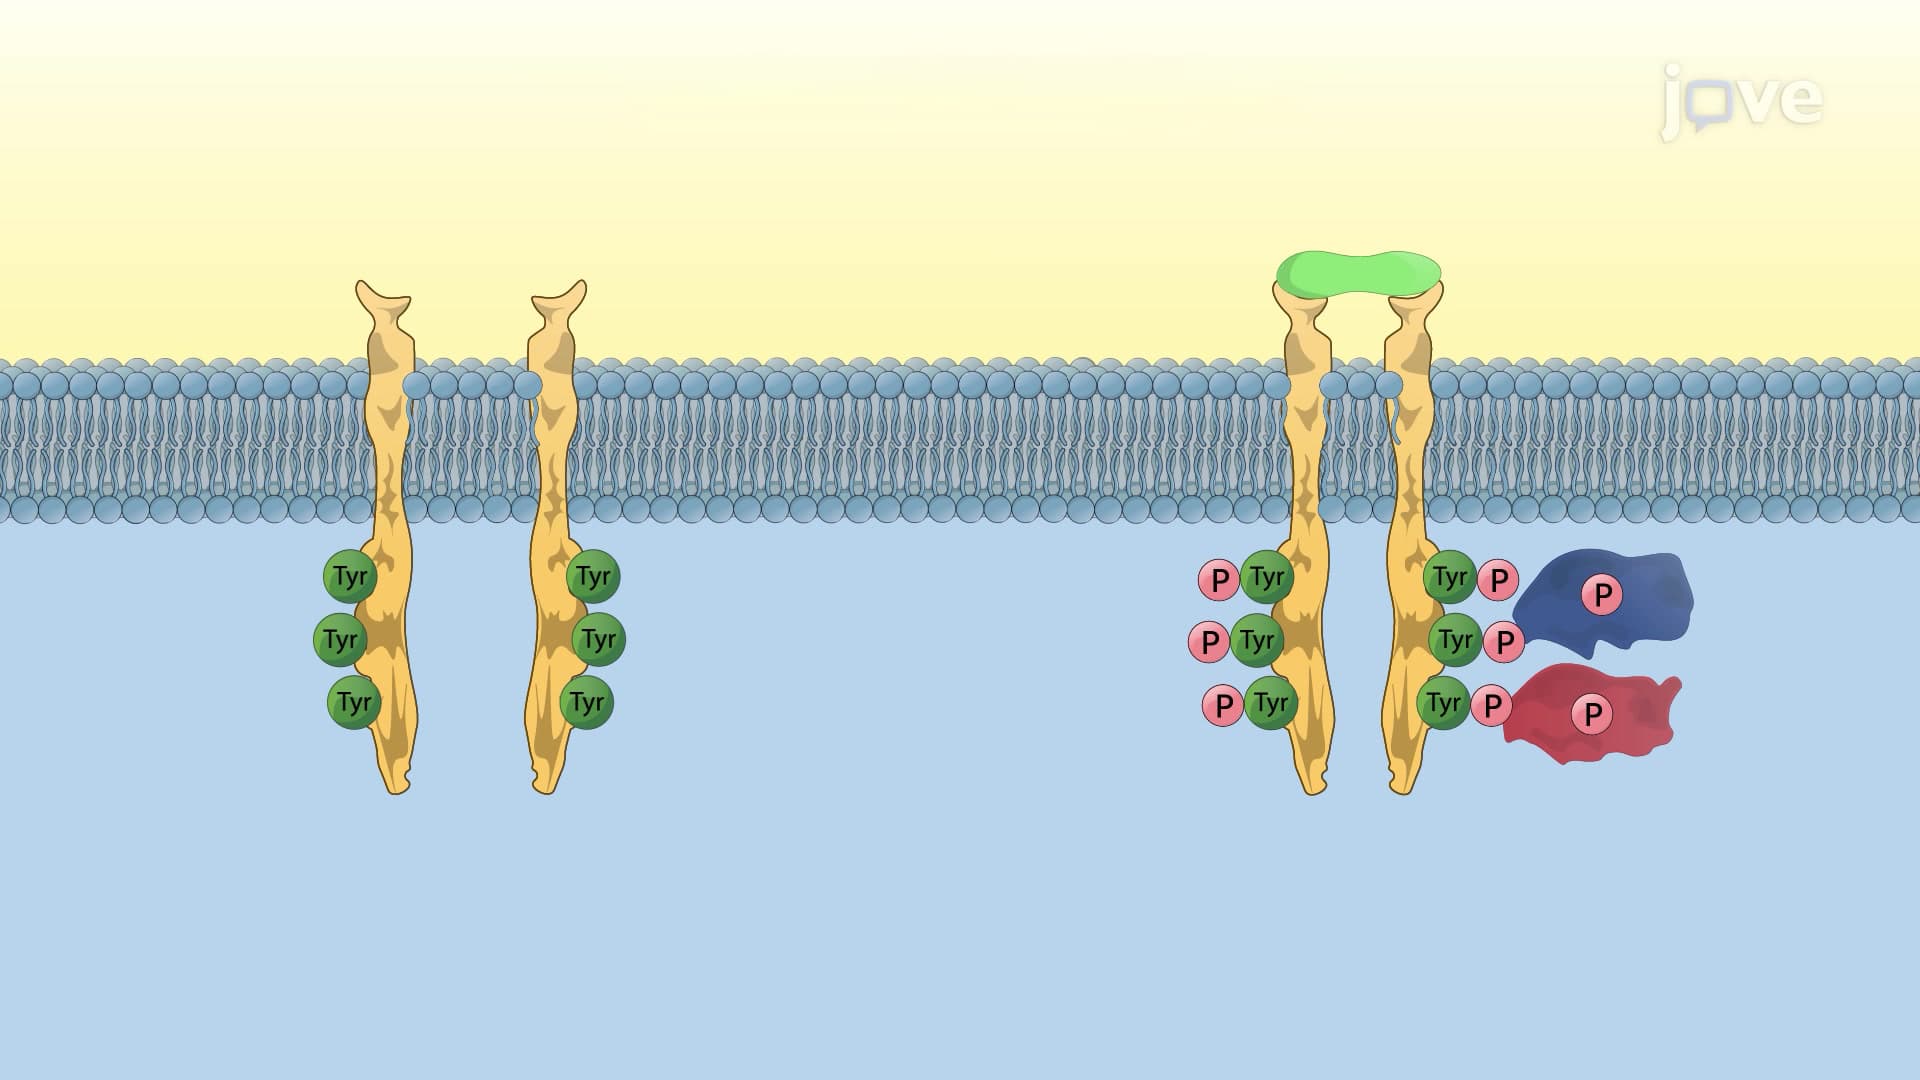
Receptor Tyrosine Kinases
 01:22
PI3K/mTOR/AKT Signaling Pathway
 02:23
Abnormal Proliferation
 03:03
mTOR Signaling and Cancer Progression
The mTOR pathway or the...
 02:57
Targeted Cancer Therapies
There are several types of targeted therapies against...
 02:38
Mitogens and the Cell Cycle