Related Experiment Video
Updated: Aug 18, 2025
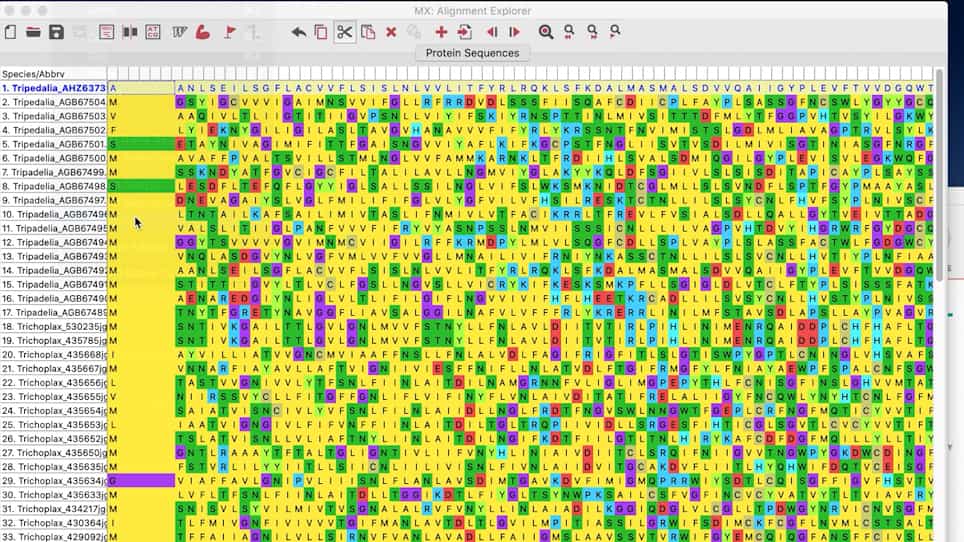
A Bioinformatics Pipeline for Investigating Molecular Evolution and Gene Expression using RNA-seq
Published on: May 28, 2021
annotate_my_genomes: an easy-to-use pipeline to improve genome annotation and uncover neglected genes by hybrid RNA
Carlos Farkas1, Antonia Recabal2, Andy Mella3,4
1Laboratorio de Investigación en Ciencias Biomédicas, Departamento de Ciencias Básicas y Morfología, Facultad de Medicina, Universidad Católica de la Santísima Concepción, Concepción, Chile.

Background:
The advancement of hybrid sequencing technologies is increasingly expanding genome assemblies that are often annotated using hybrid sequencing transcriptomics, leading to improved genome characterization and the identification of novel genes and isoforms in a wide variety of organisms.
Results:
We developed an easy-to-use genome-guided transcriptome annotation pipeline that uses assembled transcripts from hybrid sequencing data as input and distinguishes between coding and long non-coding RNAs by integration of several bioinformatic approaches, including gene reconciliation with previous annotations in GTF format. We demonstrated the efficiency of this approach by correctly assembling and annotating all exons from the chicken SCO-spondin gene (containing more than 105 exons), including the identification of missing genes in the chicken reference annotations by homology assignments.
Conclusions:
Our method helps to improve the current transcriptome annotation of the chicken brain. Our pipeline, implemented on Anaconda/Nextflow and Docker is an easy-to-use package that can be applied to a broad range of species, tissues, and research areas helping to improve and reconcile current annotations. The code and datasets are publicly available at https://github.com/cfarkas/annotate_my_genomes.
Related Concept Videos
 03:36
Genome Annotation and Assembly
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...