Related Experiment Video
Updated: Aug 18, 2025

Large Scale Energy Efficient Sensor Network Routing Using a Quantum Processor Unit
Published on: September 8, 2023
Insights from incorporating quantum computing into drug design workflows
Bayo Lau1, Prashant S Emani2,3, Jackson Chapman2,3
1HypaHealth, HypaHub Inc., San Jose, CA 95128, USA.

Motivation:
While many quantum computing (QC) methods promise theoretical advantages over classical counterparts, quantum hardware remains limited. Exploiting near-term QC in computer-aided drug design (CADD) thus requires judicious partitioning between classical and quantum calculations.
Results:
We present HypaCADD, a hybrid classical-quantum workflow for finding ligands binding to proteins, while accounting for genetic mutations. We explicitly identify modules of our drug-design workflow currently amenable to replacement by QC: non-intuitively, we identify the mutation-impact predictor as the best candidate. HypaCADD thus combines classical docking and molecular dynamics with quantum machine learning (QML) to infer the impact of mutations. We present a case study with the coronavirus (SARS-CoV-2) protease and associated mutants. We map a classical machine-learning module onto QC, using a neural network constructed from qubit-rotation gates. We have implemented this in simulation and on two commercial quantum computers. We find that the QML models can perform on par with, if not better than, classical baselines. In summary, HypaCADD offers a successful strategy for leveraging QC for CADD.
Availability And Implementation:
Jupyter Notebooks with Python code are freely available for academic use on GitHub: https://www.github.com/hypahub/hypacadd_notebook.
Supplementary Information:
Supplementary data are available at Bioinformatics online.
More Related Videos
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:06
Drug Clearance: Overview
Drug clearance is not limited to renal excretion but encompasses all organs involved in drug elimination, including...
 00:59
Model-Independent Approaches for Pharmacokinetic Data: Noncompartmental Analysis
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This...
 01:16
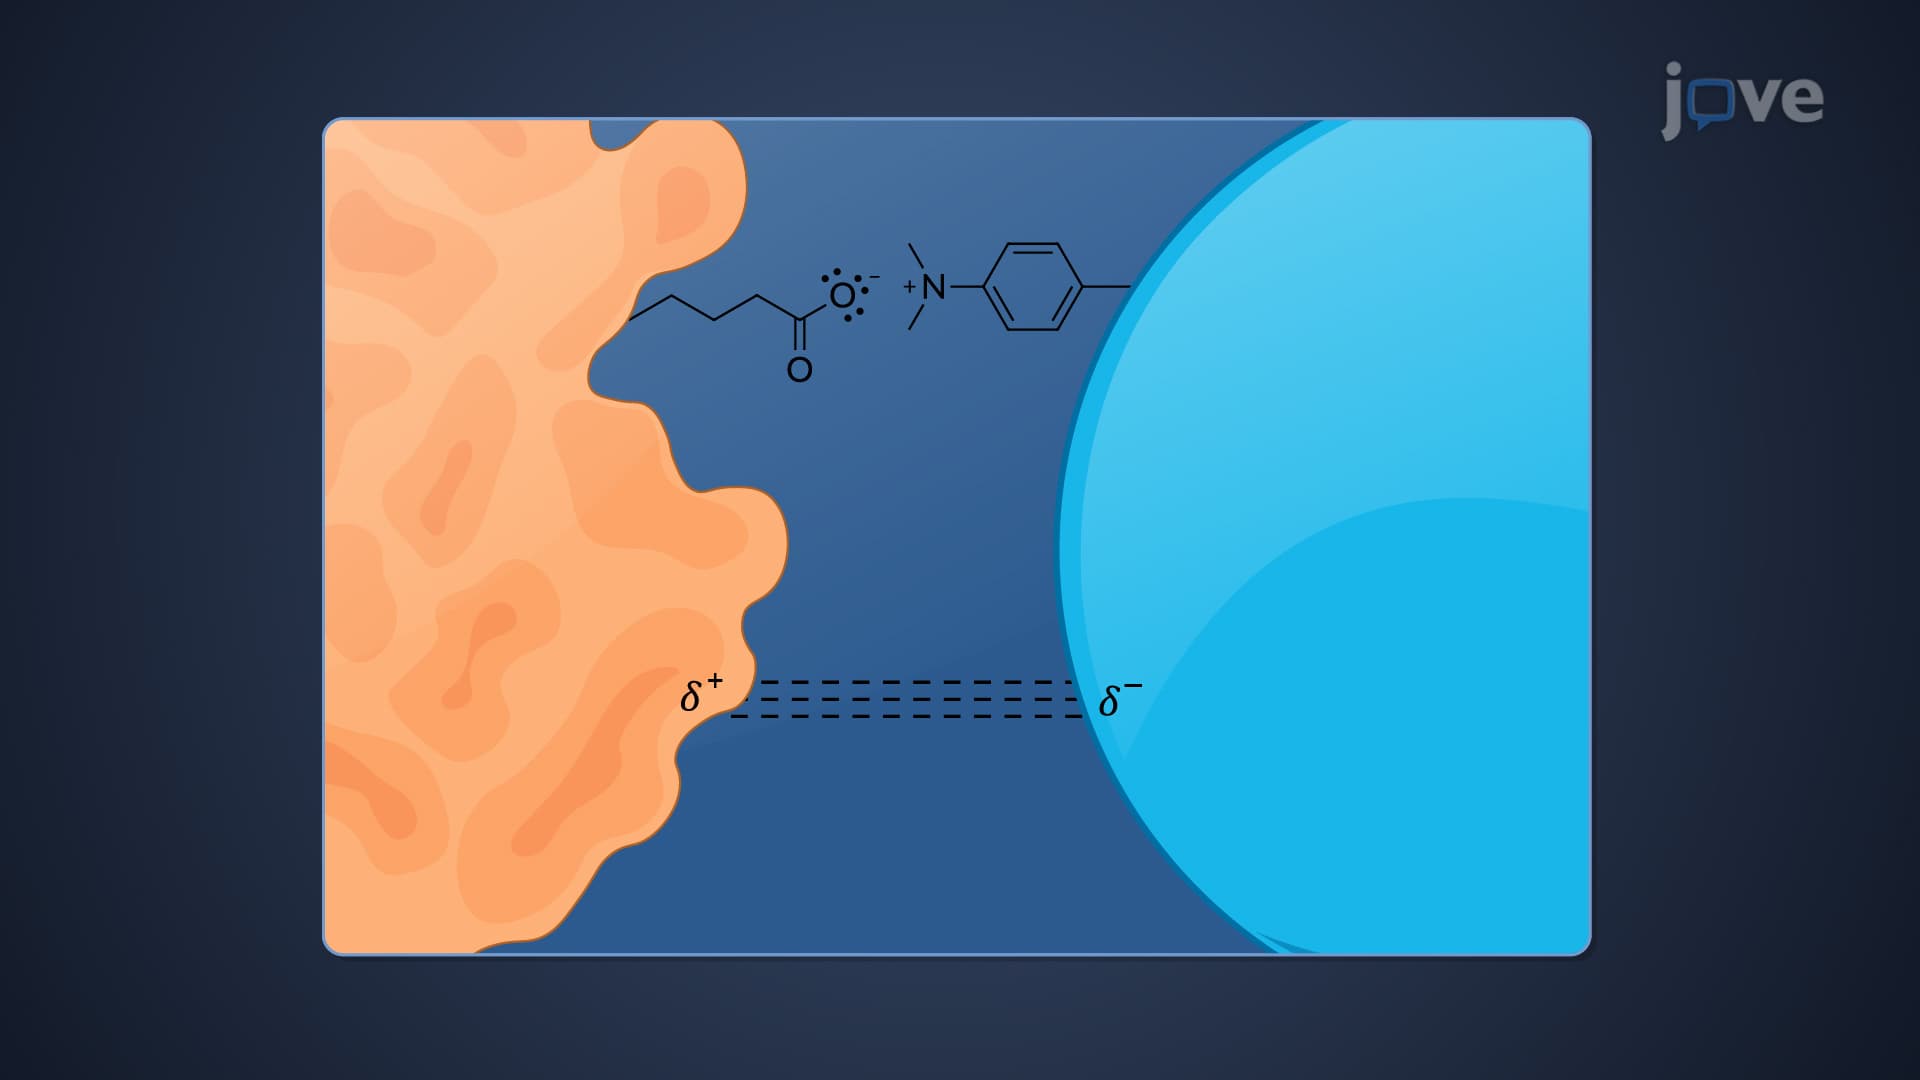
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...