Related Experiment Video
Updated: Aug 17, 2025


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Benchmarking AlphaFold2 on peptide structure prediction
Eli Fritz McDonald1, Taylor Jones1, Lars Plate2
1Department of Chemistry, Vanderbilt University, Nashville, TN 37212, USA; Center for Structural Biology, Vanderbilt University, Nashville, TN 37212, USA.

Abstract:
Recent advancements in computational tools have allowed protein structure prediction with high accuracy. Computational prediction methods have been used for modeling many soluble and membrane proteins, but the performance of these methods in modeling peptide structures has not yet been systematically investigated. We benchmarked the accuracy of AlphaFold2 in predicting 588 peptide structures between 10 and 40 amino acids using experimentally determined NMR structures as reference. Our results showed AlphaFold2 predicts α-helical, β-hairpin, and disulfide-rich peptides with high accuracy. AlphaFold2 performed at least as well if not better than alternative methods developed specifically for peptide structure prediction. AlphaFold2 showed several shortcomings in predicting Φ/Ψ angles, disulfide bond patterns, and the lowest RMSD structures failed to correlate with lowest pLDDT ranked structures. In summary, computation can be a powerful tool to predict peptide structures, but additional steps may be necessary to analyze and validate the results.


Related Concept Videos
 01:29
Protein Folding Quality Check in the RER
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....
 02:21
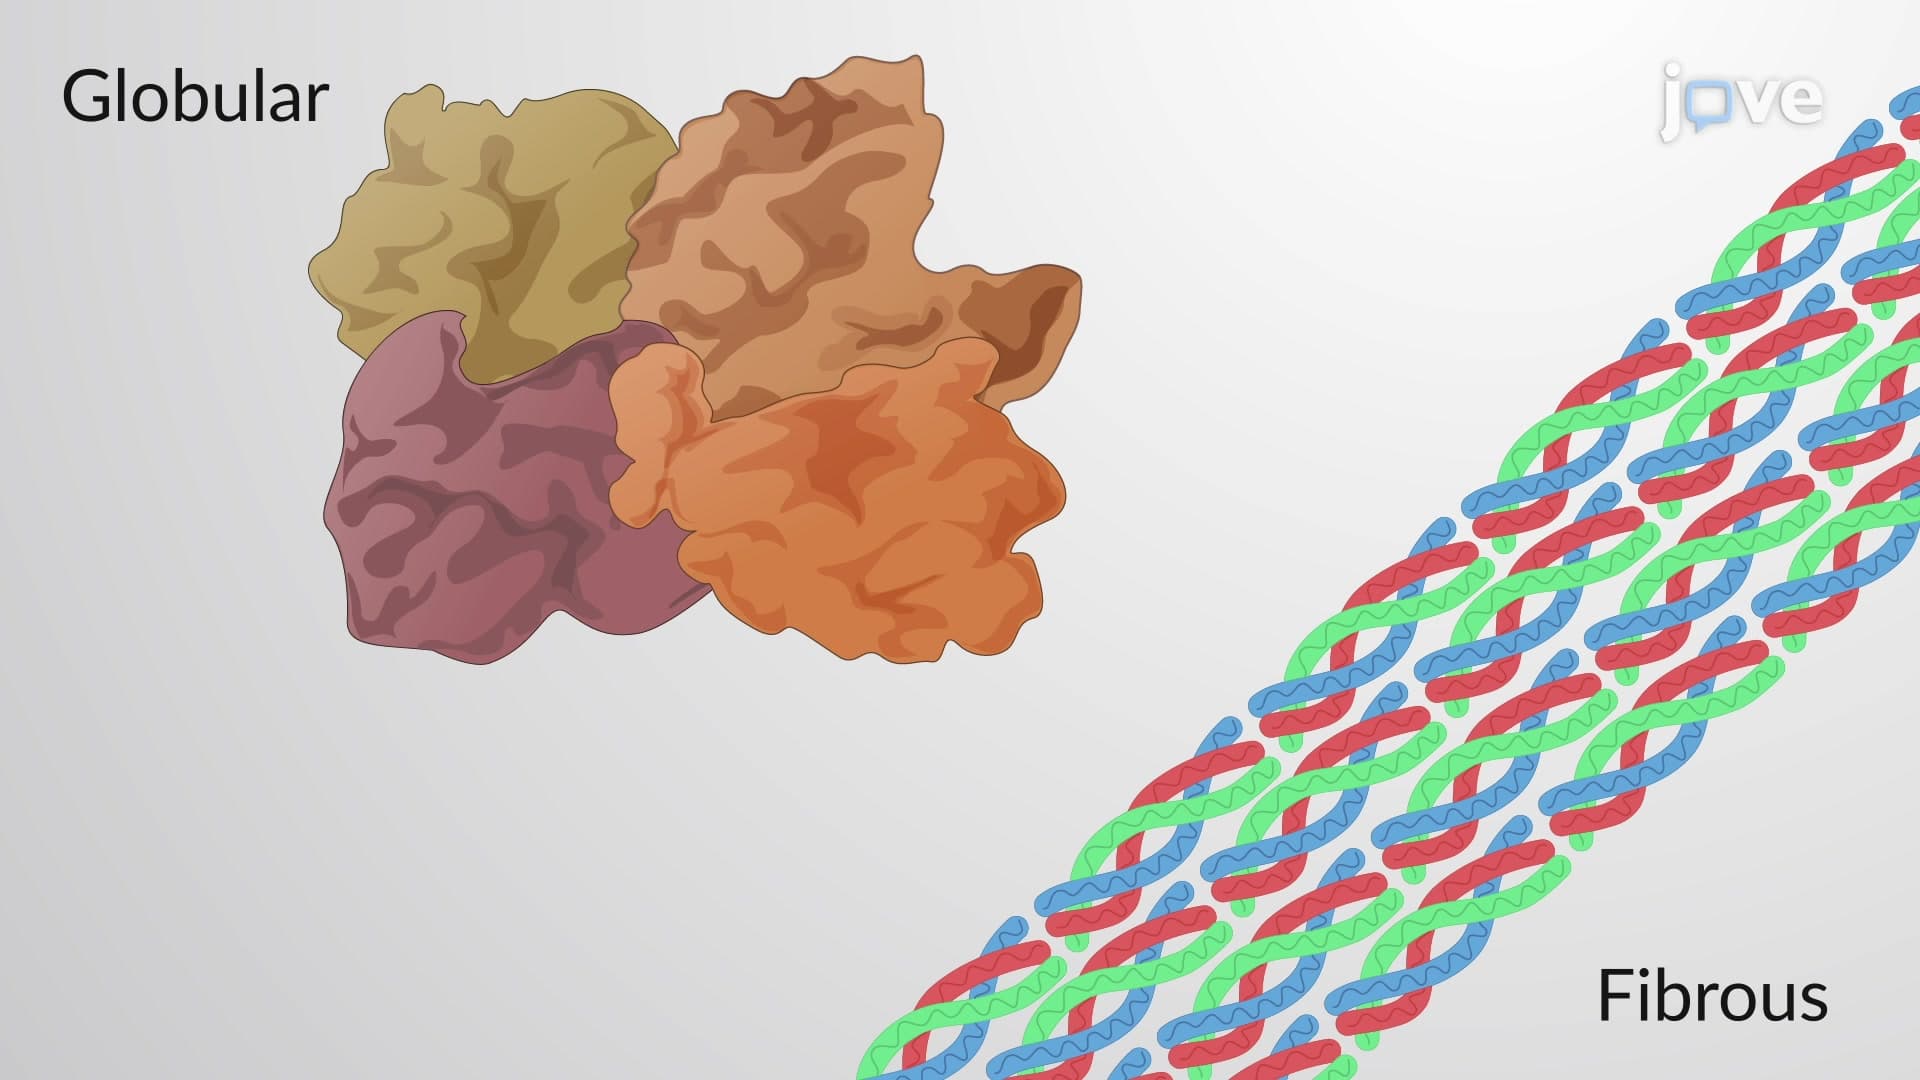
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...