Related Experiment Video
Updated: Aug 16, 2025


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Molecular dynamics simulation of adsorption and separation of xylene isomers by Cu-HKUST-1
Guo-Jian Ji1,2, Ting Xiang1, Xiao-Qing Zhou1
1Jiangsu Key Laboratory of Green Process Equipment, School of Petroleum and Gas Engineering, School of Energy, Changzhou University Changzhou 213164 P. R. China jgj@cczu.edu.cn +86-15806128724.

Abstract:
Metal-organic frameworks (MOFs) are widely used in the adsorption separation of various gases. A fundamental understanding of the effective separation of xylene isomers helps improve aromatic products' separation efficiency and reduce industrial separation costs. Grand Canonical Monte Carlo (GCMC) simulations combined with Molecular Science is widely used to predict gas adsorption and diffusion in single crystals with metal-organic frameworks. We performed a GCMC + MD combined approach to study xylene isomers' adsorption and separation in Cu-HKUST-1 to predict the permeability and selectivity of the ternary gas mixture in the MOF with the adsorption and diffusion usage data. Most current studies take into account the computational cost and difficulty. Most recent research models are limited to the adsorption of a single or specific molecule, such as hydrogen, methane, carbon dioxide, etc. For this reason, we report an attempt to study the adsorption separation of aromatic gases (p-xylene/o-xylene/m-xylene) based on Cu-HKUST-1 single-crystal materials based on some previous research methods with an appropriate increase in computational cost. To predict the adsorption selectivity and permeability of the ternary mixture of xylene isomers on the MOF surface, the model simulation calculates key parameters of gas adsorption, including gas adsorption volume (N), the heat of adsorption (Q st), Henry coefficient (K), and diffusion coefficient (D).
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:58
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:23
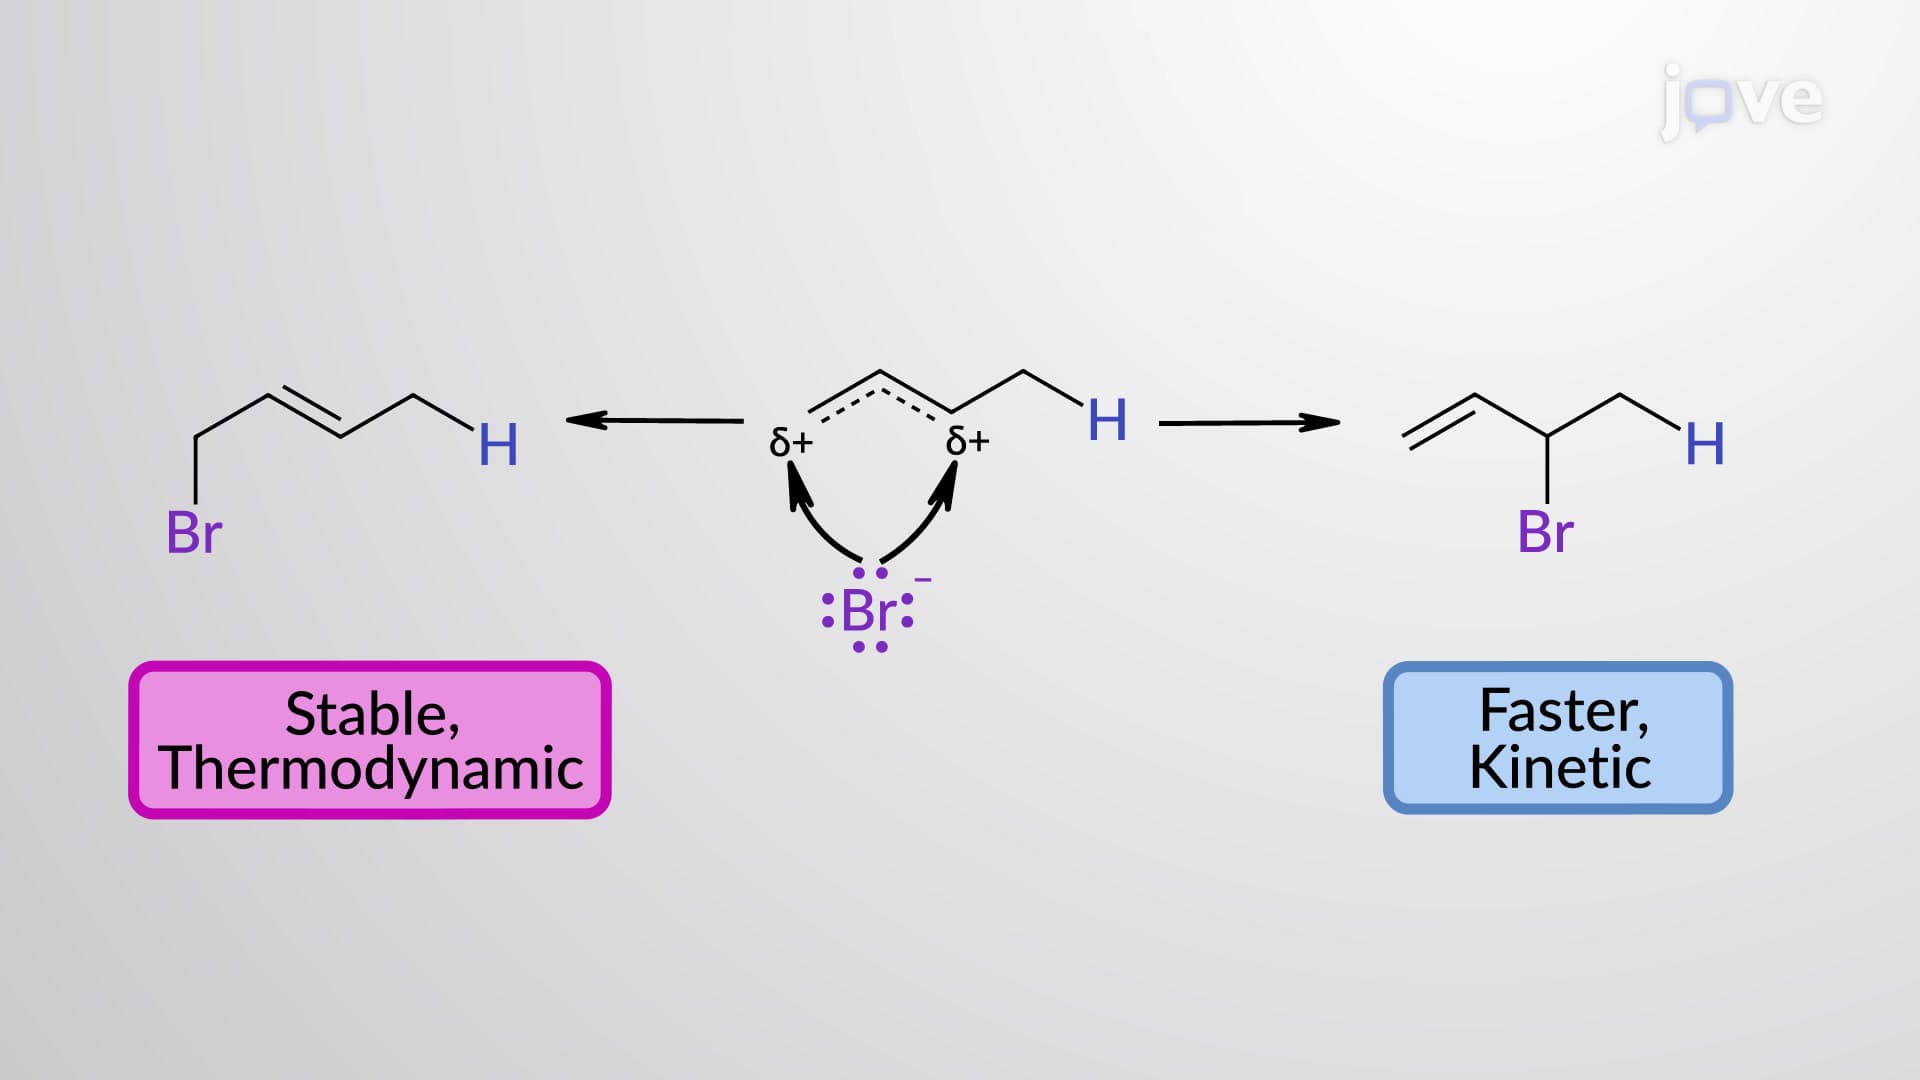
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
 02:37
Disubstituted Cyclohexanes: cis-trans Isomerism
In cyclohexane, the substituents can occupy different positions generating distinct isomers....
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this...
 01:09
Analyte Adsorption and Distribution