
Acidity and metal complexation of edge surface of birnessite-type MnO2: Insight from first principles simulations
Yingchun Zhang1, Xiandong Liu1, Xiancai Lu1
1State Key Laboratory for Mineral Deposits Research, School of Earth Sciences and Engineering, Nanjing University, Nanjing, Jiangsu 210023, People's Republic of China.

Abstract:
Birnessite-type MnO2 plays key roles in scavenging trace elements in numerous natural environments and has also been regarded as a promising energy storage material. The interfacial properties of birnessite are highly pH-dependent due to the presence of various amphoteric groups on its edges, and, therefore, the acidity constants (pKa) of these groups are vital to the understanding of its electrochemical and environmental performances. However, an accurate acidity dataset for birnessite is absent yet. In this study, we employed first-principles molecular dynamics simulations and the vertical energy gap method to calculate the pKas of groups on the birnessite (010) edge. The interfacial hydration structure was characterized with a focus on the hydrogen bonding network. The obtained pKas suggest that MnOH2 is active while Mn2OH remains inert in a common pH range. Based on these results, the incorporation of transition metals on the edge surface was investigated by taking Ni2+ and Zn2+ as the model cations. The energy changes associated with the incorporation process of Ni2+ from the outer-sphere state indicate that incorporation on the edge surface is more feasible than that on the basal surface presumed in previous studies. Overall, the results obtained provide an atomic-scale insight into the acid-base chemistry of birnessite and form a physical basis for understanding the interfacial processes of birnessite.

Related Concept Videos
 02:42
Valence Bond Theory
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:09
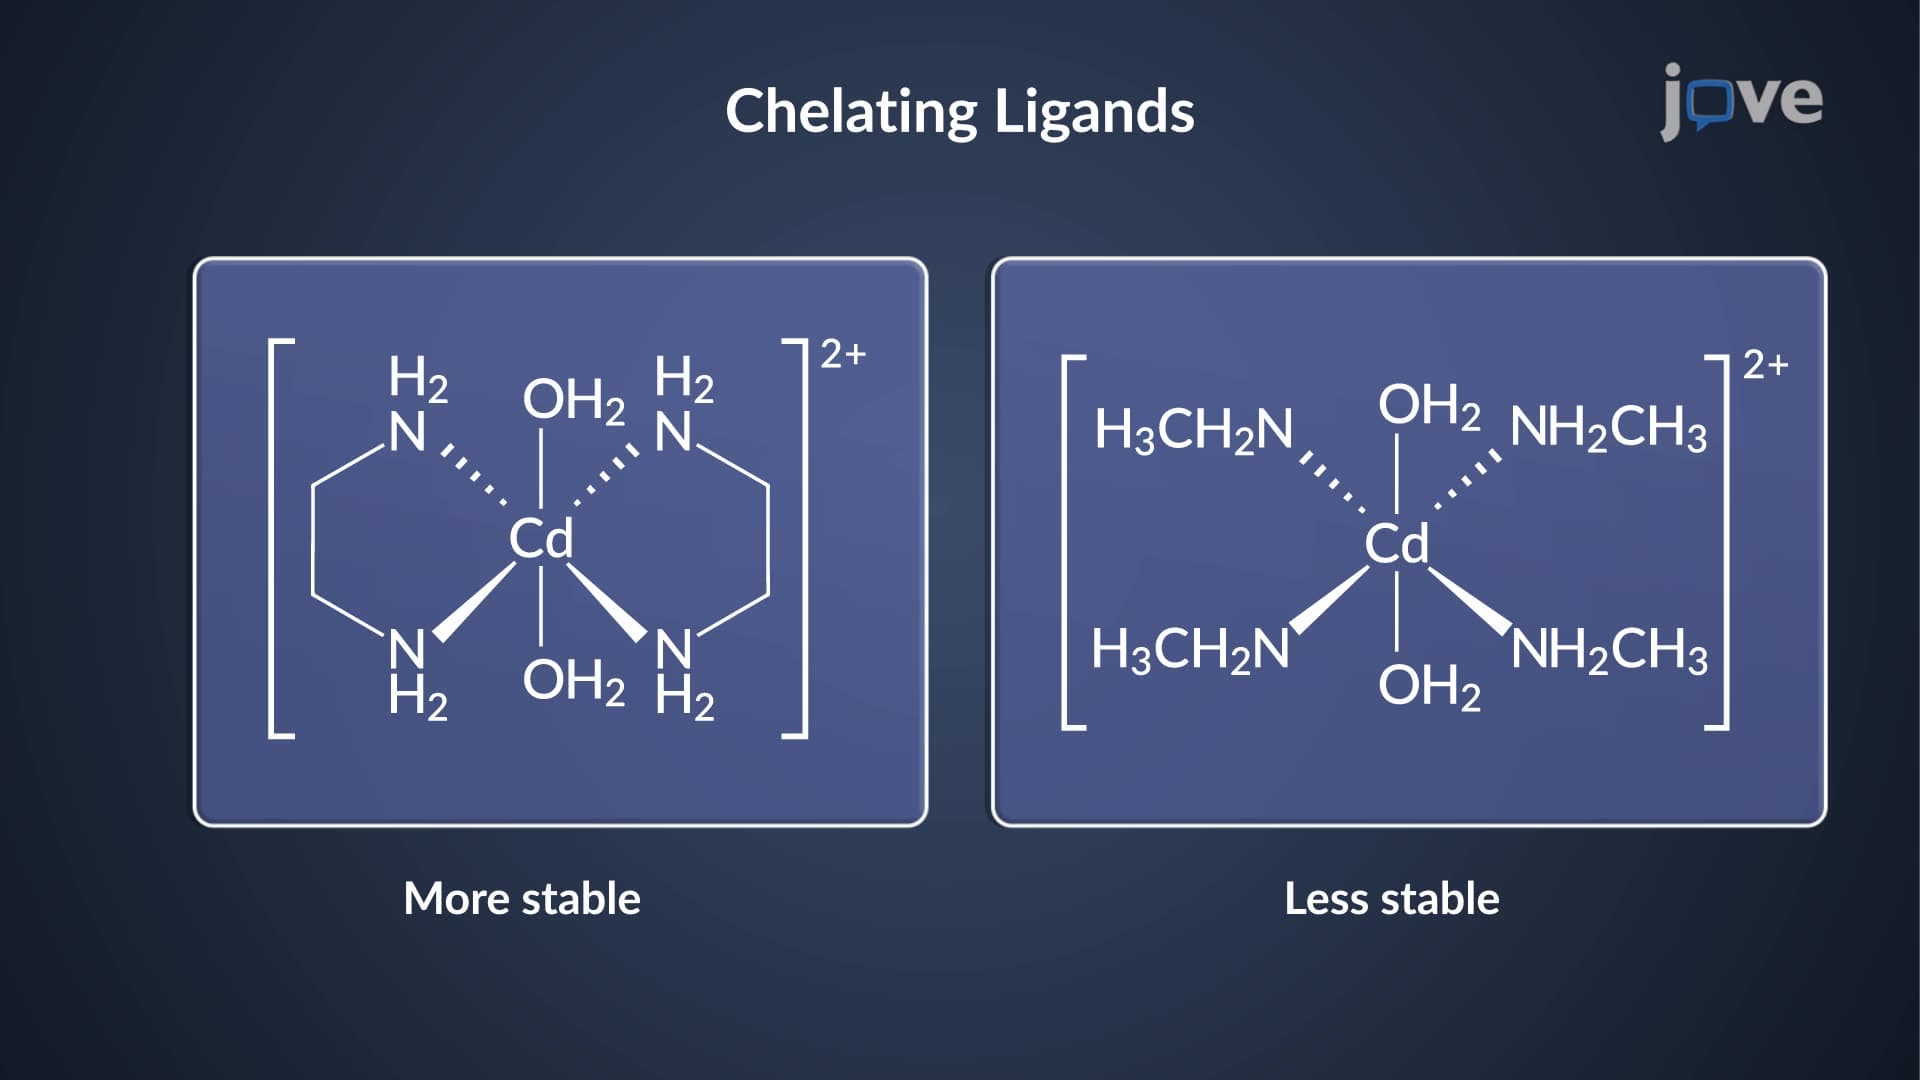
Complexation Equilibria: Factors Influencing Stability of Complexes
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 03:45
Formation of Complex Ions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...