Related Experiment Video
Updated: Aug 16, 2025

Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Evolutionary divergence in the conformational landscapes of tyrosine vs serine/threonine kinases
Joan Gizzio1,2, Abhishek Thakur1,2, Allan Haldane1,3
1Center for Biophysics and Computational Biology, Temple University, Philadelphia, United States.

Abstract:
Inactive conformations of protein kinase catalytic domains where the DFG motif has a "DFG-out" orientation and the activation loop is folded present a druggable binding pocket that is targeted by FDA-approved 'type-II inhibitors' in the treatment of cancers. Tyrosine kinases (TKs) typically show strong binding affinity with a wide spectrum of type-II inhibitors while serine/threonine kinases (STKs) usually bind more weakly which we suggest here is due to differences in the folded to extended conformational equilibrium of the activation loop between TKs vs. STKs. To investigate this, we use sequence covariation analysis with a Potts Hamiltonian statistical energy model to guide absolute binding free-energy molecular dynamics simulations of 74 protein-ligand complexes. Using the calculated binding free energies together with experimental values, we estimated free-energy costs for the large-scale (~17-20 Å) conformational change of the activation loop by an indirect approach, circumventing the very challenging problem of simulating the conformational change directly. We also used the Potts statistical potential to thread large sequence ensembles over active and inactive kinase states. The structure-based and sequence-based analyses are consistent; together they suggest TKs evolved to have free-energy penalties for the classical 'folded activation loop' DFG-out conformation relative to the active conformation, that is, on average, 4-6 kcal/mol smaller than the corresponding values for STKs. Potts statistical energy analysis suggests a molecular basis for this observation, wherein the activation loops of TKs are more weakly 'anchored' against the catalytic loop motif in the active conformation and form more stable substrate-mimicking interactions in the inactive conformation. These results provide insights into the molecular basis for the divergent functional properties of TKs and STKs, and have pharmacological implications for the target selectivity of type-II inhibitors.
Insights
Type-II inhibitors target inactive protein kinases for cancer treatment. Differences in activation loop dynamics between tyrosine kinases (TKs) and serine/threonine kinases (STKs) explain varying inhibitor binding affinities, impacting drug selectivity.
Area of Science:
- Biochemistry
- Structural Biology
- Pharmacology
Background:
- FDA-approved type-II inhibitors target inactive protein kinase conformations with a 'DFG-out' motif and folded activation loop.
- Tyrosine kinases (TKs) exhibit strong binding to type-II inhibitors, unlike serine/threonine kinases (STKs), suggesting differences in activation loop dynamics.
- Understanding these dynamics is crucial for developing selective kinase inhibitors for cancer therapy.
Purpose of the Study:
- To investigate the molecular basis for differential binding affinities of type-II inhibitors between TKs and STKs.
- To quantify the free-energy costs associated with activation loop conformational changes in kinases.
- To explore the evolutionary divergence in kinase activation loop dynamics and its pharmacological implications.
Main Methods:
- Sequence covariation analysis using a Potts Hamiltonian statistical energy model.
- Absolute binding free-energy molecular dynamics simulations for 74 protein-ligand complexes.
- Indirect estimation of activation loop conformational change free-energy costs and sequence-based analysis of kinase states.
Main Results:
- TKs possess smaller free-energy penalties for the 'DFG-out' conformation compared to STKs (4-6 kcal/mol difference).
- TK activation loops are less anchored in the active state and form more stable substrate-mimicking interactions in the inactive state.
- Structure- and sequence-based analyses reveal consistent differences in kinase activation loop dynamics.
Conclusions:
- Divergent activation loop dynamics between TKs and STKs underlie their differential responses to type-II inhibitors.
- TKs evolved to favor the active conformation less than STKs, influencing inhibitor binding.
- These findings provide molecular insights into kinase function and inform the design of selective type-II inhibitors for cancer treatment.
Related Concept Videos
 01:26
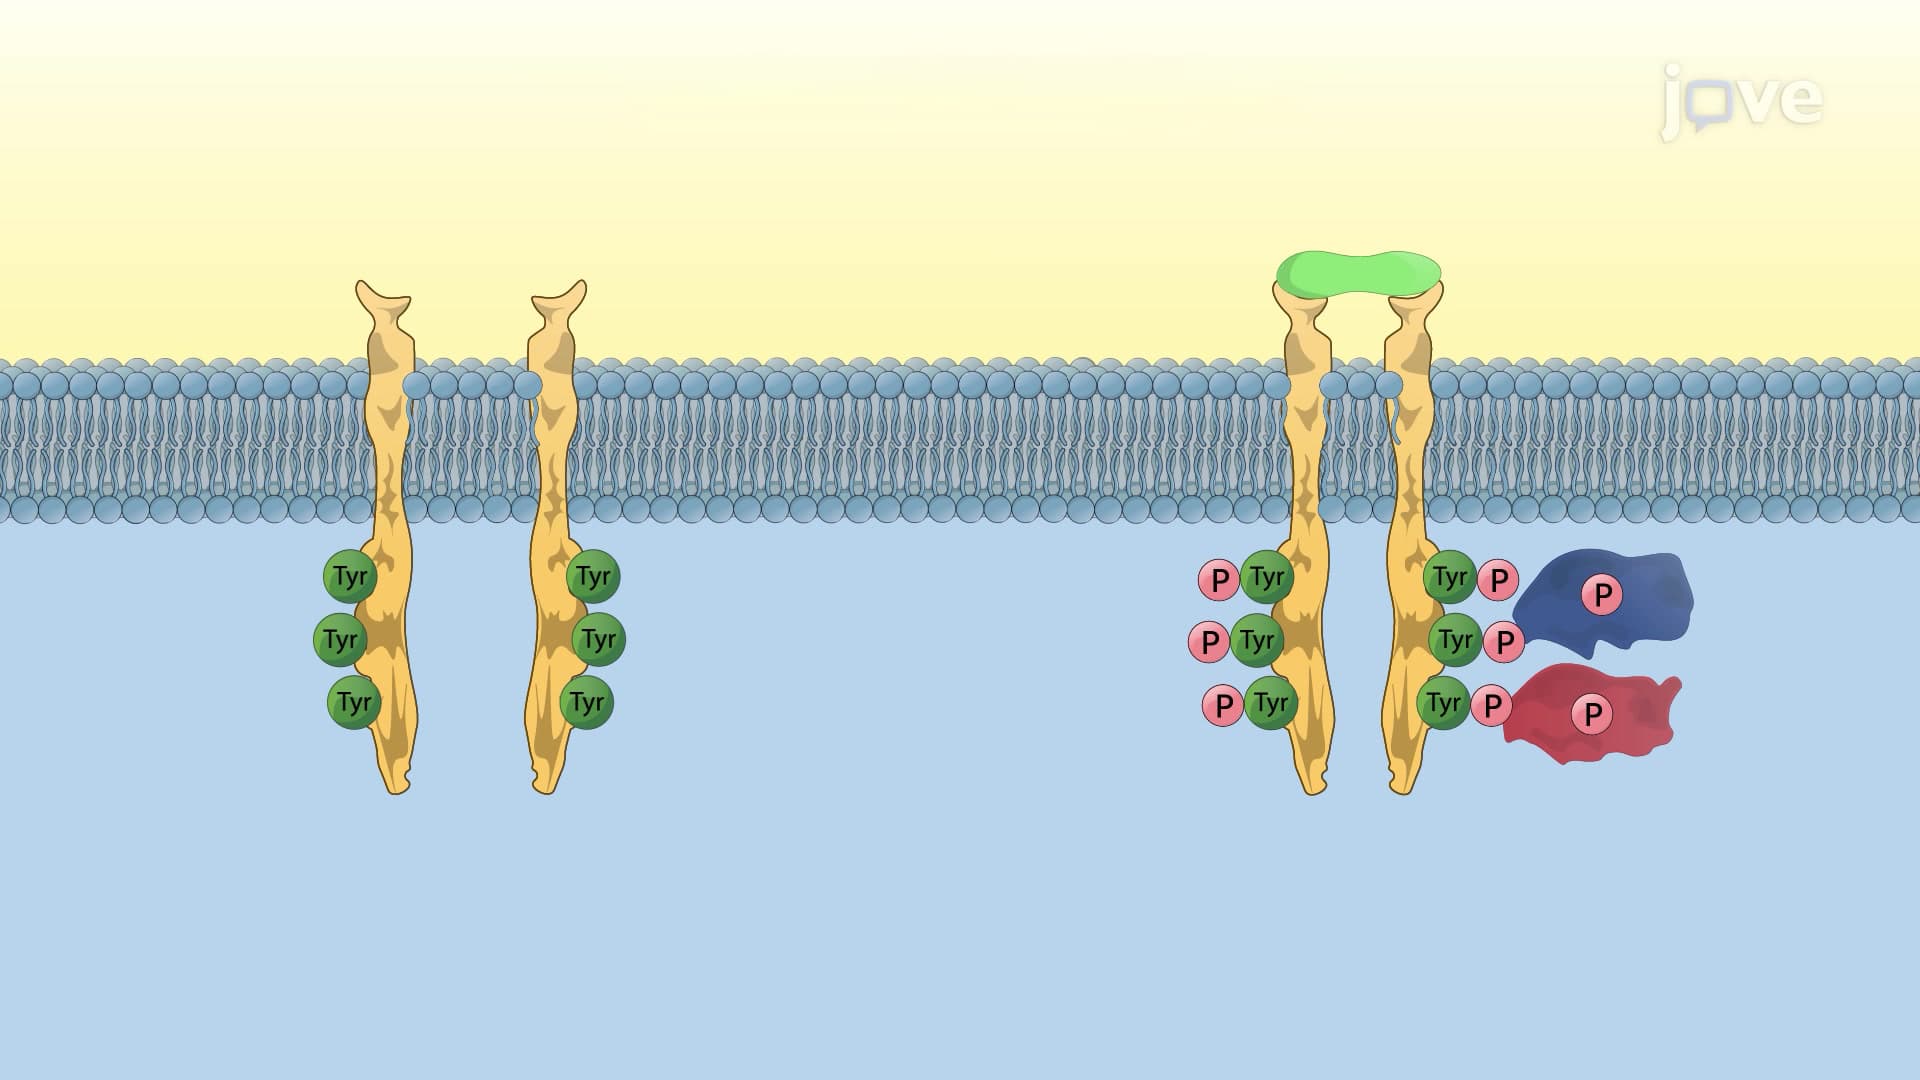
Receptor Tyrosine Kinases
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
 01:02
Phosphorylation
During phosphorylation, protein kinases transfer the terminal phosphate group of ATP to specific amino acid side chains of substrate proteins. Serine, threonine, and tyrosine are the most commonly...