Related Experiment Video
Updated: Aug 16, 2025


A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Target-template relationships in protein structure prediction and their effect on the accuracy of thermostability
Muyun Lihan1,2, Dmitry Lupyan2, Daniel Oehme2
1NIH Center for Macromolecular Modeling and Bioinformatics, Beckman Institute for Advanced Science and Technology, and Center for Biophysics and Quantitative Biology, University of Illinois Urbana-Champaign, Urbana, Illinois, USA.

Abstract:
Improving protein thermostability has been a labor- and time-consuming process in industrial applications of protein engineering. Advances in computational approaches have facilitated the development of more efficient strategies to allow the prioritization of stabilizing mutants. Among these is FEP+, a free energy perturbation implementation that uses a thoroughly tested physics-based method to achieve unparalleled accuracy in predicting changes in protein thermostability. To gauge the applicability of FEP+ to situations where crystal structures are unavailable, here we have applied the FEP+ approach to homology models of 12 different proteins covering 316 mutations. By comparing predictions obtained with homology models to those obtained using crystal structures, we have identified that local rather than global sequence conservation between target and template sequence is a determining factor in the accuracy of predictions. By excluding mutation sites with low local sequence identity (<40%) to a template structure, we have obtained predictions with comparable performance to crystal structures (R2 of 0.67 and 0.63 and an RMSE of 1.20 and 1.16 kcal/mol for crystal structure and homology model predictions, respectively) for identifying stabilizing mutations when incorporating residue scanning into a cascade screening strategy. Additionally, we identify and discuss inherent limitations in sequence alignments and homology modeling protocols that translate into the poor FEP+ performance of a few select examples. Overall, our retrospective study provides detailed guidelines for the application of the FEP+ approach using homology models for protein thermostability predictions, which will greatly extend this approach to studies that were previously limited by structure availability.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:22
Protein Folding
 01:24
Protein Organization
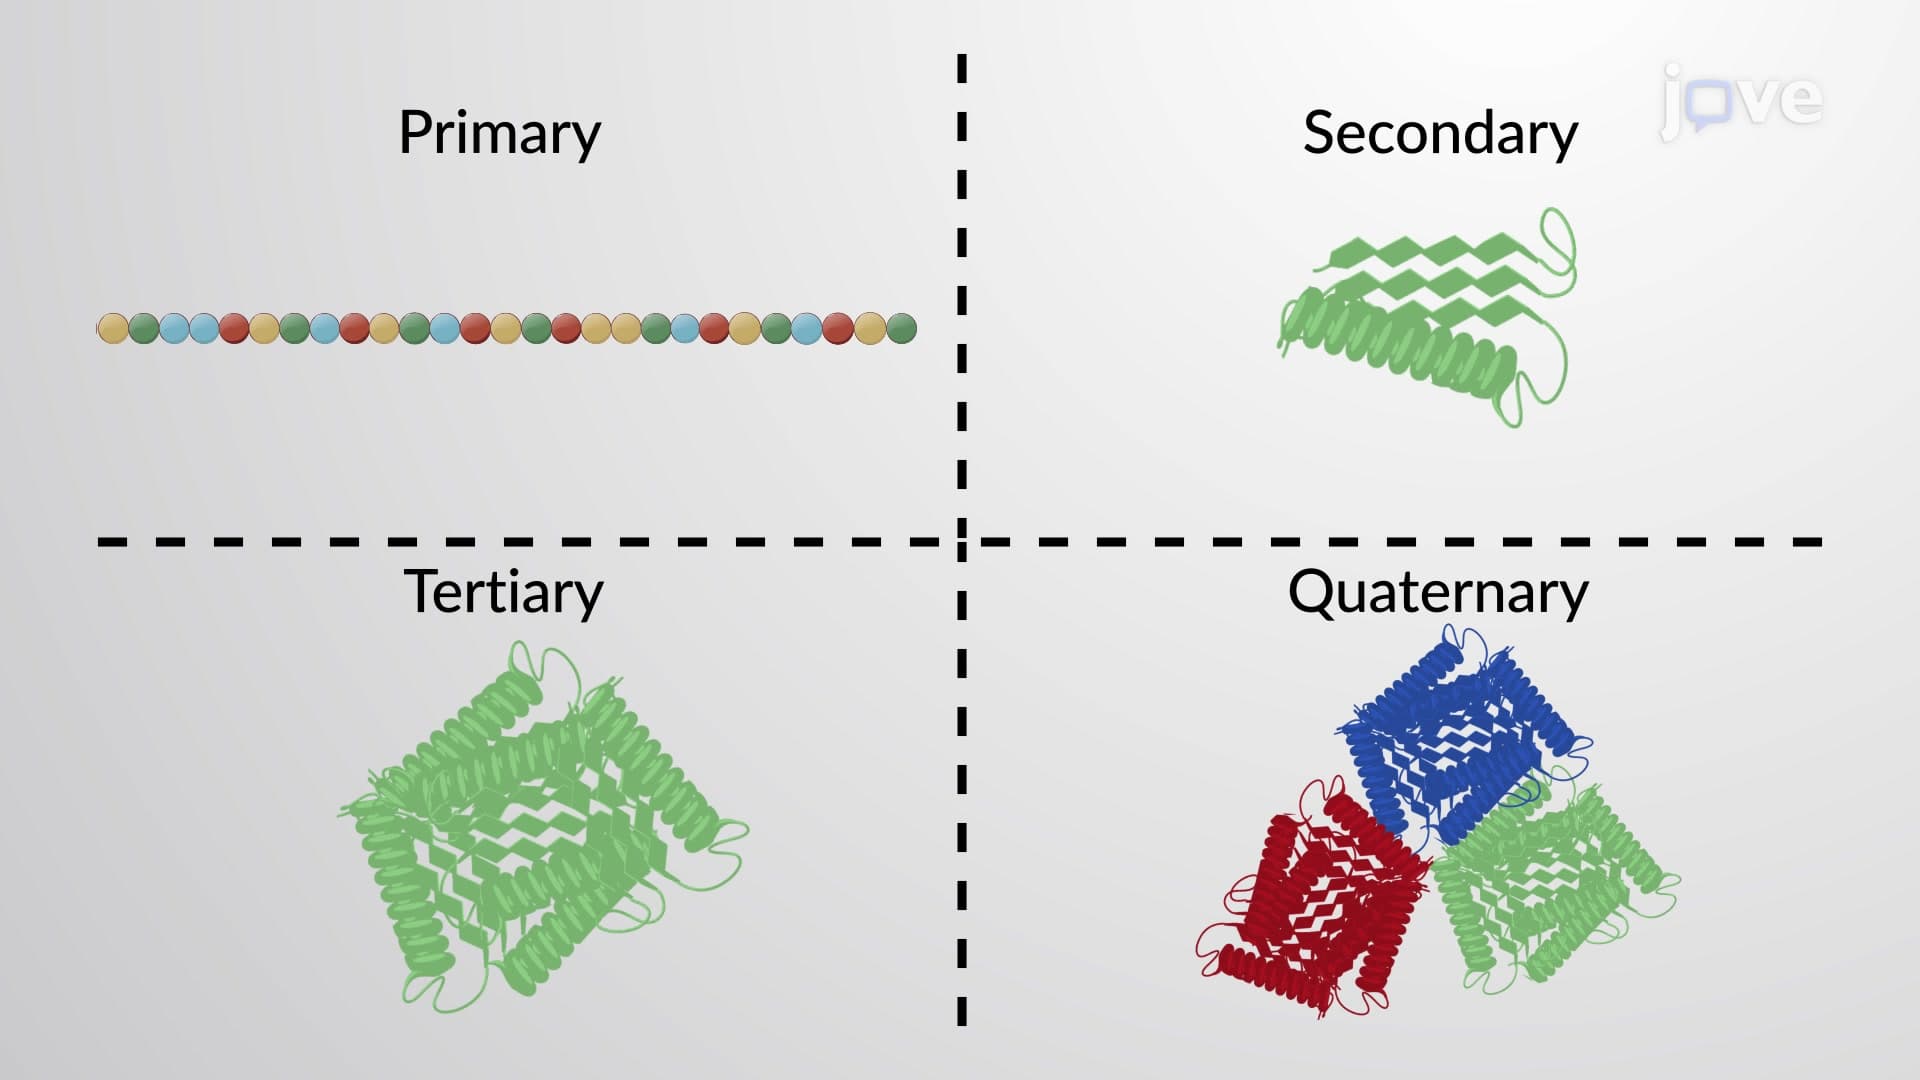
The primary structure of a protein is its amino acid sequence....
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 02:07
Improving Translational Accuracy
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...