Related Experiment Video
Updated: Aug 15, 2025
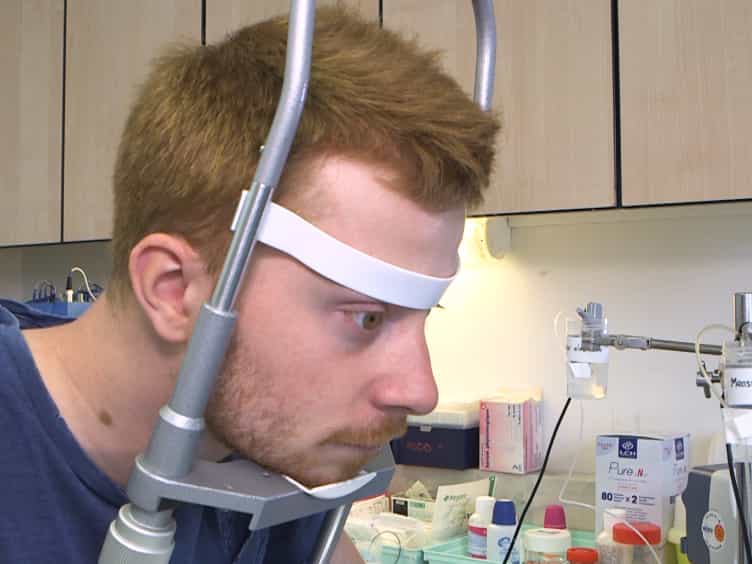

Standardized Measurement of Nasal Membrane Transepithelial Potential Difference NPD
Published on: September 13, 2018
Gradual increase in sweat chloride concentration is associated with a higher risk of CRMS/CFSPID to CF
Danieli B Salinas1, Daniella K Ginsburg1, Choo Phei Wee2
1Department of Pediatrics, Division of Pediatric Pulmonology, Children's Hospital Los Angeles, Keck School of Medicine, University of Southern California, Los Angeles, California, USA.

Insights
High sweat chloride and Pseudomonas aeruginosa colonization increase the risk of reclassifying from cystic fibrosis (CF) screen-positive, inconclusive diagnosis (CFSPID) to CF. Close monitoring for phenotypic changes is crucial for these infants.
Area of Science:
- Pediatrics
- Genetics
- Medical Diagnostics
Background:
- Universal newborn screening for cystic fibrosis (CF) has identified infants with CF screen-positive, inconclusive diagnosis (CFSPID).
- Limited data exists on prognosis and standardized care for CFSPID infants.
- Risk factors for reclassification from CFSPID to CF are not well-established.
Purpose of the Study:
- To investigate clinical characteristics associated with the risk of reclassification from CFSPID to a confirmed CF diagnosis.
- To identify risk factors that predict progression from CFSPID to CF.
Main Methods:
- Recruitment of children with positive CF newborn screening (NBS) from two California sites.
- Collection of retrospective, longitudinal, and cross-sectional data, including nasal epithelial cells for CFTR functional assessment.
- Multivariate logistic regression analysis to determine risk factors for reclassification.
Main Results:
- 112 children completed the study (53 CF, 59 CFSPID).
- Differences observed in pancreatic insufficiency, immunoreactive trypsinogen (IRT), and Pseudomonas aeruginosa (PSA) colonization between groups.
- Increasing sweat chloride concentration (sw[Cl- ]) and PSA colonization were independent risk factors for reclassification to CF.
Conclusions:
- Elevated sw[Cl- ] and a history of PSA colonization are linked to higher risk of CF reclassification in infants with high IRT and two CFTR variants.
- Continued phenotypic monitoring is essential for CFSPID infants.
- Further research is needed to clarify the role of CFTR functional assays in managing CFSPID.
Objectives:
Universal implementation of cystic fibrosis (CF) newborn screening (NBS) has led to the diagnostic dilemma of infants with CF screen-positive, inconclusive diagnosis (CFSPID), with limited guidance regarding prognosis and standardized care. Rates of reclassification from CFSPID to CF vary and risk factors for reclassification are not well established. We investigated whether clinical characteristics are associated with the risk of reclassification from CFSPID to a CF diagnosis.
Methods:
Children with a positive CF NBS were recruited from two sites in California. Retrospective, longitudinal, and cross-sectional data were collected. A subset of subjects had nasal epithelial cells collected for CF transmembrane conductance regulator (CFTR) functional assessment. Multivariate logistic regression was used to assess the risk of reclassification.
Results:
A total of 112 children completed the study (CF = 53, CFSPID = 59). Phenotypic characteristics between groups showed differences in pancreatic insufficiency prevalence, immunoreactive trypsinogen (IRT) levels, and Pseudomonas aeruginosa (PSA) colonization. Spirometry measures were not different between groups. Nasal epithelial cells from 10 subjects showed 7%-30% of wild-type (WT)-CFTR (wtCFTR) function in those who reclassified and 27%-67% of wtCFTR function in those who retained the CFSPID designation. Modeling revealed that increasing sweat chloride concentration (sw[Cl- ]) and PSA colonization were independent risk factors for reclassification to CF.
Conclusion:
Increasing sw[Cl- ] and a history of PSA colonization are associated with the risk of reclassification from CFSPID to CF in a population with high IRT and two CFTR variants. A close follow-up to monitor phenotypic changes remains critical in this population. The role of CFTR functional assays in this population requires further exploration.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...