Related Experiment Video
Updated: Aug 15, 2025
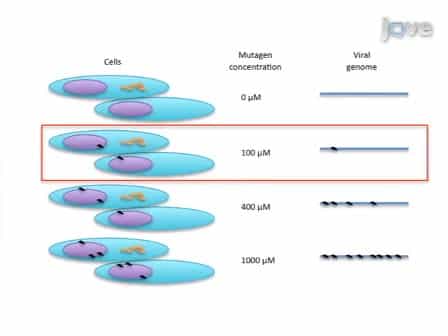

Isolation of Fidelity Variants of RNA Viruses and Characterization of Virus Mutation Frequency
Published on: June 16, 2011
Viral genome sequence datasets display pervasive evidence of strand-specific substitution biases that are best
Rita Sianga-Mete1, Penelope Hartnady1, Wimbai Caroline Mandikumba1
1Division of Computational Biology, Institute of Infectious Diseases and Molecular Medicine, Department of Integrative Biomedical Sciences, Faculty of Health Sciences, University of Cape Town, Anzio Road Observatory 7549, Cape Town, South Africa.

Abstract:
Most phylogenetic trees are inferred using time-reversible evolutionary models that assume that the relative rates of substitution for any given pair of nucleotides are the same regardless of the direction of the substitutions. However, there is no reason to assume that the underlying biochemical mutational processes that cause substitutions are similarly symmetrical. We consider two non-reversible nucleotide substitution models: (1) a 6-rate non-reversible model (NREV6) that is applicable to analyzing mutational processes in double-stranded genomes in that complementary substitutions occur at identical rates; and (2) a 12-rate non-reversible model (NREV12) that is applicable to analyzing mutational processes in single-stranded (ss) genomes in that all substitution types are free to occur at different rates. Using likelihood ratio and Akaike Information Criterion-based model tests, we show that, surprisingly, NREV12 provided a significantly better fit than the General Time Reversible (GTR) and NREV6 models to 21/31 dsRNA and 20/30 dsDNA datasets. As expected, however, NREV12 provided a significantly better fit to 24/33 ssDNA and 40/47 ssRNA datasets. We tested how non-reversibility impacts the accuracy with which phylogenetic trees are inferred. As simulated degrees of non-reversibility (DNR) increased, the tree topology inferences using both NREV12 and GTR became more accurate, whereas inferred tree branch lengths became less accurate. We conclude that while non-reversible models should be helpful in the analysis of mutational processes in most virus species, there is no pressing need to use these models for routine phylogenetic inference.
Related Concept Videos
 00:36
Viral Mutations
 02:28
Leaky Scanning
 01:26
Size and Structure of Viral Genomes
 00:57
Viral Recombination
 01:18
Mutations in Microorganisms
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...