Related Experiment Video
Updated: Aug 15, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
ωB97X-3c: A composite range-separated hybrid DFT method with a molecule-optimized polarized valence double-ζ basis
Marcel Müller1, Andreas Hansen1, Stefan Grimme1
1Mulliken Center for Theoretical Chemistry, Clausius-Institut für Physikalische und Theoretische Chemie, Rheinische Friedrich-Wilhelms Universität Bonn, Beringstraße 4, 53115 Bonn, Germany.

Abstract:
A new composite density functional theory (DFT) method is presented. It is based on ωB97X-V as one of the best-performing density functionals for the GMTKN55 thermochemistry database and completes the family of "3c" methods toward range-separated hybrid DFT. This method is consistently available for all elements up to Rn (Z = 1-86). Its further key ingredients are a polarized valence double-ζ (vDZP) Gaussian basis set, which was fully optimized in molecular DFT calculations, in combination with large-core effective core potentials and a specially adapted D4 dispersion correction. Unlike most existing double-ζ atomic orbital sets, vDZP shows only small basis set superposition errors (BSSEs) and can compete with standard sets of triple-ζ quality. Small residual BSSE effects are efficiently absorbed by the D4 damping scheme, which overall eliminates the need for an explicit treatment or empirical corrections for BSSE. Thorough tests on a variety of thermochemistry benchmark sets show that the new composite method, dubbed ωB97X-3c, is on par with or even outperforms standard hybrid DFT methods in a quadruple-zeta basis set at a small fraction of the computational cost. Particular strengths of this method are the description of non-covalent interactions and barrier heights, for which it is among the best-performing density functionals overall.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 03:35
Hybridization of Atomic Orbitals II
 03:24
Hybridization of Atomic Orbitals I
 02:38
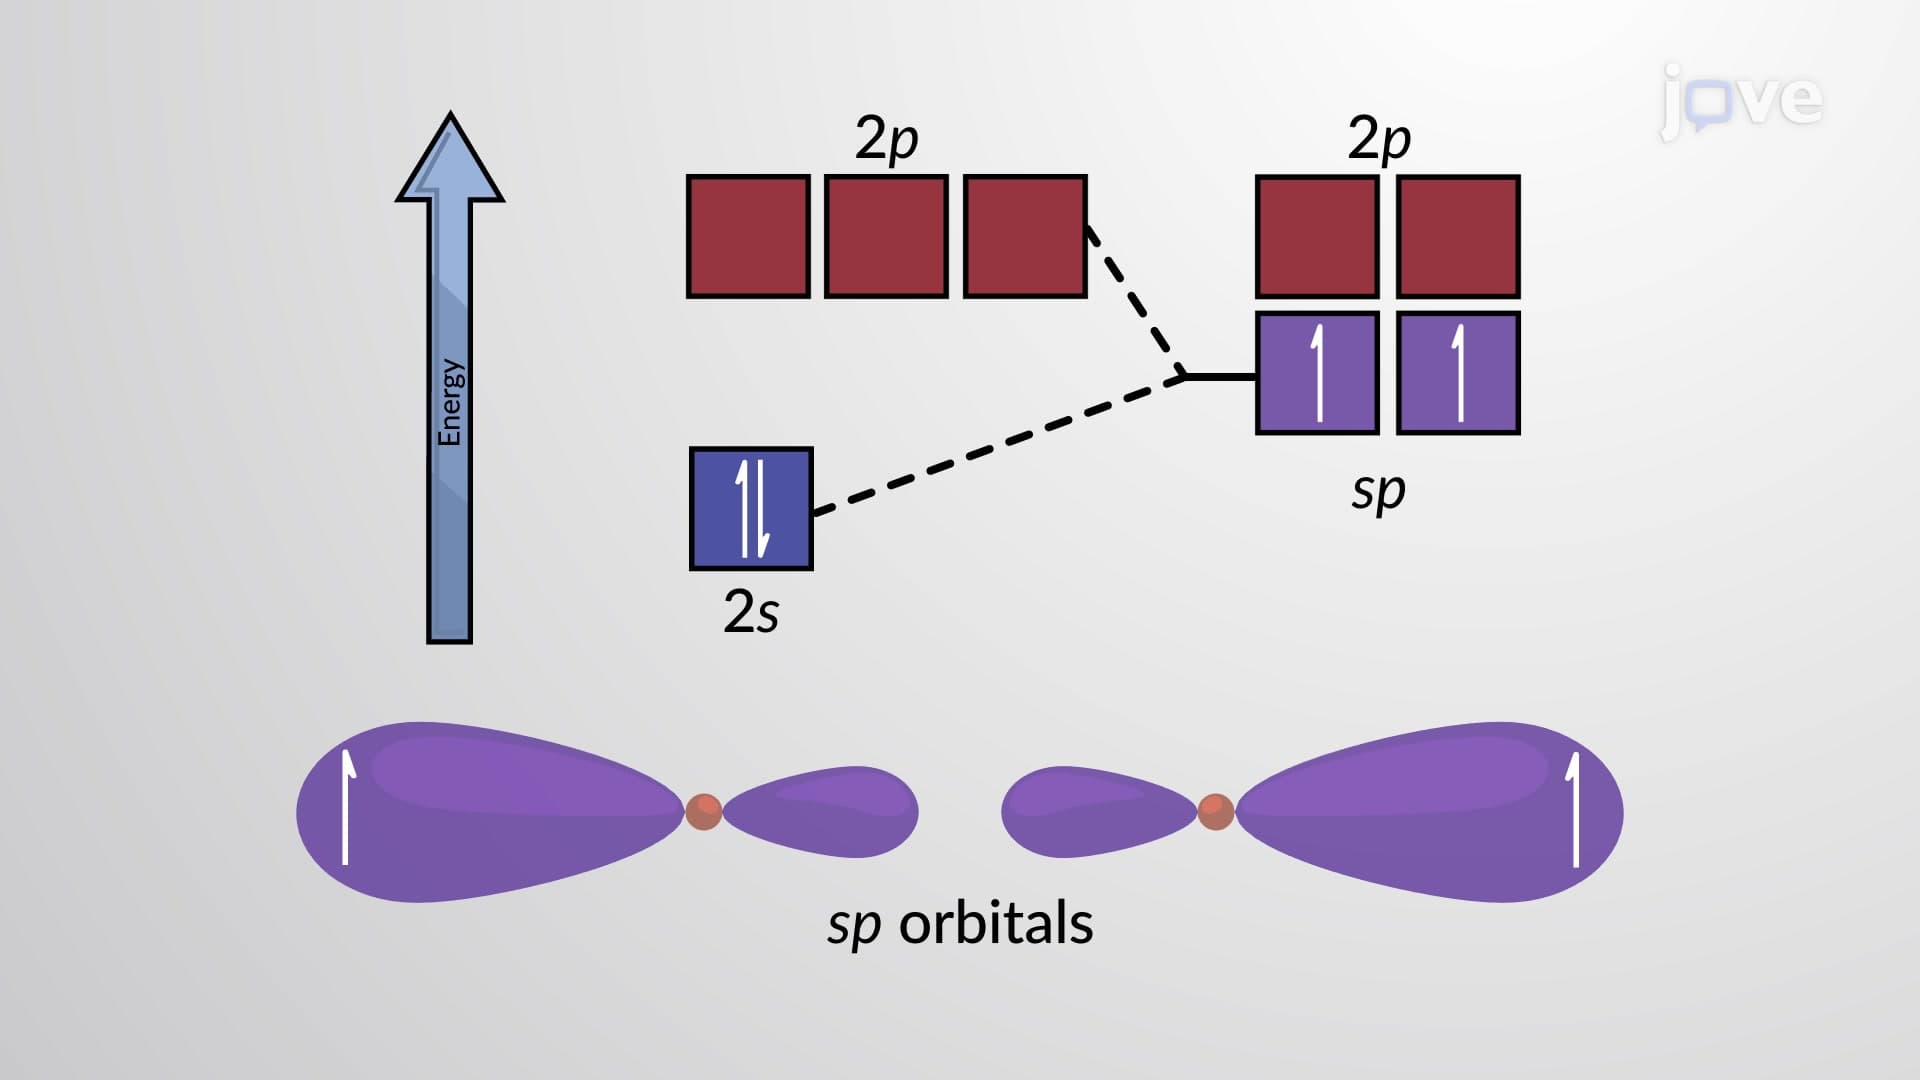
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:21
IR Absorption Frequency: Hybridization
Among the sp, sp2, and sp3 hybridized orbitals, sp orbitals have the maximum s character (50%). Consequently, the electrons are held more closely to the nucleus, resulting in stronger and shorter C–H bonds that...
 01:20
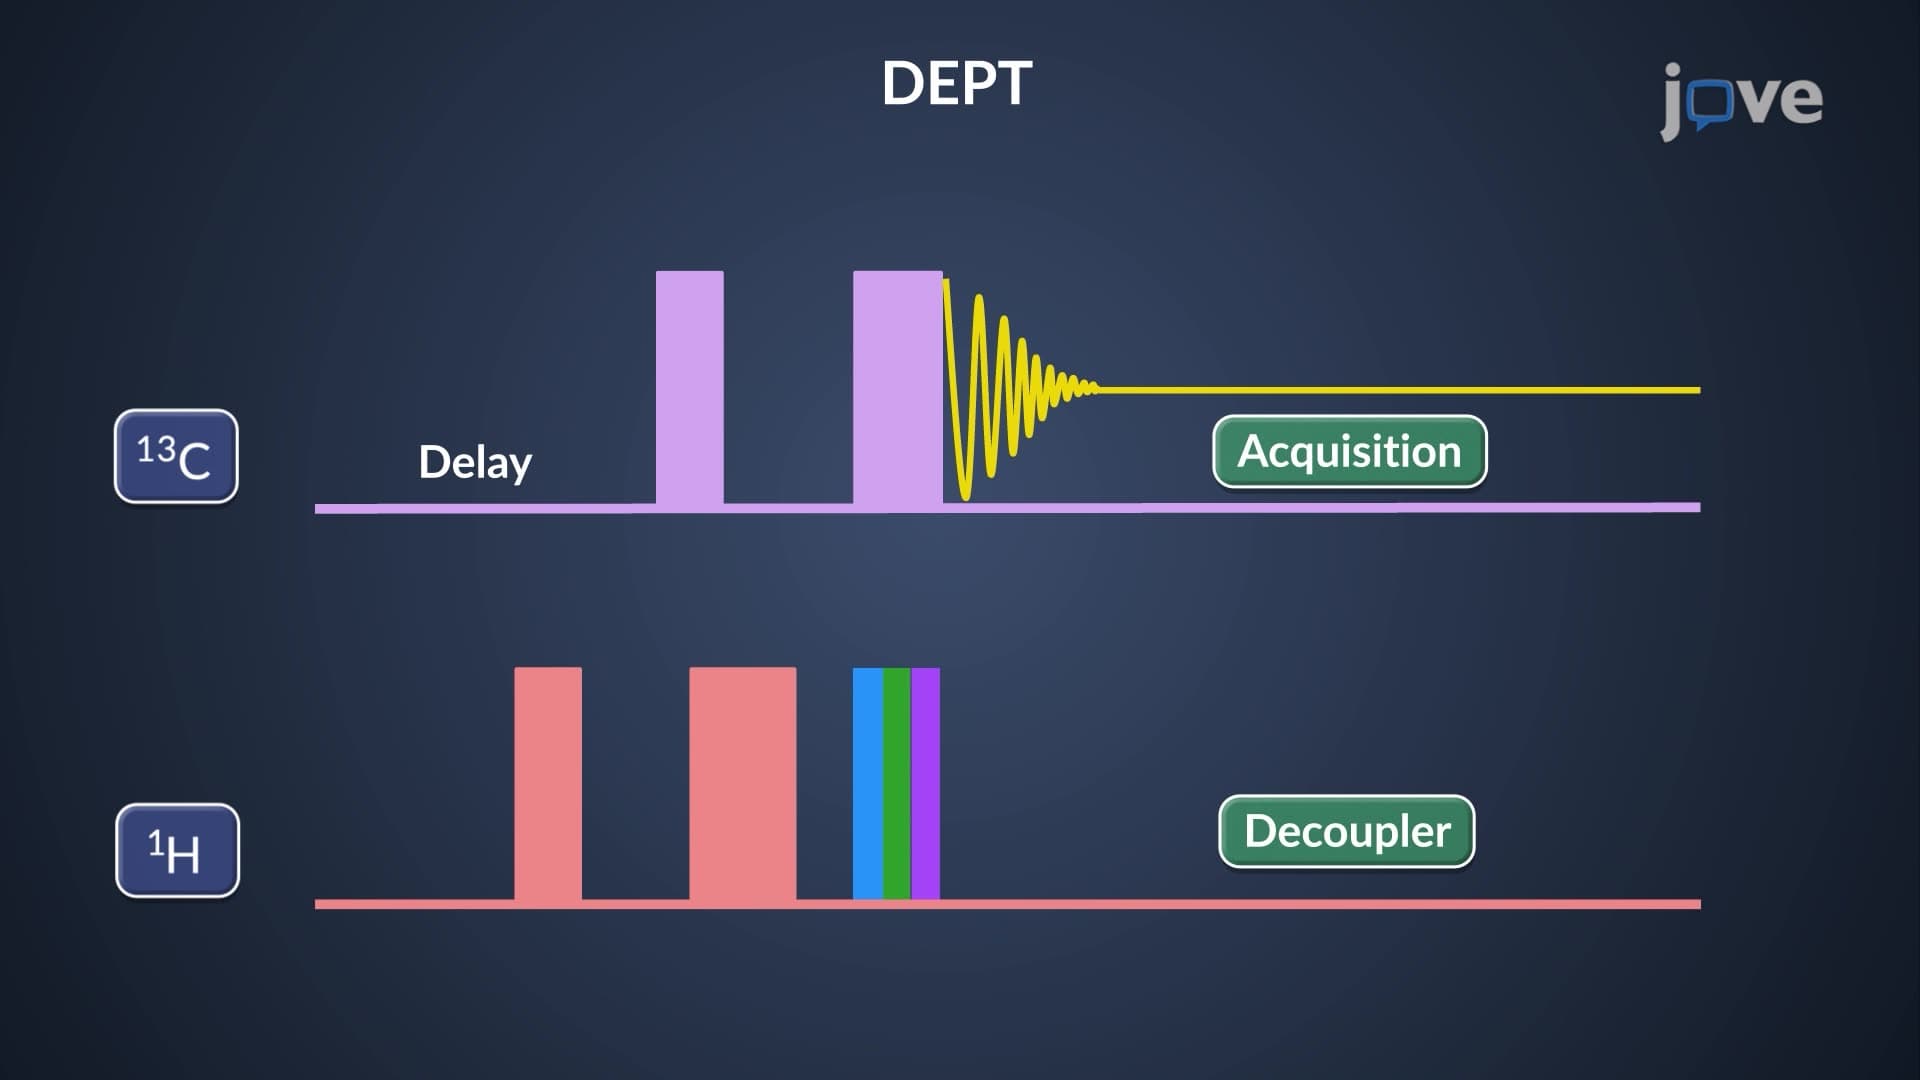
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 03:51
Molecular Orbital Theory II