Rosetta's Predictive Ability for Low-Affinity Ligand Binding in Fragment-Based Drug Discovery
Related Experiment Video
Updated: Aug 14, 2025

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Elleansar Okwei1,2, Shannon T Smith2,3, Brian J Bender2,4
1Department of Chemistry, Vanderbilt University, Nashville, Tennessee37235, United States.

Abstract:
Fragment-based drug discovery begins with the identification of small molecules with a molecular weight of usually less than 250 Da which weakly bind to the protein of interest. This technique is challenging for computational docking methods as binding is determined by only a few specific interactions. Inaccuracies in the energy function or slight deviations in the docking pose can lead to the prediction of incorrect binding or difficulties in ranking fragments in in silico screening. Here, we test RosettaLigand by docking a series of fragments to a cysteine-depleted variant of the TIM-barrel protein, HisF (UniProtKB Q9X0C6). We compare the computational results with experimental NMR spectroscopy screens. NMR spectroscopy gives details on binding affinities of individual ligands, which allows assessment of the ligand-ranking ability using RosettaLigand and also provides feedback on the location of the binding pocket, which serves as a reliable test of RosettaLigand's ability to identify plausible binding poses. From a library screen of 3456 fragments, we identified a set of 31 ligands with intrinsic affinities to HisF with dissociation constants as low as 400 μM. The same library of fragments was blindly screened in silico. RosettaLigand was able to rank binders before non-binders with an area under the curve of the receiver operating characteristics of 0.74. The docking poses observed for binders agreed with the binding pocket identified by NMR chemical shift perturbations for all fragments. Taken together, these results provide a baseline performance of RosettaLigand in a fragment-based drug discovery setting.

Related Concept Videos
 02:40
Ligand Binding Sites
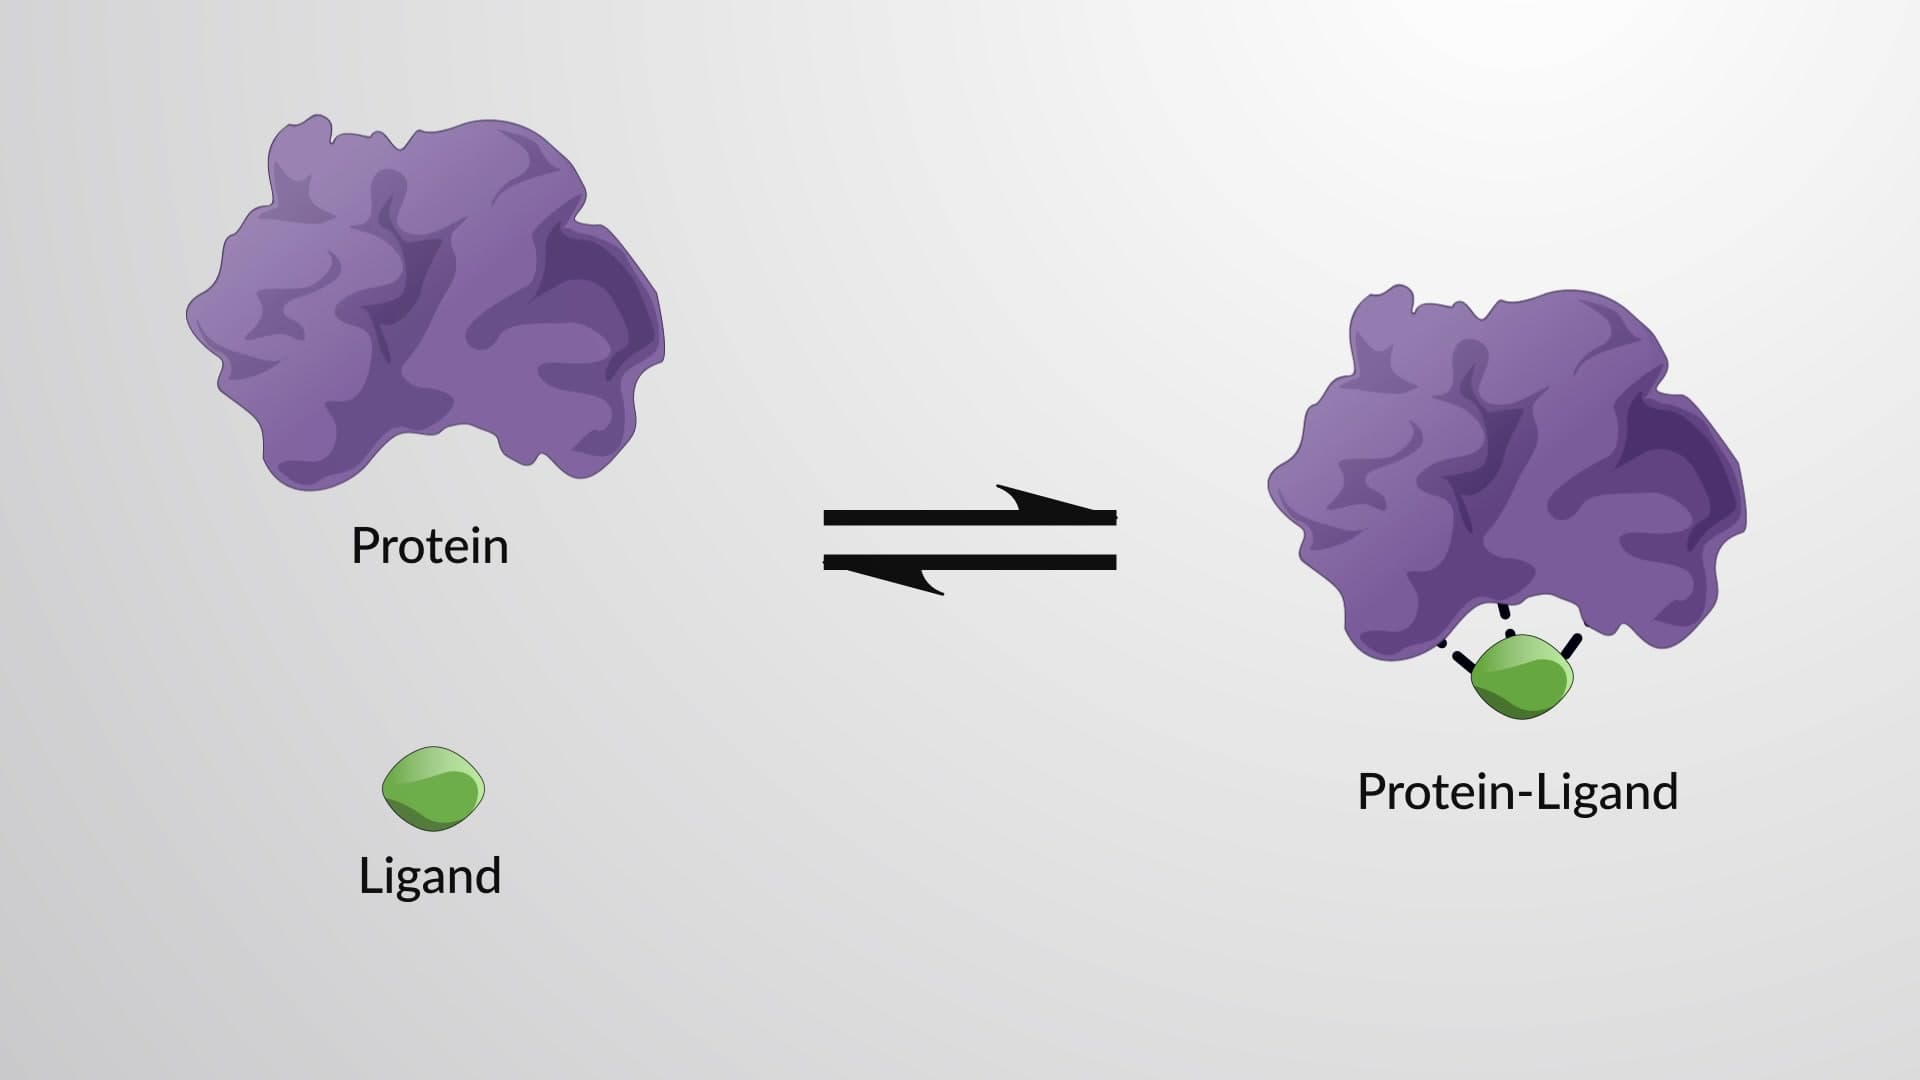
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...