Related Experiment Video
Updated: Aug 14, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Efficient implementation of time-dependent auxiliary density functional theory
Luis I Hernández-Segura1, Andreas M Köster1
1Departamento de Química, Cinvestav, Avenida Instituto Politécnico Nacional 2508, A.P. 14-740, CDMX C.P. 07360, Mexico.

Abstract:
The random phase approximation of time-dependent auxiliary density functional theory (TDADFT) is rederived from auxiliary density perturbation theory. Our exhaustive validation of TDADFT reveals an upshift of the excitation energies by ∼0.1 eV with respect to standard time-dependent density functional theory. For the computationally efficient implementation of TDADFT, floating point operation optimized three-center electron repulsion integral recurrence relations and their double asymptotic expansions are implemented into the Davidson solver. The computational efficiency of TDADFT is benchmarked with four sets of molecules comprising alkanes, fullerenes, DNA fragments, and zeolites. The results show that TDADFT has a computational scaling between 1.3 and 1.9 with respect to the number of basis functions, which is lower than the scaling of standard time-dependent density functional theory. Due to its computational simplifications, TDADFT is particularly well suited for Born-Oppenheimer molecular dynamics simulations. As illustrative examples, we present the temperature effects on the gas-phase absorption spectra of benzene, naphthalene, and anthracene.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:23
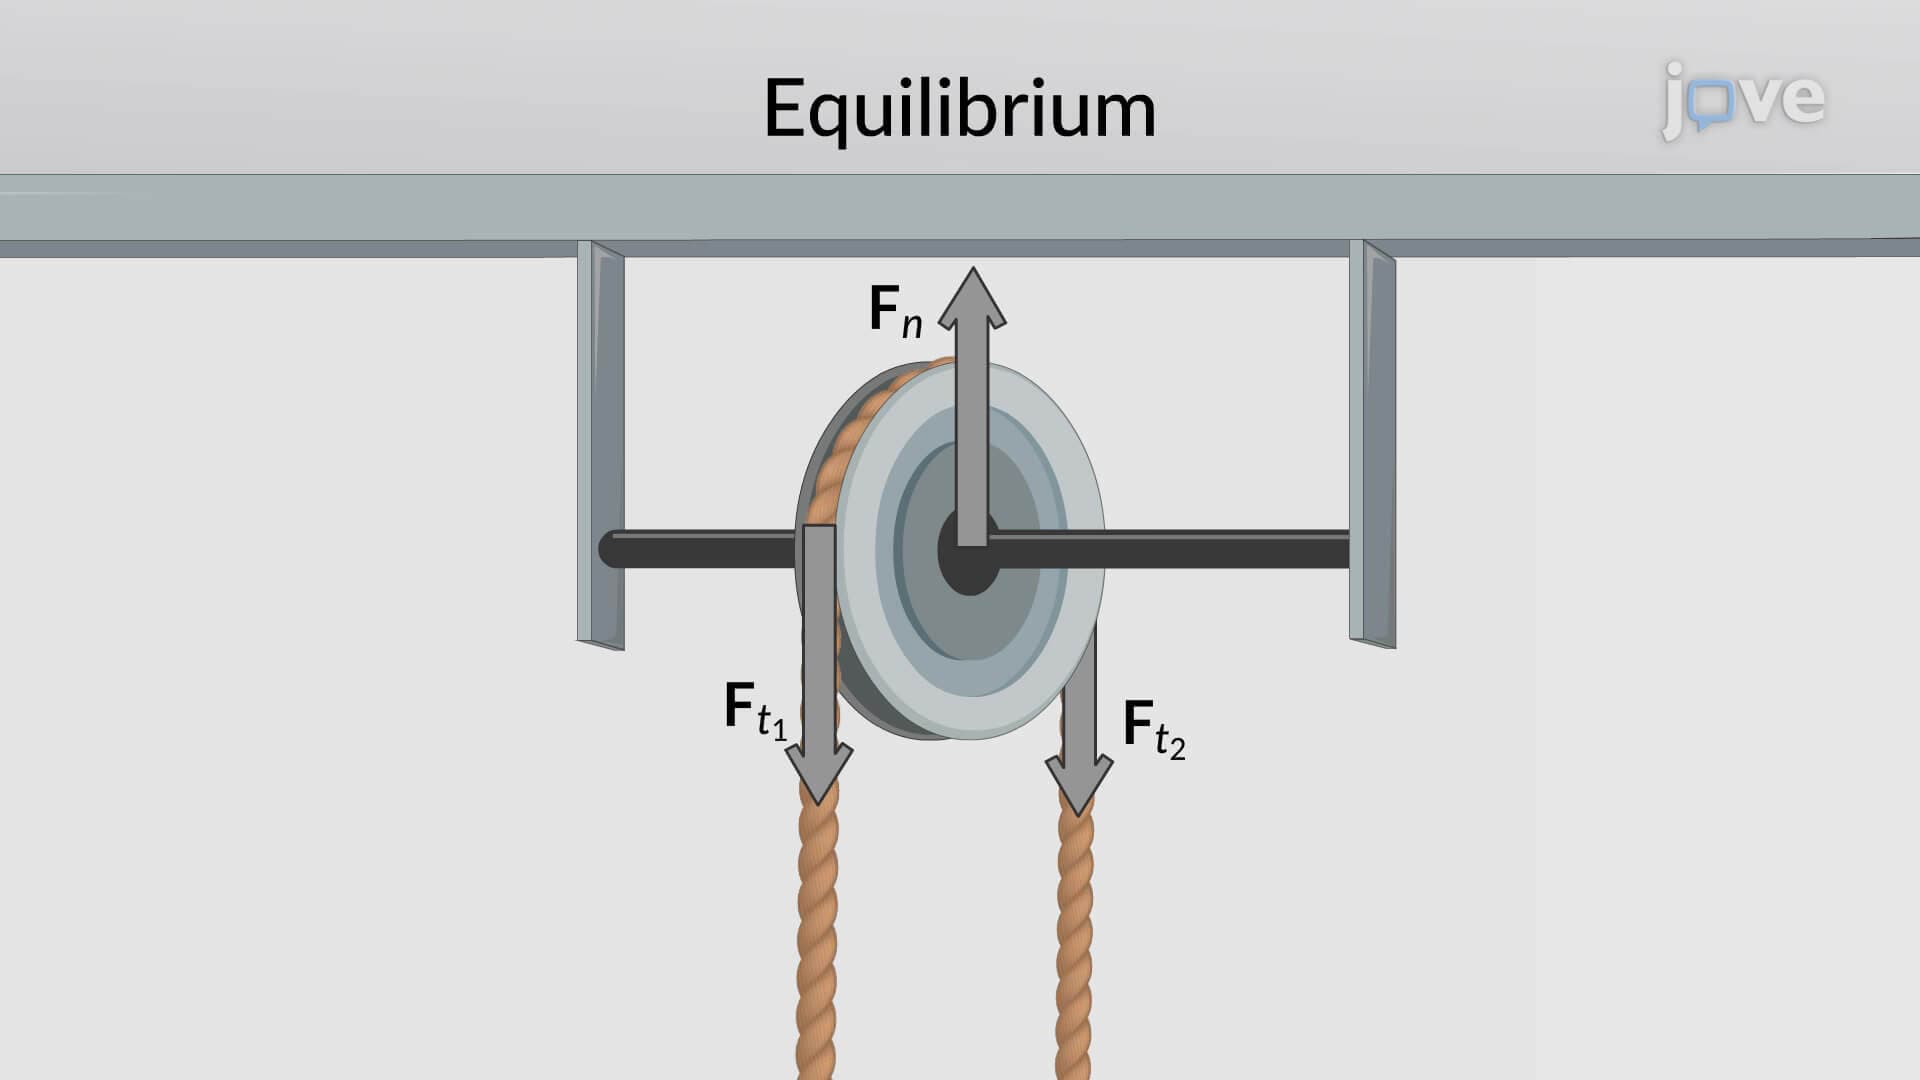
Equilibrium Conditions for a Particle
To understand the concept of equilibrium, let us first consider the forces acting on an object. When different forces act on an object, they can...
 02:39
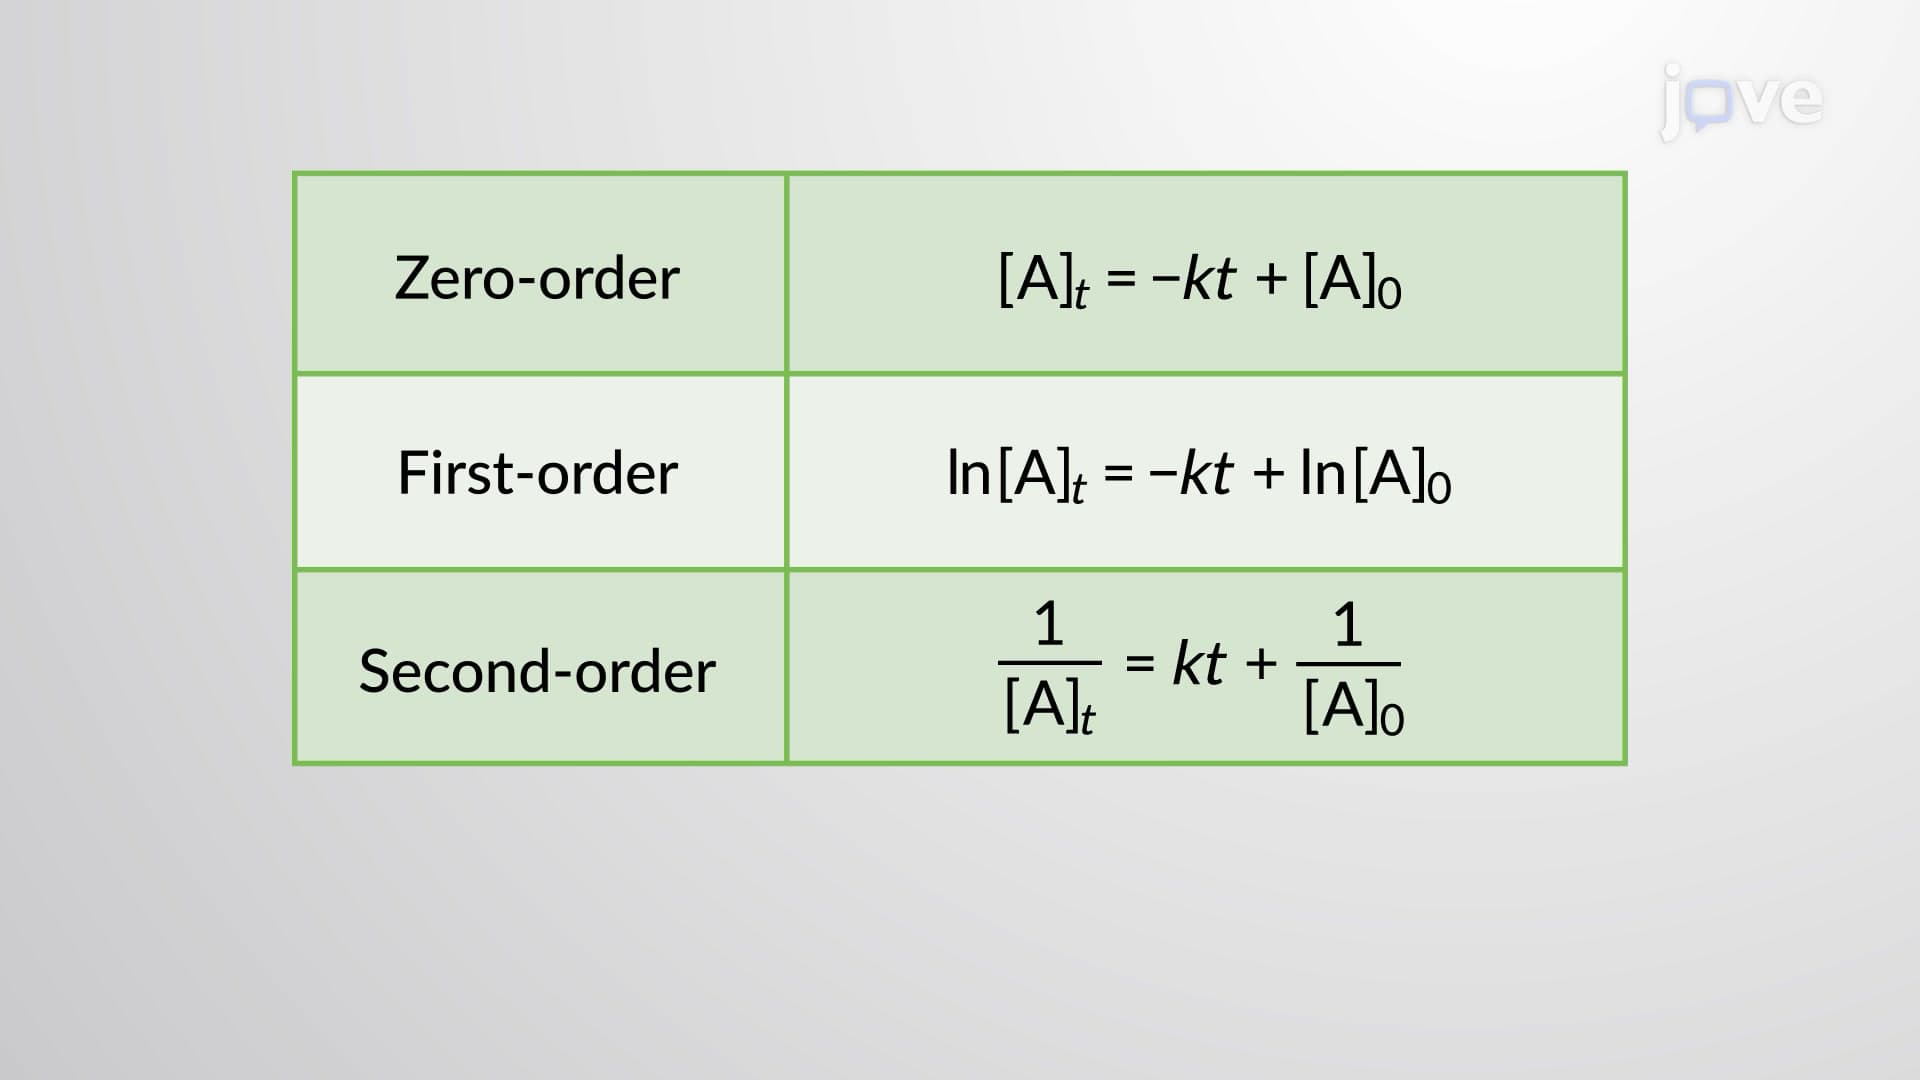
The Integrated Rate Law: The Dependence of Concentration on Time
 01:12
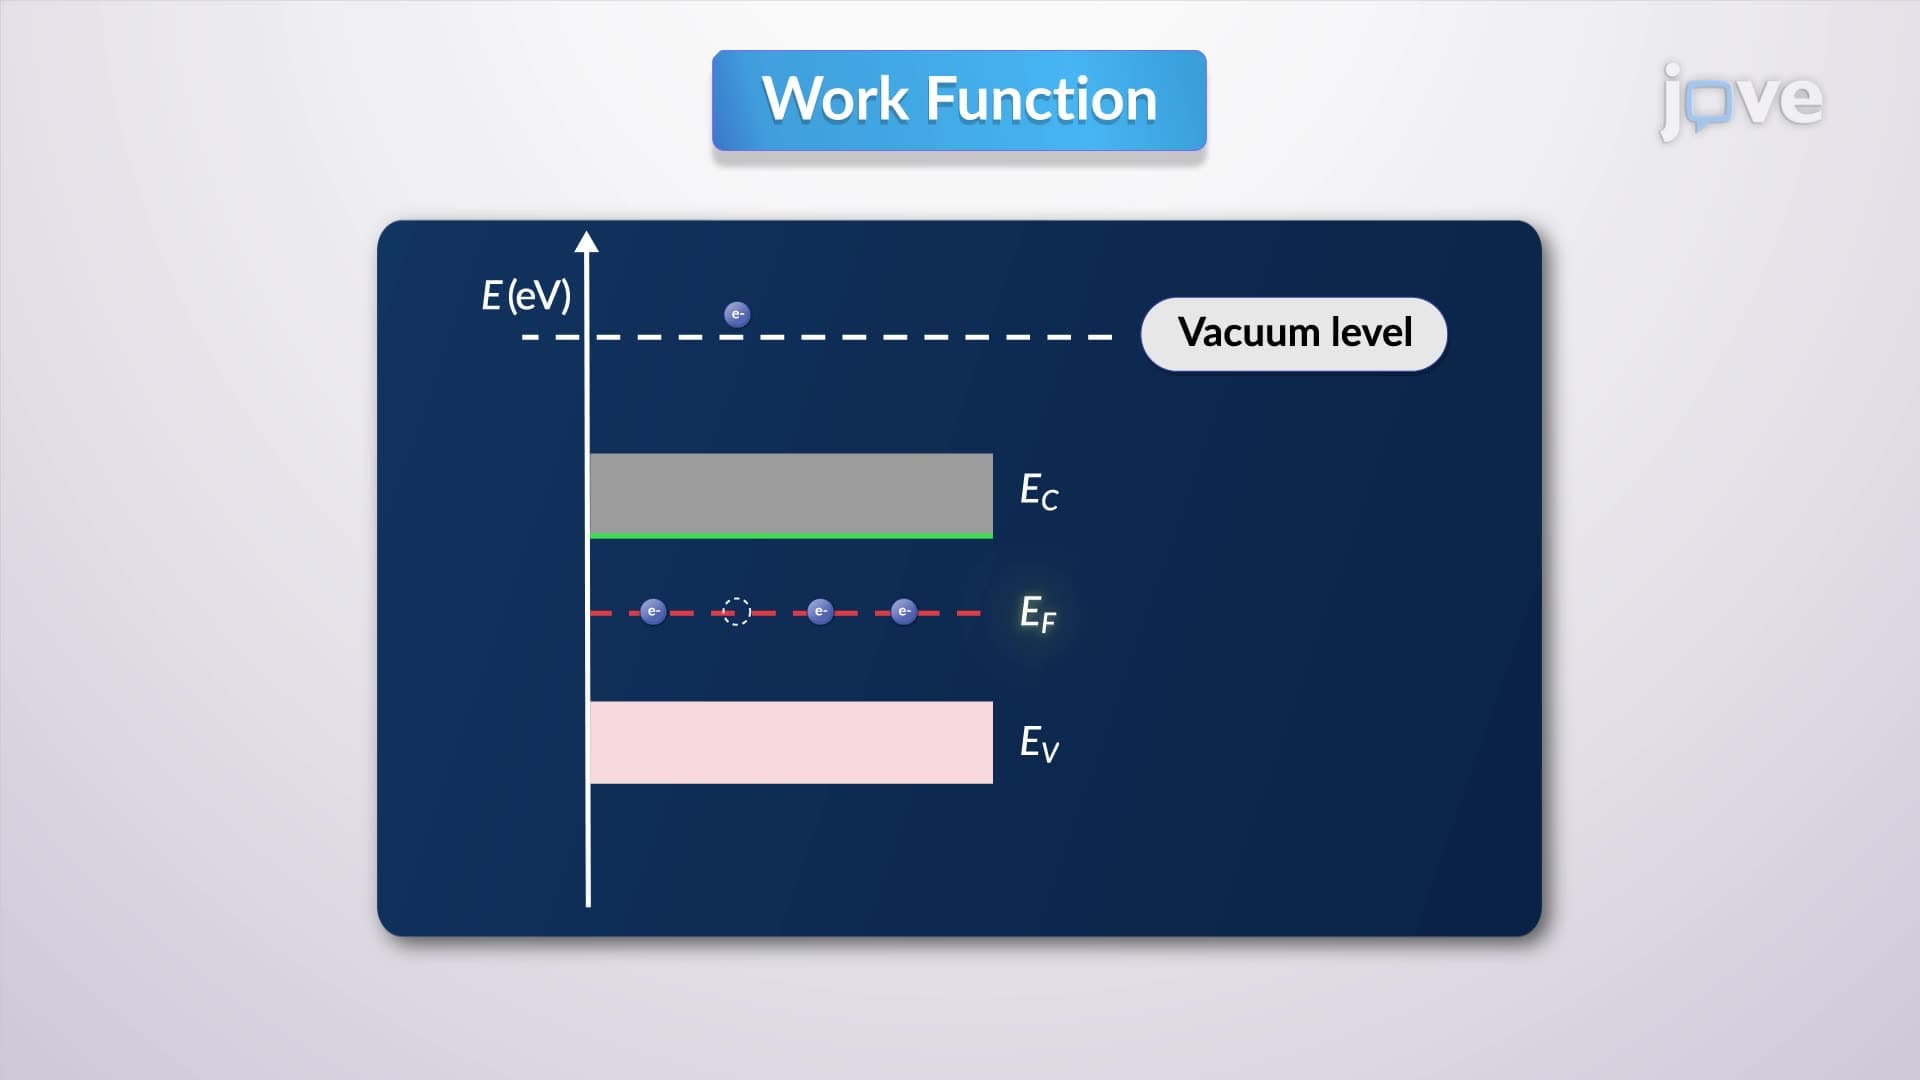
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:26
Thermodynamic Potentials
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
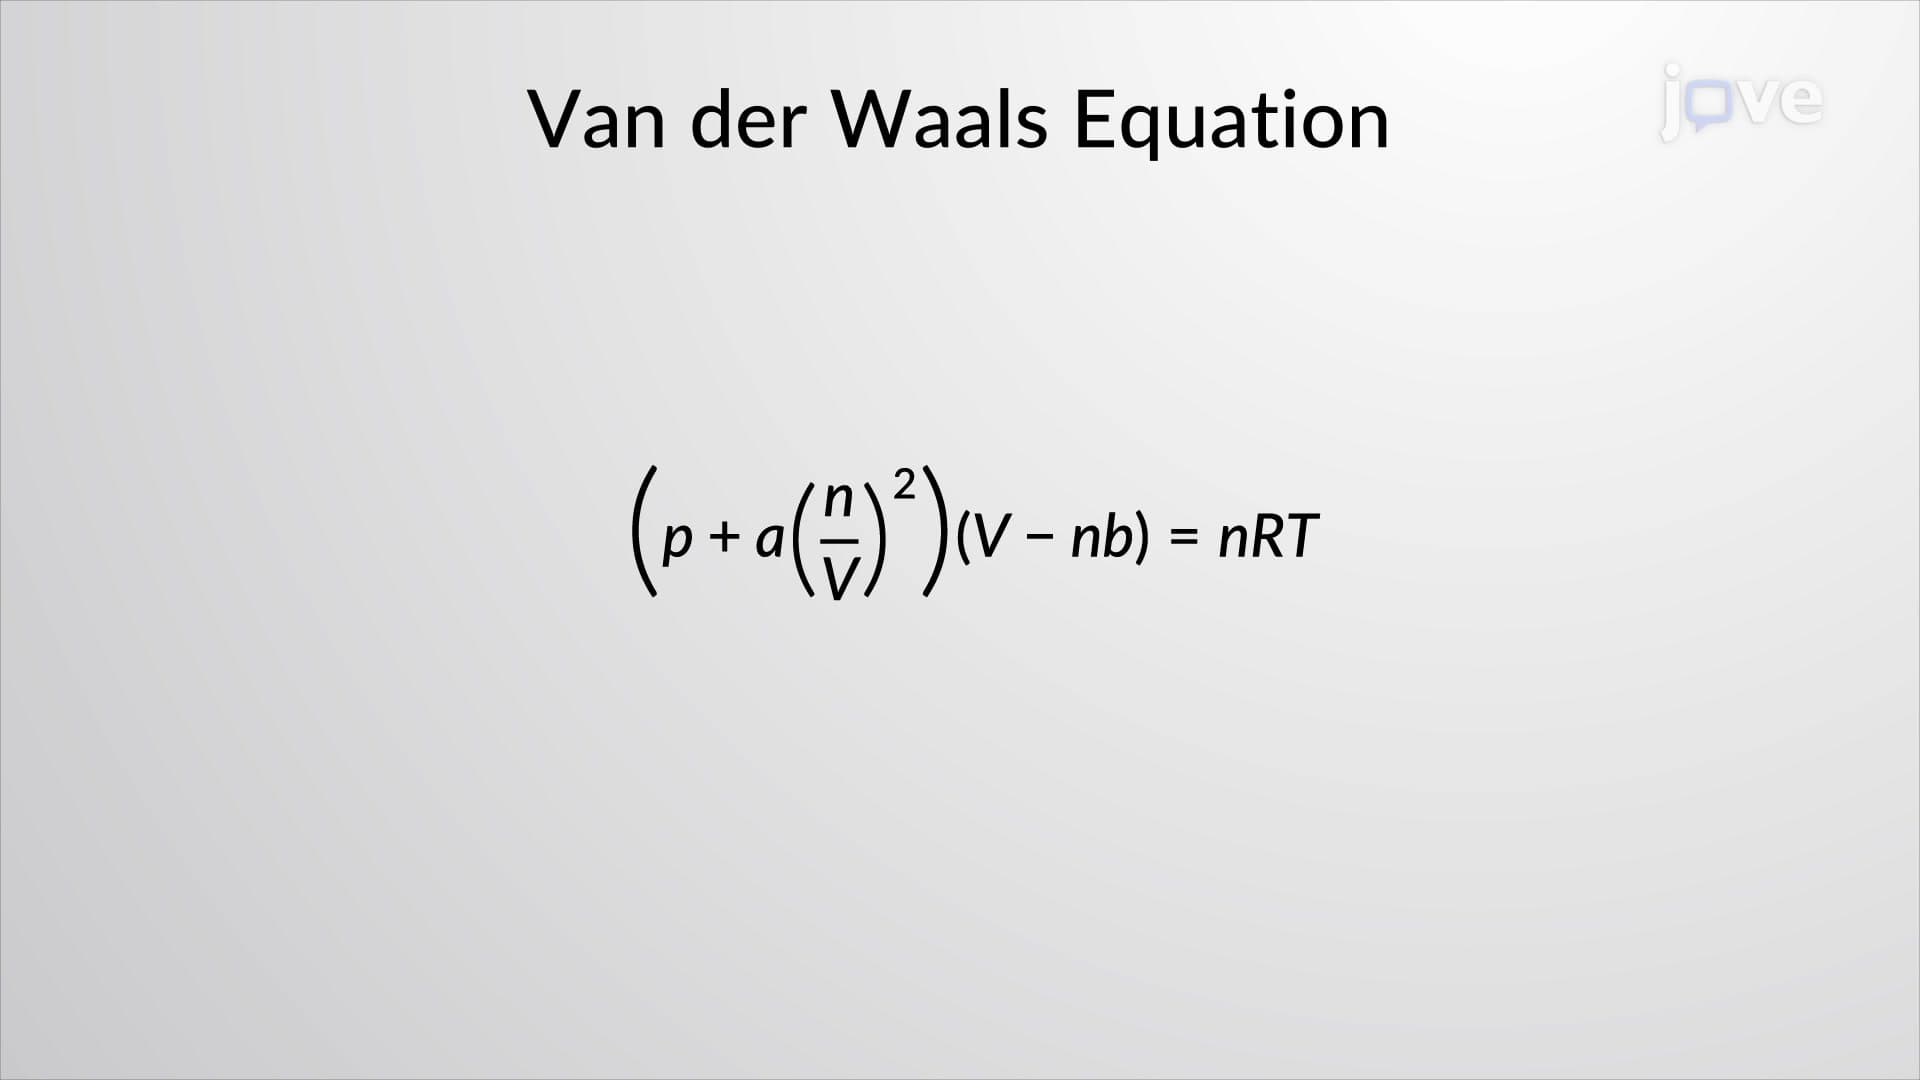
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...