Advancing the field of computational drug design using multicanonical molecular dynamics-based dynamic docking
Related Experiment Video
Updated: Aug 13, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Gert-Jan Bekker1, Narutoshi Kamiya2
1Institute for Protein Research, Osaka University, 3-2 Yamadaoka, Suita, Osaka 565-0871 Japan.

Abstract:
Multicanonical molecular dynamics (McMD)-based dynamic docking is a powerful tool to not only predict the native binding configuration between two flexible molecules, but it can also be used to accurately simulate the binding/unbinding pathway. Furthermore, it can also predict alternative binding sites, including allosteric ones, by employing an exhaustive sampling approach. Since McMD-based dynamic docking accurately samples binding/unbinding events, it can thus be used to determine the molecular mechanism of binding between two molecules. We developed the McMD-based dynamic docking methodology based on the powerful, but woefully underutilized McMD algorithm, combined with a toolset to perform the docking and to analyze the results. Here, we showcase three of our recent works, where we have applied McMD-based dynamic docking to advance the field of computational drug design. In the first case, we applied our method to perform an exhaustive search between Hsp90 and one of its inhibitors to successfully predict the native binding configuration in its binding site, as we refined our analysis methods. For our second case, we performed an exhaustive search of two medium-sized ligands and Bcl-xL, which has a cryptic binding site that differs greatly between the apo and holo structures. Finally, we performed a dynamic docking simulation between a membrane-embedded GPCR molecule and a high affinity ligand that binds deep within its receptor's pocket. These advanced simulations showcase the power that the McMD-based dynamic docking method has, and provide a glimpse of the potential our methodology has to unravel and solve the medical and biophysical issues in the modern world.
Supplementary Information:
The online version contains supplementary material available at 10.1007/s12551-022-01010-z.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
 08:15
Author Spotlight: Network Pharmacology and Molecular Docking to Decipher the Action of Jiawei Shengjiang San Against Diabetic Kidney Disease
Published on: May 10, 2024
Related Concept Videos
 01:16
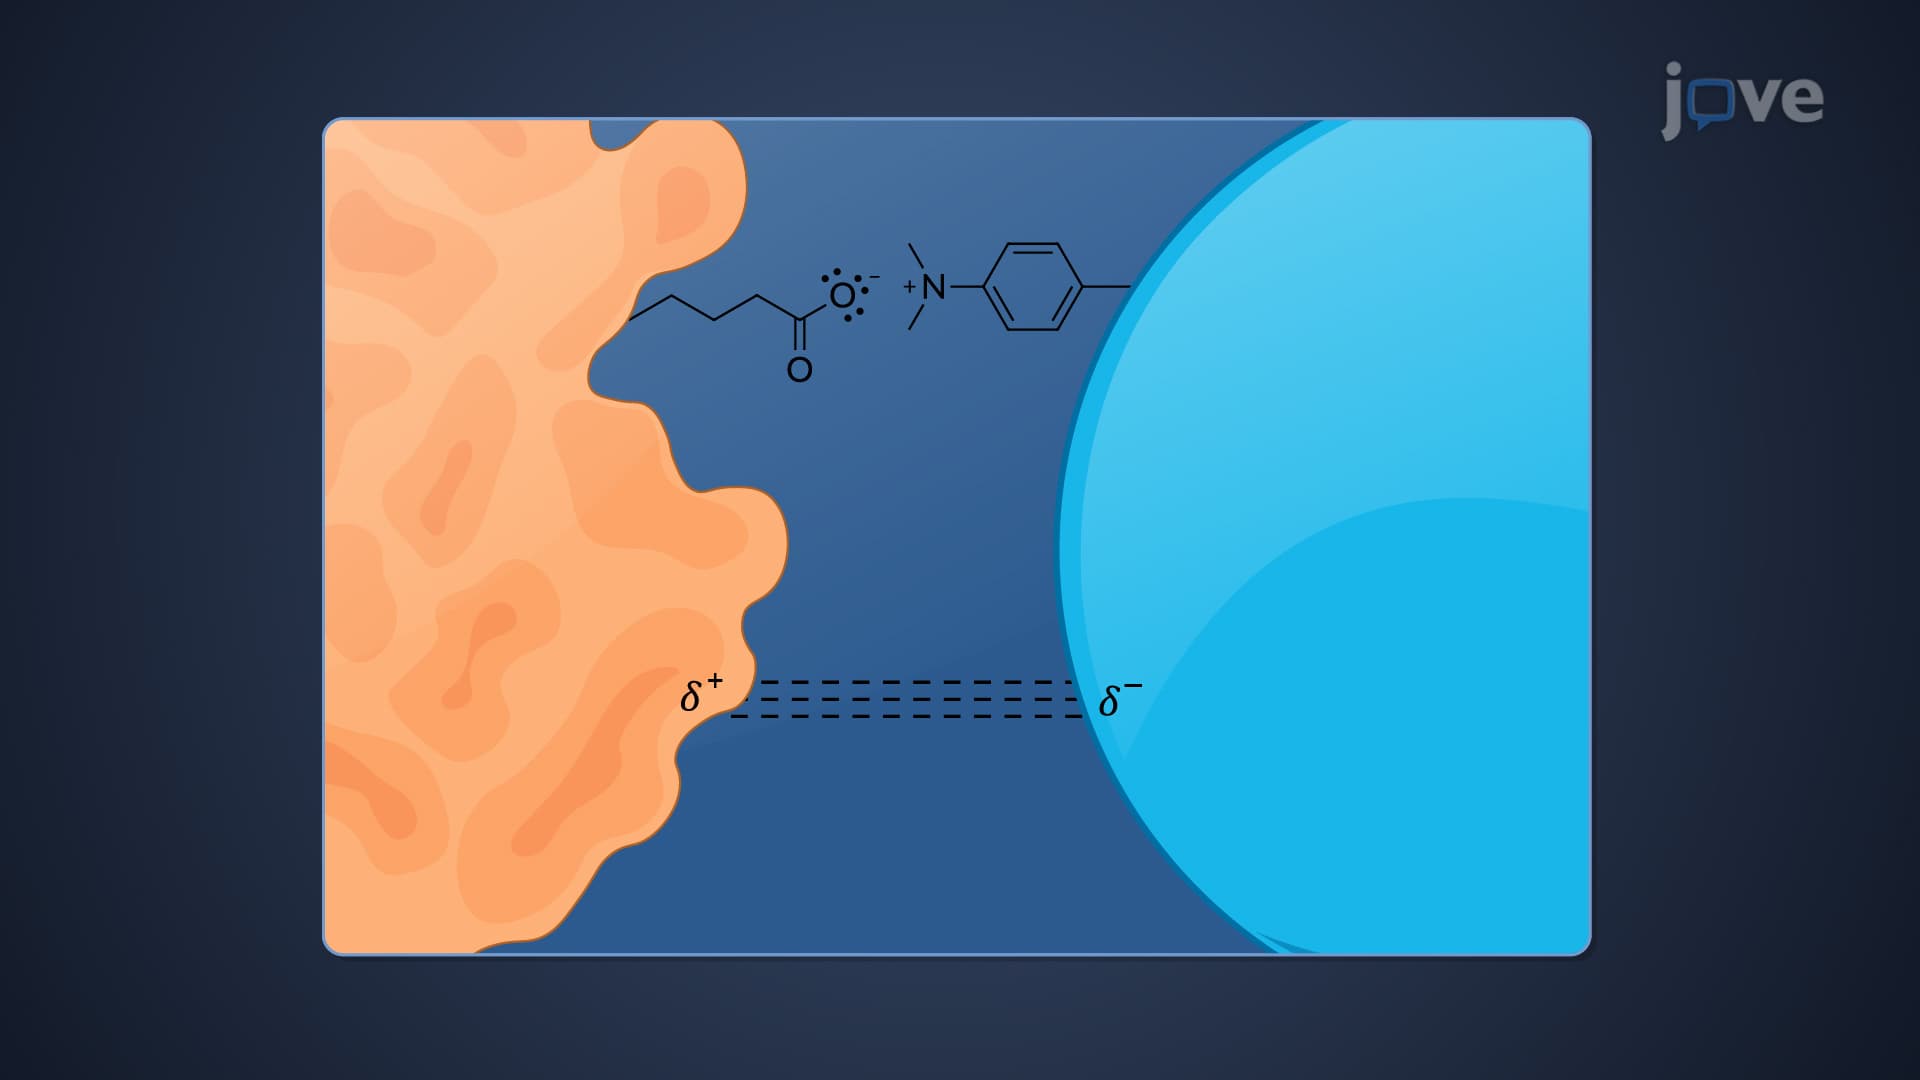
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:22
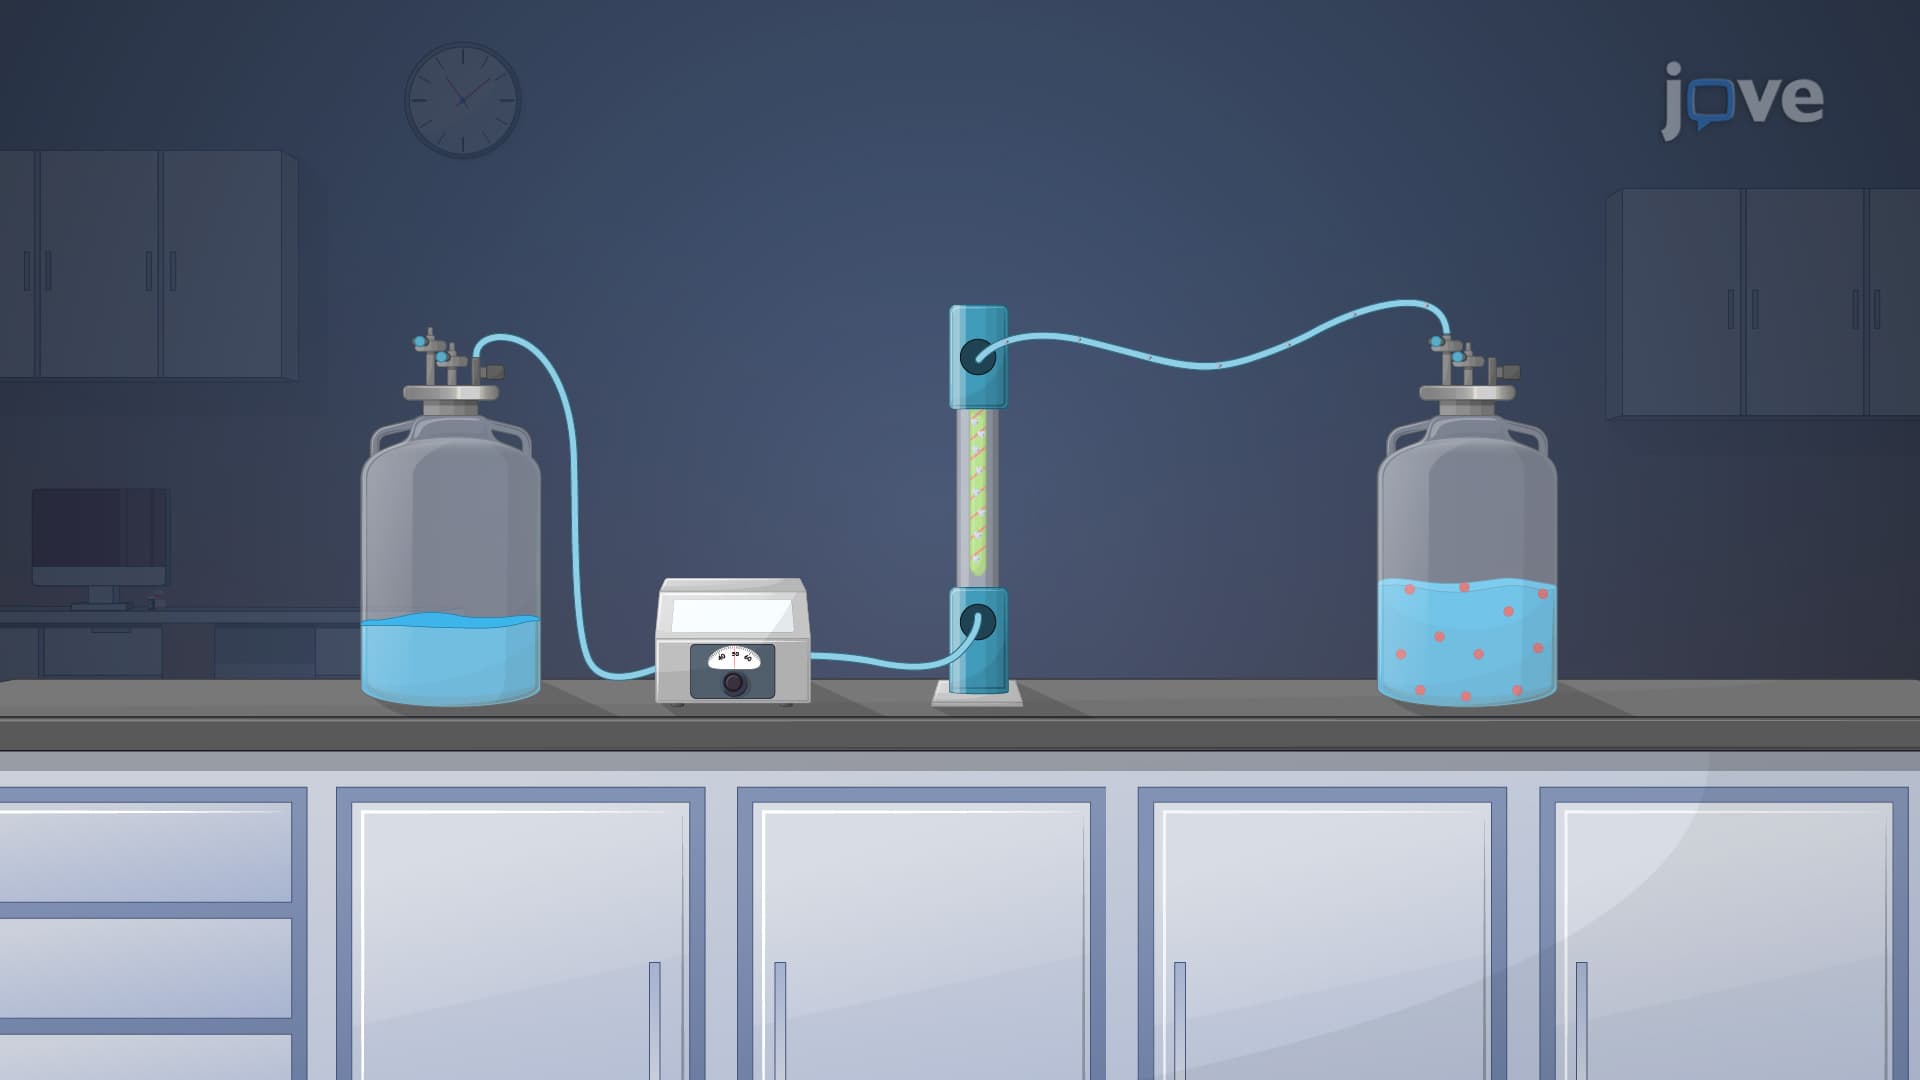
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:00
Molecular Models
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength