Related Experiment Video
Updated: Jun 19, 2026

Rapid Analysis of Chromosome Aberrations in Mouse B Lymphocytes by PNA-FISH
Published on: August 19, 2014
A cloud-based resource for genome coordinate-based exploration and large-scale analysis of chromosome aberrations and
Janet Wang1, Jeanne Zheng2, Elaine E Lee3
1Department of Genetics and Genome Sciences, Case Western Reserve University, Cleveland, Ohio, USA.

Abstract:
Cytogenetic analysis provides important information on the genetic mechanisms of cancer. The Mitelman Database of Chromosome Aberrations and Gene Fusions in Cancer (Mitelman DB) is the largest catalog of acquired chromosome aberrations, presently comprising >70 000 cases across multiple cancer types. Although this resource has enabled the identification of chromosome abnormalities leading to specific cancers and cancer mechanisms, a large-scale, systematic analysis of these aberrations and their downstream implications has been difficult due to the lack of a standard, automated mapping from aberrations to genomic coordinates. We previously introduced CytoConverter as a tool that automates such conversions. CytoConverter has now been updated with improved interpretation of karyotypes and has been integrated with the Mitelman DB, providing a comprehensive mapping of the 70 000+ cases to genomic coordinates, as well as visualization of the frequencies of chromosomal gains and losses. Importantly, all CytoConverter-generated genomic coordinates are publicly available in Google BigQuery, a cloud-based data warehouse, facilitating data exploration and integration with other datasets hosted by the Institute for Systems Biology Cancer Gateway in the Cloud (ISB-CGC) Resource. We demonstrate the use of BigQuery for integrative analysis of Mitelman DB with other cancer datasets, including a comparison of the frequency of imbalances identified in Mitelman DB cases with those found in The Cancer Genome Atlas (TCGA) copy number datasets. This solution provides opportunities to leverage the power of cloud computing for low-cost, scalable, and integrated analysis of chromosome aberrations and gene fusions in cancer.
Related Concept Videos
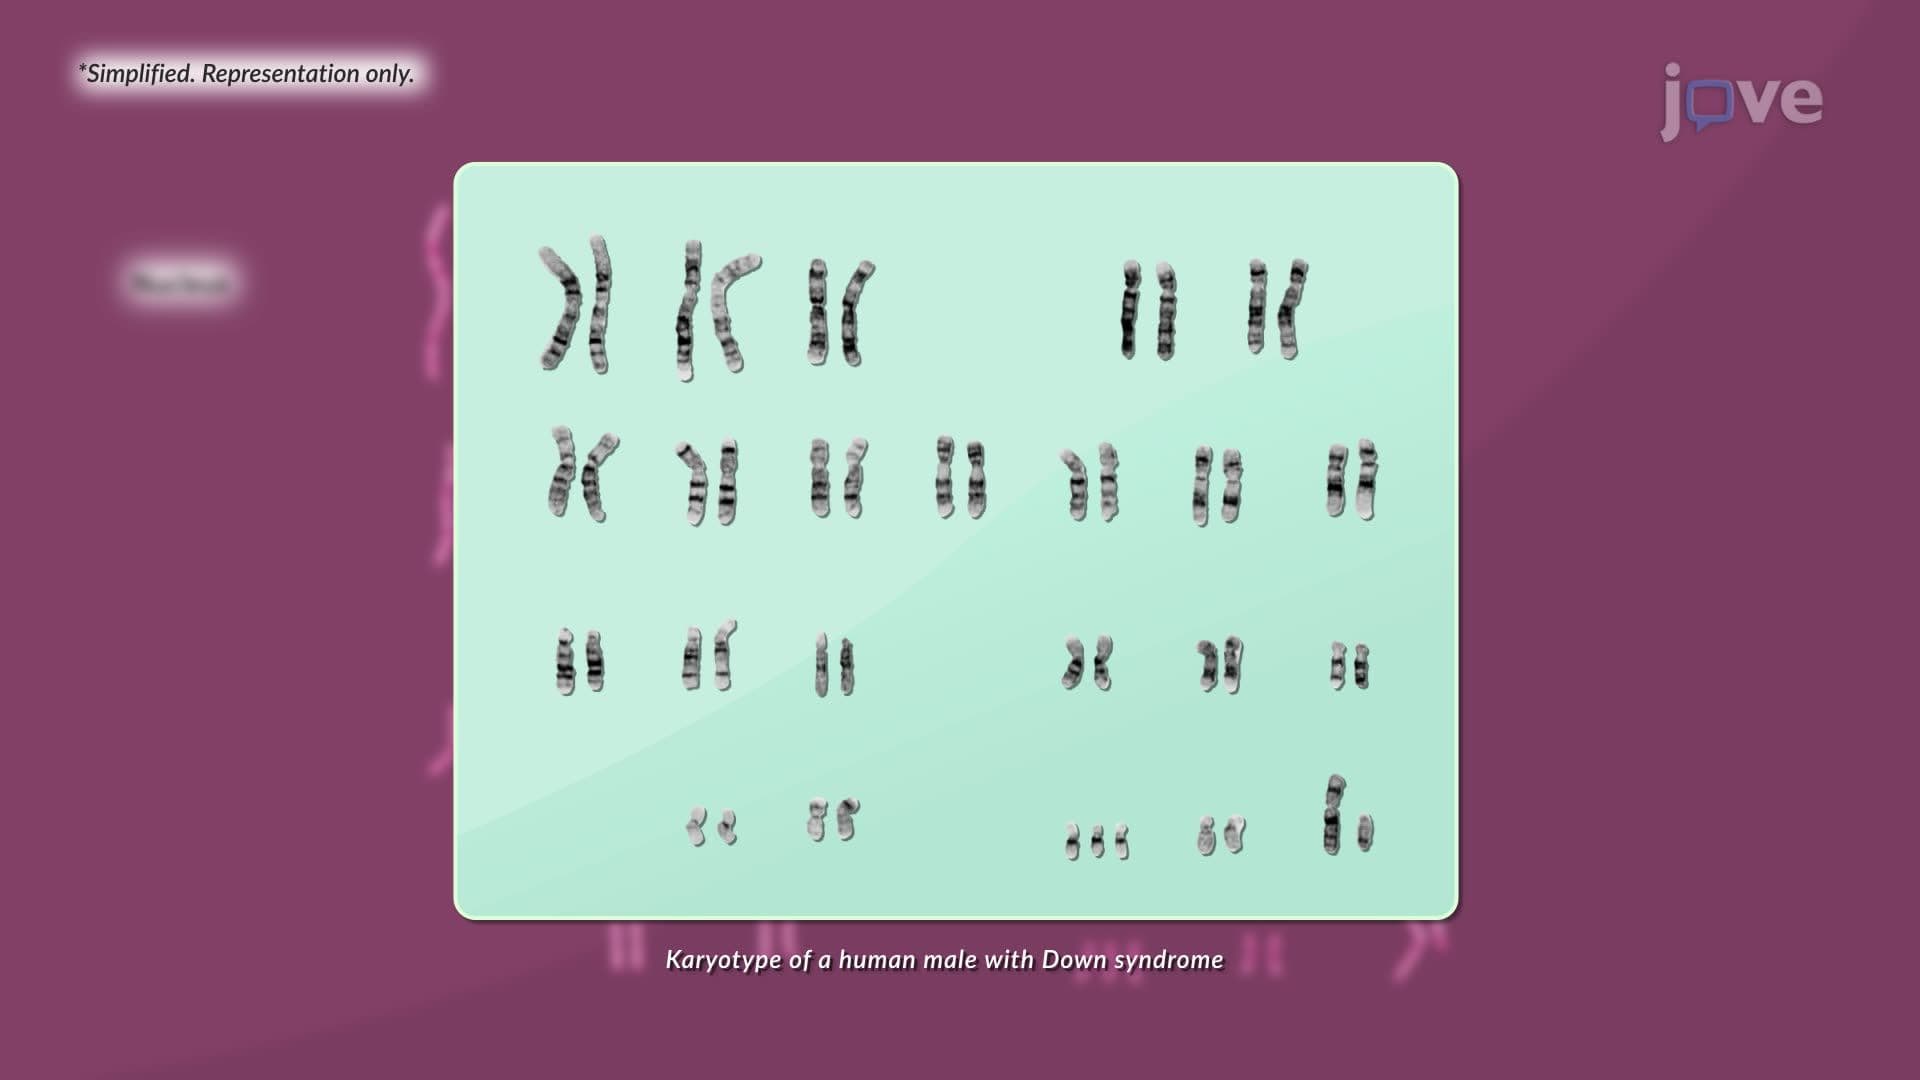 01:17
Karyotyping
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while microarray-based...