Related Experiment Video
Updated: Aug 11, 2025

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Molecular docking, 3D-QSAR and simulation studies for identifying pharmacophoric features of indole derivatives as
Swanand Kulkarni1, Yogesh Singh1, Avadh Biharee1
1Department of Pharmaceutical Sciences and Natural Products, Central University of Punjab, Bathinda, Punjab, India.

Abstract:
Excess of androgens leads to various diseases such as Poly-Cystic Ovarian Syndrome, Prostate Cancer, Hirsutism, Obesity and Acne. 17β-Hydroxysteroid Dehydrogenase type 5 (17β-HSD5) converts androstenedione into testosterone peripherally, thereby significantly contributing to the development of these diseases. Indole-bearing scaffolds are reported as potential 17β-HSD5 inhibitors for the manifestation of diseases arising due to androgen excess. In the present work, we have extensively performed a combination of molecular docking, Gaussian field-based 3D-QSAR, Pharmacophore mapping and MD-simulation studies (100 ns) to identify the pharmacophoric features of indole-based compounds as potent 17β-HSD5 inhibitors. Molecular simulation studies of the most potent compound in the binding pocket of enzyme revealed that the compound 11 was stable in the binding pocket and showed good binding affinity through interactions with various residues of active site pocket. The Molecular mechanics Generalized Born surface area continuum solvation (MM/GBSA) and Molecular mechanics Poisson-Boltzmann surface area (MM/PBSA) calculations revealed that the compound 11 possessed a free binding energy of -36.36 kcal/mol and -7.00 kcal/mol, respectively, which was better as compared to reference compound Desmethyl indomethacin (DES). The developed pharmacophore will be helpful to design novel indole-based molecules as potent 17β-HSD5 inhibitors for the treatment of various androgenic disorders.Communicated by Ramaswamy H. Sarma.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:26
Drug Discovery: Overview
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:16
Protein-Drug Binding: Mechanism and Kinetics
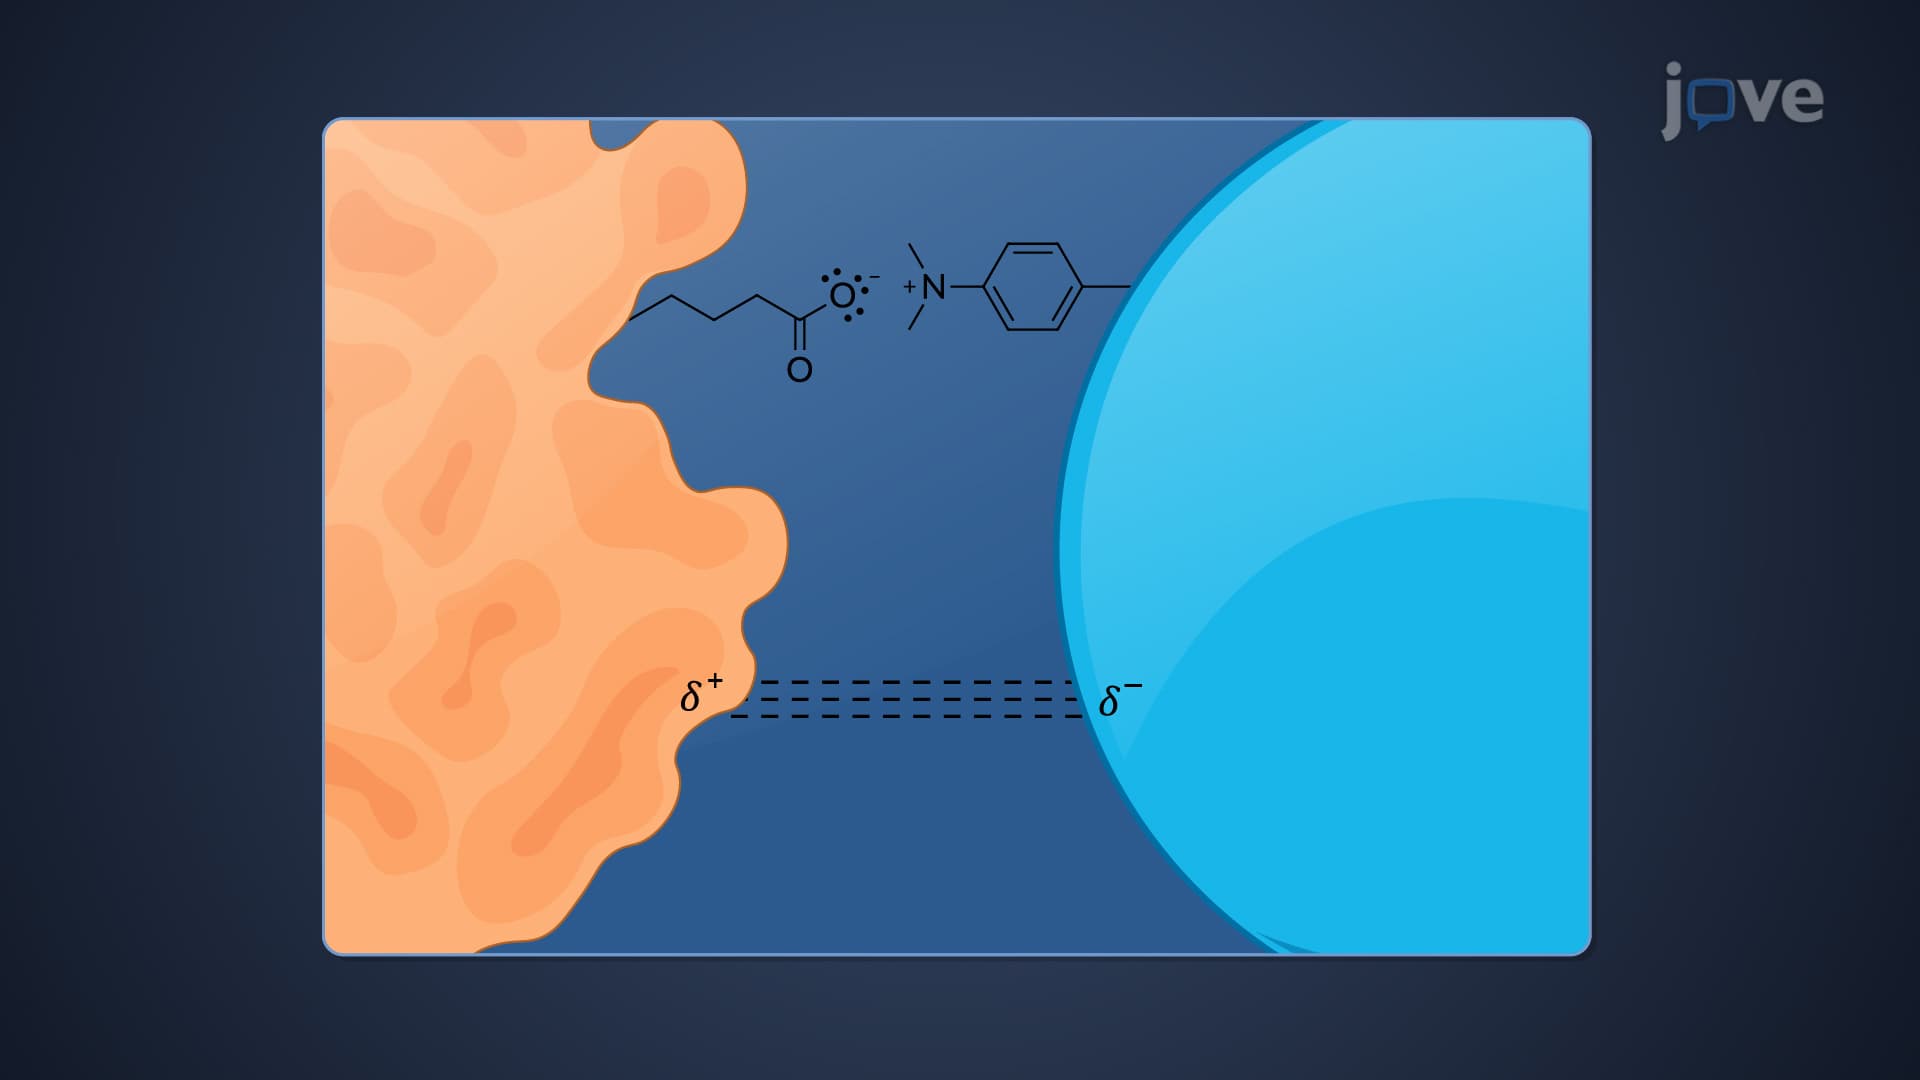
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
Protein-Drug Binding: Determination Methods
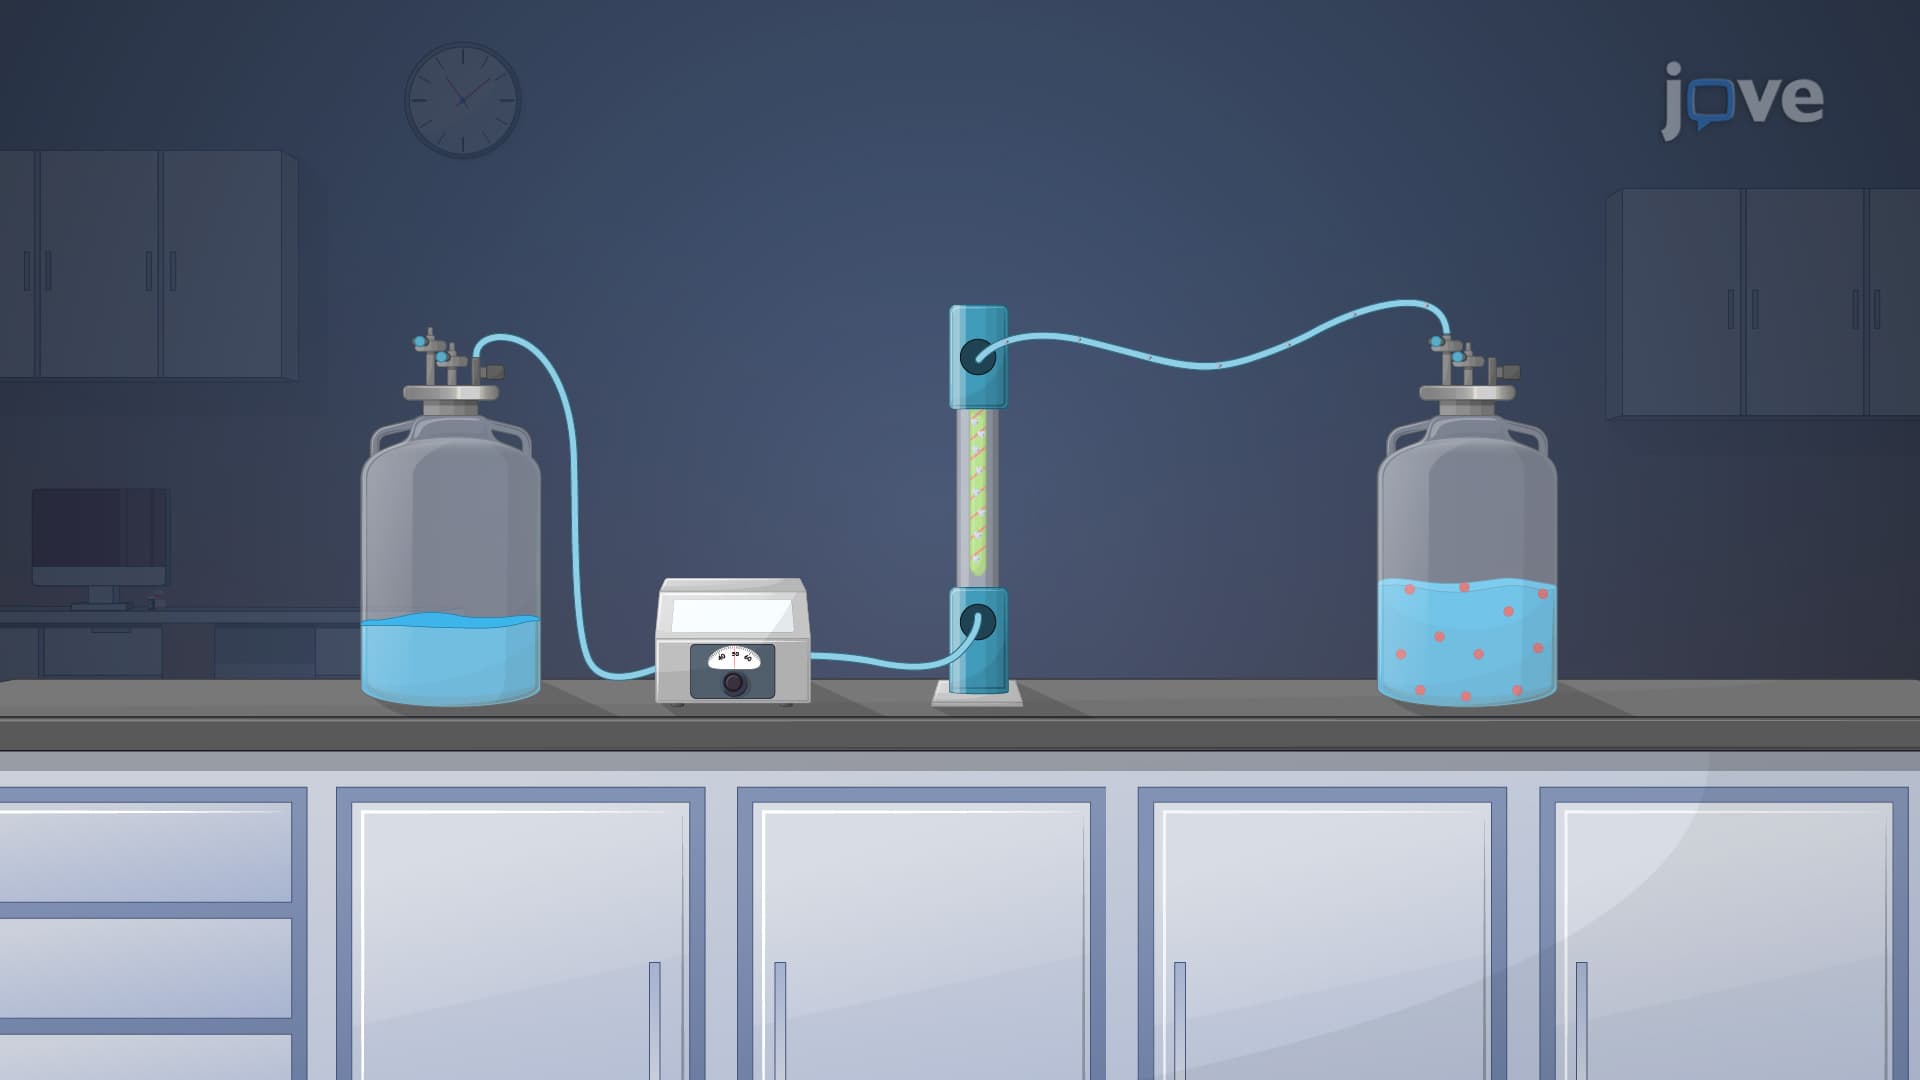
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...