Related Experiment Video
Updated: Aug 11, 2025

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Toward an efficient f-in-core/f-in-valence switchable description for DFTB calculations of Ce 4f states in ceria
Bojana Kocmaruk1, Akshay Krishna Ammothum Kandy1, Kersti Hermansson1
1Department of Chemistry - Ångström Laboratory, Uppsala University, Box 538, 751 21 Uppsala, Sweden.

Abstract:
A computational protocol is developed for efficient studies of partially reduced redox-active oxides using the self-consistent charge density functional tight-binding method. The protocol is demonstrated for ceria, which is a prototypical reducible oxide material. The underlying idea is to achieve a consistent (and harmonized) set of Slater-Koster (SK) tables with connected repulsive potentials that enable switching on and off the in-valence description of the Ce 4f states without serious loss of accuracy in structure and energetics. The implicit treatment of the Ce 4f states, with the use of f-in-core SK-tables, is found to lead to a significant decrease in computational time. More importantly, it allows for explicit control of the oxidation states of individual Ce atoms. This makes it possible to "freeze" the electronic configuration, thereby allowing the exploration of the energetics for various meta-stable configurations. We anticipate that the outlined strategy can help to shed light on the interplay between the size, shape, and redox activity for nanoceria and other related materials.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 06:53
Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Related Concept Videos
 03:26
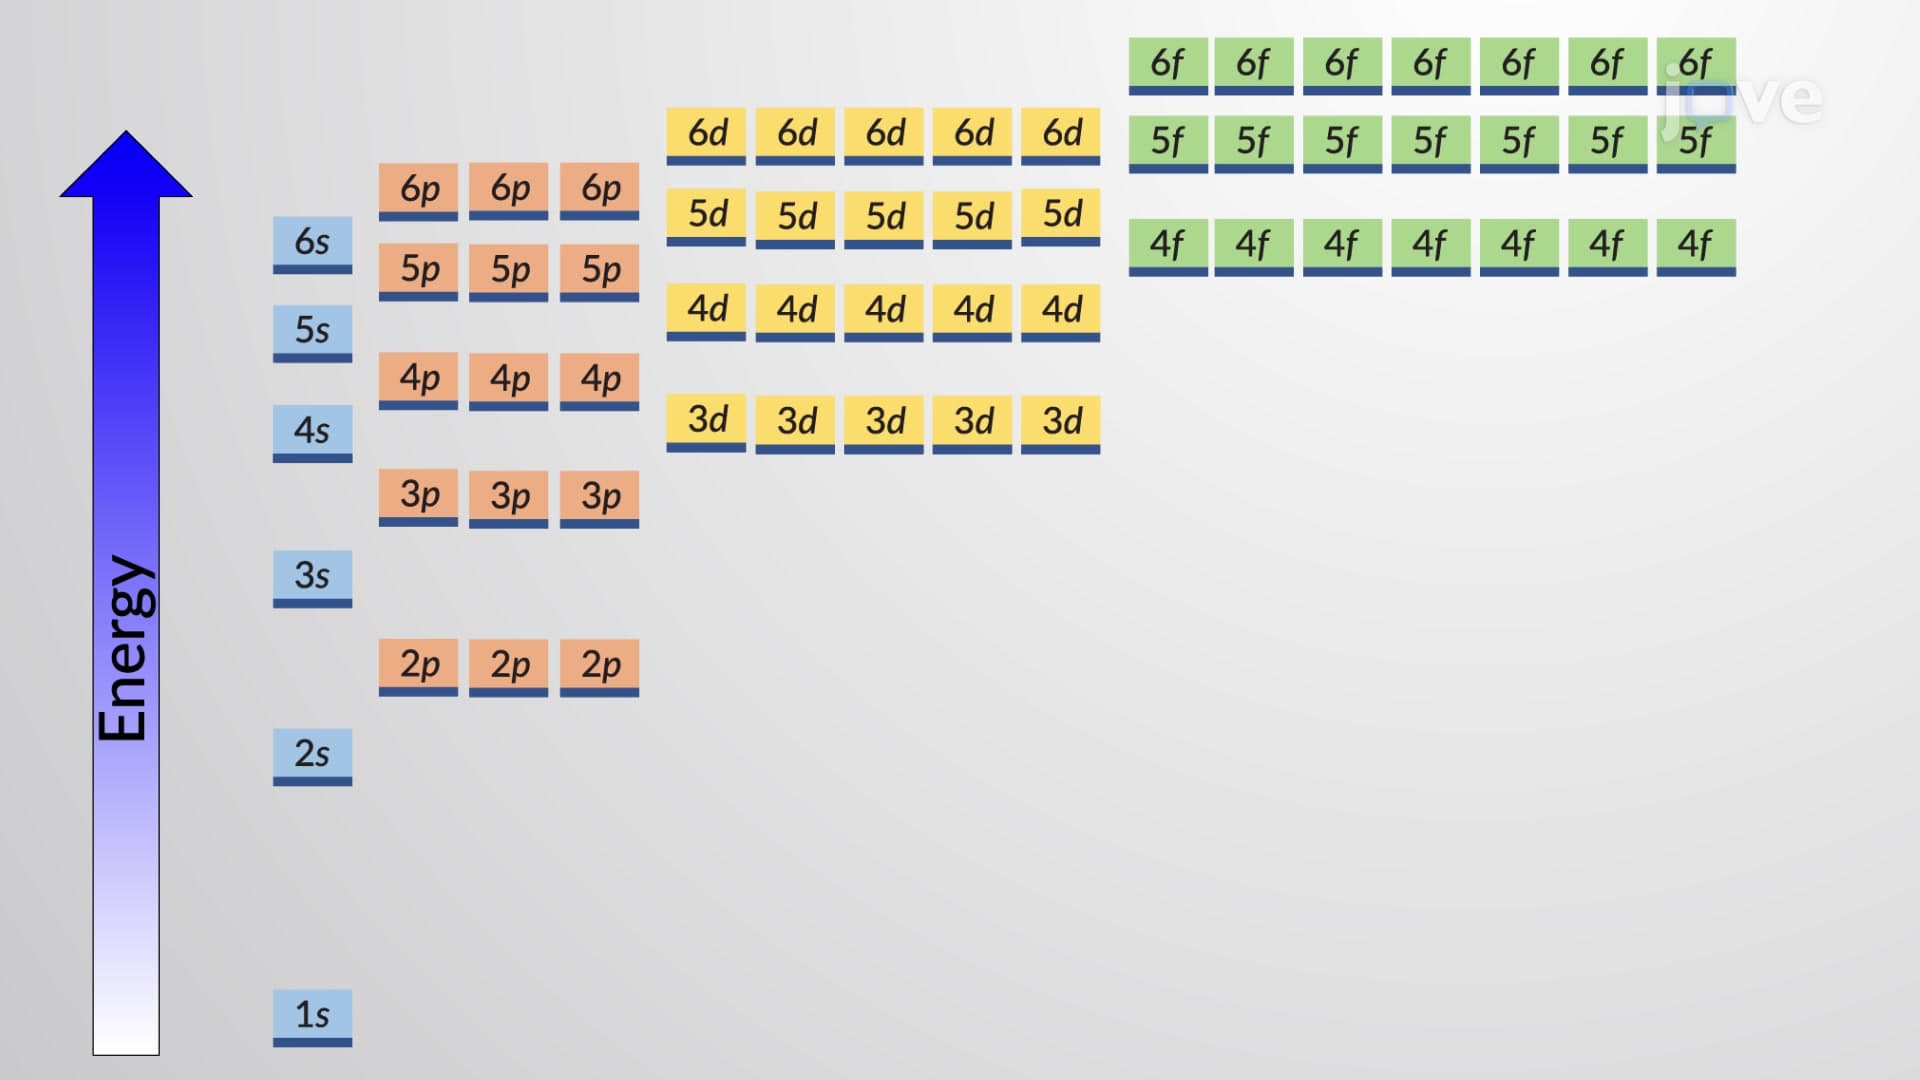
Electron Configuration of Multielectron Atoms
 01:12
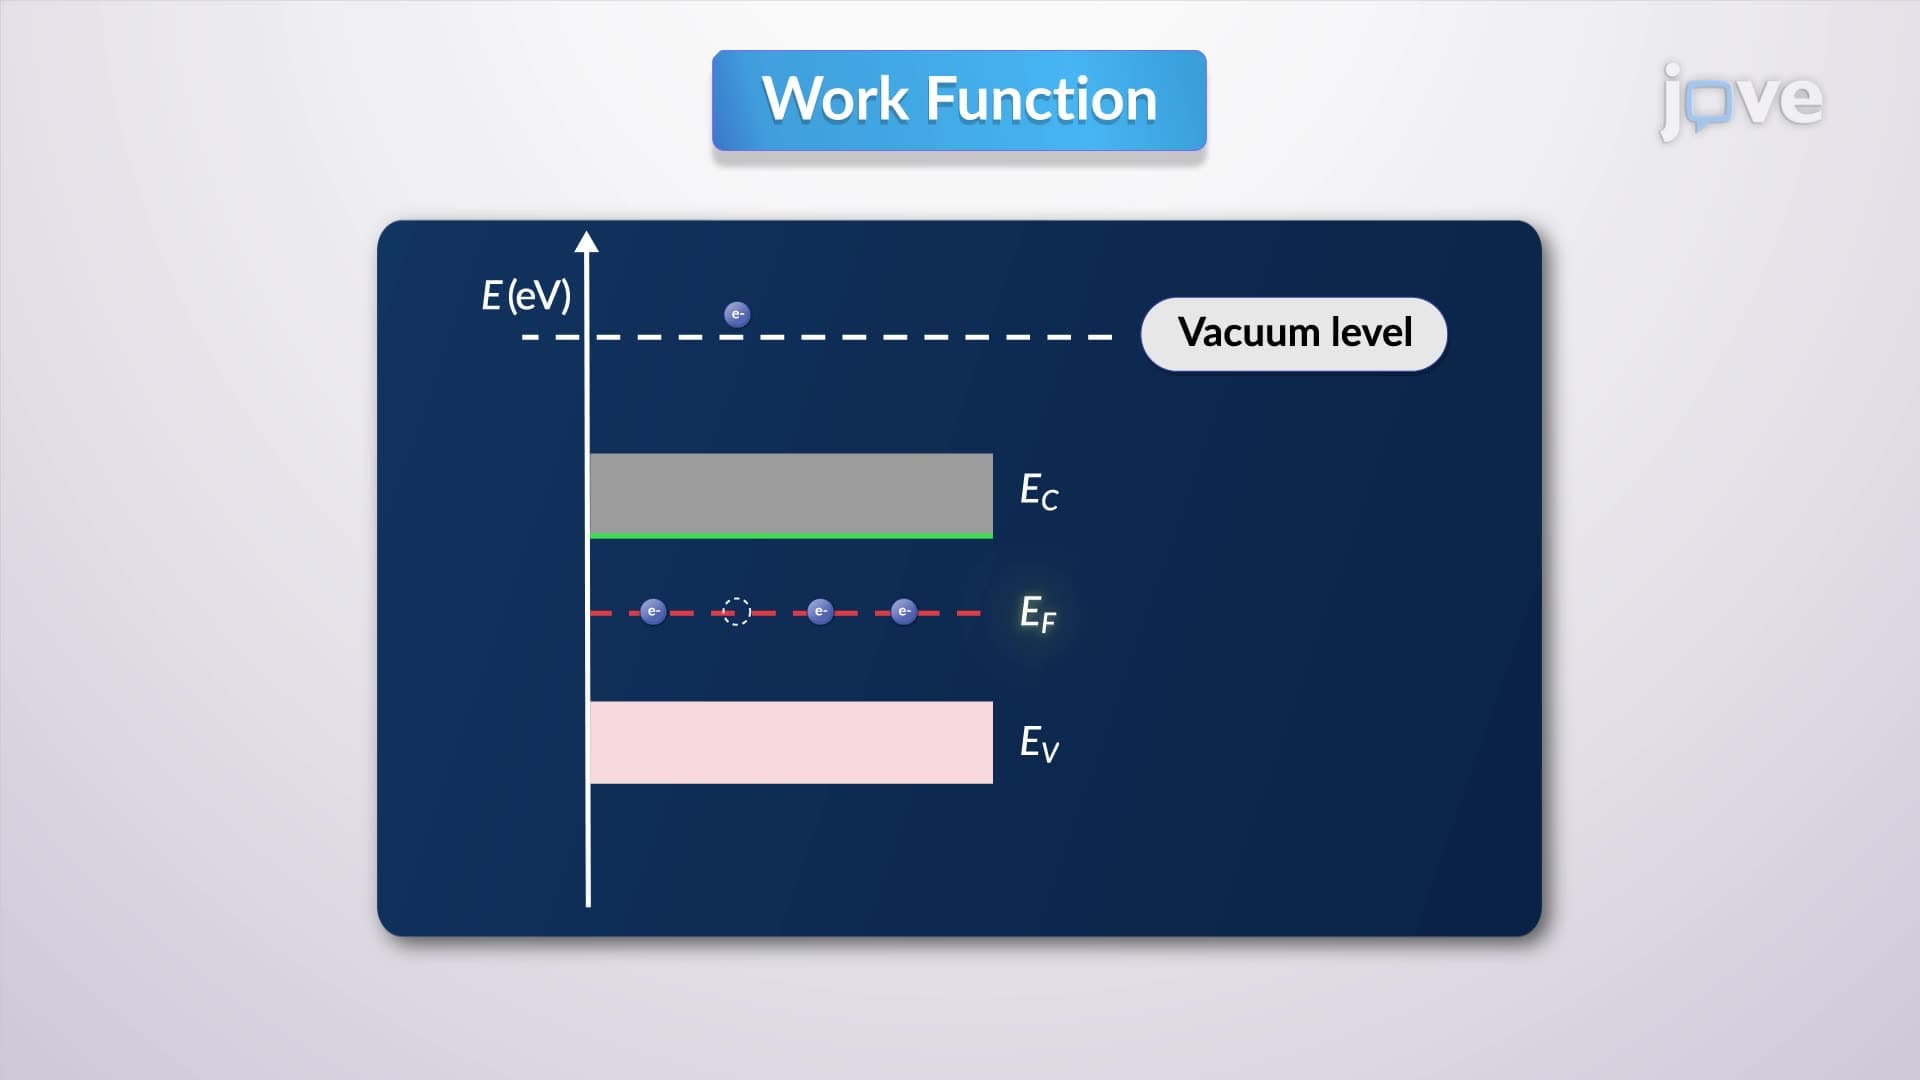
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:44
The Born-Haber Cycle