

XDM-corrected hybrid DFT with numerical atomic orbitals predicts molecular crystal lattice energies with
Alastair J A Price1, Alberto Otero-de-la-Roza2, Erin R Johnson1
1Department of Chemistry, Dalhousie University 6274 Coburg Rd Halifax B3H 4R2 Nova Scotia Canada erin.johnson@dal.ca.

Abstract:
Molecular crystals are important for many applications, including energetic materials, organic semiconductors, and the development and commercialization of pharmaceuticals. The exchange-hole dipole moment (XDM) dispersion model has shown good performance in the calculation of relative and absolute lattice energies of molecular crystals, although it has traditionally been applied in combination with plane-wave/pseudopotential approaches. This has limited XDM to use with semilocal functional approximations, which suffer from delocalization error and poor quality conformational energies, and to systems with a few hundreds of atoms at most due to unfavorable scaling. In this work, we combine XDM with numerical atomic orbitals, which enable the efficient use of XDM-corrected hybrid functionals for molecular crystals. We test the new XDM-corrected functionals for their ability to predict the lattice energies of molecular crystals for the X23 set and 13 ice phases, the latter being a particularly stringent test. A composite approach using a XDM-corrected, 25% hybrid functional based on B86bPBE achieves a mean absolute error of 0.48 kcal mol-1 per molecule for the X23 set and 0.19 kcal mol-1 for the total lattice energies of the ice phases, compared to recent diffusion Monte-Carlo data. These results make the new XDM-corrected hybrids not only far more computationally efficient than previous XDM implementations, but also the most accurate density-functional methods for molecular crystal lattice energies to date.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:18
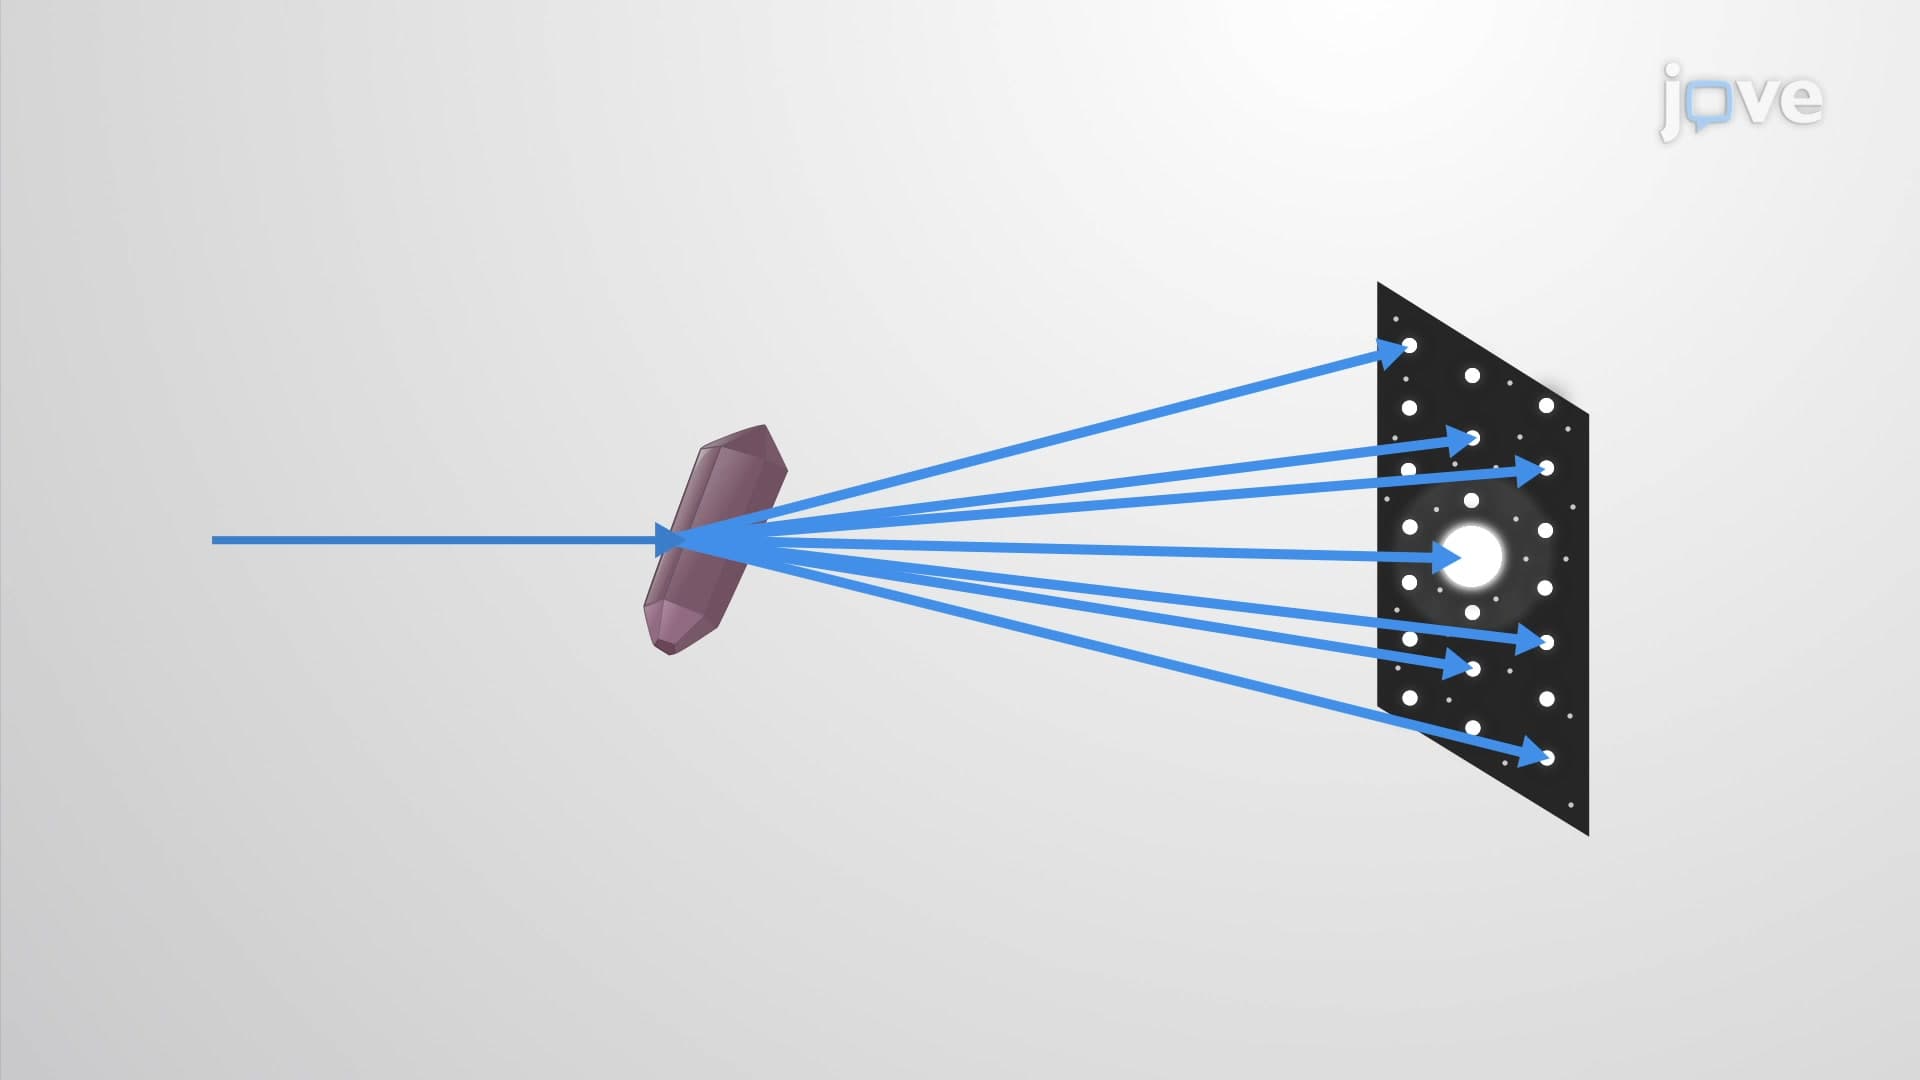
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 03:35
Hybridization of Atomic Orbitals II
 03:51
Molecular Orbital Theory II