Related Experiment Video
Updated: Aug 9, 2025

Using Scaffold Liposomes to Reconstitute Lipid-proximal Protein-protein Interactions In Vitro
Published on: January 11, 2017
Disease-associated mutations in Drp1 have fundamentally different effects on the mitochondrial fission machinery
Brianna L Bauer1, Kristy Rochon1, Jasmine C Liu1
1Department of Pharmacology, Case Western Reserve University School of Medicine, Cleveland, OH 44106, USA.

Insights
Disease-causing mutations in dynamin-related protein 1 (Drp1) cause severe neurological defects. This study reveals that Drp1 mutations variably impair mitochondrial fission by affecting self-assembly and membrane remodeling, offering insights into disease mechanisms.
Area of Science:
- Biochemistry
- Cell Biology
- Genetics
Background:
- Dynamin-related protein 1 (Drp1) is crucial for mitochondrial fission, a process vital for cellular function.
- Mutations in Drp1 are linked to severe neurological disorders in children, but their functional consequences remain unclear.
Purpose of the Study:
- To investigate the functional impact of six disease-associated Drp1 mutations on its self-assembly, GTPase activity, and membrane remodeling capabilities.
- To elucidate the molecular mechanisms underlying Drp1-related neurological diseases.
Main Methods:
- Analysis of Drp1 mutants in solution and on lipid membranes.
- Assays for GTP hydrolysis and oligomerization.
- Liposome-based membrane remodeling experiments.
Main Results:
- Mutations in the middle domain (MD) variably affected Drp1 oligomerization and membrane remodeling.
- The F370C mutation impaired liposome membrane remodeling, highlighting Drp1's role in membrane curvature generation.
- GTPase domain mutations (G32A, G223V) showed reduced GTPase activity and impaired membrane remodeling, indicating the GTPase domain's role in self-assembly and curvature.
Conclusions:
- Functional defects caused by Drp1 mutations are diverse, even within the same domain.
- Drp1's GTPase domain contributes to self-assembly and membrane curvature.
- This study provides a framework for understanding Drp1 mutations and their role in neurological diseases.
Abstract:
Patient mutations have been identified throughout dynamin-related protein 1 (Drp1), the key protein mediator of mitochondrial fission. These changes generally impact young children and often result in severe neurological defects and, in some instances, death. Until now, the underlying functional defect leading to patient phenotypes has been largely speculative. We therefore analyzed six disease-associated mutations throughout the GTPase and middle domains (MD) of Drp1. The MD plays a role in Drp1 oligomerization, and three mutations in this region were predictably impaired in self-assembly. However, another mutant in this region (F370C) retained oligomerization capability on pre-curved membranes despite being assembly-limited in solution. Instead, this mutation impaired membrane remodeling of liposomes, which highlights the importance of Drp1 in generating local membrane curvature before fission. Two GTPase domain mutations were also observed in different patients. The G32A mutation was impaired in GTP hydrolysis both in solution and in the presence of lipid but remains capable of self-assembly on these lipid templates. The G223V mutation also exhibited decreased GTPase activity and was able to assemble on pre-curved lipid templates; however, this change impaired membrane remodeling of unilamellar liposomes similar to F370C. This demonstrates that the Drp1 GTPase domain also contributes to self-assembly interactions that drive membrane curvature. Overall, the functional defects caused by mutations in Drp1 are highly variable even for mutations that reside within the same functional domain. This study provides a framework for characterizing additional Drp1 mutations to provide a comprehensive understanding of functional sites within this essential protein.
Related Concept Videos
 01:19
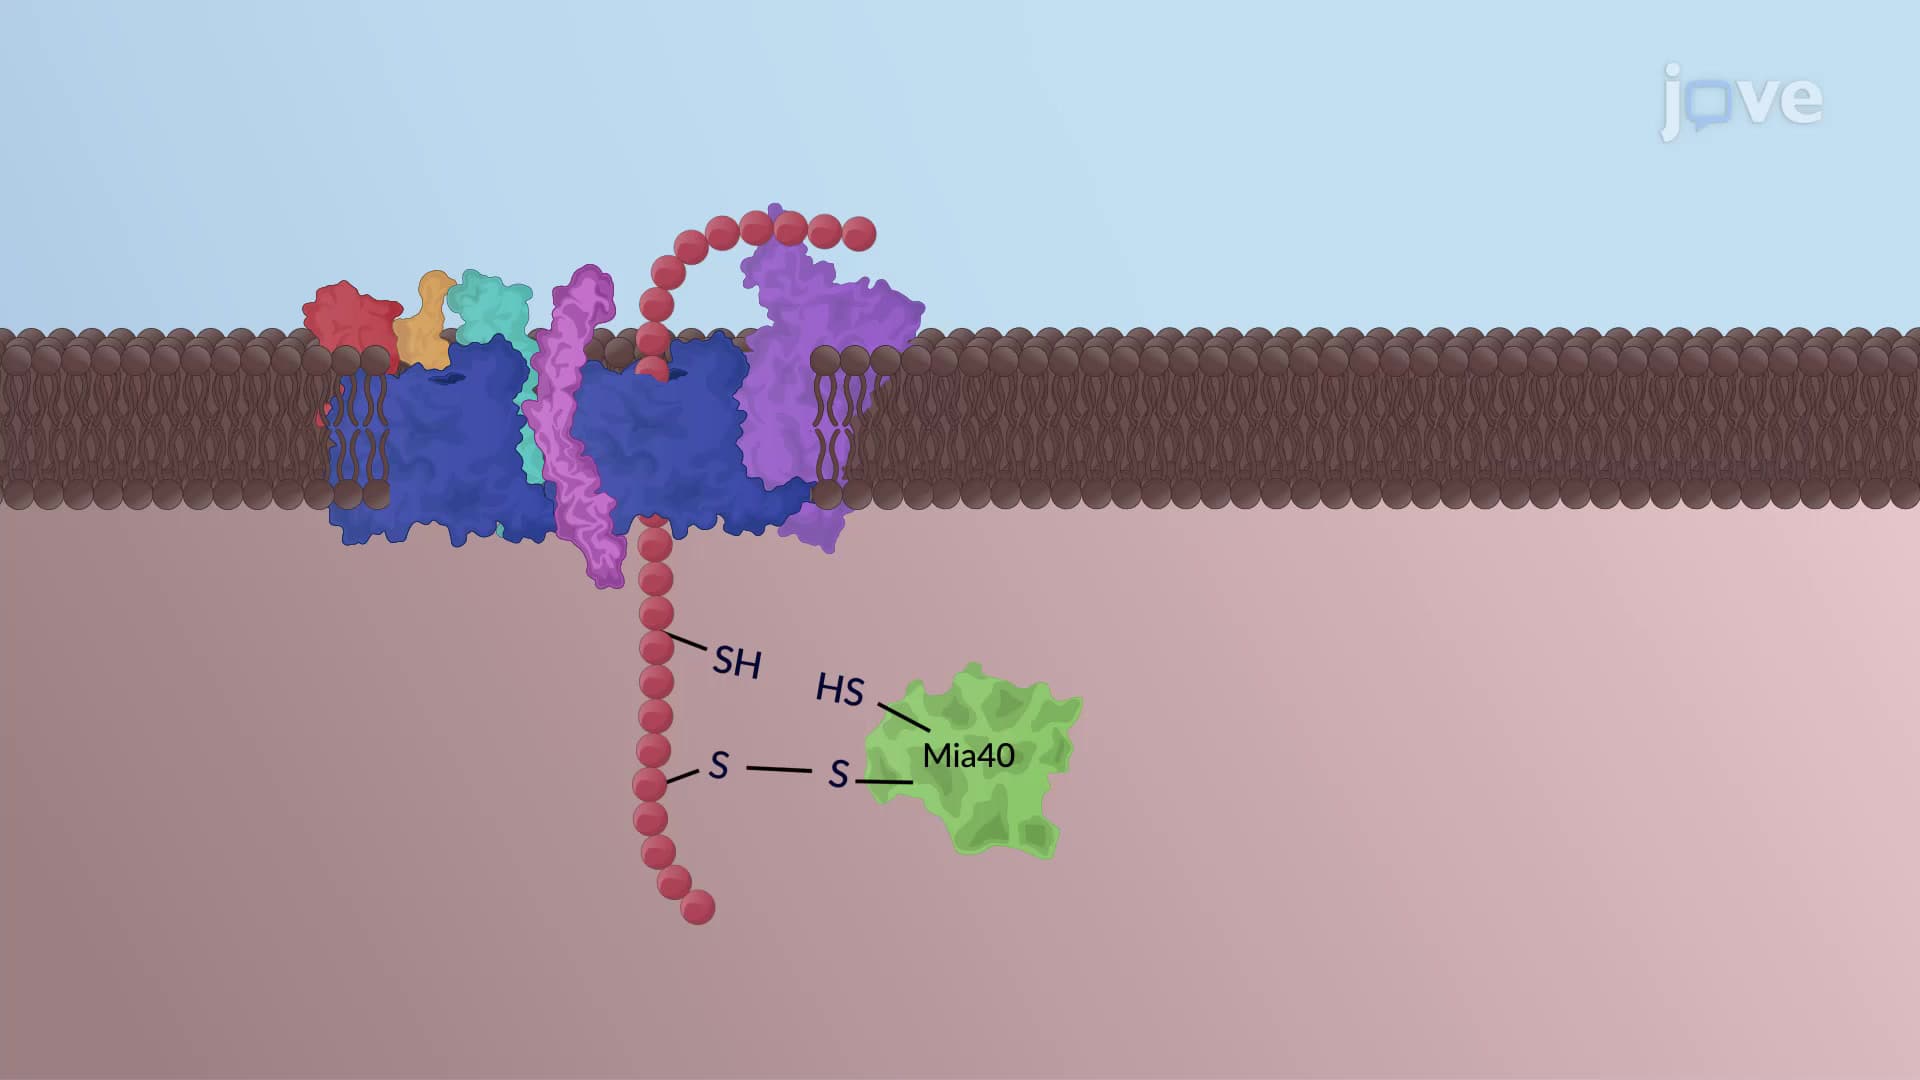
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 02:37
Restarting Stalled Replication Forks