Related Experiment Video
Updated: Aug 9, 2025
Tuning Oxide Properties by Oxygen Vacancy Control During Growth and Annealing
Published on: June 9, 2023
In-gap states and strain-tuned band convergence in layered structure trivalent iridate K0.75Na0.25IrO2
Xujia Gong1, Carmine Autieri2, Huanfu Zhou3
1College of Science, Guilin University of Technology, Guilin 541004, People's Republic of China. mingxing@glut.edu.cn.

Abstract:
Iridium oxides (iridates) provide a good platform to study the delicate interplay between spin-orbit coupling (SOC) interactions, electron correlation effects, Hund's coupling and lattice degrees of freedom. An overwhelming number of investigations primarily focus on tetravalent (Ir4+, 5d5) and pentavalent (Ir5+, 5d4) iridates, and far less attention has been paid to iridates with other valence states. Here, we pay our attention to a less-explored trivalent (Ir3+, 5d6) iridate, K0.75Na0.25IrO2, crystallizing in a triangular lattice with edge-sharing IrO6 octahedra and alkali metal ion intercalated [IrO2]- layers, offering a good platform to explore the interplay between different degrees of freedom. We theoretically determine the preferred occupied positions of the alkali metal ions from energetic viewpoints and reproduce the experimentally observed semiconducting behavior and nonmagnetic (NM) properties of K0.75Na0.25IrO2. The SOC interactions play a critical role in the band dispersion, resulting in NM Jeff = 0 states. More intriguingly, our electronic structure not only uncovers the presence of intrinsic in-gap states and nearly free electron character for the conduction band minimum, but also explains the abnormally low activation energy in K0.75Na0.25IrO2. Particularly, the band edge can be effectively modulated by mechanical strain, and the in-gap states feature enhanced band-convergence characteristics by 6% compressive strain, which will greatly enhance the electrical conductivity of K0.75Na0.25IrO2. The present work sheds new light on the unconventional electronic structures of trivalent iridates, indicating their promising application as a nanoelectronic and thermoelectric material, which will attract extensive interest and stimulate experimental works to further understand the unprecedented electronic structures and exploit potential applications of the triangular trivalent iridate.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 09:06
Visualizing Uniaxial-strain Manipulation of Antiferromagnetic Domains in Fe1+YTe Using a Spin-polarized Scanning Tunneling Microscope
Published on: March 24, 2019
Related Concept Videos
 01:01
Energy Bands in Solids
Band Formation:
When atoms are brought close together, as in a solid, these discrete energy levels begin to split due to the overlap of electron orbitals from adjacent atoms. This split occurs because of the Pauli exclusion principle, which states...
 02:35
Band Theory
The energy difference between these bands is known as the band gap.
Conductor, Semiconductor,...
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 02:42
Ionic Crystal Structures
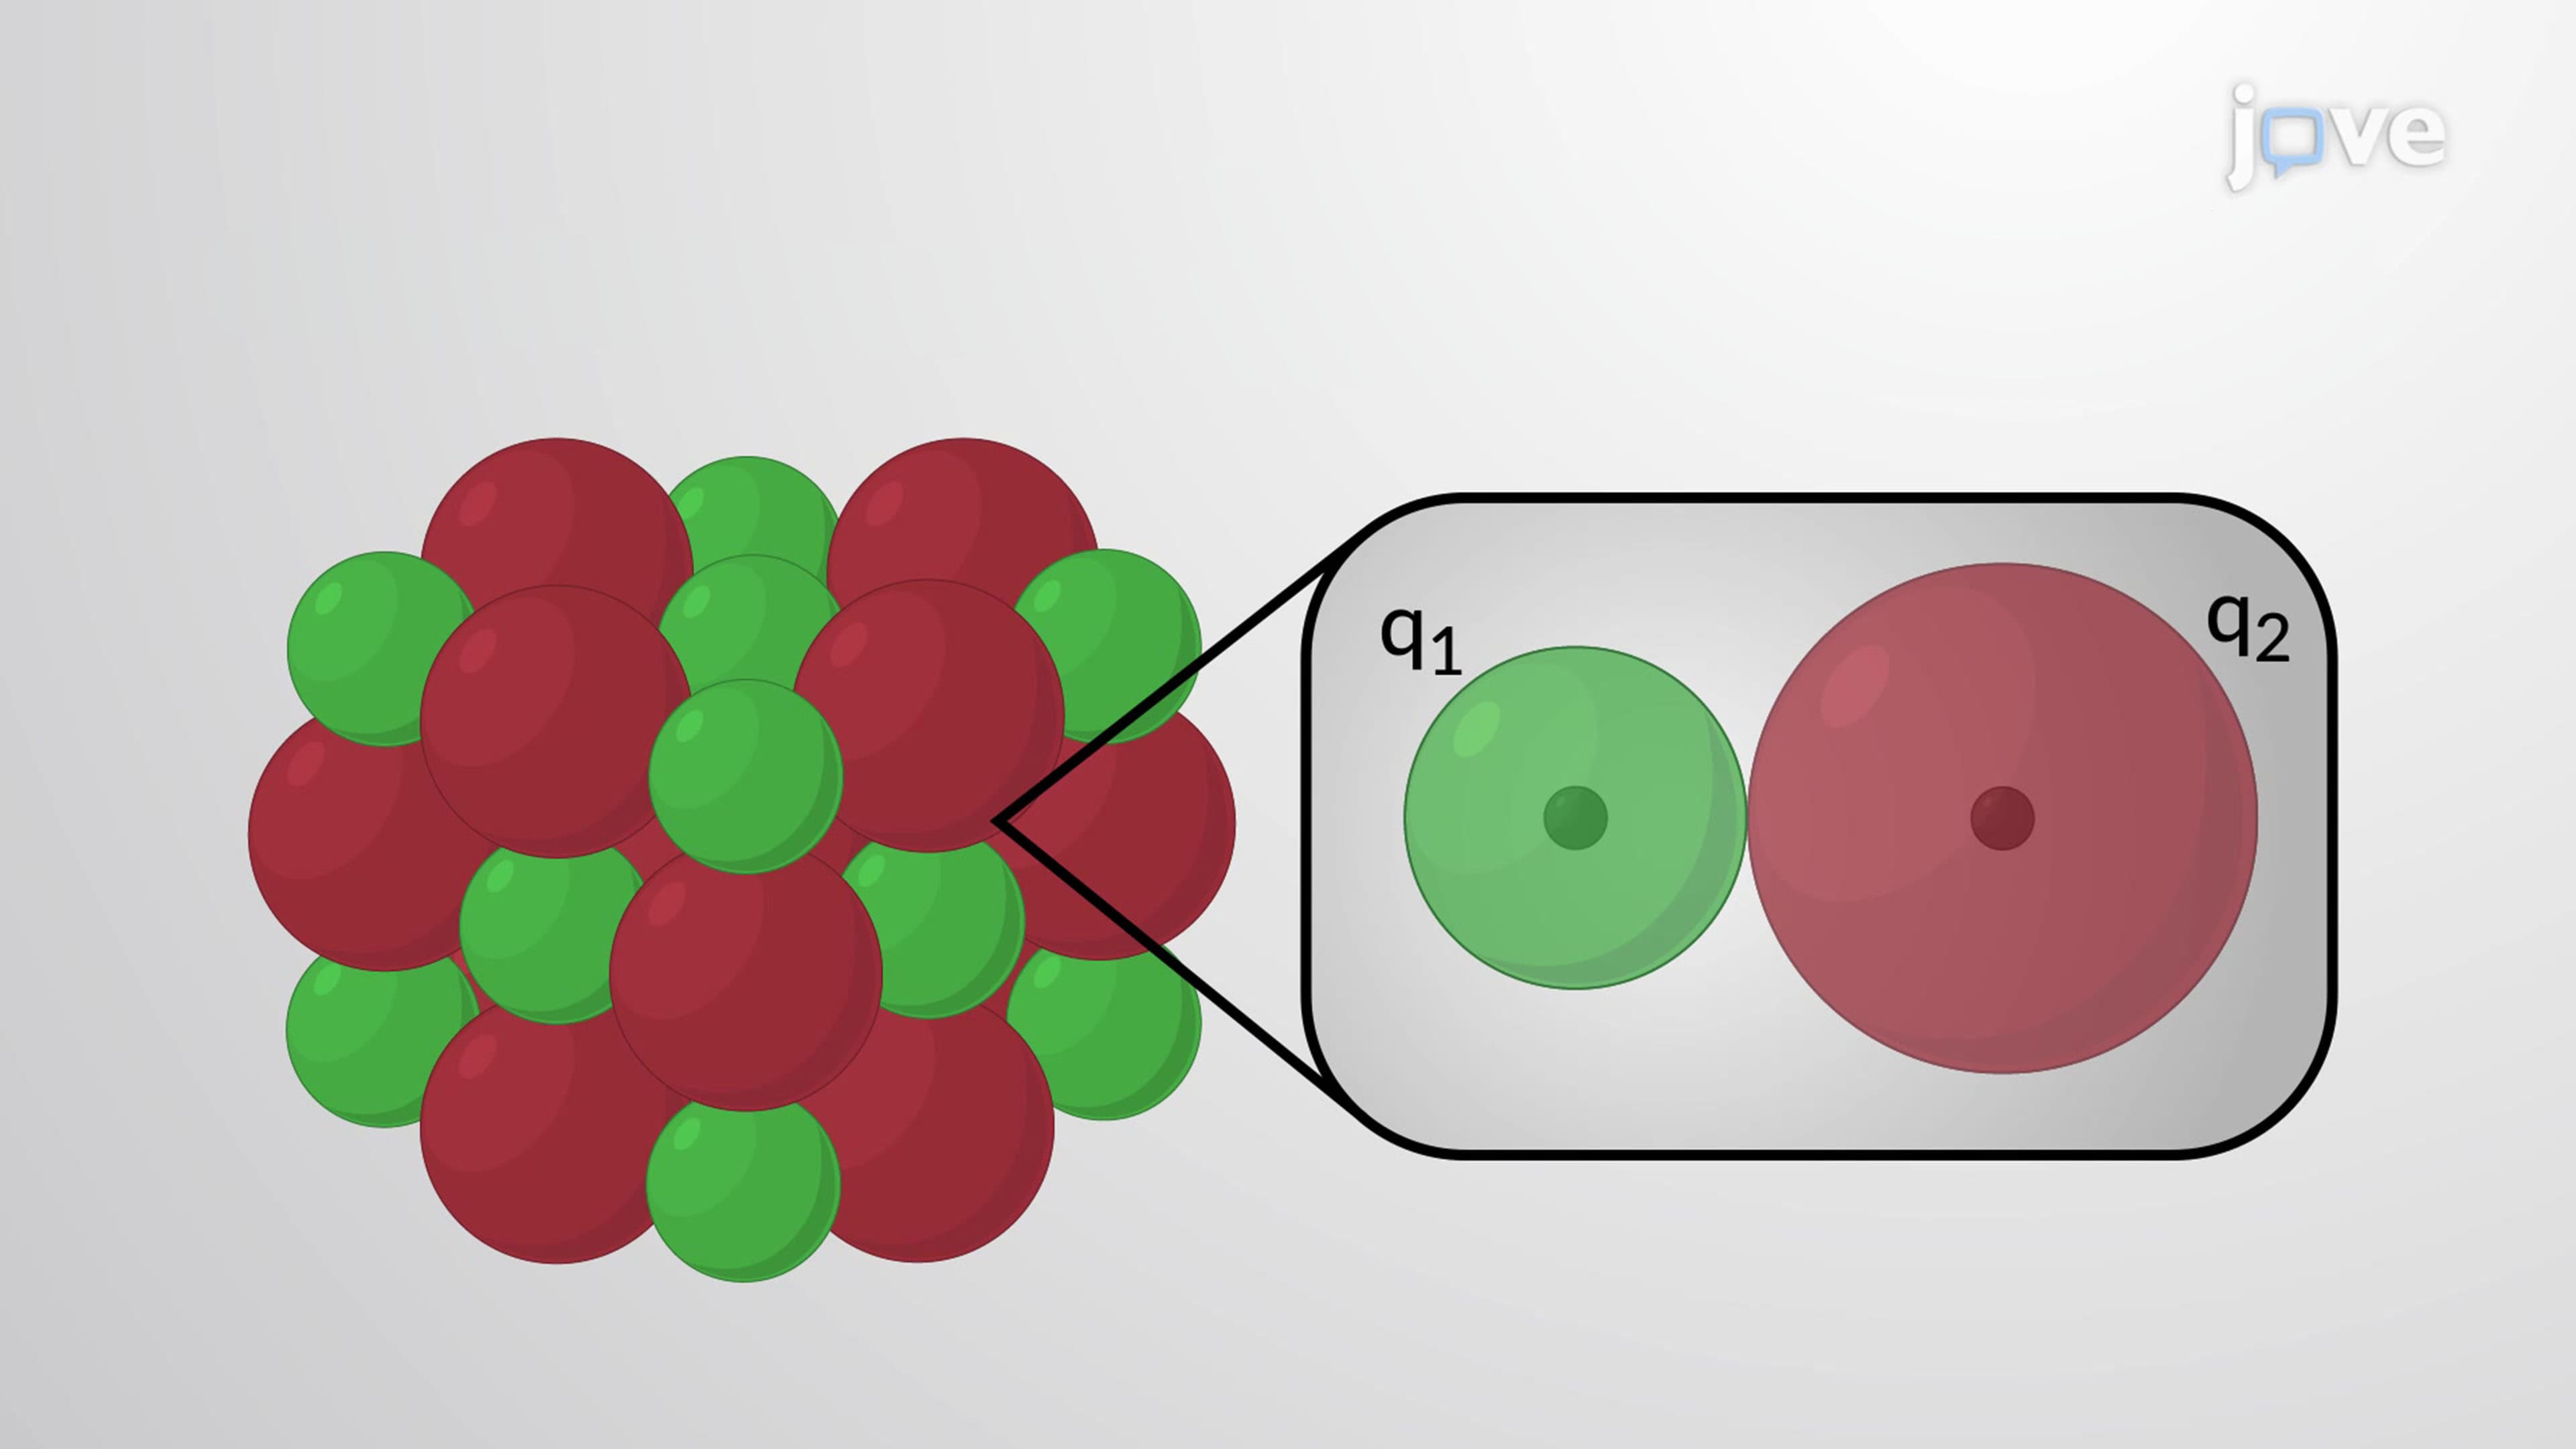
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:54
Trends in Lattice Energy: Ion Size and Charge