Related Experiment Video
Updated: Aug 8, 2025


Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
PSnpBind-ML: predicting the effect of binding site mutations on protein-ligand binding affinity
Ammar Ammar1, Rachel Cavill2, Chris Evelo3
1Department of Bioinformatics-BiGCaT, NUTRIM, Maastricht University, Maastricht, The Netherlands. a.ammar@maastrichtuniversity.nl.

Abstract:
Protein mutations, especially those which occur in the binding site, play an important role in inter-individual drug response and may alter binding affinity and thus impact the drug's efficacy and side effects. Unfortunately, large-scale experimental screening of ligand-binding against protein variants is still time-consuming and expensive. Alternatively, in silico approaches can play a role in guiding those experiments. Methods ranging from computationally cheaper machine learning (ML) to the more expensive molecular dynamics have been applied to accurately predict the mutation effects. However, these effects have been mostly studied on limited and small datasets, while ideally a large dataset of binding affinity changes due to binding site mutations is needed. In this work, we used the PSnpBind database with six hundred thousand docking experiments to train a machine learning model predicting protein-ligand binding affinity for both wild-type proteins and their variants with a single-point mutation in the binding site. A numerical representation of the protein, binding site, mutation, and ligand information was encoded using 256 features, half of them were manually selected based on domain knowledge. A machine learning approach composed of two regression models is proposed, the first predicting wild-type protein-ligand binding affinity while the second predicting the mutated protein-ligand binding affinity. The best performing models reported an RMSE value within 0.5 [Formula: see text] 0.6 kcal/mol-1 on an independent test set with an R2 value of 0.87 [Formula: see text] 0.90. We report an improvement in the prediction performance compared to several reported models developed for protein-ligand binding affinity prediction. The obtained models can be used as a complementary method in early-stage drug discovery. They can be applied to rapidly obtain a better overview of the ligand binding affinity changes across protein variants carried by people in the population and narrow down the search space where more time-demanding methods can be used to identify potential leads that achieve a better affinity for all protein variants.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 00:49
Ligand Binding and Linkage
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
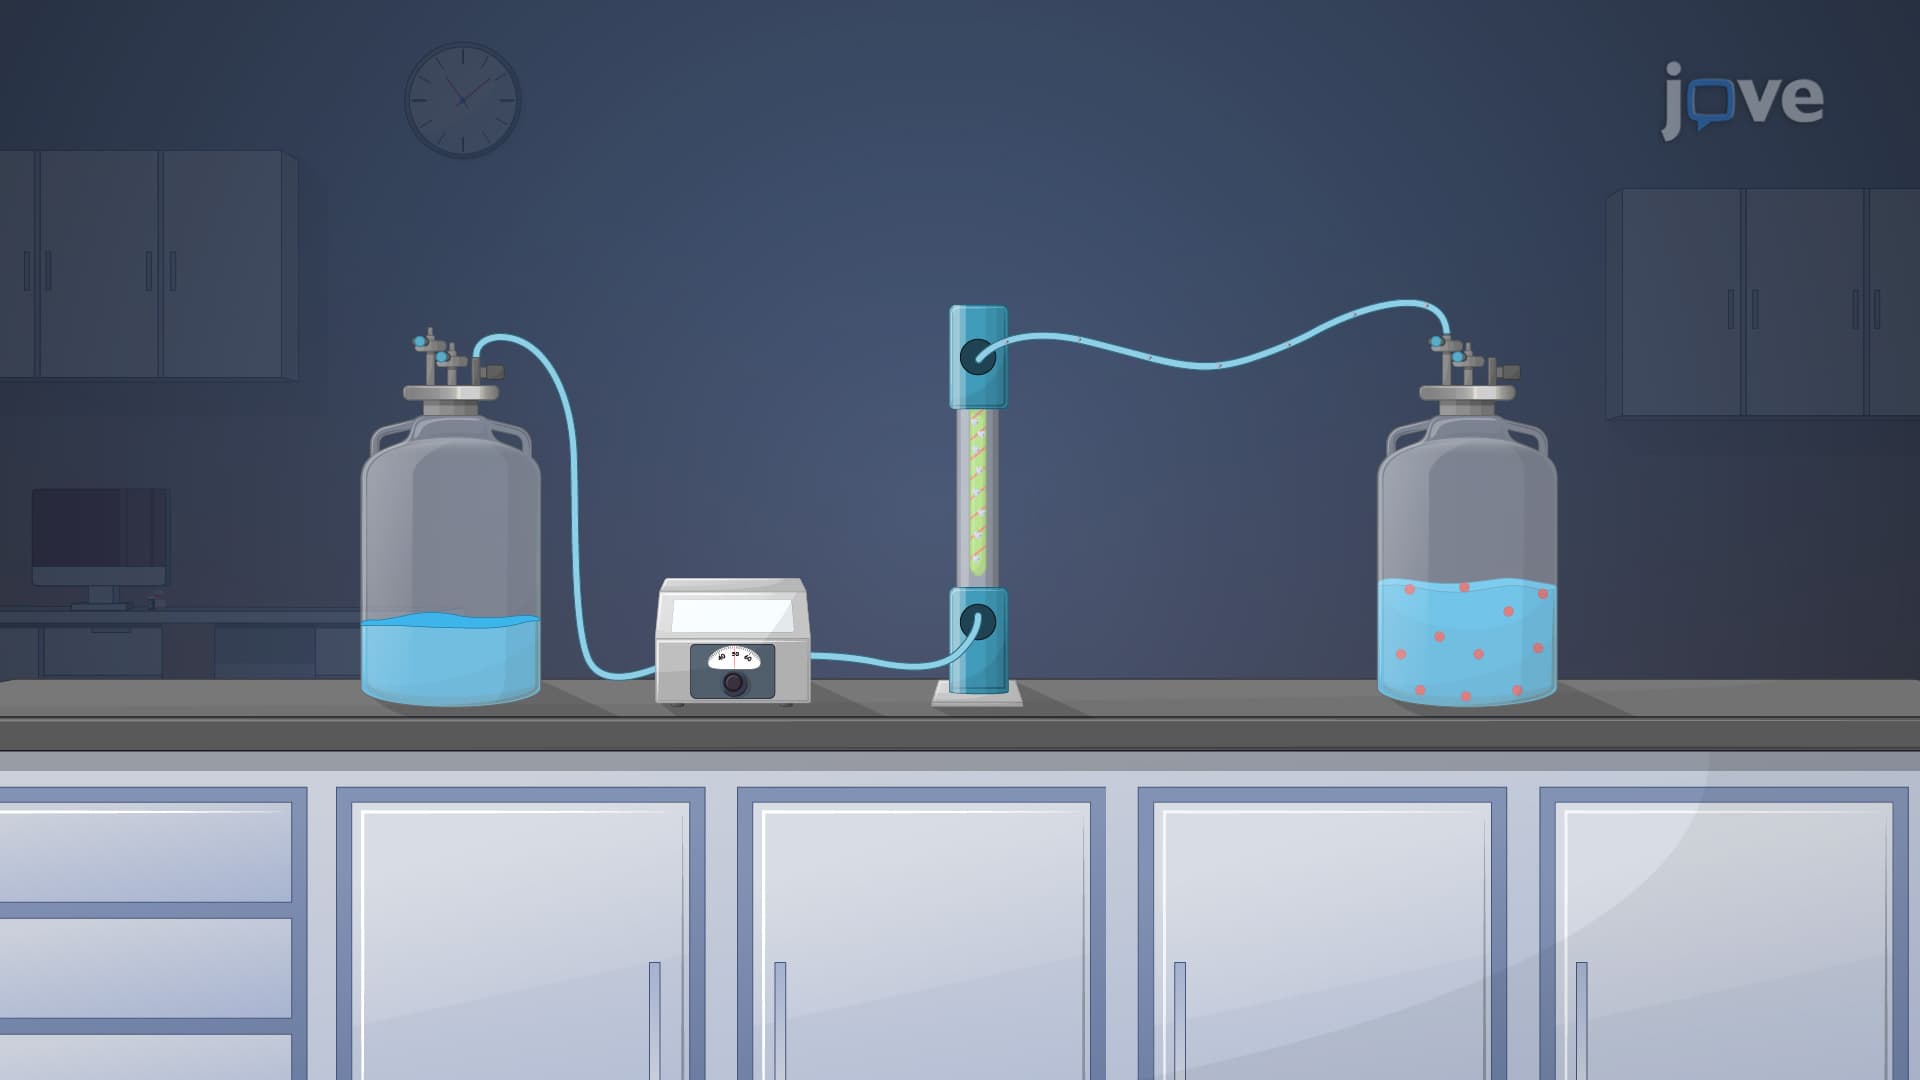
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis...