Assessment of three-body dispersion models against coupled-cluster benchmarks for crystalline benzene, carbon dioxide, and triazine
Related Experiment Video
Updated: Aug 7, 2025

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Yi Xie1, Zachary L Glick1, C David Sherrill1
1Center for Computational Molecular Science and Technology, School of Chemistry and Biochemistry, School of Computational Science and Engineering, Georgia Institute of Technology, Atlanta, Georgia 30332-0400, USA.

Abstract:
To study the contribution of three-body dispersion to crystal lattice energies, we compute the three-body contributions to the lattice energies for crystalline benzene, carbon dioxide, and triazine using various computational methods. We show that these contributions converge quickly as the intermolecular distances between the monomers grow. In particular, the smallest value among the three pairwise intermonomer closest-contact distances, Rmin, shows a strong correlation with the three-body contribution to the lattice energy, and, here, the largest of the closest-contact distances, Rmax, serves as a cutoff criterion to limit the number of trimers to be considered. We considered all trimers up to Rmax=15Å. The trimers with Rmin<4Å contribute 90.4%, 90.6%, and 93.9% of the total three-body contributions for crystalline benzene, carbon dioxide, and triazine, respectively, for the coupled-cluster singles, doubles, and perturbative triples [CCSD(T)] method. For trimers with Rmin>4Å, the second-order Møller-Plesset perturbation theory (MP2) supplemented with the Axilrod-Teller-Muto (ATM) three-body dispersion correction reproduces the CCSD(T) values for the cumulative three-body contributions with errors of less than 0.1 kJ mol-1. Moreover, three-body contributions are converged within 0.15 kJ mol-1 by Rmax=10Å. From these results, it appears that in molecular crystals where dispersion dominates the three-body contribution to the lattice energy, the trimers with Rmin>4Å can be computed with the MP2+ATM method to reduce the computational cost, and those with Rmax>10Å appear to be basically negligible.
More Related Videos
 05:47
In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
 06:56
1,3,5-Triphenylbenzene and Corannulene as Electron Receptors for Lithium Solvated Electron Solutions
Published on: October 10, 2016
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:34
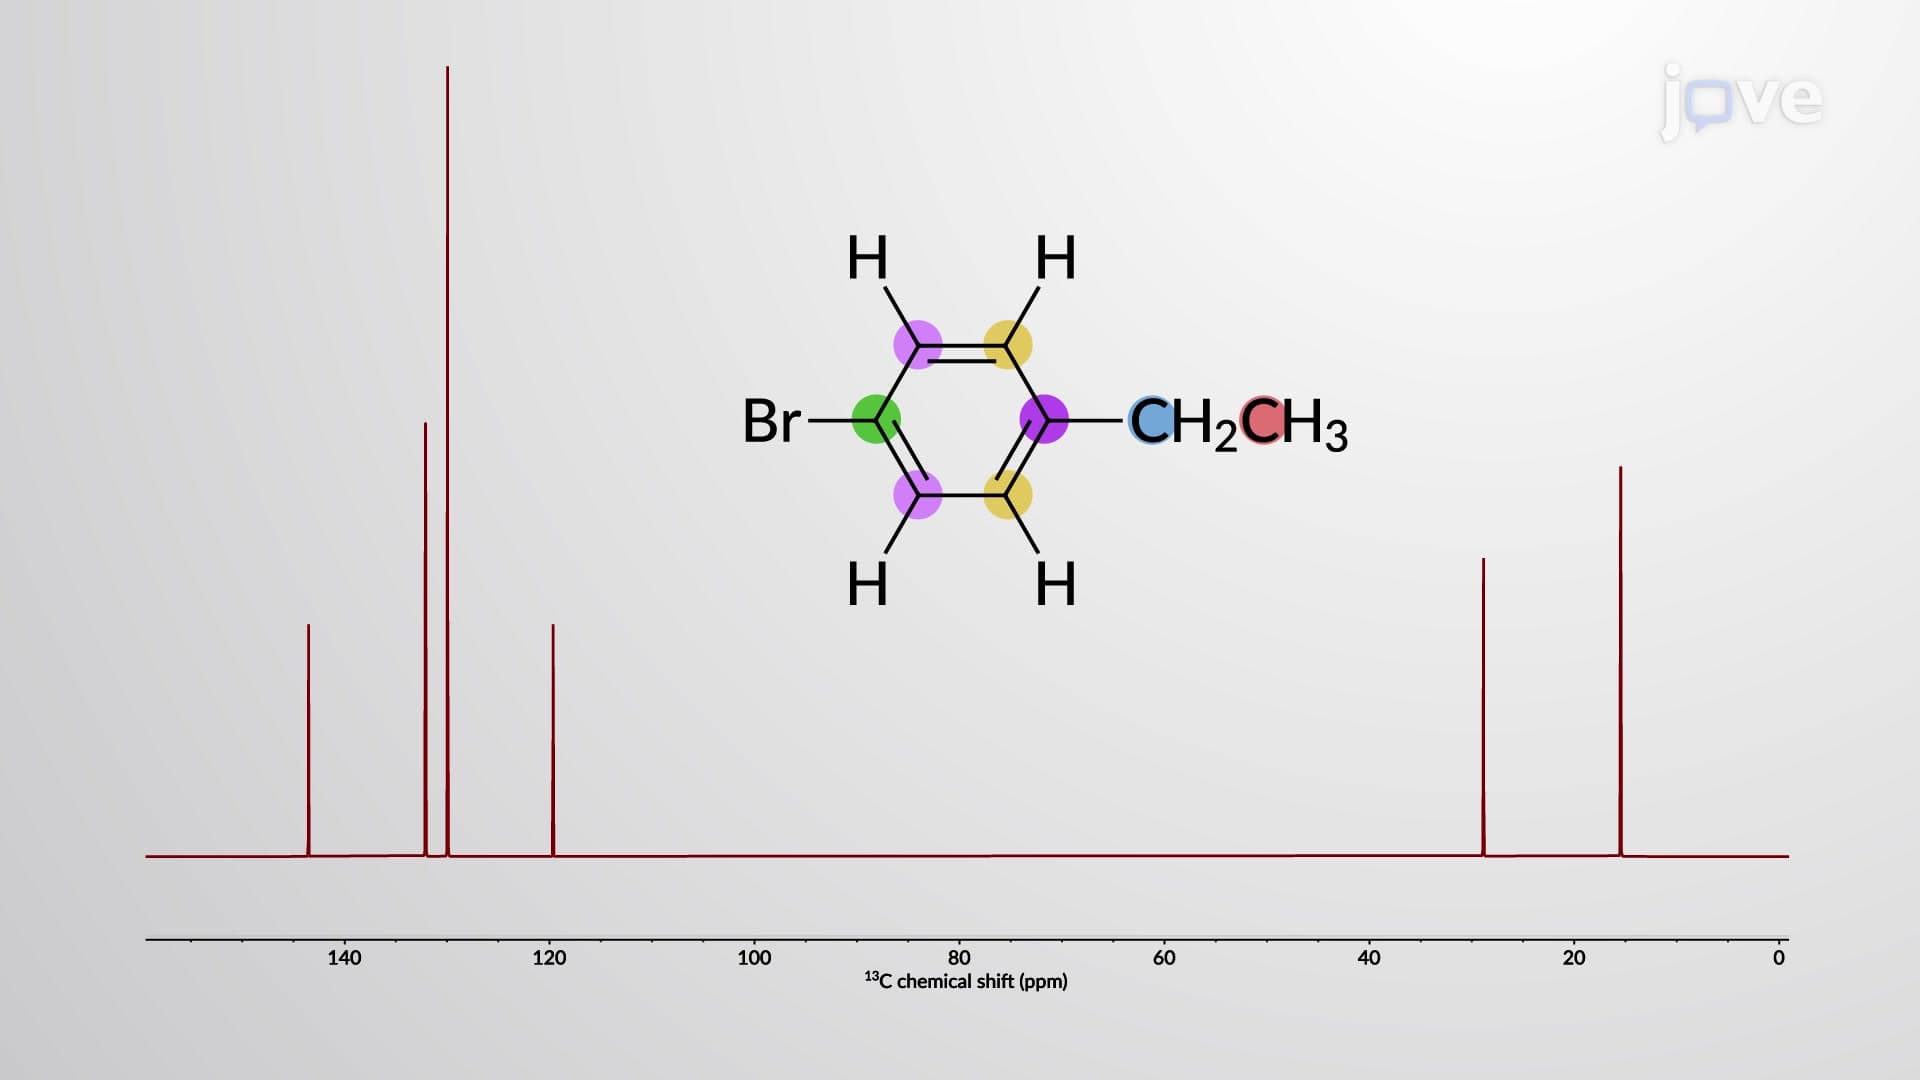
NMR Spectroscopy of Benzene Derivatives
 01:07
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:18
Frost Circles for Different Conjugated Systems
 03:51
Molecular Orbital Theory II
 01:18
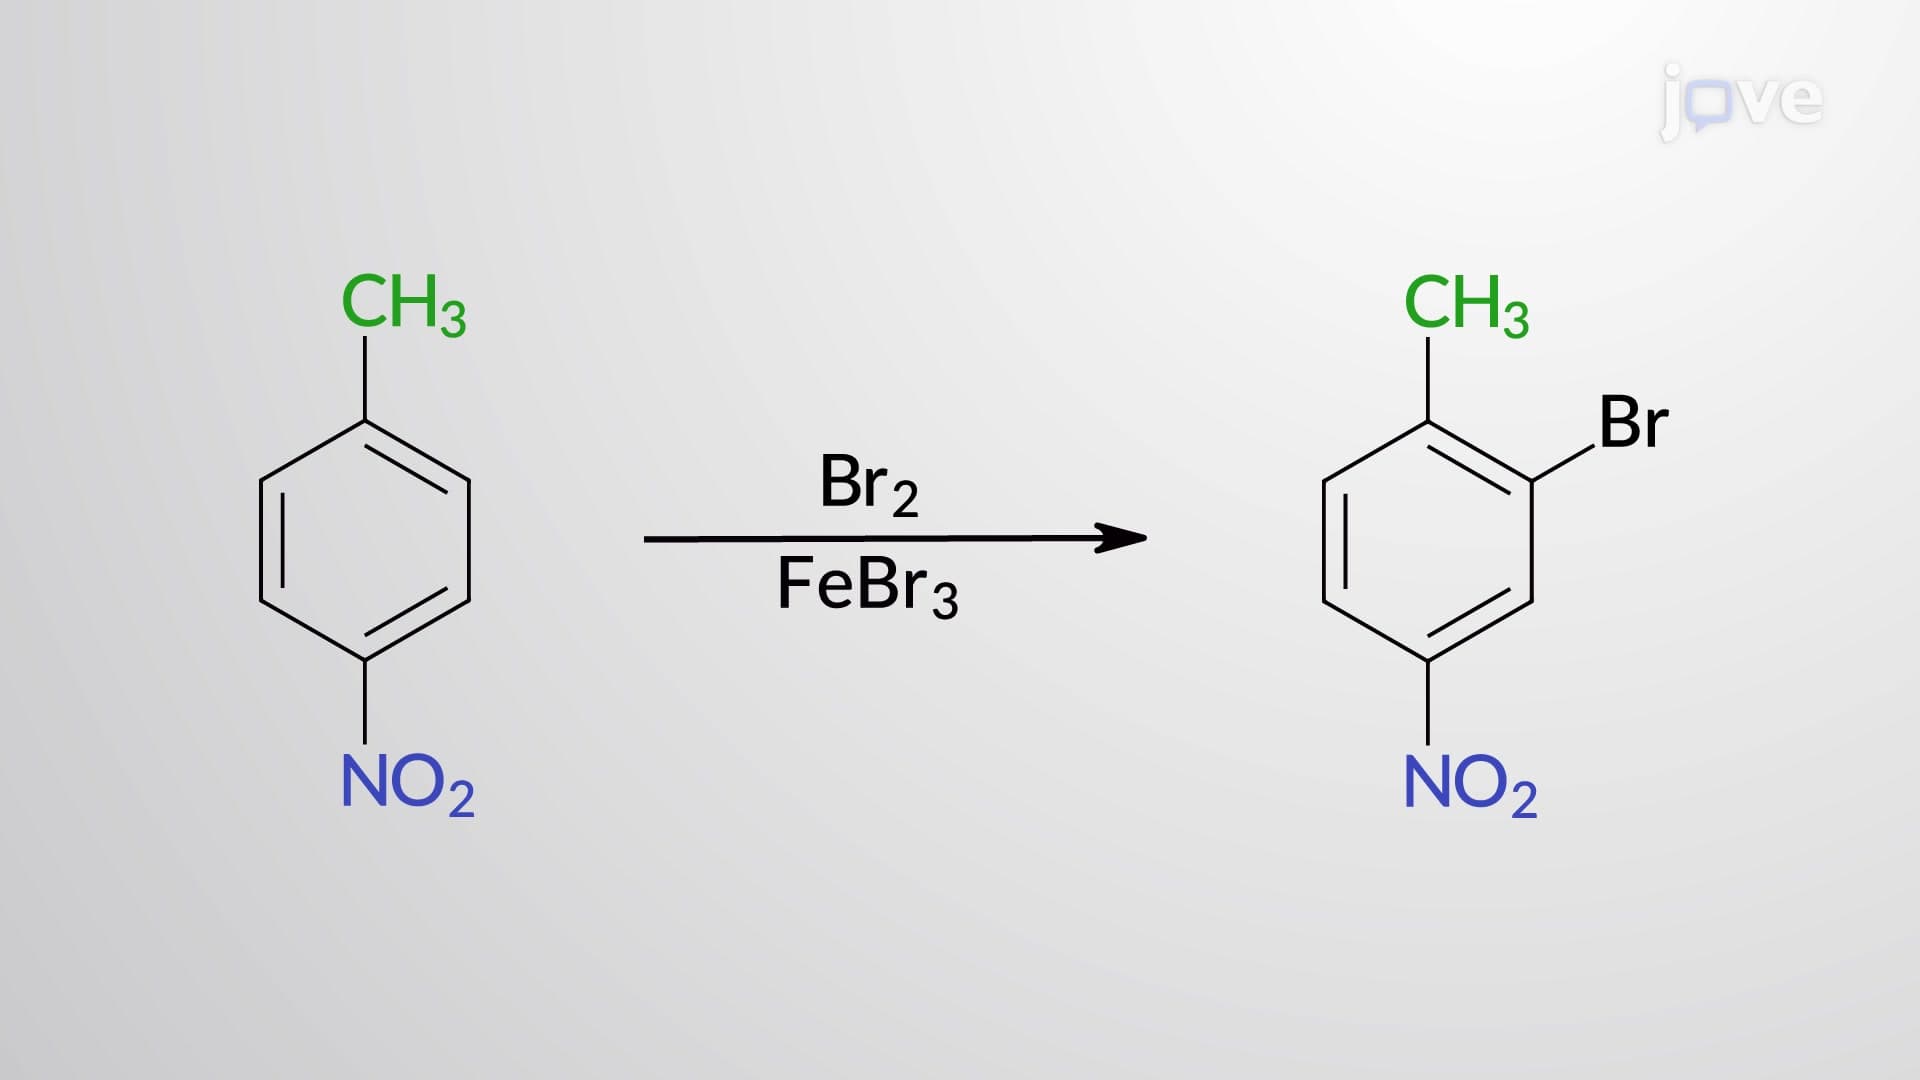
Directing and Steric Effects in Disubstituted Benzene Derivatives