Related Experiment Video
Updated: Aug 7, 2025
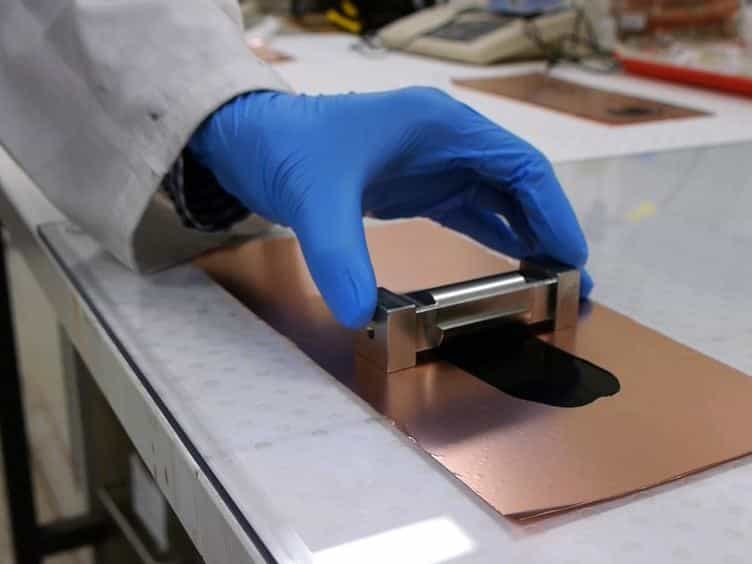

In Situ Neutron Powder Diffraction Using Custom-made Lithium-ion Batteries
Published on: November 10, 2014
Density Functional Tight-Binding Model for Lithium-Silicon Alloys
María Belén Oviedo1,2, Francisco Fernandez3, Manuel Otero3
1Universidad Nacional de Córdoba, Facultad de Ciencias Químicas, Departamento de Química Teórica y Computacional, Córdoba, X5000HUA, Argentina.

Abstract:
The predictive power of molecular dynamic simulations is mainly restricted by the time scale and model accuracy. Many systems of current relevance are of such complexity that they require addressing both issues simultaneously. This is the case of silicon electrodes in Li-ion batteries, where different LiSi alloys are formed during charge/discharge cycles. While first-principles treatments for this system are seriously limited by the computational cost of exploring its large conformational space, classical force fields are not transferable enough to represent it accurately. Density Functional Tight Binding (DFTB) is an intermediate complexity approach capable of capturing the electronic nature of different environments with a relatively low computational cost. In this work, we present a new set of DFTB parameters suited to model amorphous LiSi alloys. LiSi is the usual finding upon cycling the Si electrodes in the presence of Li ions. The model parameters are constructed with a particular emphasis on their transferability for the entire LiSi composition range. This is achieved by introducing a new optimization procedure that weights stoichiometries differently to improve the prediction of their formation energies. The resulting model is shown to be robust for predicting crystal and amorphous structures for the different compositions, giving excellent agreement with DFT calculations and outperforming state-of-the-art ReaxFF potentials.
Related Concept Videos
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:51
Molecular Orbital Theory II
 01:27
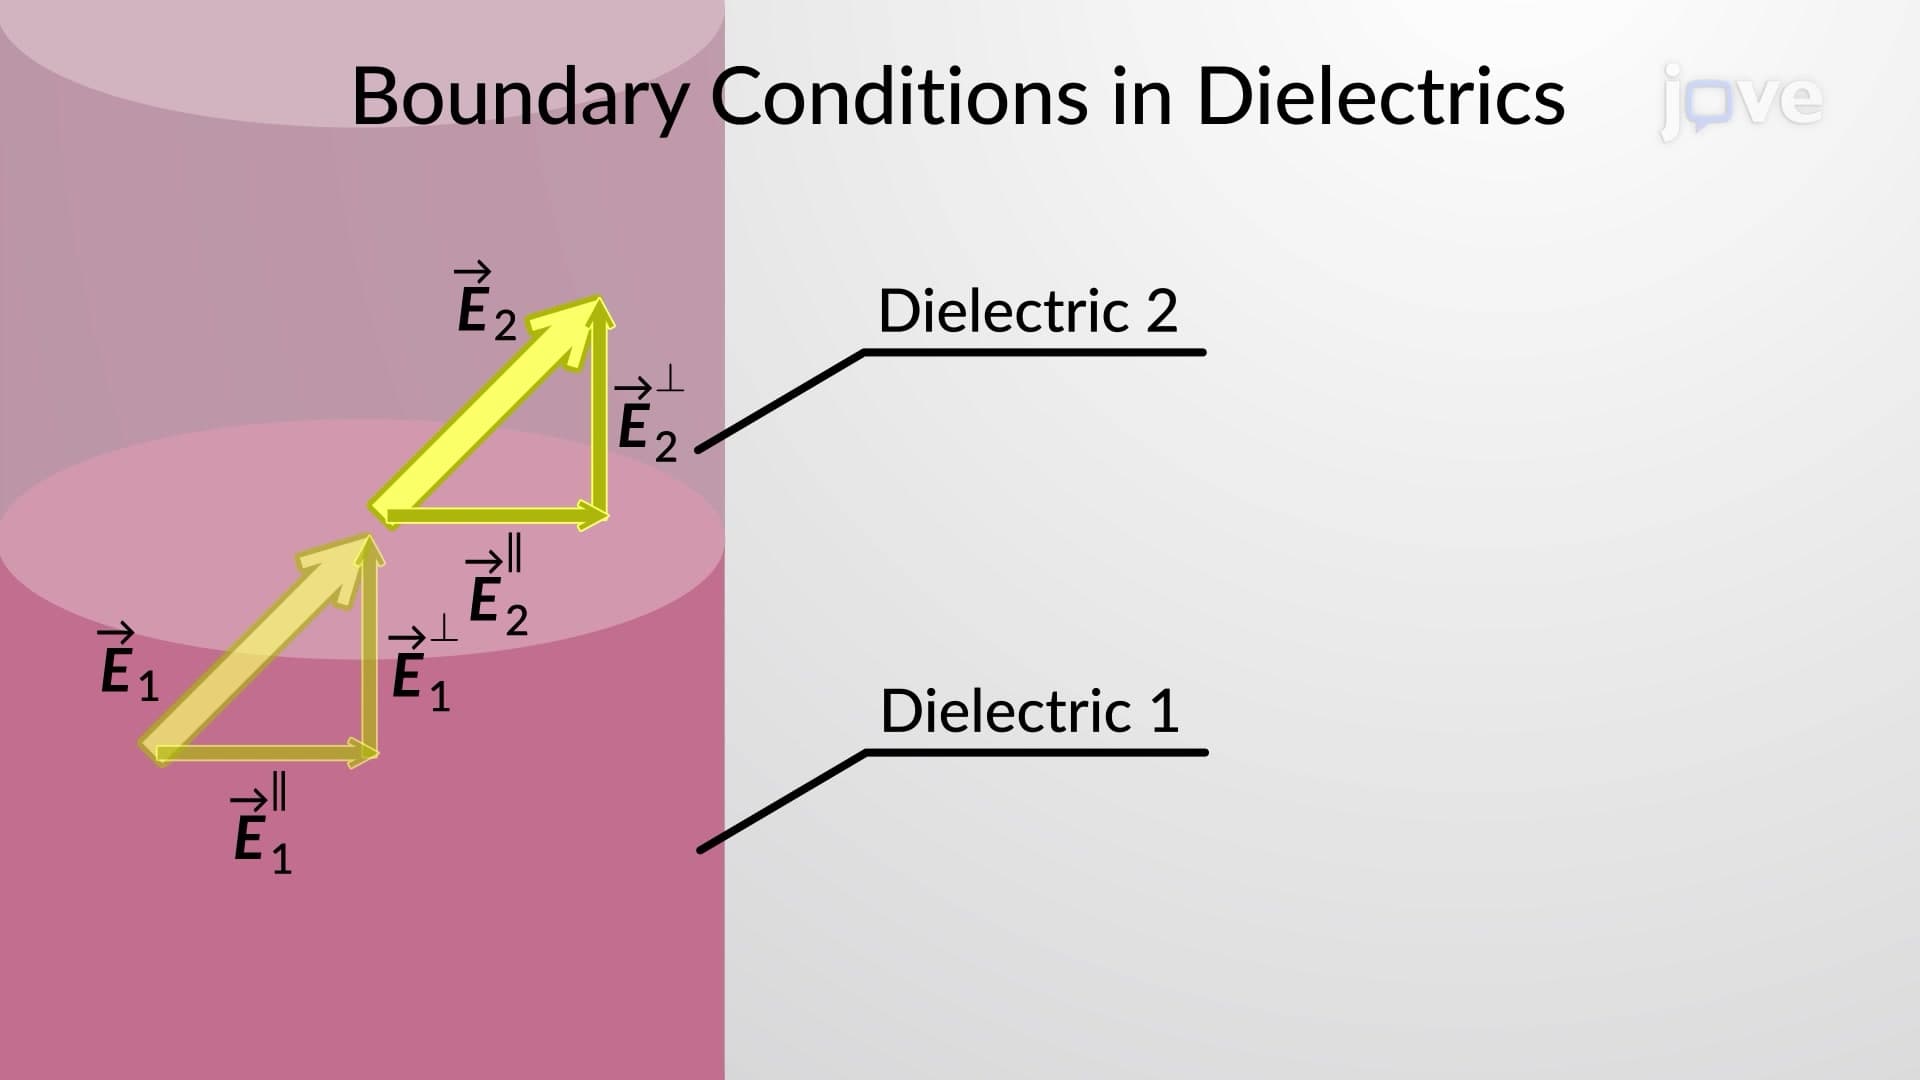
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's...