Related Experiment Video
Updated: Aug 7, 2025
Visualization of Bacterial Resistance using Fluorescent Antibiotic Probes
Published on: March 2, 2020
Synthesis of 4-Amino-N-[2 (diethylamino)Ethyl]Benzamide Tetraphenylborate Ion-Associate Complex: Characterization,
Gamal A E Mostafa1, Ahmed H Bakheit1, Mohamed H Al-Agamy2
1Department of Pharmaceutical Chemistry, College of Pharmacy, King Saud University, P.O. Box 2457, Riyadh 11451, Saudi Arabia.

Abstract:
The 4-amino-N-[2 (diethylamino) ethyl] benzamide (procainamide)-tetraphenylborate complex was synthesized by reacting sodium tetraphenyl borate with 4-amino-N-[2 (diethylamino) ethyl] benzamide, chloride salt, and procainamide in deionized water at room temperature through an ion-associate reaction (green chemistry) at room temperature, and characterized by several physicochemical methods. The formation of ion-associate complex between bio-active molecules and/or organic molecules is crucial to comprehending the relationships between bioactive molecules and receptor interactions. The solid complex was characterized by infrared spectra, NMR, elemental analysis, and mass spectrometry, indicating the formation of ion-associate or ion-pair complex. The complex under study was examined for antibacterial activity. The ground state electronic characteristics of the S1 and S2 complex configurations were computed using the density functional theory (DFT) approach, using B3LYP level 6-311 G(d,p) basis sets. R2 = 0.9765 and 0.9556, respectively, indicate a strong correlation between the observed and theoretical 1H-NMR, and the relative error of vibrational frequencies for both configurations was acceptable, as well. HOMO and LUMO frontier molecular orbitals and molecular electrostatics using the optimized were used to obtain a potential map of the chemical. The n → π* UV absorption peak of the UV cutoff edge was detected for both configurations of the complex. Spectroscopic methods were structures used to characterize the structure (FT-IR and 1HNMR). In the ground state, DFT/B3LYP/6-311G(d,p) basis sets were used to determine the electrical and geometric properties of the S1 and S2 configurations of the title complex. Comparing the observed and calculated values for the S1 and S2 forms, the HOMO-LUMO energy gap of compounds was 3182 and 3231 eV, respectively. The small energy gap between HOMO and LUMO indicated that the compound was stable. In addition, the MEP reveals that positive potential sites were around the PR molecule, whereas negative potential sites were surrounding the TPB site of atoms. The UV absorption of both arrangements is comparable to the experimental UV spectrum.
More Related Videos
 07:20
Amide Coupling Reaction for the Synthesis of Bispyridine-based Ligands and Their Complexation to Platinum as Dinuclear Anticancer Agents
Published on: May 28, 2014
 08:56
Synthesis of 1,2-Azaborines and the Preparation of Their Protein Complexes with T4 Lysozyme Mutants
Published on: March 25, 2017
Related Concept Videos
 01:20
Electrophilic Aromatic Substitution: Nitration of Benzene
 01:19
Diazonium Group Substitution: –OH and –H
 01:19
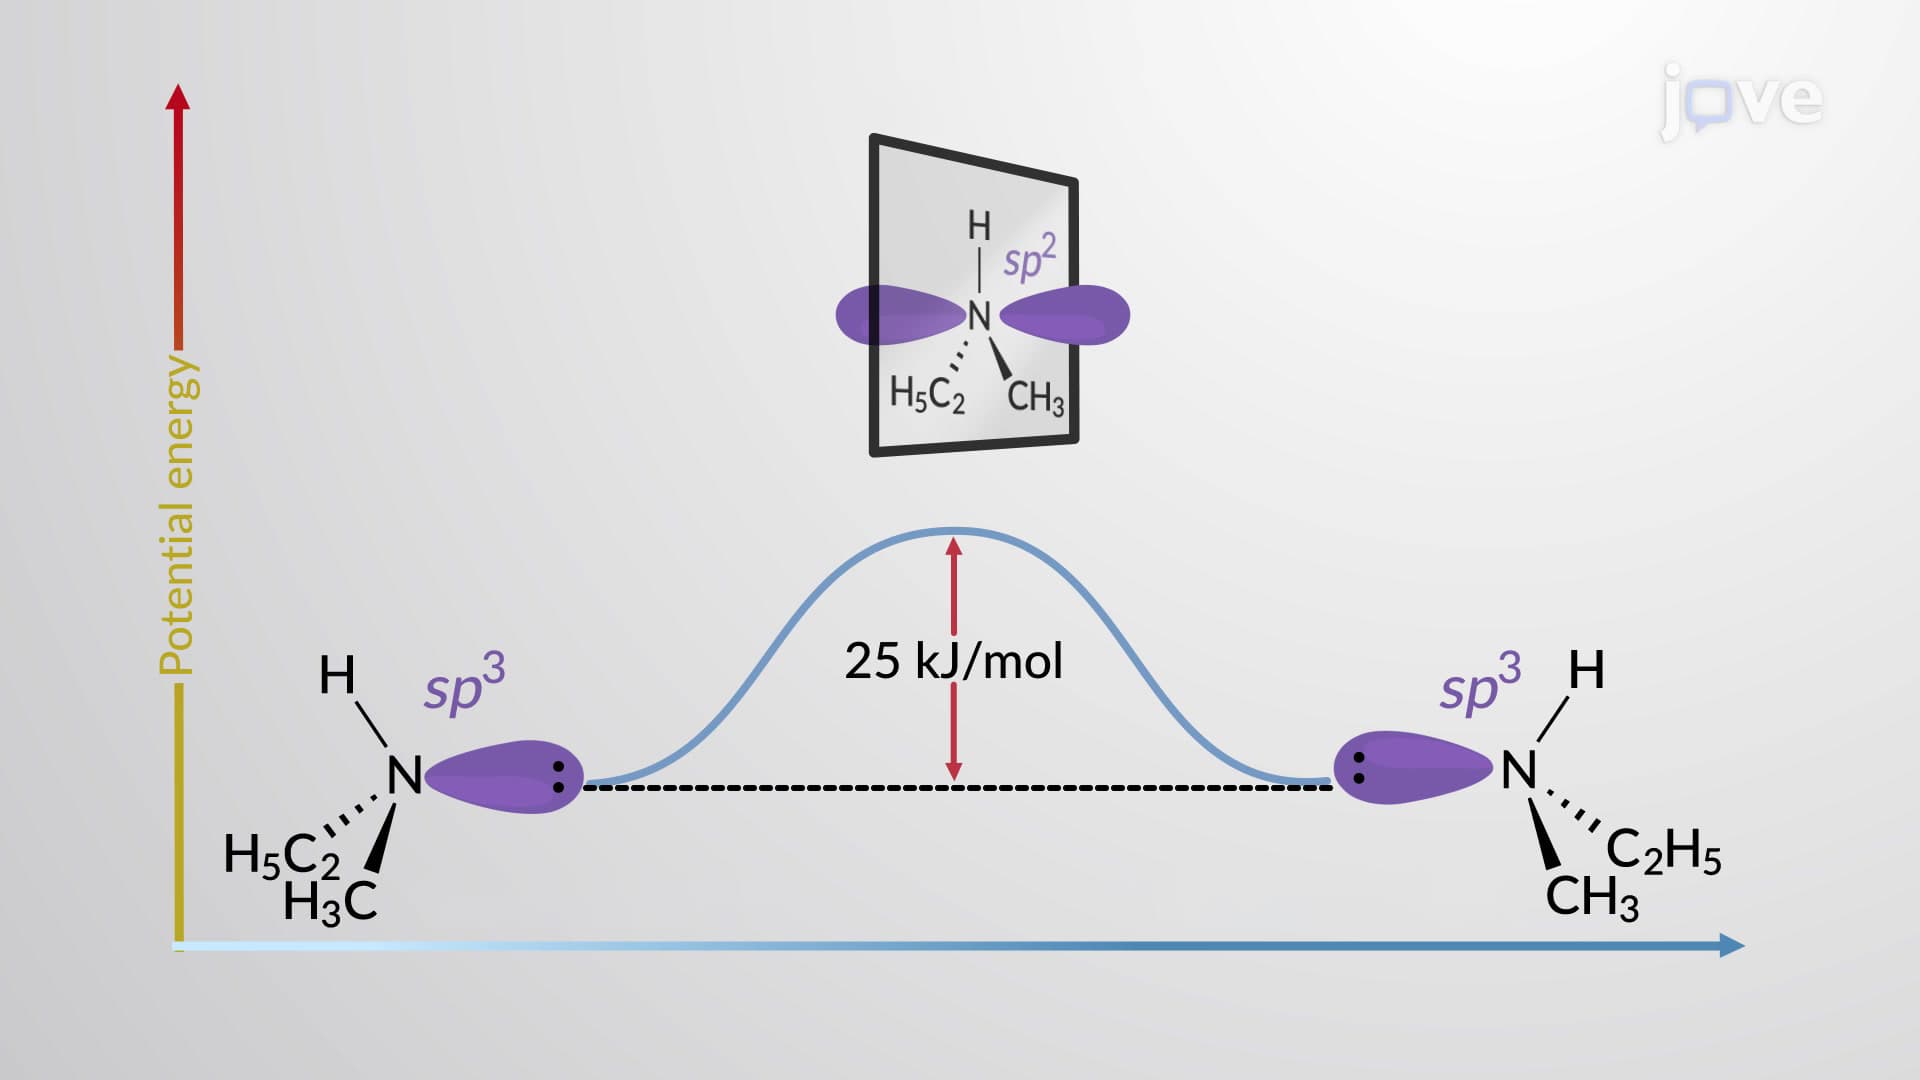
Structure of Amines
 01:25
Basicity of Heterocyclic Aromatic Amines
 02:42
Valence Bond Theory
 01:26
Physical Properties of Amines