Related Experiment Video
Updated: Aug 6, 2025

Screening and Identification of Small Peptides Targeting Fibroblast Growth Factor Receptor2 using a Phage Display Peptide Library
Published on: September 30, 2019
Discovery of Orally Bioavailable FGFR2/FGFR3 Dual Inhibitors via Structure-Guided Scaffold Repurposing Approach
Minh H Nguyen1, Hai-Fen Ye1, Yao Xu1
1Incyte Research Institute, Incyte Corporation, 1801 Augustine Cut-Off, Wilmington, Delaware 19803, United States.

Abstract:
Fibroblast growth factor receptors (FGFRs) are transmembrane receptor tyrosine kinases that regulate multiple physiological processes. Aberrant activation of FGFR2 and FGFR3 has been linked to the pathogenesis of many tumor types, including cholangiocarcinoma and bladder cancer. Current therapies targeting the FGFR2/3 pathway exploiting small-molecule kinase inhibitors are associated with adverse events due to undesirable inhibition of FGFR1 and FGFR4. Isoform-specific FGFR2 and FGFR3 inhibitors that spare FGFR1 and FGFR4 could offer a favorable toxicity profile and improved therapeutic window to current treatments. Herein we disclose the discovery of dual FGFR2/FGFR3 inhibitors exploiting scaffold repurposing of a previously reported ALK2 tool compound. Structure-based drug design and structure-activity relationship studies were employed to identify selective and orally bioavailable inhibitors with equipotent activity toward wild-type kinases and a clinically observed gatekeeper mutant.
Insights
New dual Fibroblast Growth Factor Receptor (FGFR) 2/3 inhibitors were discovered. These selective inhibitors offer a potentially improved safety profile by sparing FGFR1 and FGFR4, crucial for cancer therapy.
Area of Science:
- Oncology
- Pharmacology
- Molecular Biology
Background:
- Fibroblast Growth Factor Receptors (FGFRs) are key regulators of cellular processes.
- Aberrant FGFR2 and FGFR3 signaling drives cancers like cholangiocarcinoma and bladder cancer.
- Current FGFR inhibitors cause adverse events by inhibiting FGFR1 and FGFR4.
Purpose of the Study:
- To discover selective FGFR2/FGFR3 inhibitors with reduced off-target activity.
- To develop novel cancer therapeutics with improved safety profiles.
- To identify orally bioavailable inhibitors targeting wild-type and mutant FGFRs.
Main Methods:
- Scaffold repurposing of an ALK2 inhibitor.
- Structure-based drug design.
- Structure-activity relationship (SAR) studies.
Main Results:
- Discovery of potent dual FGFR2/FGFR3 inhibitors.
- Achieved selectivity, sparing FGFR1 and FGFR4.
- Identified orally bioavailable compounds active against wild-type and gatekeeper mutant kinases.
Conclusions:
- Developed novel, selective FGFR2/FGFR3 inhibitors.
- These compounds show promise for improved cancer treatment with reduced toxicity.
- The findings support further development of these inhibitors for clinical application.
More Related Videos
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
 06:26
Nano-Differential Scanning Fluorimetry for Screening in Fragment-based Lead Discovery
Published on: May 16, 2021
Related Concept Videos
 01:15
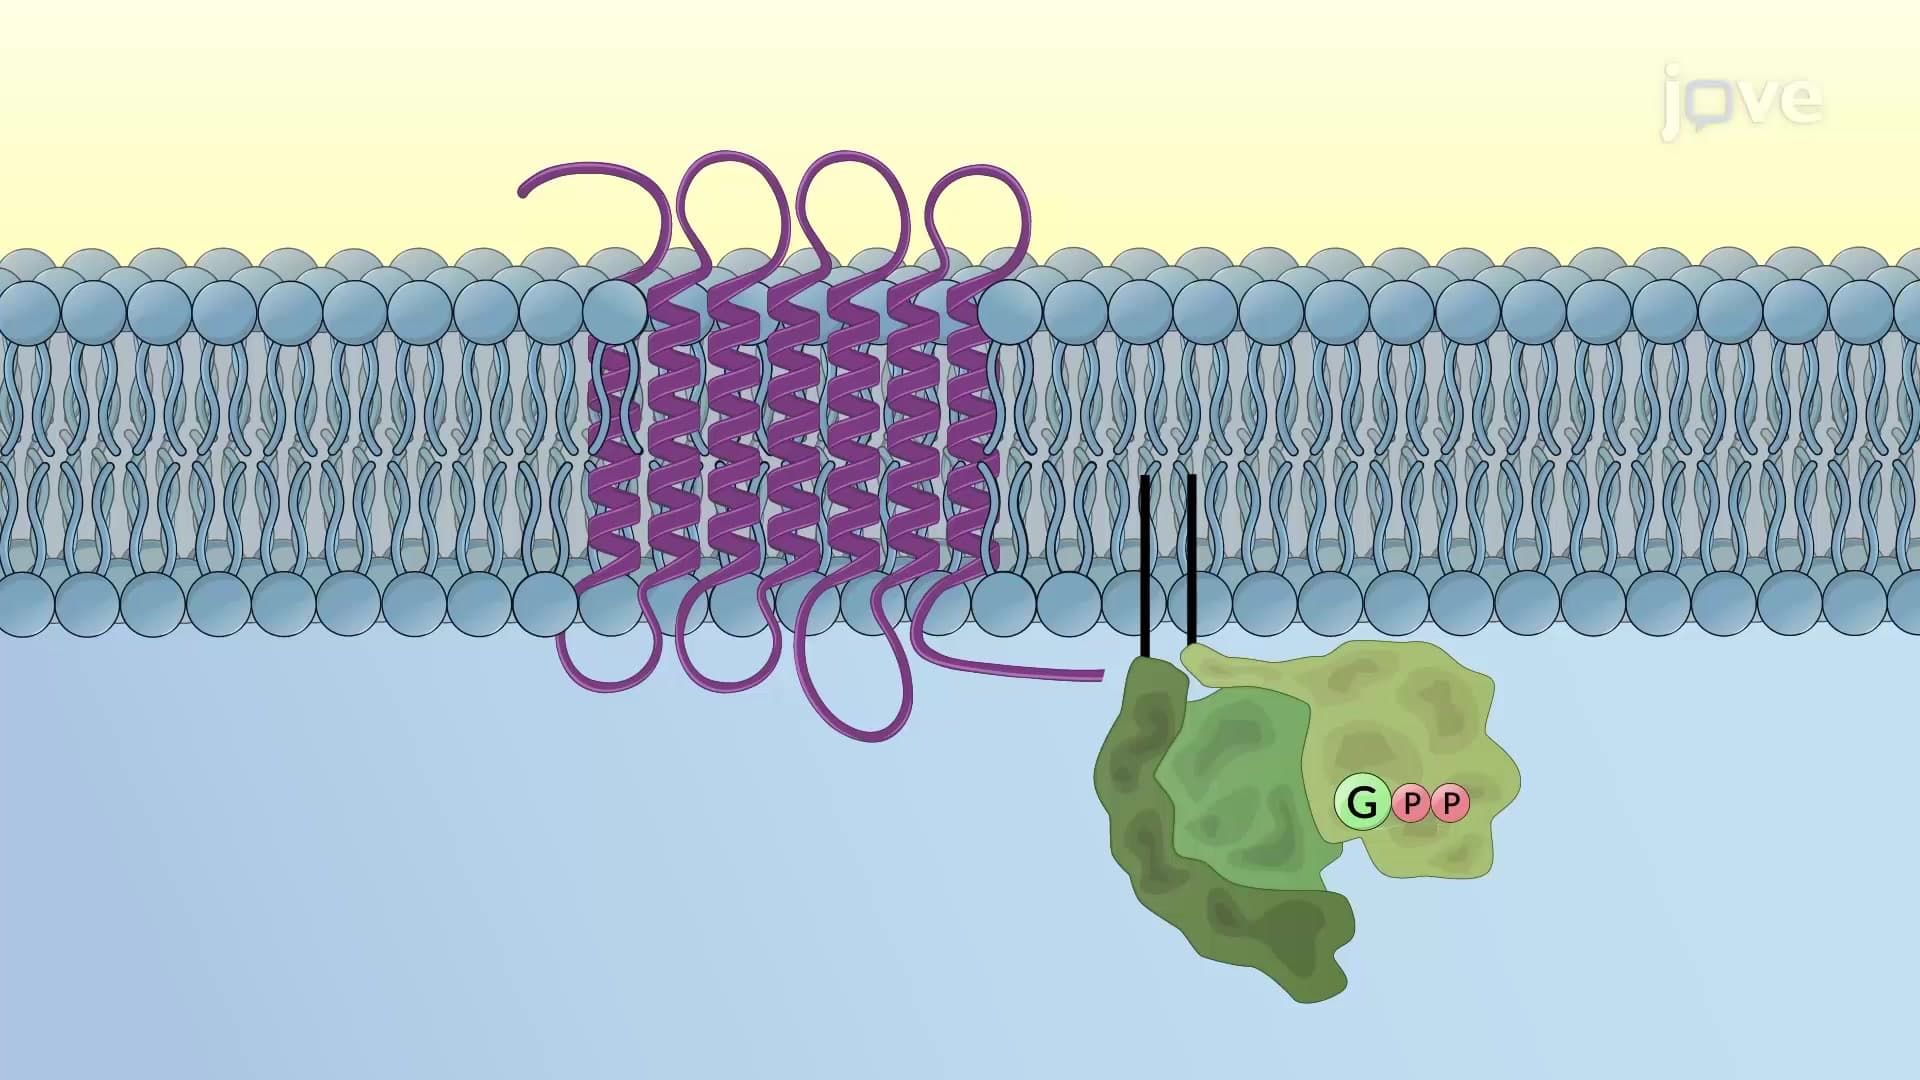
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...