Related Experiment Video
Updated: Aug 6, 2025

An In Vitro Approach to Study Mitochondrial Dysfunction: A Cybrid Model
Published on: March 9, 2022
Restoration of mitochondrial function through activation of hypomodified tRNAs with pathogenic mutations associated
Ena Tomoda1, Asuteka Nagao1, Yuki Shirai1
1Department of Chemistry and Biotechnology, Graduate School of Engineering, University of Tokyo, 7-3-1 Hongo, Bunkyo-ku, Tokyo 113-8656, Japan.

Abstract:
Mutations in mitochondrial (mt-)tRNAs frequently cause mitochondrial dysfunction. Mitochondrial myopathy, encephalopathy, lactic acidosis, and stroke-like episodes (MELAS), and myoclonus epilepsy associated with ragged red fibers (MERRF) are major clinical subgroups of mitochondrial diseases caused by pathogenic point mutations in tRNA genes encoded in mtDNA. We previously reported a severe reduction in the frequency of 5-taurinomethyluridine (τm5U) and its 2-thiouridine derivative (τm5s2U) in the anticodons of mutant mt-tRNAs isolated from the cells of patients with MELAS and MERRF, respectively. The hypomodified tRNAs fail to decode cognate codons efficiently, resulting in defective translation of respiratory chain proteins in mitochondria. To restore the mitochondrial activity of MELAS patient cells, we overexpressed MTO1, a τm5U-modifying enzyme, in patient-derived myoblasts. We used a newly developed primer extension method and showed that MTO1 overexpression almost completely restored the τm5U modification of the MELAS mutant mt-tRNALeu(UUR). An increase in mitochondrial protein synthesis and oxygen consumption rate suggested that the mitochondrial function of MELAS patient cells can be activated by restoring the τm5U of the mutant tRNA. In addition, we confirmed that MTO1 expression restored the τm5s2U of the mutant mt-tRNALys in MERRF patient cells. These findings pave the way for epitranscriptomic therapies for mitochondrial diseases.
Insights
Restoring tRNA modifications in mitochondrial diseases like MELAS and MERRF can reactivate mitochondrial function. Overexpressing the MTO1 enzyme successfully repaired mutant mt-tRNAs, improving cellular respiration and protein synthesis.
Area of Science:
- Mitochondrial biology
- Molecular genetics
- Biochemistry
Background:
- Mutations in mitochondrial transfer RNAs (mt-tRNAs) are a common cause of mitochondrial dysfunction.
- Diseases like MELAS and MERRF result from pathogenic point mutations in mtDNA-encoded tRNA genes, leading to hypomodified tRNAs and impaired mitochondrial protein synthesis.
- Previously, a reduction in 5-taurinomethyluridine (τm5U) and 2-thiouridine (τm5s2U) modifications in mutant mt-tRNAs from MELAS and MERRF patients was reported.
Purpose of the Study:
- To investigate the potential of restoring tRNA modifications to reactivate mitochondrial function in MELAS and MERRF patient cells.
- To evaluate the efficacy of MTO1 enzyme overexpression in correcting hypomodified mt-tRNAs and improving mitochondrial activity.
Main Methods:
- Overexpression of MTO1, a τm5U-modifying enzyme, in patient-derived myoblasts.
- Utilizing a novel primer extension method to assess tRNA modification levels.
- Measuring mitochondrial protein synthesis and oxygen consumption rates.
Main Results:
- MTO1 overexpression restored τm5U modification in the MELAS mutant mt-tRNALeu(UUR) to near-normal levels.
- This restoration led to increased mitochondrial protein synthesis and oxygen consumption rates in MELAS patient cells.
- MTO1 expression also successfully restored the τm5s2U modification in the MERRF mutant mt-tRNALys.
Conclusions:
- Restoring essential tRNA modifications, specifically τm5U, can reactivate mitochondrial function in cells affected by MELAS.
- The findings support the potential of MTO1 as a therapeutic target for mitochondrial diseases caused by tRNA mutations.
- This study opens avenues for developing epitranscriptomic therapies for a range of mitochondrial disorders.
Related Concept Videos
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:48
ATP Synthase: Mechanism
 01:19
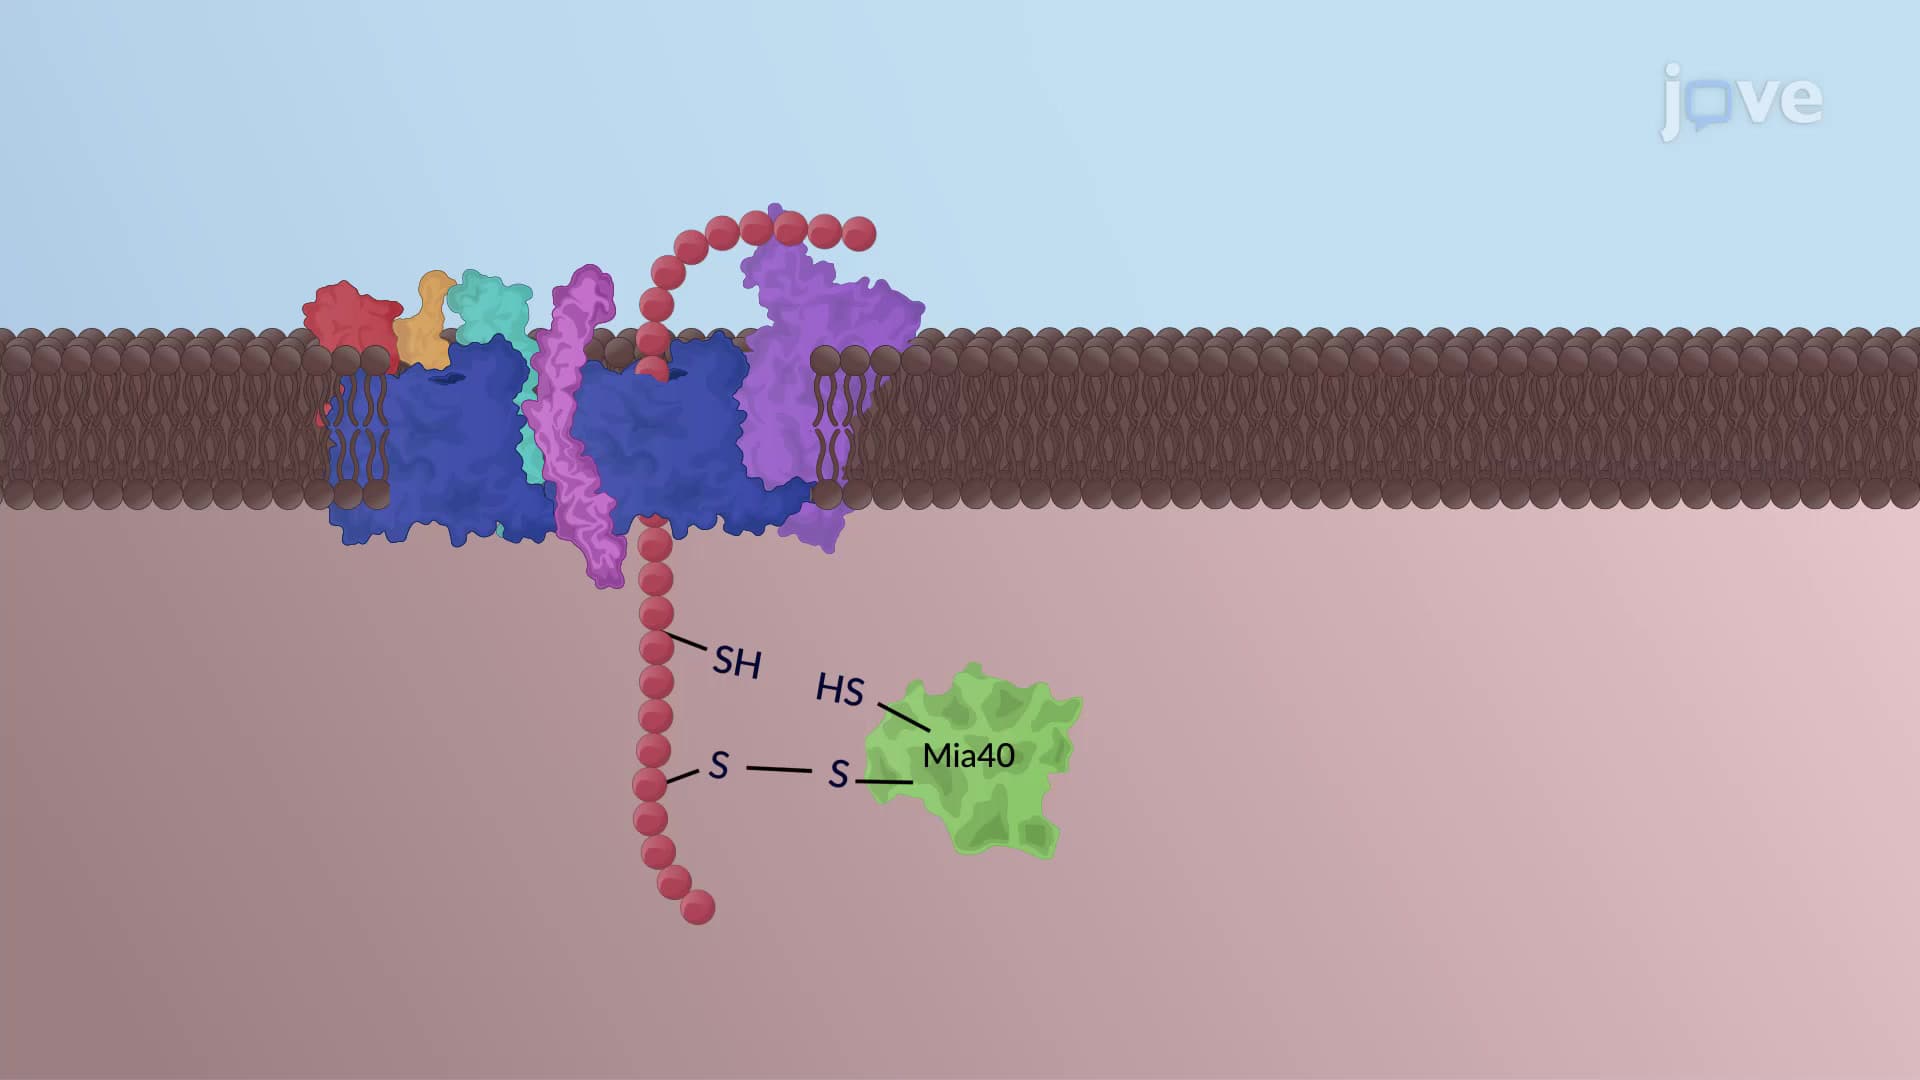
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
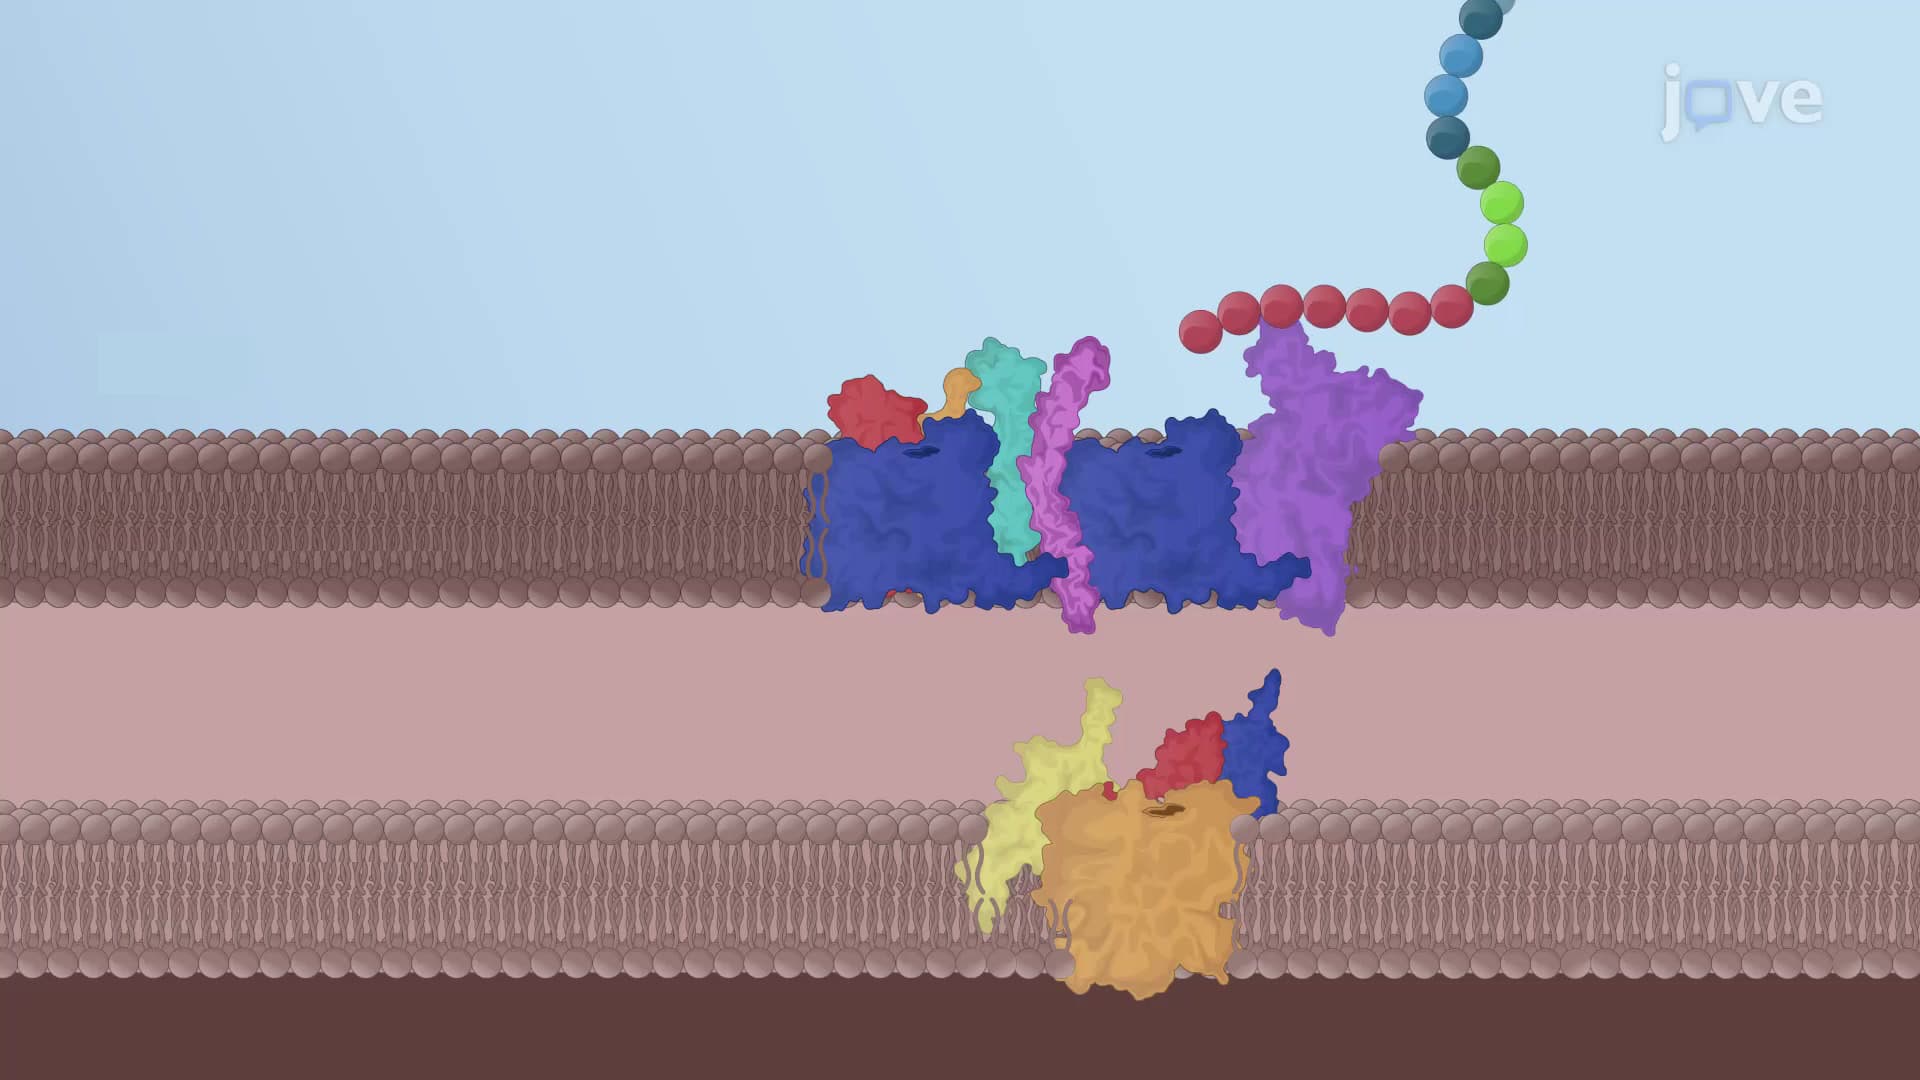
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:37
Mitochondria