Related Experiment Video
Updated: Aug 6, 2025


The Synthesis of [Sn10SiSiMe334]2- Using a Metastable SnI Halide Solution Synthesized via a Co-condensation Technique
Published on: November 28, 2016
Hollow polyhedral structures and properties of Ag2n-1Sn- (n = 2-11) clusters: A theoretical study
Zhimei Tian1, Chongfu Song2, Chang Wang1
1School of Chemistry and Materials Engineering, Fuyang Normal University, Fuyang, 236037, Anhui, China.

Context:
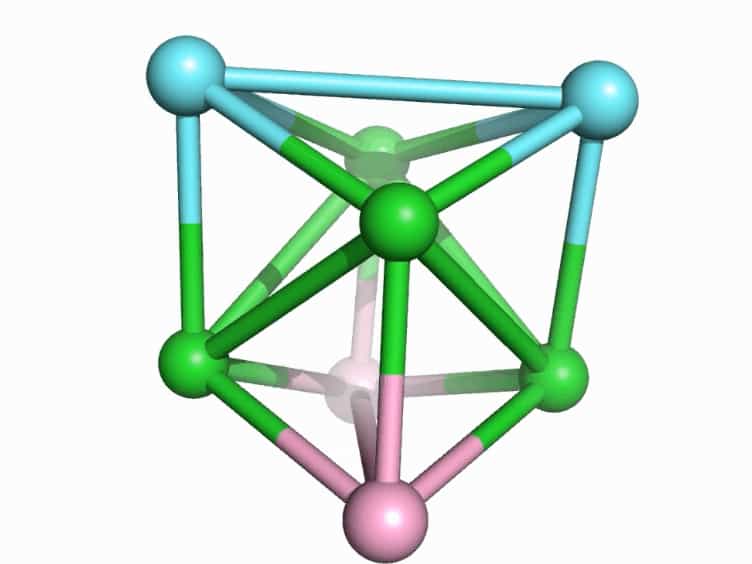
The structures of Ag2n-1Sn- (n = 2-11) clusters are obtained by the combination of genetic algorithm (GA) and density functional theory (DFT). All the global minimum structures prefer hollow polyhedral structures, in which S-Ag-S element, triangular Ag3S3 and tetragonal Ag4S4 units present to stabilize the structures. The S atoms in the structures appear in μ3-S or μ4-S form. Adiabatic and vertical electron affinities of the clusters have been obtained, which reveals that they increases as cluster size. Stability analysis shows that Ag9S5- and Ag19S10- have special stability. The HOMO, LUMO orbitals of the clusters are obtained and the orbital components of them are calculated. The HOMO orbitals are mainly from the p orbitals of S atoms, whereas the s, p and d orbitals of Ag atoms contribute much bigger than the p orbitals of S atoms for LUMO orbitals. The orbital delocalization indexes (ODI) of the HOMOs and LUMOs are calculated, and the small ODIs of the HOMOs and LUMOs for n = 4-10 reveal that these orbitals are highly delocalized. By studying the projected density of states and molecular orbitals of Ag9S5- and Ag19S10- clusters, it is found that their molecular orbitals have superatomic properties. Superatomic properties play an important role in stabilizing clusters.
Methods:
This work used combined genetic algorithm and density functional theory (GA-DFT), and PBE0/Lanl2tz(Ag)/6-311G(d,p)(S) method to optimize the structures. Gaussian 16 program, Gauss view 6.0.16 program and Multiwfn 3.8 code are the softwares used.
Related Concept Videos
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
 02:57
Coordination Number and Geometry
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:22
Structures of Solids
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...