Related Experiment Video
Updated: Aug 6, 2025

An Electrochemical Cholesteric Liquid Crystalline Device for Quick and Low-Voltage Color Modulation
Published on: February 27, 2019
Computational study of physicochemical, optical, and thermodynamic properties of 2,2-dimethylchromene derivatives
J R Eone1, Y Tadjouteu Assatse1, R A Yossa Kamsi1
1Faculty of Science, Department of Physics, University of Yaoundé I, P.O. Box 812, Yaoundé, Cameroon.

Context:
A large number of heterocyclic compounds are used as drugs, mainly due to the duality of lipophilicity playing in hydrophobic interactions and solubility with at least one hydrogen bond acceptor. The study of electronic properties is then important to better understand not only these charge distribution effects but also some other physicochemical properties involved in bioactivity to directly assess the bioavailability of these compounds and a possible classification in related applications. Phytomolecules such as chromenes are very accessible molecules exhibiting a bioactivity. Our study is focused on the impact of a number of functional groups acting on some 2,2-dimethylchromene derivatives, namely their global reactivity from the frontier molecular orbitals and local reactivity from the Fukui functions, where the carbonyl group acting as an electron withdrawal group has the most relevant effect, the solubility from the partition coefficient Log P strongly depending on the charge distribution and electronegative sites, the optical effects from the delocalization in the vinyl group, as well as the evaluation of the entropy associated with the molecular flexibility also acting on the bioactivity. Despite the effects of the wave function or density methods on the order of magnitude of these properties, these compounds are consistent with the rules for a potential oral drug candidate.
Methods:
The calculations of the electronic properties were performed through two levels of theory: Hartree-Fock level as a wave function-based method as an ab initio reference including some physically consistent eigenvalues and density functional theory DFT as a correlation consistent method using different functionals: hybrid or with a long-range correction. The basis set used is a 6-311++G(d,p) Pople basis set including diffuse and polarization basis functions. The basis set is adapted to the size of the molecules and consequently to the degree of electronic localization. Gaussian 09 software was used for the computation.
More Related Videos
 06:24
High-Contrast and Fast Photorheological Switching of a Twist-Bend Nematic Liquid Crystal
Published on: October 31, 2019
 12:07
Microwave-assisted Intramolecular Dehydrogenative Diels-Alder Reactions for the Synthesis of Functionalized Naphthalenes/Solvatochromic Dyes
Published on: April 1, 2013
Related Concept Videos
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:33
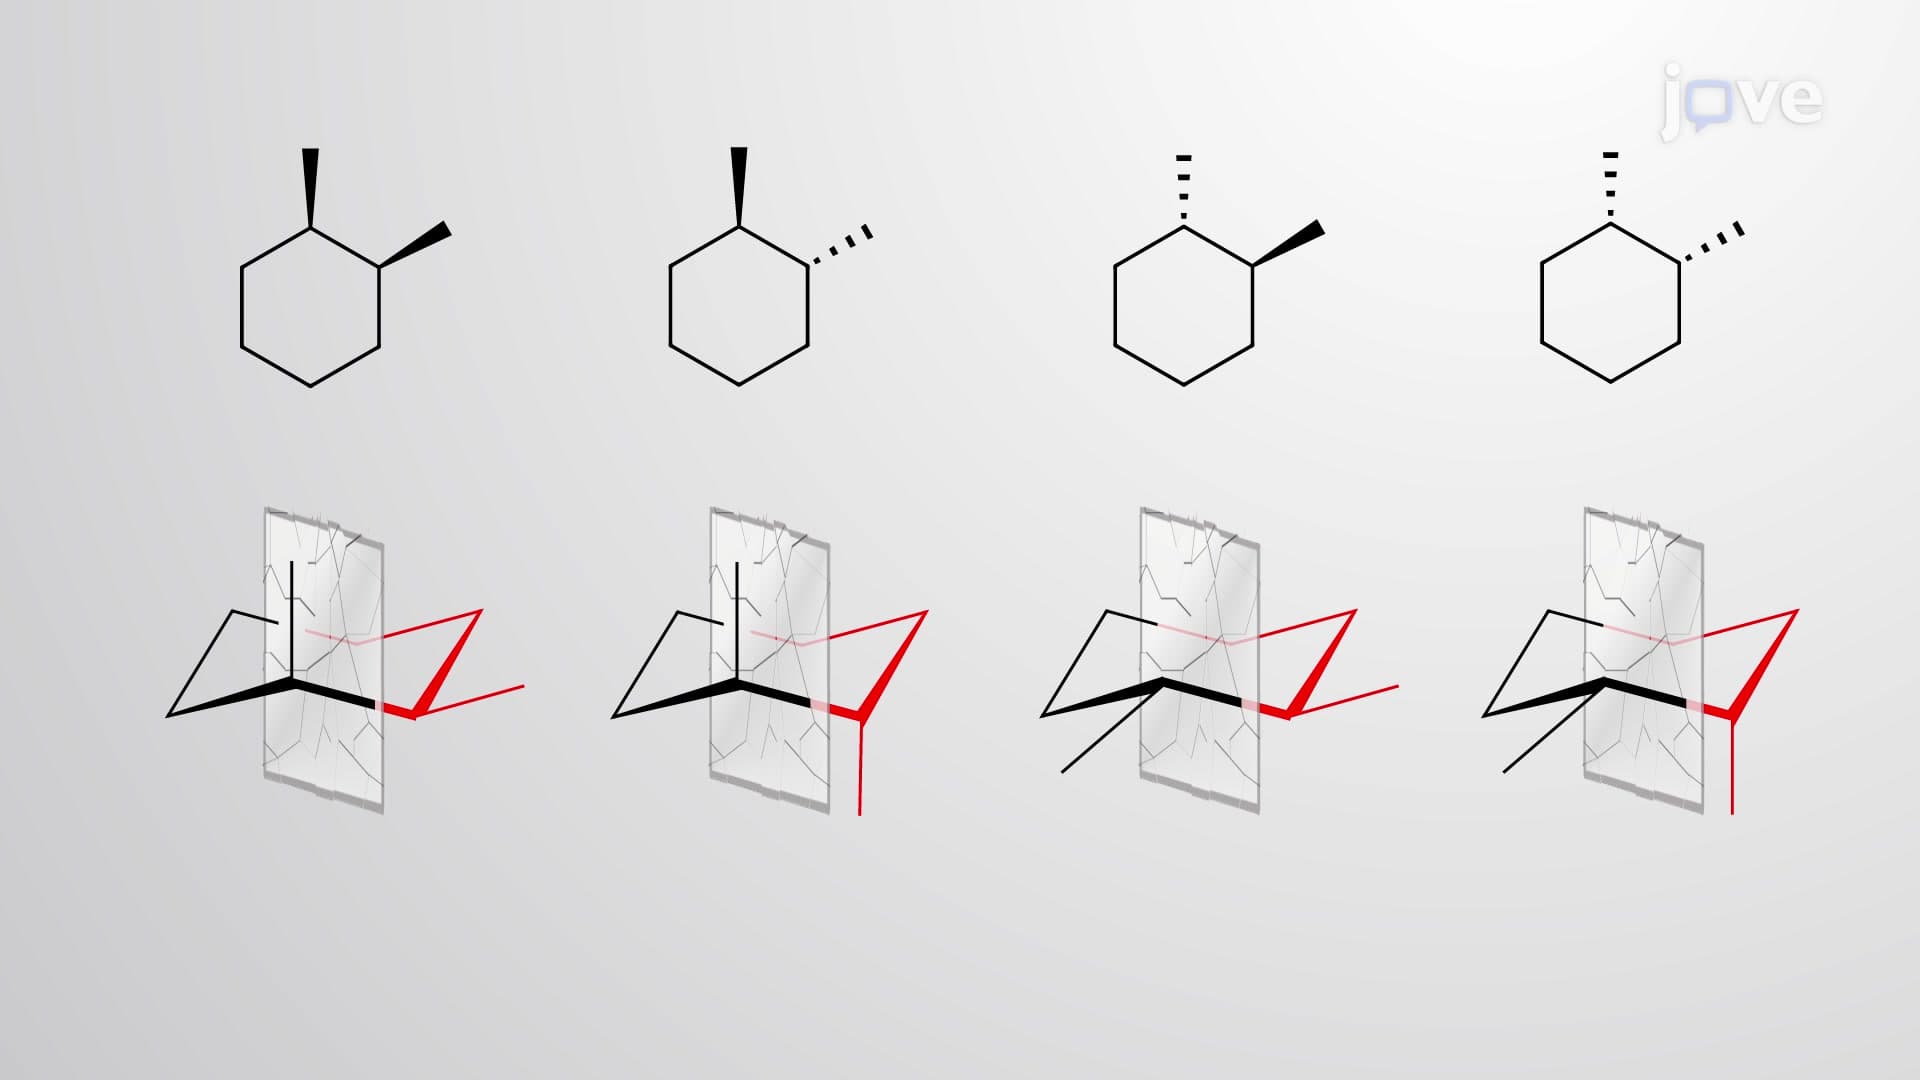
Stereoisomerism of Cyclic Compounds
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:24
Properties of Enantiomers and Optical Activity
 01:15
Optimizing Chromatographic Separations
Band broadening refers to spreading solute bands as they travel through the column. This broadening can impact resolution. Plate height (H) represents the length required for one theoretical plate. A lower plate height corresponds to...