Related Experiment Video
Updated: Aug 5, 2025
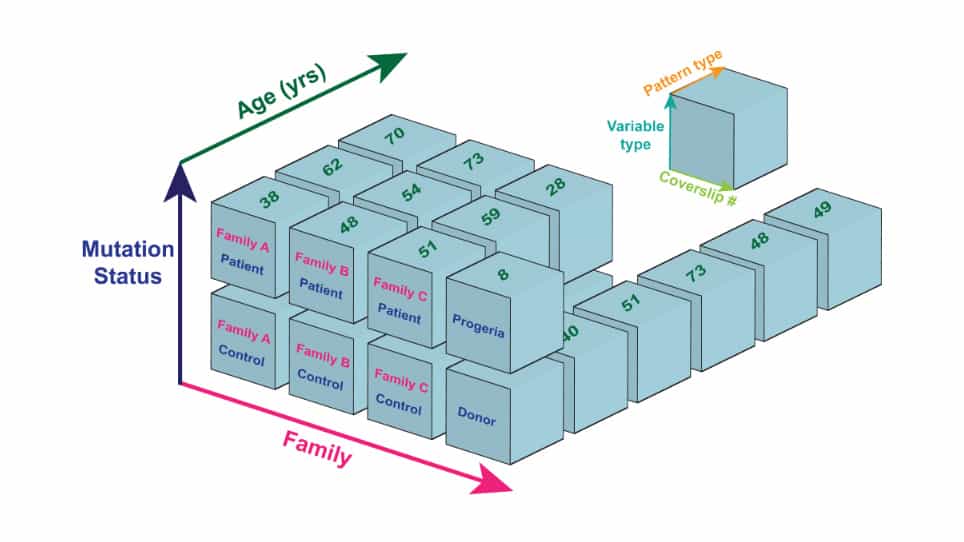

Databases to Efficiently Manage Medium Sized, Low Velocity, Multidimensional Data in Tissue Engineering
Published on: November 22, 2019
The Molecular and Cellular Basis of Hutchinson-Gilford Progeria Syndrome and Potential Treatments
Noelle J Batista1, Sanket G Desai1, Alexis M Perez1
1Department of Biomedical Sciences, College of Osteopathic Medicine, New York Institute of Technology, Old Westbury, NY 11568, USA.

Abstract:
Hutchinson-Gilford progeria syndrome (HGPS) is a rare, autosomal-dominant, and fatal premature aging syndrome. HGPS is most often derived from a de novo point mutation in the LMNA gene, which results in an alternative splicing defect and the generation of the mutant protein, progerin. Progerin behaves in a dominant-negative fashion, leading to a variety of cellular and molecular changes, including nuclear abnormalities, defective DNA damage response (DDR) and DNA repair, and accelerated telomere attrition. Intriguingly, many of the manifestations of the HGPS cells are shared with normal aging cells. However, at a clinical level, HGPS does not fully match normal aging because of the accelerated nature of the phenotypes and its primary effects on connective tissues. Furthermore, the epigenetic changes in HGPS patients are of great interest and may play a crucial role in the pathogenesis of HGPS. Finally, various treatments for the HGPS patients have been developed in recent years with important effects at a cellular level, which translate to symptomatic improvement and increased lifespan.
Related Concept Videos
 02:18
Alternative RNA Splicing
There are five types of alternative RNA splicing that vary in the ways the pre-mRNA segments are removed or retained in the mature mRNA. The first...
 01:22
iPS Cell Differentiation
 01:21
Satellite Stem Cells and Muscular Dystrophy